Related Experiment Video
Updated: Aug 20, 2026
Construction and Systematical Symmetric Studies of a Series of Supramolecular Clusters with Binary or Ternary Ammonium Triphenylacetates
Published on: February 15, 2016
A density-functional theory investigation of 3-nitro-1,2,4-triazole-5-one dimers and crystal
He-Ming Xiao1, Xue-Hai Ju, Li-Na Xu
1Department of Chemistry, Nanjing University of Science and Technology, Nanjing 210094, People's Republic of China. xiao@mail.njust.edu.cn

Abstract:
Density-functional method with different basis sets was applied to the study of the highly efficient and low sensitive explosive 3-nitro-1,2,4-triazole-5-one (NTO) in both gaseous dimer and its bulk state. The binding energies have been corrected for the basis set superposition errors. Six stable dimers (II-VII) were located. The corrected binding energy of the most stable dimer VII is predicted to be -53.66 kJ/mol at the B3LYP/6-311++G(**) level. It was found that the structures of the more stable dimers (V-VII) are through the hydrogen bonding interaction between the carbonyl oxygen and the azole hydrogen of 3-nitro-1,2,4-triazole-5-one. The changes of Gibbs free energies (DeltaG) in the processes from the monomer to the dimers at 298.15 K are 8.51, 0.90, 0.35, -8.74, -10.67, and -11.06 kJ/mol for dimers from II to VII, respectively. Dimers V-VII, possessing cyclic structures, can be spontaneously produced from the isolated monomer at room temperature. The lattice energy is -156.14 kJ/mol, and this value becomes to -150.43 kJ/mol when a 50% correction of the basis set superposition error was adopted. The frontier bands are quite flat. Judged from the value of band gap of 4.0 eV, it may be predicted that 3-nitro-1,2,4-triazole-5-one is an insulator. Most atoms in NTO, with the exception of C(5) atom and the nitro atoms, make up the upper valence bands. In contrast, the lower conduction bands mainly consist of the nitro N and O atoms. The population of the C-NO(2) bond is much less than those of the other bonds and the detonation may be initiated by the breakdown of this bond.
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:19
Crystal Density
 03:24
Hybridization of Atomic Orbitals I
 01:19
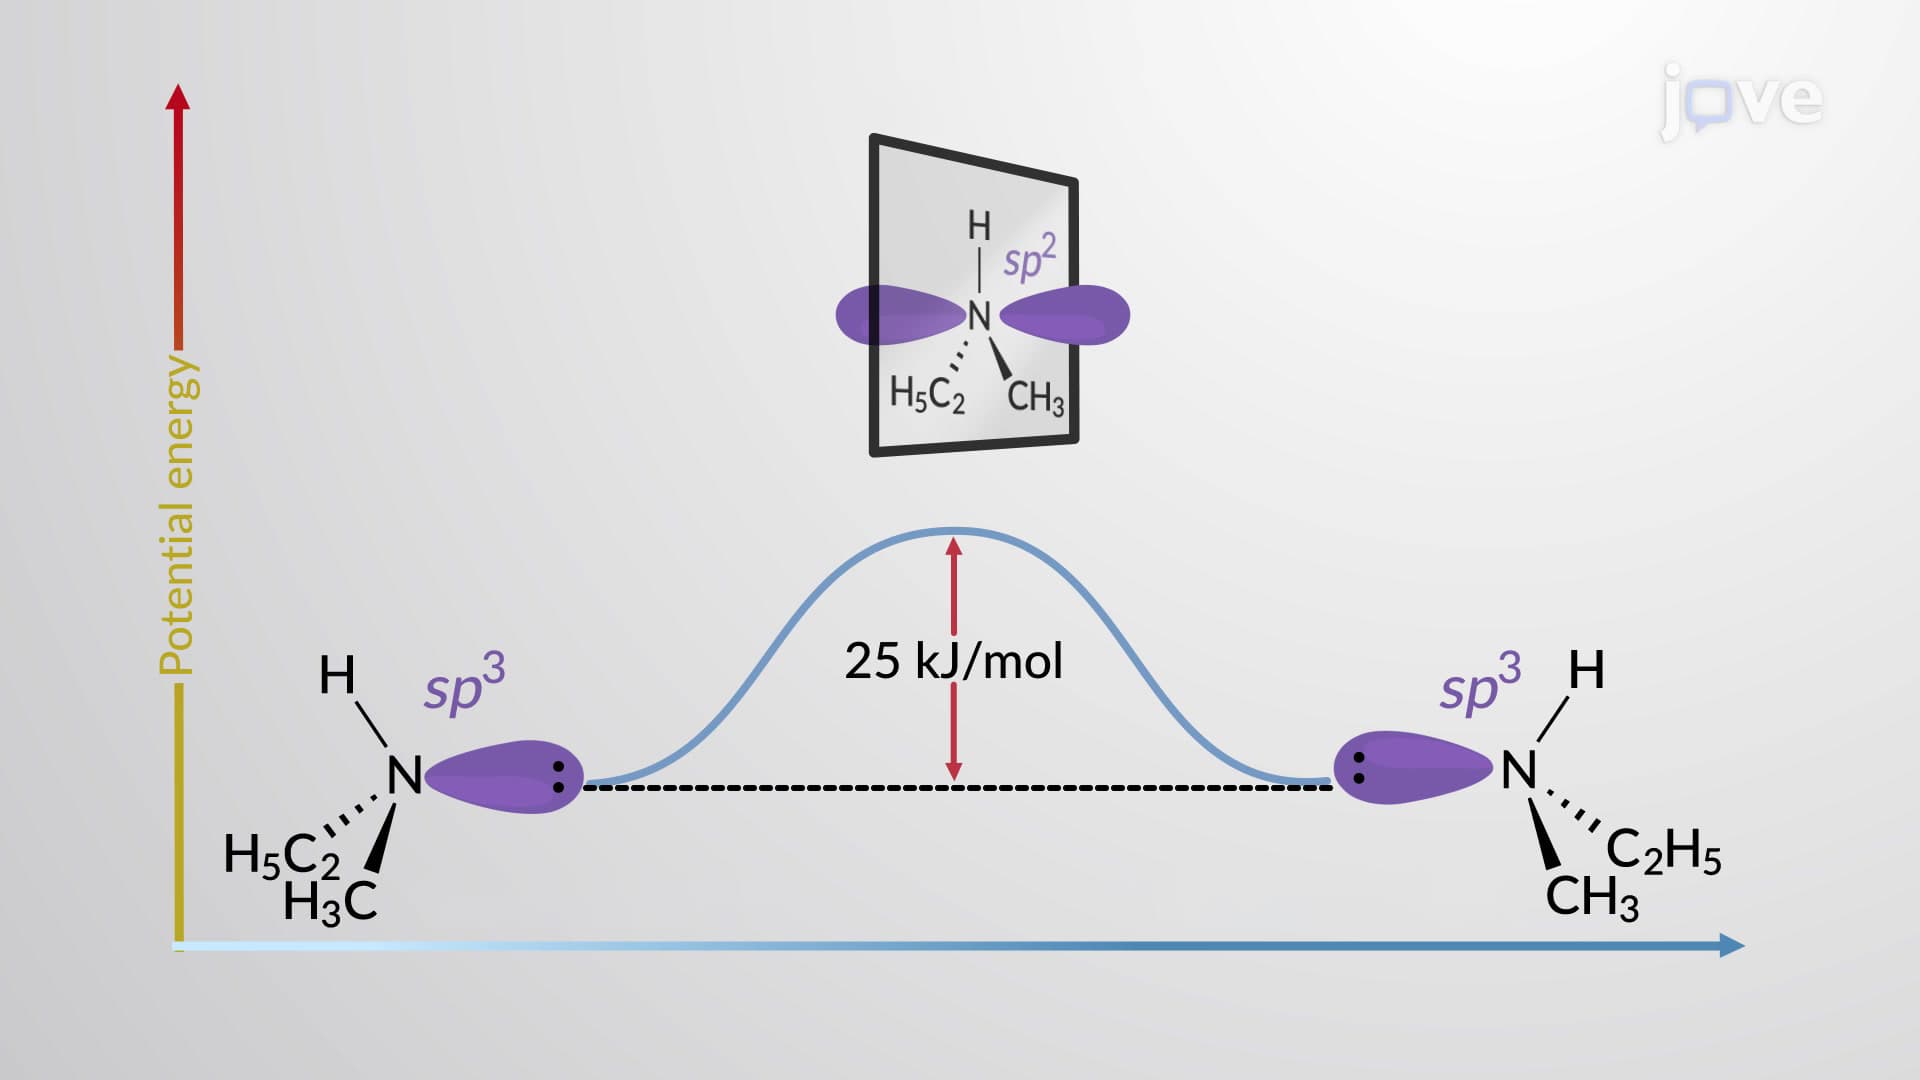
Structure of Amines
 02:42
Valence Bond Theory