Predicting binding sites of hydrolase-inhibitor complexes by combining several methods
Related Experiment Video
Updated: Aug 20, 2026
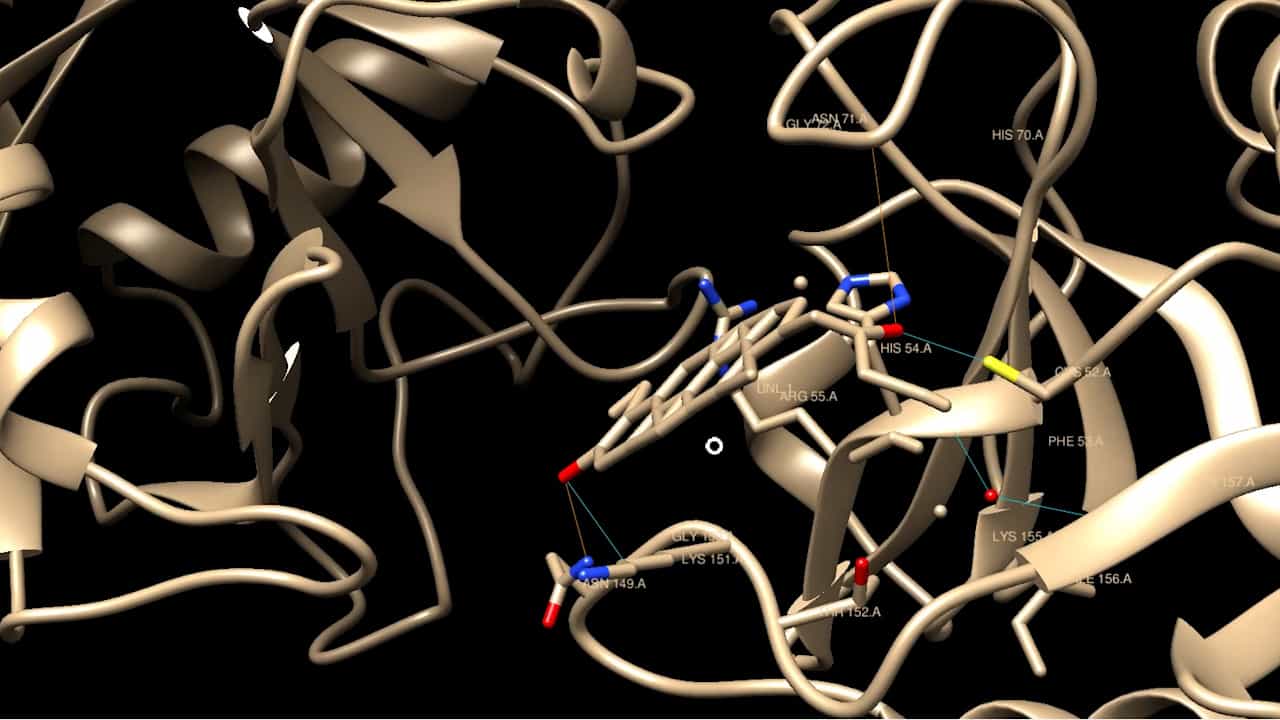
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Taner Z Sen1, Andrzej Kloczkowski, Robert L Jernigan
1L.H. Baker Center for Bioinformatics and Biological Statistics, Iowa State University, Ames, IA 50011, USA. taner@iastate.edu

Background:
Protein-protein interactions play a critical role in protein function. Completion of many genomes is being followed rapidly by major efforts to identify interacting protein pairs experimentally in order to decipher the networks of interacting, coordinated-in-action proteins. Identification of protein-protein interaction sites and detection of specific amino acids that contribute to the specificity and the strength of protein interactions is an important problem with broad applications ranging from rational drug design to the analysis of metabolic and signal transduction networks.
Results:
In order to increase the power of predictive methods for protein-protein interaction sites, we have developed a consensus methodology for combining four different methods. These approaches include: data mining using Support Vector Machines, threading through protein structures, prediction of conserved residues on the protein surface by analysis of phylogenetic trees, and the Conservatism of Conservatism method of Mirny and Shakhnovich. Results obtained on a dataset of hydrolase-inhibitor complexes demonstrate that the combination of all four methods yield improved predictions over the individual methods.
Conclusions:
We developed a consensus method for predicting protein-protein interface residues by combining sequence and structure-based methods. The success of our consensus approach suggests that similar methodologies can be developed to improve prediction accuracies for other bioinformatic problems.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:04
Protein-protein Interfaces
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 00:49
Ligand Binding and Linkage
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis pathway,...