Related Experiment Video
Updated: Aug 20, 2026

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Density functional calculation of the electronic absorption spectrum of Cu+ and Ag+ aqua ions
Leonardo Bernasconi1, Jochen Blumberger, Michiel Sprik
1Department of Chemistry, University of Cambridge, Lensfield Road, Cambridge CB2 1EW, United Kingdom.

Abstract:
The UV absorption of aqueous Cu+ and Ag+ has been studied using Time Dependent Density Functional Theory (TDDFT) response techniques. The TDDFT electronic spectrum was computed from finite temperature dynamical trajectories in solution generated using the Density Functional Theory (DFT) based Ab Initio Molecular Dynamics (AIMD) method. The absorption of the two ions is shown to arise from similar excitation mechanisms, namely transitions from d orbitals localized on the metal center to a rather delocalized state originating from hybridization of the metal s orbital to the conduction band edge of the solvent. The ions differ in the way the spectral profile builds up as a consequence of solvent thermal motion. The Cu+ absorption is widely modulated, both in transition energies and intensities by fluctuations in the coordination environment which is characterized by the formation of strong coordination bonds to two water molecules in an approximately linear geometry. Though, on average, absorption intensities are typical of symmetry forbidden transitions of metal ions in the solid state, occasionally very short (<100 fs) bursts in intensity are observed, associated with anomalous Cu-H interactions. Absorption by the Ag+ complex is in comparison relatively stable in time, and can be interpreted in terms of the energy splitting of the metal 4d manifold in an average crystal field corresponding to a fourfold coordination in a distorted tetrahedral arrangement. Whereas the spectral profile of the Ag+ aqua ion is in good agreement with experiment, the overall position of the band is underestimated by 2 eV in the BLYP approximation to DFT. The discrepancy with experiment is reduced to 1.3 eV when a hybrid functional (PBE0) is used. The remaining inaccuracy of TDDFT in this situation is related to the delocalized character of the target state in d-->s transitions.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:30
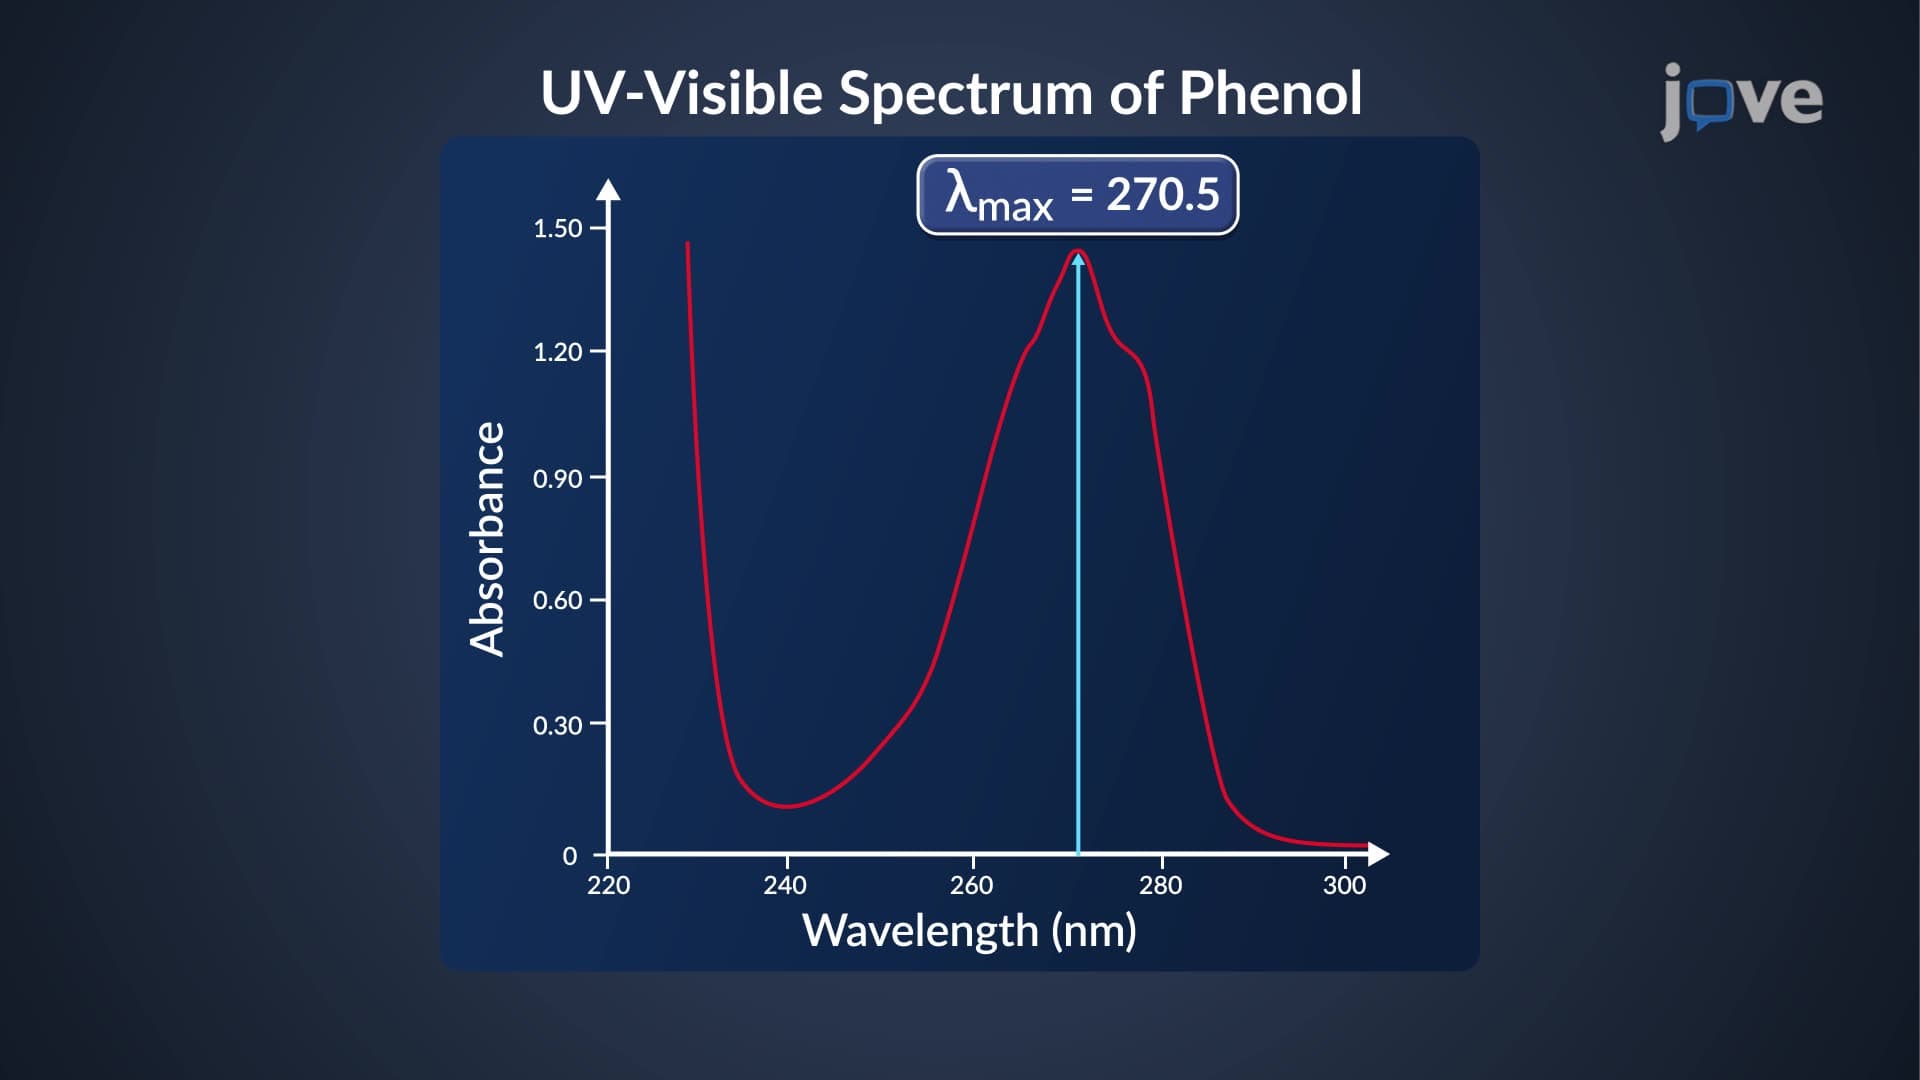
UV–Vis Spectrum
The UV–Vis spectrum of a molecule is the plot of its absorbance versus wavelength. The plot is drawn by taking molar absorptivity (ε) or log ε on the y-axis (ordinate)...
 01:28
Debye–Huckel–Onsager Conductance Equation
 01:14
UV–Vis Spectrometers
 00:56
Extraction: Advanced Methods
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions