Related Experiment Video
Updated: Aug 20, 2026

Investigating the Spreading and Toxicity of Prion-like Proteins Using the Metazoan Model Organism C. elegans
Published on: January 8, 2015
Pathogenesis of prion diseases
Ursula Unterberger1, Till Voigtländer, Herbert Budka
1Institute of Neurology, Medical University of Vienna, AKH, Austria.

Abstract:
Prion diseases are rare neurological disorders that may be of genetic or infectious origin, but most frequently occur sporadically in humans. Their outcome is invariably fatal. As the responsible pathogen, prions have been implicated. Prions are considered to be infectious particles that represent mainly, if not solely, an abnormal, protease-resistant isoform of a cellular protein, the prion protein or PrP(C). As in other neurodegenerative diseases, aggregates of misfolded protein conformers are deposited in the CNS of affected individuals. Pathogenesis of prion diseases comprises mainly two equally important, albeit essentially distinct, topics: first, the mode, spread, and amplification of infectivity in acquired disease, designated as peripheral pathogenesis. In this field, significant advances have implicated an essential role of lymphoid tissues for peripheral prion replication, before a likely neural spread to the CNS. The second is the central pathogenesis, dealing, in addition to spread and replication of prions within the CNS, with the mechanisms of nerve cell damage and death. Although important roles for microglial neurotoxicity, oxidative stress, and complement activation have been identified, we are far from complete understanding, and therapeutic applications in prion diseases still need to be developed.
Insights
Prion diseases are fatal neurodegenerative disorders caused by misfolded prion proteins. Research is advancing understanding of prion spread and nerve cell damage, but therapies are still needed.
Area of Science:
- Neurology
- Neuroscience
- Pathology
Background:
- Prion diseases are rare, fatal neurological disorders.
- They can be genetic, infectious, or sporadic.
- Prions are abnormal, protease-resistant prion proteins (PrP(C)) that aggregate in the central nervous system (CNS).
Purpose of the Study:
- To review the pathogenesis of prion diseases.
- To discuss peripheral and central mechanisms of prion disease.
- To highlight the need for therapeutic development.
Main Methods:
- Literature review of prion disease pathogenesis.
- Analysis of prion replication and spread.
- Examination of mechanisms of neurodegeneration in prion diseases.
Main Results:
- Prion diseases involve aggregates of misfolded prion proteins in the CNS.
- Peripheral pathogenesis involves prion replication in lymphoid tissues before CNS spread.
- Central pathogenesis includes prion spread within the CNS and mechanisms of nerve cell damage.
Conclusions:
- Understanding prion disease pathogenesis requires studying both peripheral and central mechanisms.
- Lymphoid tissues play a key role in peripheral prion replication.
- While microglial neurotoxicity, oxidative stress, and complement activation are implicated, further research is needed to develop effective therapies for prion diseases.
Related Concept Videos
 01:29
Subviral Agents
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
03:03Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:24
Parkinson Disease ll: Pathophysiology
 01:26
Encephalitis ll: Pathophysiology
 01:23
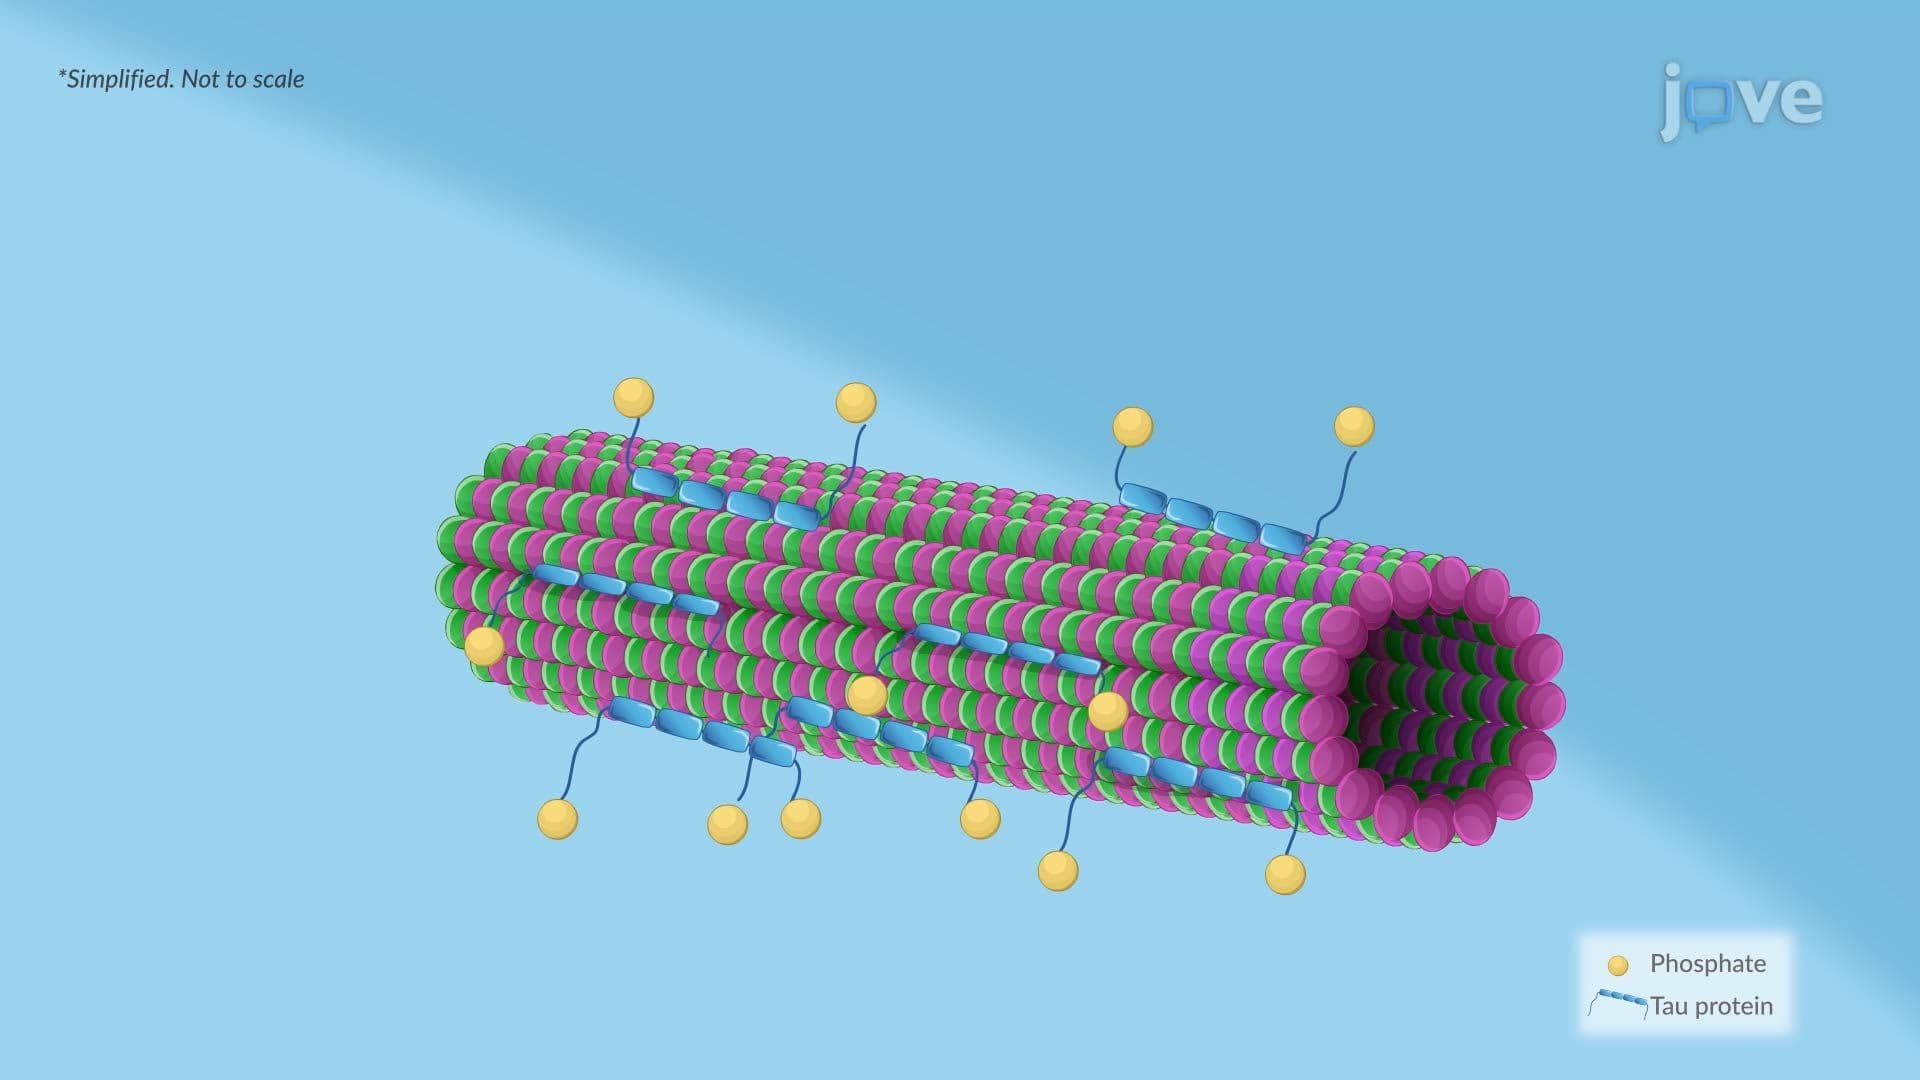
Alzheimer Disease ll: Pathophysiology