Related Experiment Video
Updated: Aug 12, 2026
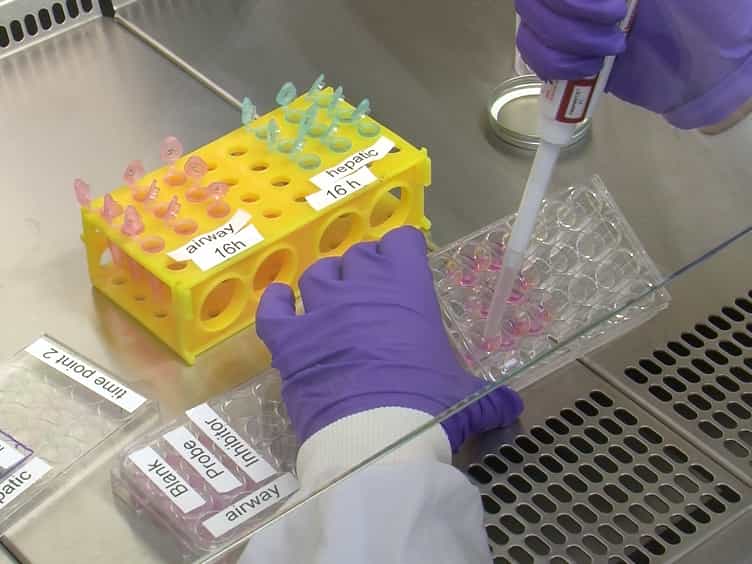

Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
Inhibition of human and rat CYP1A2 by TCDD and dioxin-like chemicals
D F Staskal1, J J Diliberto, M J Devito
1UNC Curriculum in Toxicology, US EPA, MD B143-01, 109 TW Alexander Dr., Research Triangle Park, North Carolina 27711, USA. staskal.daniel@epa.gov

Abstract:
Dioxins have been shown to bind and induce rodent CYP1A2, producing a dose-dependent hepatic sequestration in vivo. The induction of CYP1A2 activity has been used as a noninvasive biomarker for human exposure to dioxins; while there is a consistent relationship between exposure and hepatic CYP1A2 induction in rodents, this relationship has only been observed in some of the highest exposed human populations. This may be explained by inhibition of CYP1A2 activity by dioxins as some rodent studies demonstrate that rodent CYP1A2 activity can in fact be inhibited by dioxins in vitro. CYP1A2 activity was examined using a series of dioxins to inhibit human and rat CYP1A2 activity in species-specific CYP1A2 SUPERSOMES using three common CYP1A2 substrates. Methoxyresorufin was a more efficient substrate than acetanalide or caffeine in this in vitro system. Rat and human CYP1A2 enzymatic activity is inhibited by TCDD, PCDD, TCDF, 4-PeCDF, and PCBs 126, 169, 105, 118, and 156 in a concentration-dependent manner. These data demonstrate that the in vitro metabolism of prototype substrates is similar between the rat and human CYP1A2 SUPERSOME preparations and that dioxins inhibit CYP1A2 activity in both species. Because of the potential for inhibition of CYP1A2 activity by TCDD and other dioxins, studies examining CYP1A2 induction in dioxin-exposed populations using these substrates should be viewed cautiously.
Related Concept Videos
 02:34
Inhibition of Cdk Activity
 01:25
Pharmacokinetics: Drug–Drug Interactions
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:24
Drug Toxicity: Dose-Dependent Reactions
 01:16
Drug toxicity: Idiosyncratic Reactions
 01:23
Toxicity Testing in Animals