Related Experiment Video
Updated: Aug 20, 2026
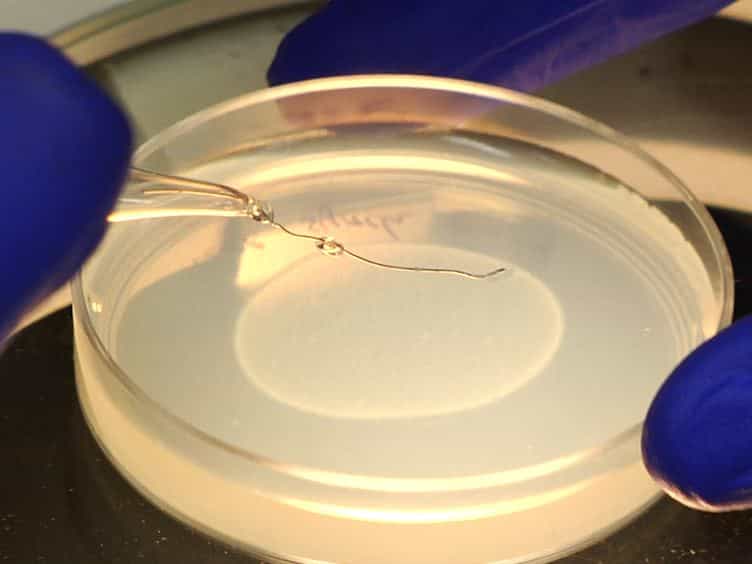
Investigating the Spreading and Toxicity of Prion-like Proteins Using the Metazoan Model Organism C. elegans
Published on: January 8, 2015
Discovering the mechanisms of neurodegeneration in prion diseases
1Department of Pathology, Institute for Neurodegenerative Diseases, University of California, San Francisco, 94143-0511, USA. sdearmo@itsa.ucsf.edu

Abstract:
The purpose of my chapter in this issue of Neuroscience Reviews dedicated to Dr. Lawrence Eng is to summarize my contributions to understanding the mechanisms of neurodegeneration in prion diseases. I explain that I was able to advance the field of prion disease neuropathology largely because of the foundation of neurochemistry and immunohistochemistry that I learned while working 5 years in Dr. Eng's laboratory. In my review, I relate how my Neuropathology Research Laboratory began as a collaboration with Dr. Stanley Prusiner 20 years ago that led from immunohistochemical staining of amyloid plaques in rodent and human brains using prion protein-specific antibodies to molecular evidence that the abnormal prion protein, PrP(Sc), is the cause of the clinically relevant neuropathological changes in animal and human prion diseases.
Related Concept Videos
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:24
Parkinson Disease ll: Pathophysiology
 01:23
Alzheimer Disease ll: Pathophysiology
 01:15
Parkinson's Disease: Overview
 01:29
Subviral Agents