Related Experiment Video
Updated: Aug 19, 2026

A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Improved greedy algorithm for protein structure reconstruction
Pierre Tuffery1, Frédéric Guyon, Philippe Derreumaux
1Equipe de Bioinformatique Génomique et Moléculaire, INSERM E0346, Université Paris 7, Tour 53-54, 2 place Jussieu, 75251 Paris Cedex 05, France. tuffery@ebgm.jussieu.fr

Abstract:
This article concerns the development of an improved greedy algorithm for protein structure reconstruction. Our stochastic greedy algorithm, which attempts to locate the ground state of an approximate energy function, exploits the fact that protein structures consist of overlapping structural building blocks that are not independent. Application of this approach to a series of 16 proteins with 50-250 amino acids leads to predicted models deviating from the experimental structures by 0.5 A RMSD using an RMSD-based energy function and within 1.5 to 4.8 A RMSD using a Go-based energy function. The Go-based results are significant because they illustrate the strength of combining structural fragments and stochastic greedy algorithms in capturing the native structures of proteins stabilized by long-range interactions separated by more than 30 amino acids. These results clearly open the door to less computationally demanding solutions to predict structures from sequences.
More Related Videos

Related Concept Videos
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence.
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 02:15
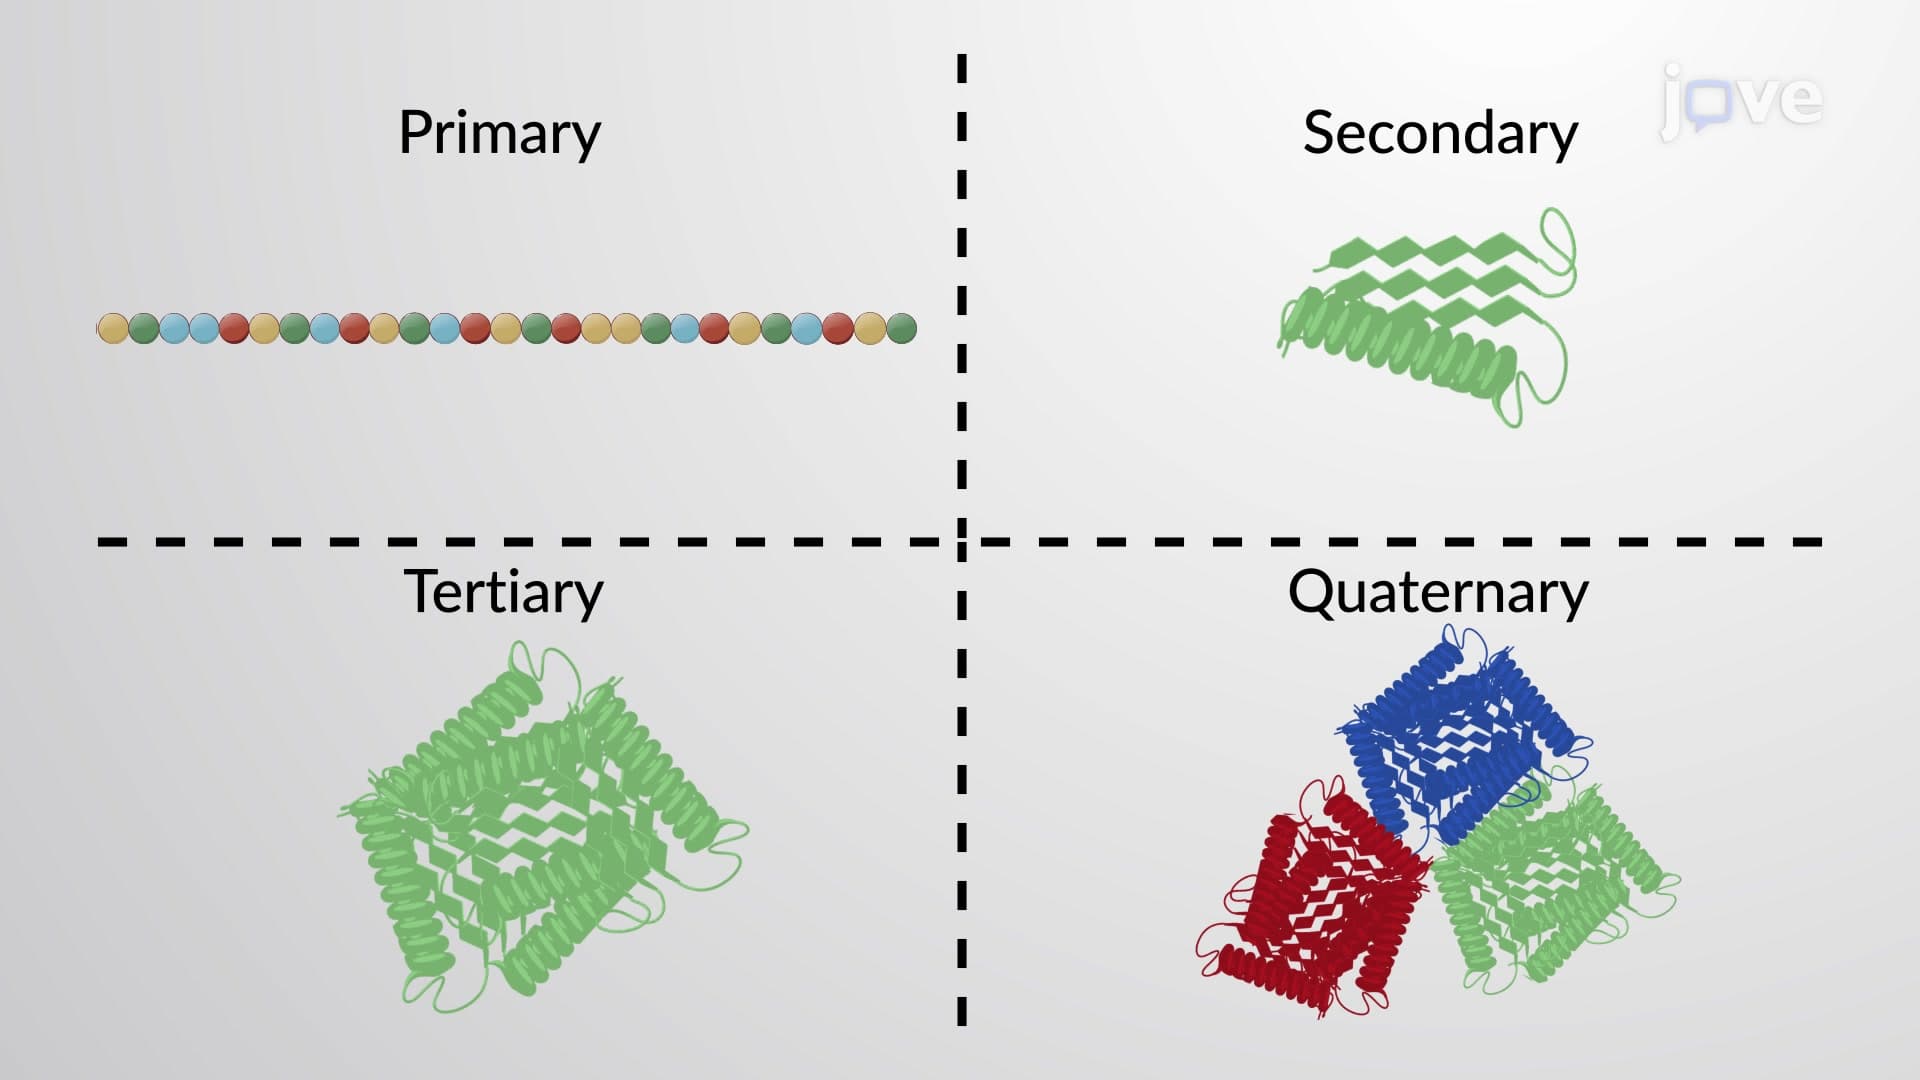
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme can...
02:26Conservation of Protein Domains
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:22
Protein Folding