Related Experiment Video
Updated: Aug 19, 2026
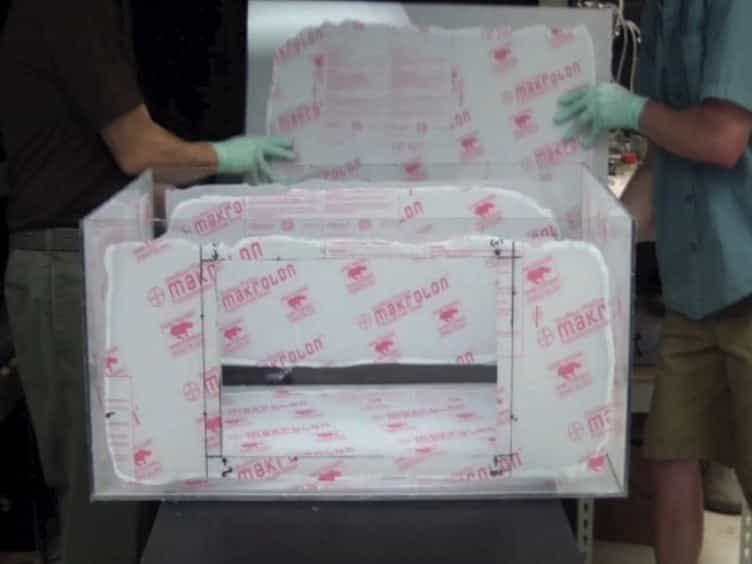

Construction of Vapor Chambers Used to Expose Mice to Alcohol During the Equivalent of all Three Trimesters of Human Development
Published on: July 13, 2014
Calcium-stimulated adenylyl cyclases modulate ethanol-induced neurodegeneration in the neonatal brain
James W Maas1, Ricardo A Indacochea, Lisa M Muglia
1Department of Pediatrics, Washington University School of Medicine, St. Louis, Missouri 63110, USA.

Abstract:
Fetal alcohol exposure results in cognitive and neurobehavioral deficits, but the effects of modifying genetic loci on the severity of these sequelas have not been well characterized. Although the cAMP signaling pathway has been shown to be an important modulator of ethanol sensitivity in adult mice, its potential role in modulating ethanol-induced neurodegeneration has not been examined. Adenylyl cyclases (ACs) 1 and 8 produce cAMP in response to intracellular calcium elevation and modulate several aspects of neuronal function, including ethanol sensitivity. AC1 and AC8 are expressed widely throughout the brain of neonatal mice, and genetic deletion of both AC1 and AC8 in double-knock-out (DKO) mice enhances ethanol-induced neurodegeneration in the brains of neonatal mice. In addition, ethanol treatment induces significantly greater levels of caspase-3 activation in the brains of DKO mice compared with wild-type (WT) mice, reflecting higher numbers of apoptotic neurons. Administration of the NMDA receptor antagonist MK801 [(+)-5-methyl-10,11-dihydro-5H-dibenzo [a,d] cyclohepten-5,10-imine hydrogen maleate] or the GABA(A) receptor potentiator phenobarbital, which mimics components of the effects of ethanol on neurons, results in significantly greater neurodegeneration in the brains of neonatal DKO mice than WT mice. Furthermore, loss of a single calcium-stimulated AC isoform potentiates neurodegeneration after administration of ethanol, MK801, or phenobarbital. In contrast, the levels of physiological cell death, death after hypoxia/ischemia, and excitotoxic cell death are not increased in the brains of DKO mice. Thus, AC1 and AC8 are critical modulators of neurodegeneration induced by activity blockade in the neonatal brain and represent genetic loci that may potentially modify the severity of fetal alcohol syndrome.
Insights
Genetic deletion of Adenylyl Cyclases 1 and 8 (AC1 and AC8) in mice worsens ethanol-induced neurodegeneration. These enzymes are critical for modulating brain cell death from fetal alcohol exposure.
Area of Science:
- Neuroscience
- Developmental Biology
- Pharmacology
Background:
- Fetal alcohol exposure causes neurodevelopmental deficits.
- The cAMP signaling pathway's role in ethanol-induced neurodegeneration is unexamined.
- Adenylyl Cyclases 1 and 8 (AC1 and AC8) are calcium-stimulated enzymes involved in cAMP production and ethanol sensitivity.
Purpose of the Study:
- To investigate the role of AC1 and AC8 in modulating ethanol-induced neurodegeneration in the neonatal brain.
- To determine if genetic deletion of AC1 and AC8 affects ethanol's impact on neuronal apoptosis.
Main Methods:
- Utilized double-knock-out (DKO) mice lacking both AC1 and AC8, and wild-type (WT) littermates.
- Administered ethanol, NMDA receptor antagonist MK801, or GABA(A) receptor agonist phenobarbital to neonatal mice.
- Assessed neurodegeneration and caspase-3 activation as indicators of apoptosis.
Main Results:
- DKO mice exhibited significantly enhanced ethanol-induced neurodegeneration compared to WT mice.
- Ethanol treatment led to greater caspase-3 activation and neuronal apoptosis in DKO neonatal brains.
- MK801 and phenobarbital also caused more severe neurodegeneration in DKO mice, indicating AC1/AC8's role in activity-blockade-induced cell death.
Conclusions:
- AC1 and AC8 are critical modulators of neurodegeneration induced by ethanol and related neuroactive drugs in the neonatal brain.
- These calcium-stimulated adenylyl cyclase isoforms represent potential genetic loci influencing fetal alcohol syndrome severity.
- Targeting AC1 and AC8 may offer therapeutic strategies for mitigating alcohol-related neurodevelopmental disorders.