Creutzfeldt-Jakob disease associated with the R208H mutation in the prion protein gene
Related Experiment Video
Updated: Aug 19, 2026
Prion Safety Laboratory Swipe Test
Published on: February 14, 2025
S Capellari1, F Cardone, S Notari
1Dipartimento di Scienze Neurologiche, Università di Bologna, Bologna, Italy.

Abstract:
The authors investigated a patient who died of apparent sporadic Creutzfeldt-Jakob disease (CJD) but carried a R208H substitution in the prion protein (PrP). The patient phenotype was indistinguishable from typical sporadic CJD (i.e., MM1 subtype). In addition, pathologic PrP, PrP(Sc), originated from both the normal and the mutated PRNP allele and had the same characteristics as PrP(Sc) type 1. The authors propose that the R208H mutation influences disease susceptibility without significantly affecting PrP(Sc) properties or disease phenotype.
Insights
Researchers studied a Creutzfeldt-Jakob disease (CJD) patient with a R208H prion protein (PrP) mutation. The mutation appears to increase susceptibility to CJD without altering the disease
Area of Science:
- Neurodegenerative diseases
- Prion biology
- Human genetics
Background:
- Sporadic Creutzfeldt-Jakob disease (sCJD) is a fatal neurodegenerative disorder.
- The prion protein gene (PRNP) encodes the prion protein (PrP).
- Genetic variations in PRNP can influence CJD susceptibility and phenotype.
Observation:
- A patient presented with clinical and pathological features typical of the MM1 subtype of sporadic CJD.
- This patient, however, possessed a heterozygous R208H substitution in the PRNP gene.
- Pathologic prion protein (PrPSc) was detected originating from both the wild-type and the mutated PRNP alleles.
Findings:
- The PrPSc in the patient exhibited characteristics consistent with PrPSc type 1, irrespective of its allele of origin.
- The R208H mutation did not alter the fundamental properties of the pathogenic PrPSc.
- The patient's phenotype remained indistinguishable from common forms of sporadic CJD.
Implications:
- The R208H mutation may confer susceptibility to developing sporadic CJD.
- This genetic variation appears to influence disease risk rather than modifying the core characteristics of the prion agent.
- Further research is needed to fully elucidate the role of PRNP mutations in prion disease pathogenesis.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:24
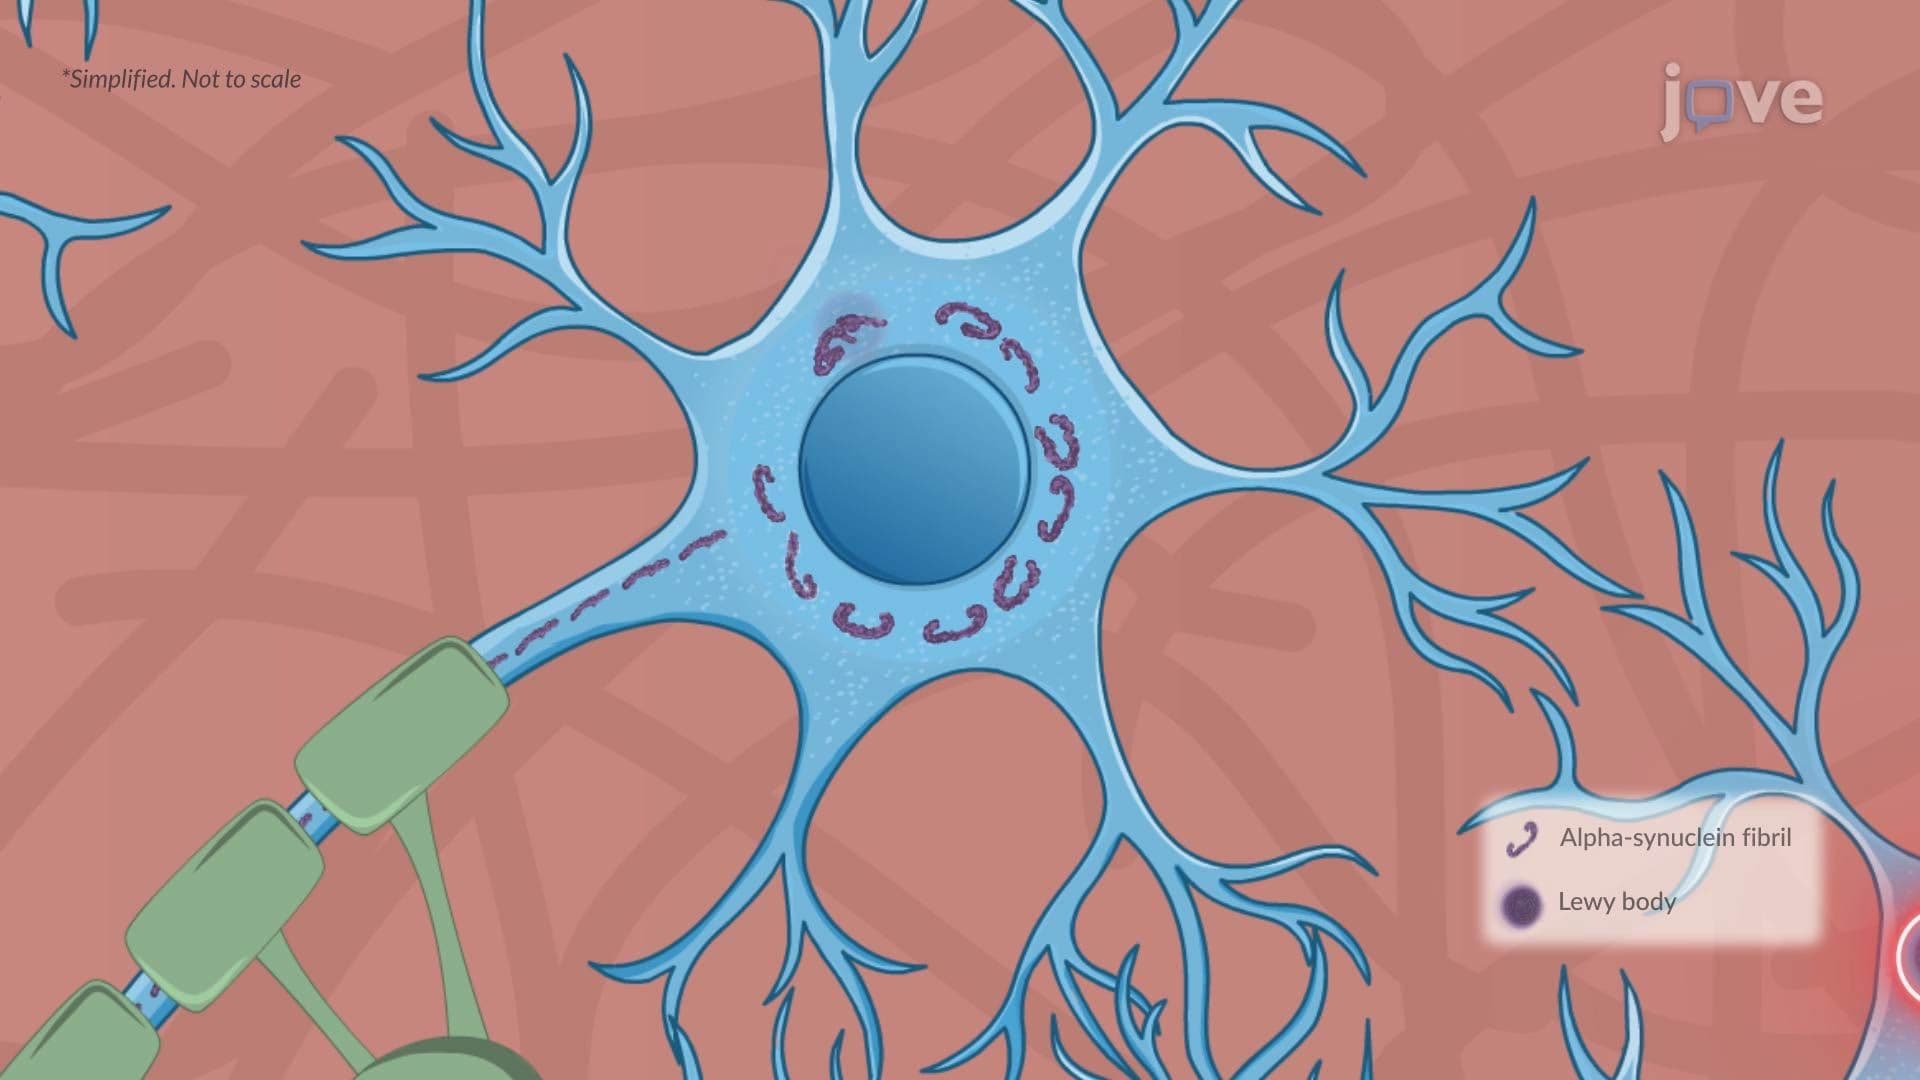
Parkinson Disease ll: Pathophysiology
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of Life
01:31Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of Life
 01:23
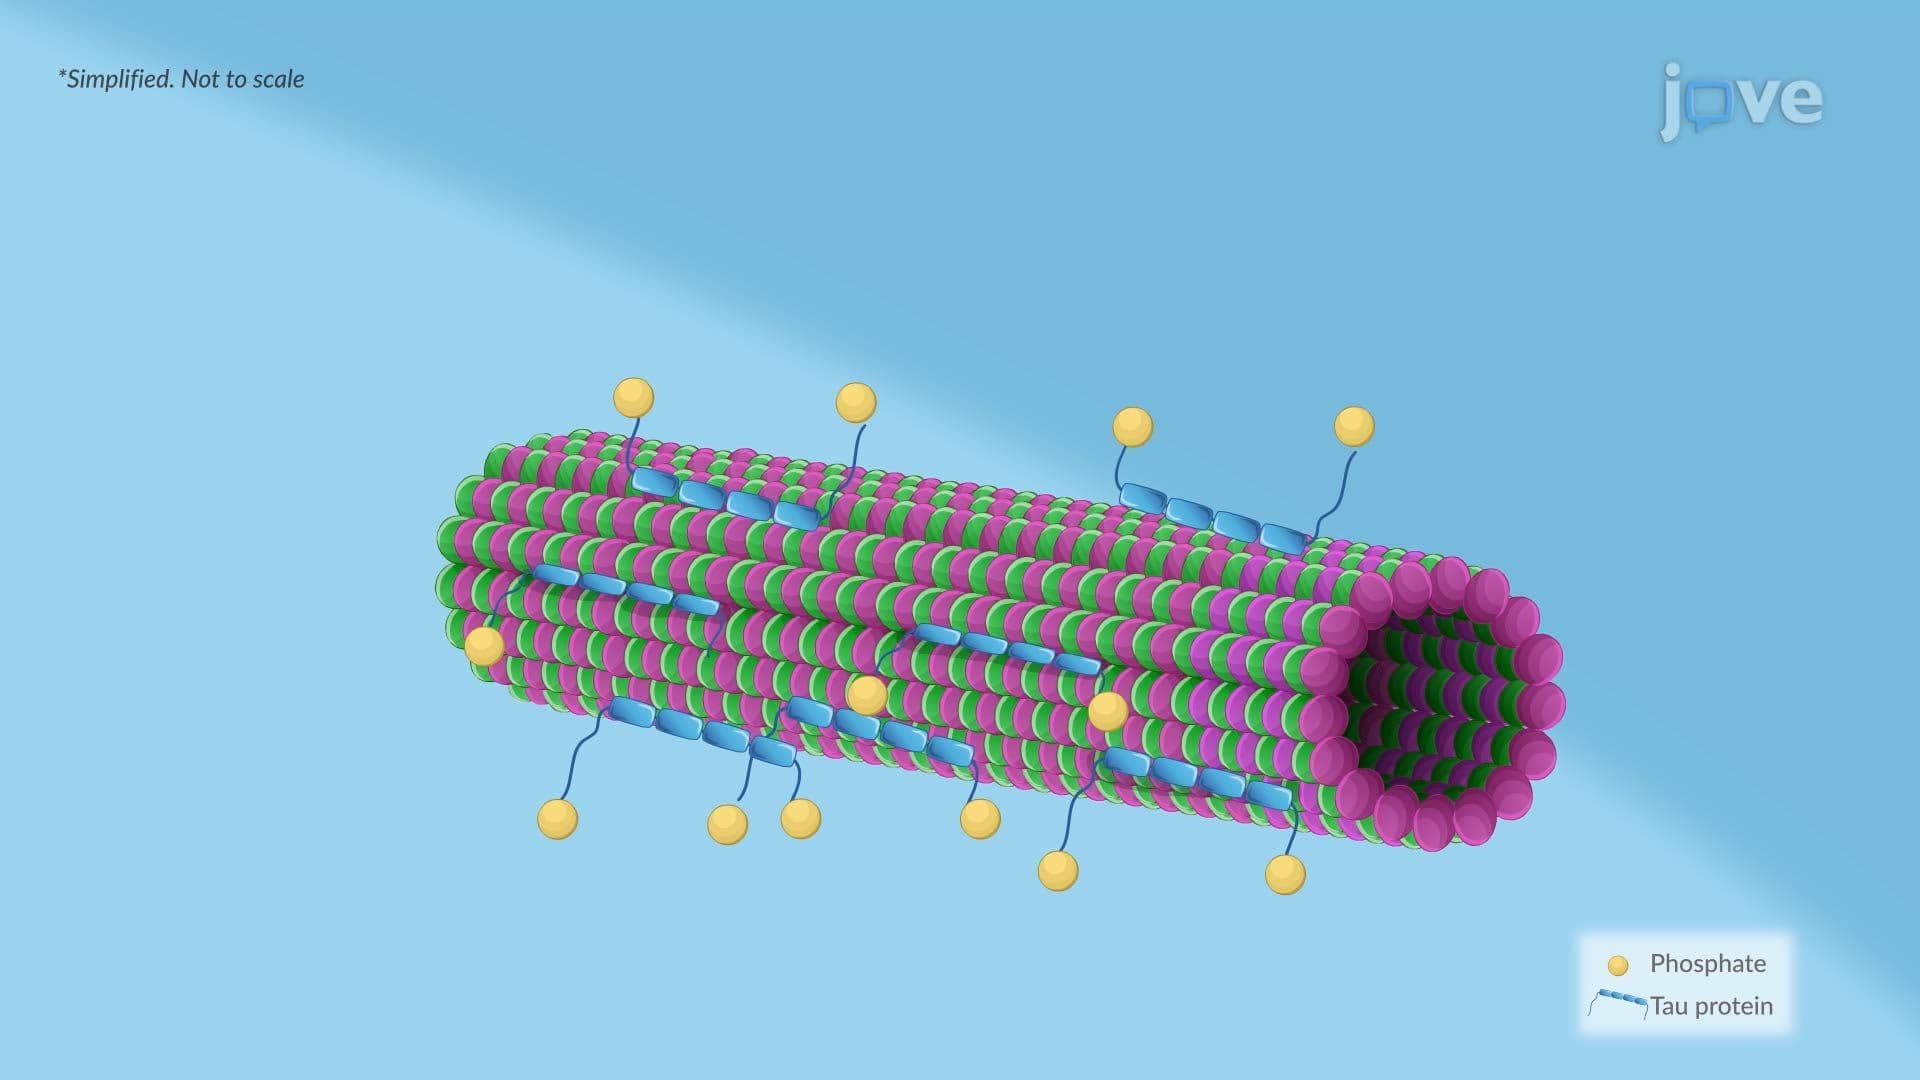
Alzheimer Disease ll: Pathophysiology