Related Experiment Video
Updated: Aug 18, 2026

Implementation of Non-invasive Point of Care Transient Elastography for Evaluation of Liver Disease in Pediatric Populations with Cystic Fibrosis
Published on: August 29, 2025
Lung function in adults with cystic fibrosis at altitude: impact on air travel
R Fischer1, S M Lang, K Brückner
1Dept of Medicine, Division of Pneumology, Clinic for Internal Medicine, Dr. von Hauner's Children's Hospital, University of Munich, Munich, Germany. rainald.fischer@med.uni-muenchen.de

Abstract:
Current guidelines for air travel state that patients with chronic respiratory diseases are required to use oxygen if their in-flight arterial oxygen tensions (Pa,O2) drop below 6.6 kPa. This recommendation may not be strictly applicable to cystic fibrosis patients, who may tolerate lower Pa,O2 for several hours without clinical symptoms. Lung function, symptoms, blood gas levels and signs of pulmonary hypertension were studied in 36 cystic fibrosis patients at altitudes of 530 m and, after 7 h, 2,650 m. A hypoxia inhalation test (inspiratory oxygen fraction 0.15) was performed at low altitude in order to predict high-altitude hypoxaemia. Median Pa,O2 dropped from 9.8 kPa at low altitude to 7.0 kPa at high altitude. Mild exercise at a workload of 30 W further decreased Pa,O2. Two-thirds of all patients exhibited Pa,O2 of <6.6 kPa during exercise and, except for one patient, were asymptomatic. Patients were significantly less obstructed at an altitude of 2,650 m. Low forced expiratory volume in one second at baseline was associated with a low Pa,O2 at altitude. It is concluded that cystic fibrosis patients with baseline arterial oxygen tensions of >8.0 kPa safely tolerate an altitude of 2,650 m for several hours under resting conditions. The risk assessment of low in-flight oxygenation should encompass the whole clinical situation of cystic fibrosis patients, with special attention being paid to the presence of severe airway obstruction.
More Related Videos
 07:38
Design and Development of a Model to Study the Effect of Supplemental Oxygen on the Cystic Fibrosis Airway Microbiome
Published on: August 3, 2021
 10:18
Real-Time, Semi-Automated Fluorescent Measurement of the Airway Surface Liquid pH of Primary Human Airway Epithelial Cells
Published on: June 13, 2019
Related Concept Videos
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:19
Factors Affecting Pulmonary Ventilation
Alveolar Surface Tension
The alveolar fluid lines the luminal surface of the alveoli and exerts a force called surface tension. This force is caused by the polar water molecules in the liquid being more strongly attracted to each...
 01:25
Pulmonary Function Tests
Pulmonary Function Tests are crucial diagnostic tools for assessing respiratory function, particularly in patients with chronic respiratory disorders. They comprehensively evaluate lung volumes, ventilatory function, breathing mechanics, diffusion, and gas exchange. These tests help diagnose pulmonary diseases and play a significant role in monitoring disease progression, evaluating disability, and assessing response to therapy.
PFTs involve using a spirometer, a...
 01:24
Pressure Relationships in Thoracic Cavity
Breathing Mechanisms
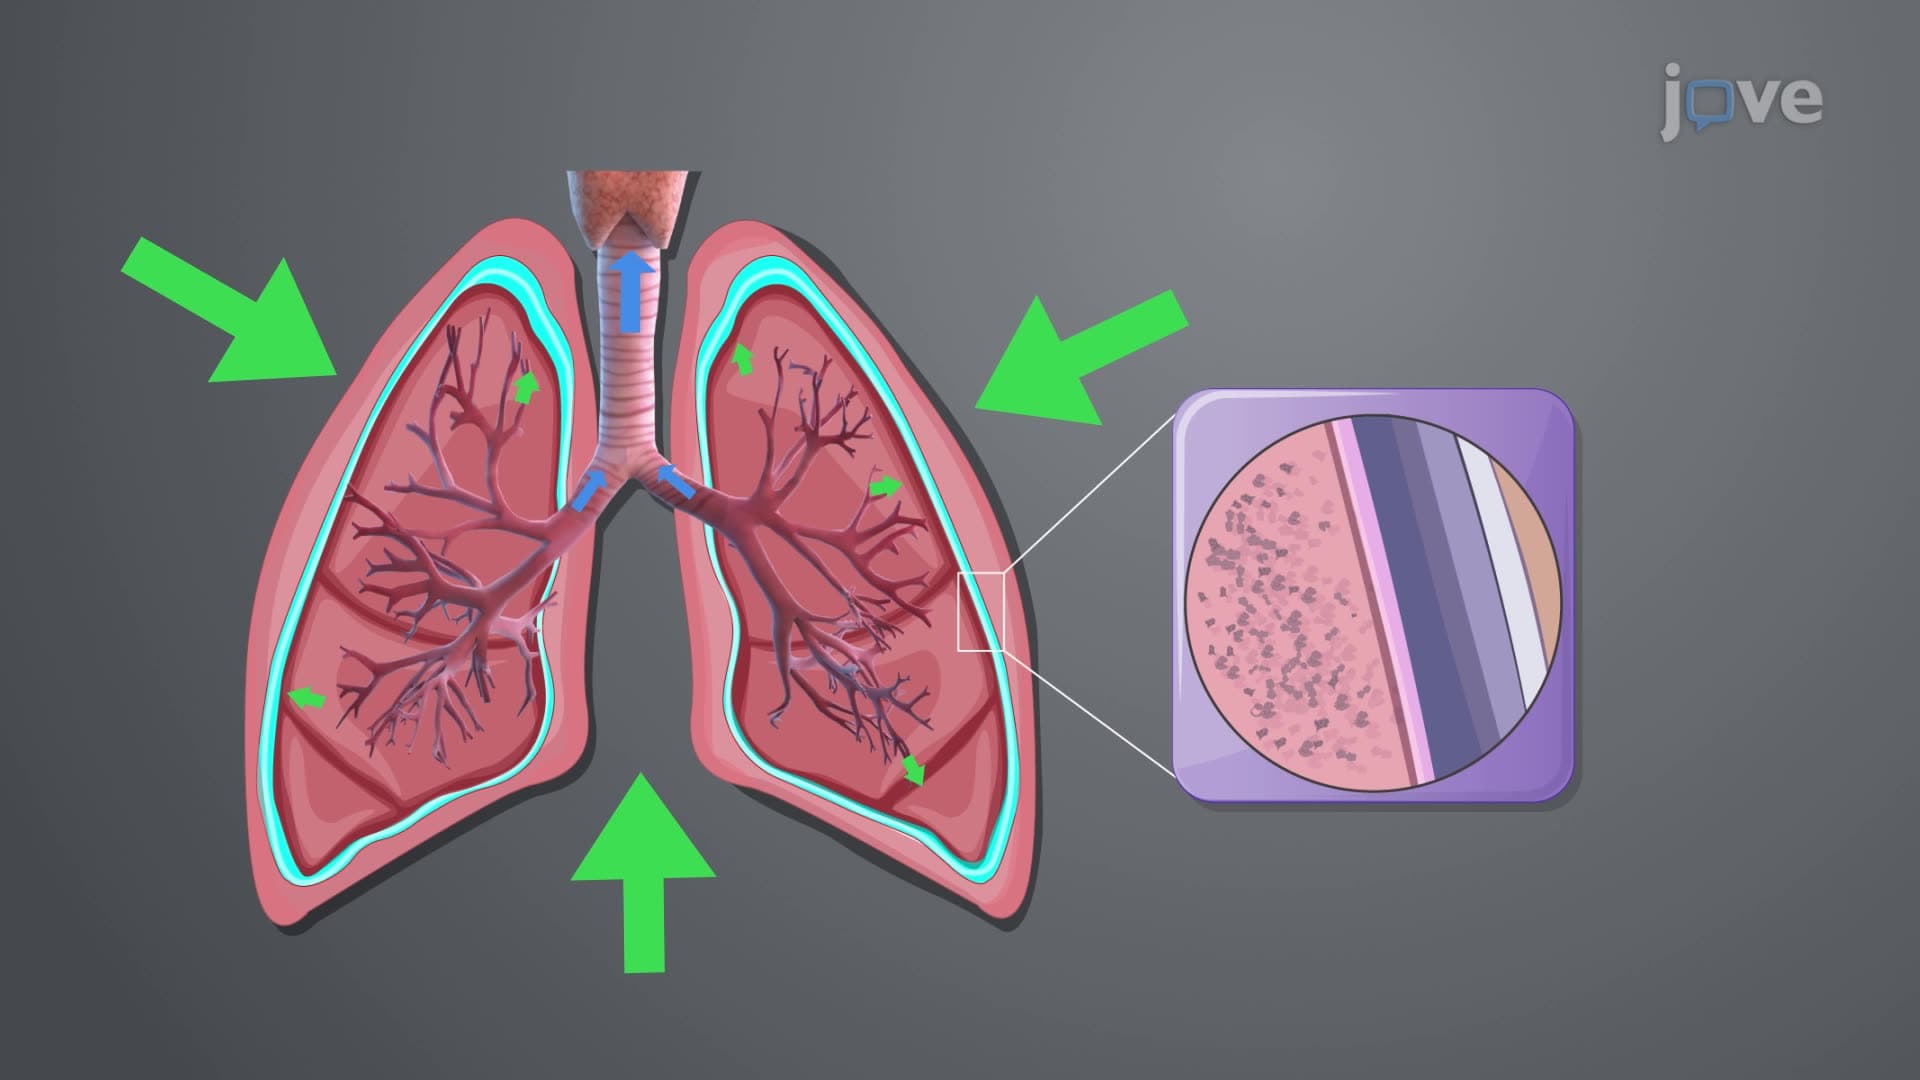
Both intra-alveolar and intrapleural pressures rely on specific lung properties. The ability to breathe—allowing air to enter the lungs during...
 01:24
Factors Affecting Respiration