Related Experiment Video
Updated: Aug 18, 2026

Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Visualizing conflicting evolutionary hypotheses in large collections of trees: using consensus networks to study the
Barbara Holland1, Frédéric Delsuc, Vincent Moulton
1The Allan Wilson Centre for Molecular Ecology and Evolution, Massey University, Private Bag 11222, Palmerston North, New Zealand. b.r.holland@massey.ac.nz

Abstract:
Many phylogenetic methods produce large collections of trees as opposed to a single tree, which allows the exploration of support for various evolutionary hypotheses. However, to be useful, the information contained in large collections of trees should be summarized; frequently this is achieved by constructing a consensus tree. Consensus trees display only those signals that are present in a large proportion of the trees. However, by their very nature consensus trees require that any conflicts between the trees are necessarily disregarded. We present a method that extends the notion of consensus trees to allow the visualization of conflicting hypotheses in a consensus network. We demonstrate the utility of this method in highlighting differences amongst maximum likelihood bootstrap values and Bayesian posterior probabilities in the placental mammal phylogeny, and also in comparing the phylogenetic signal contained in amino acid versus nucleotide characters for hexapod monophyly.
Related Concept Videos
 02:54
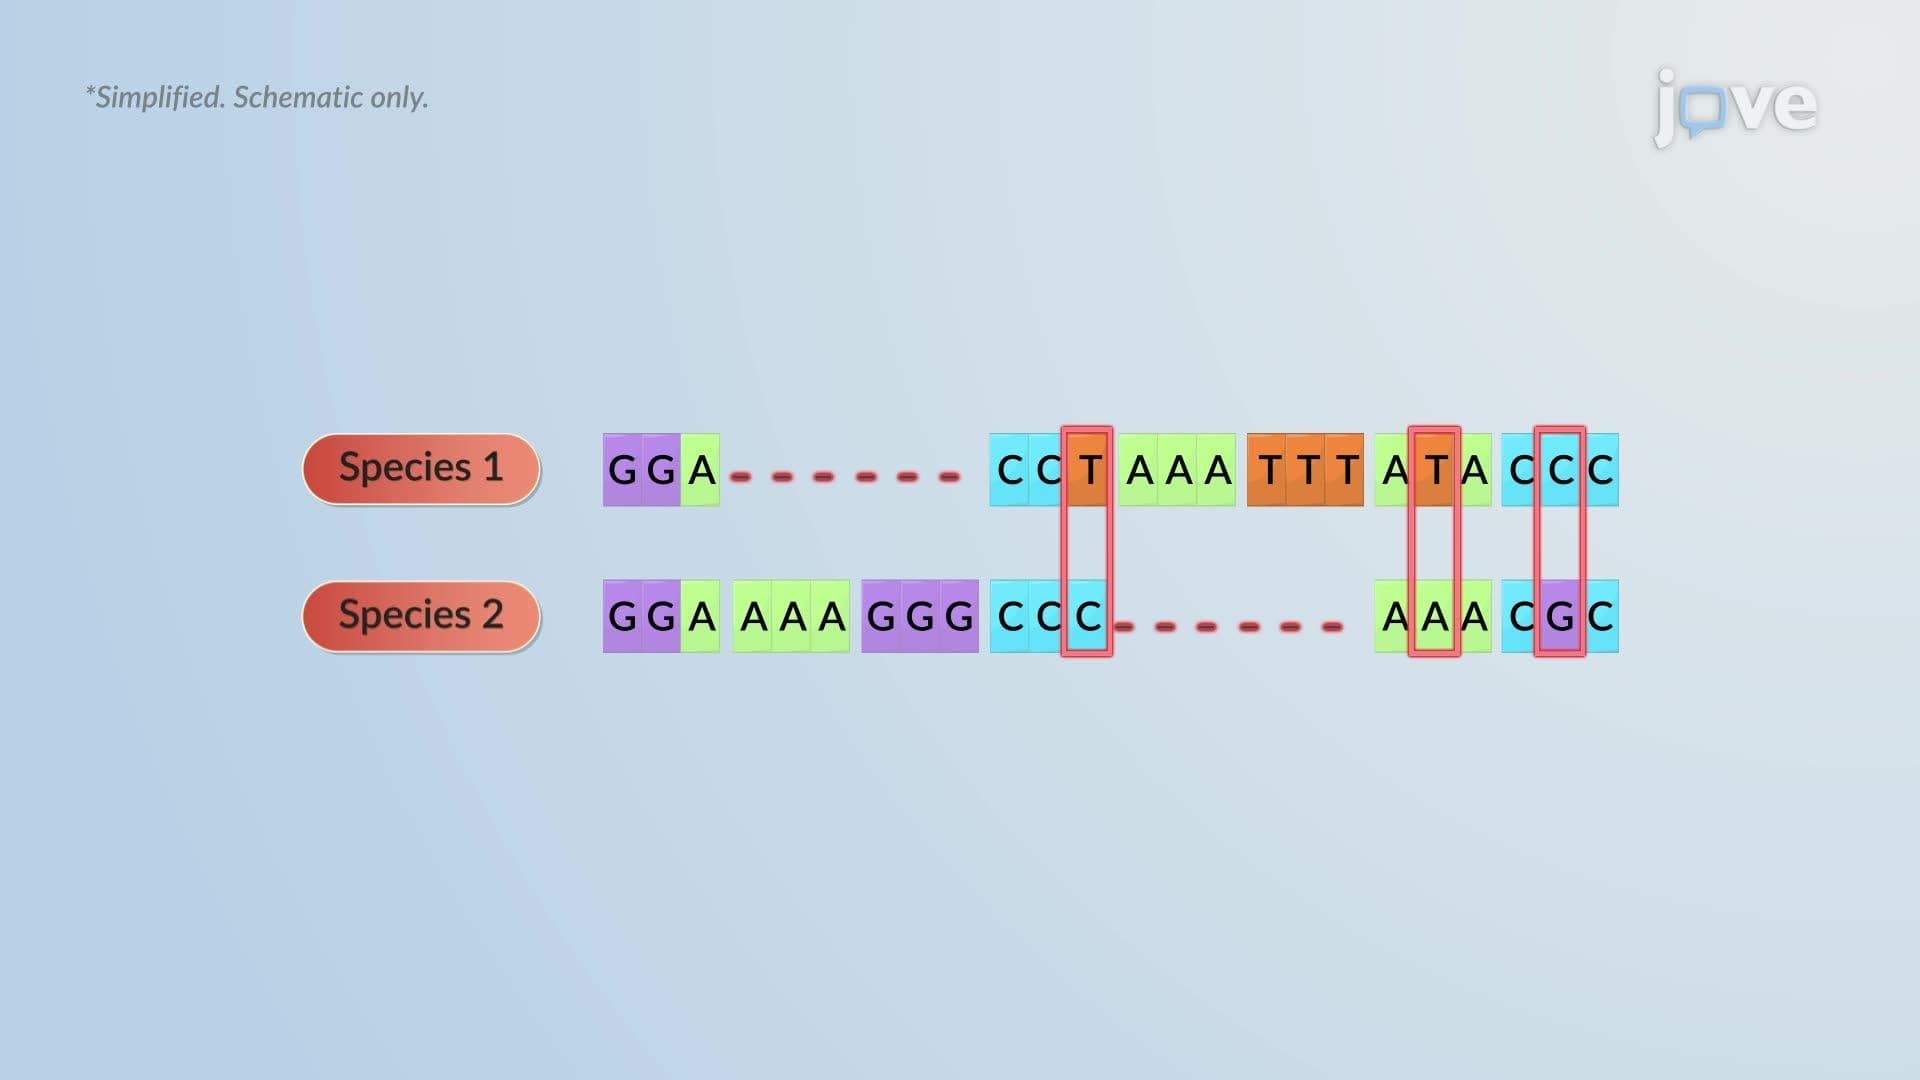
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
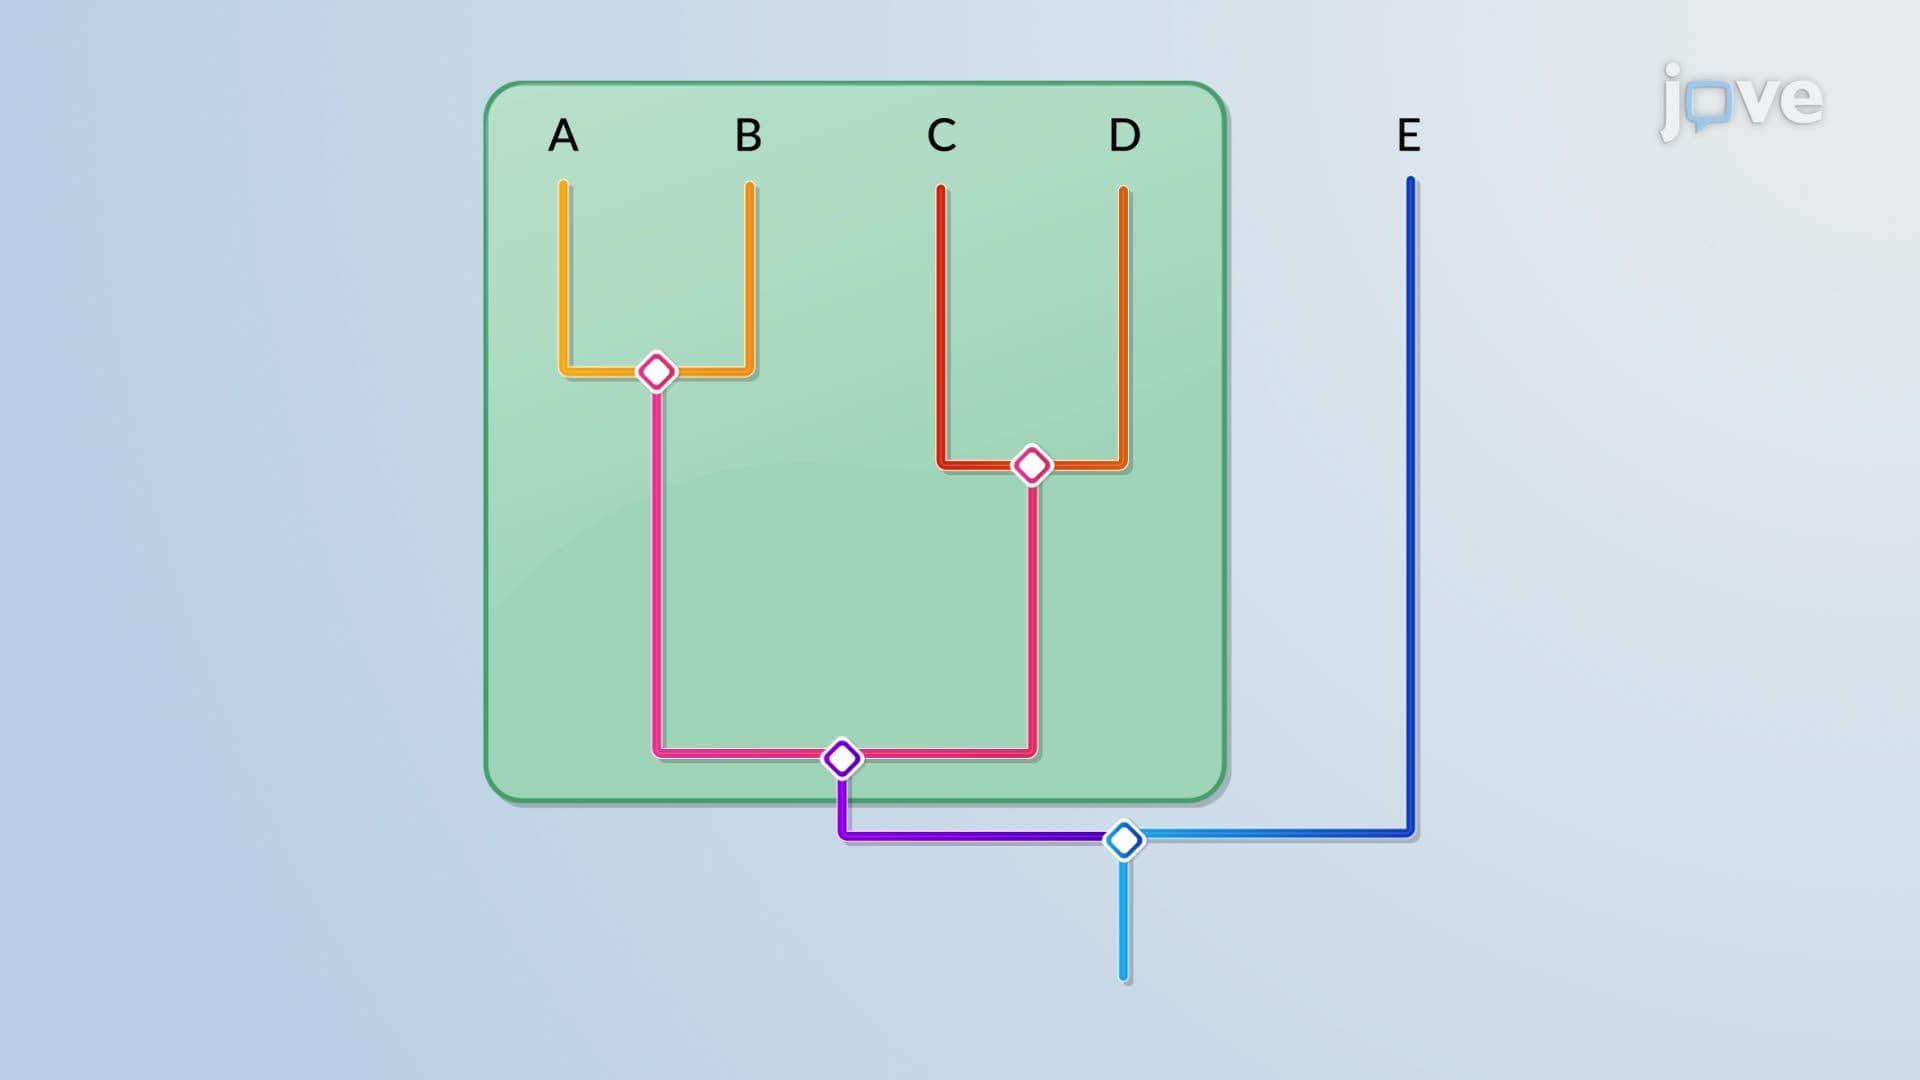 03:21
Phylogenetic Trees
03:21Phylogenetic Trees
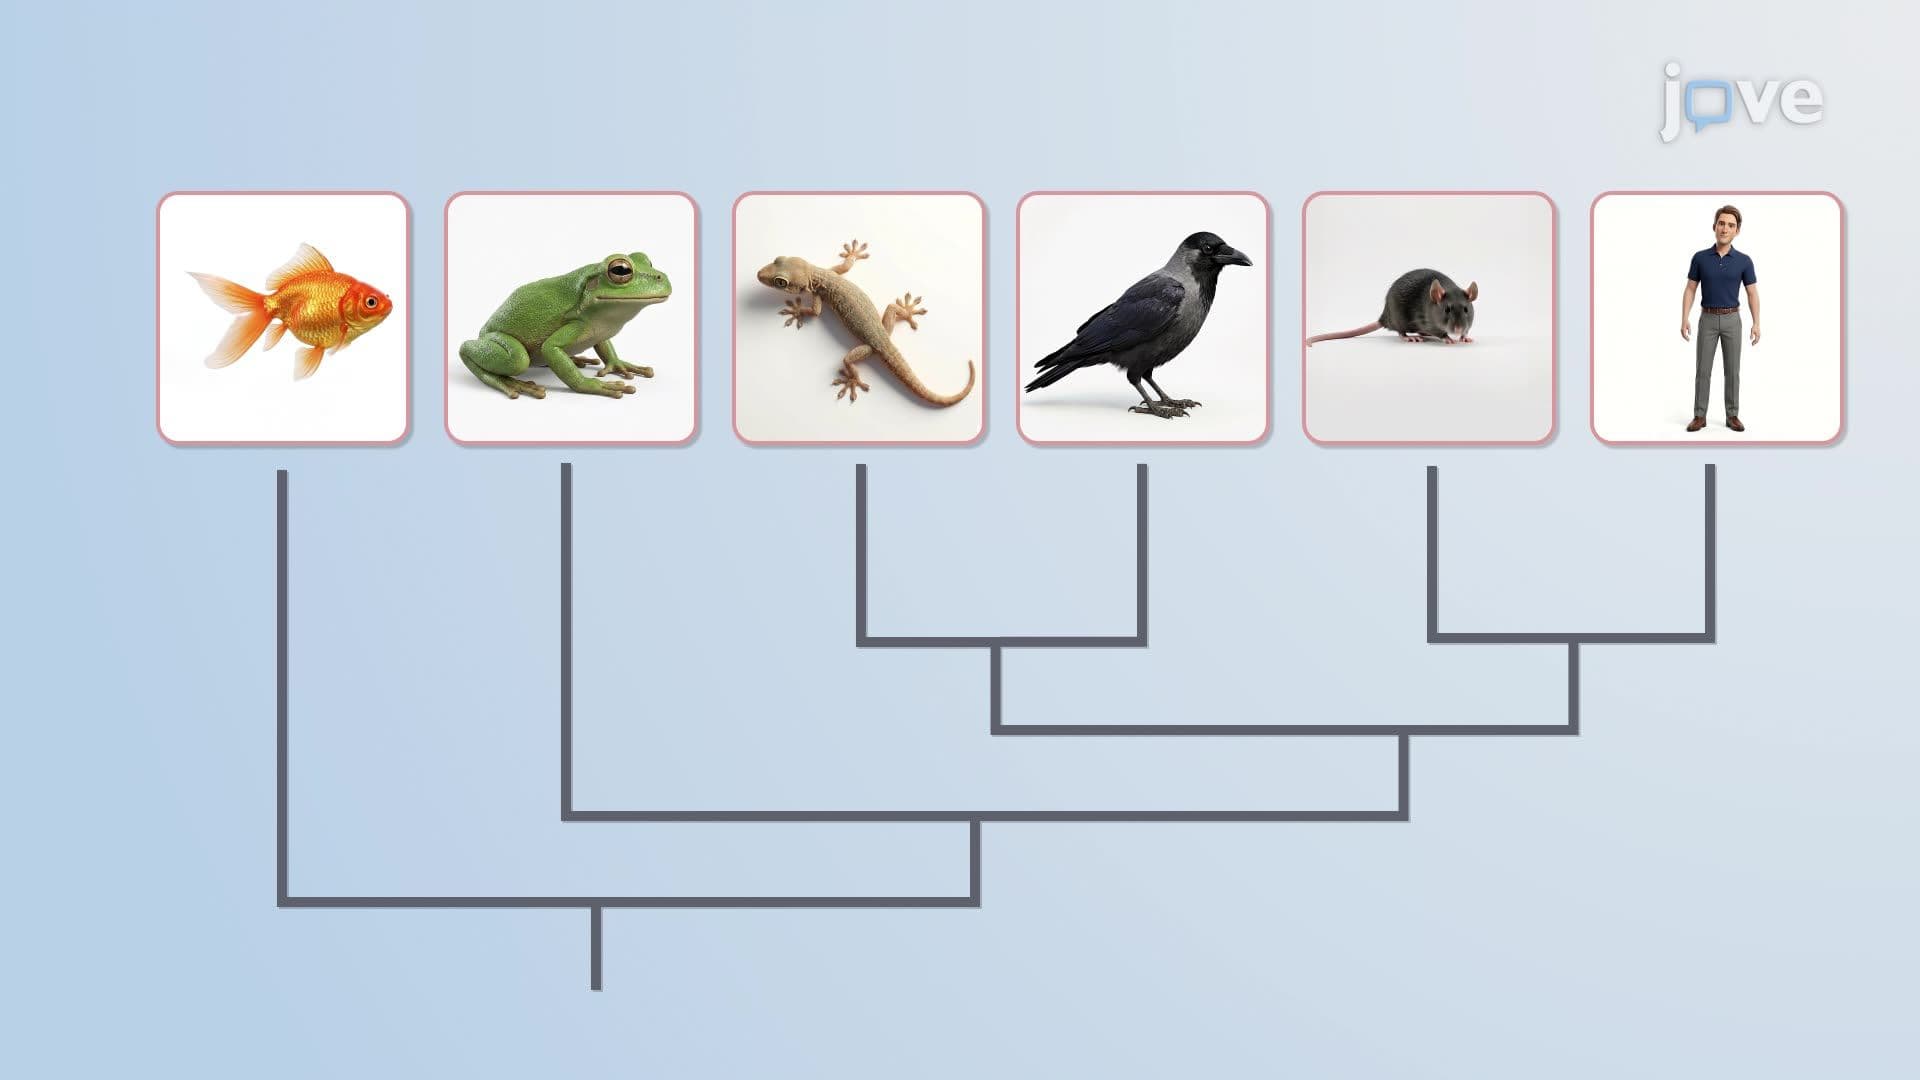 01:23
Phylogeny
 01:54
Convergent Evolution