
[INDO/CI studies on the electronic spectra of tetraphenylporphyrin and its derivatives]

Abstract:
MNDO calculations and group theory were carried out on tetraphenylporphyrine (TPP) to obtain its optimum molecular geometry. We employed an INDO/CI method to study the electronic structures of tetraphenylporphyrine and its derivatives and identify their main electonic absorption bands. Five-orbital model was advanced to explain the typical characteristics of electronic spectra of H2TPP and its derivatives. The calculated wavelengths were in good agreement with the experimental values. The effects of the substitution of variant group were discussed and some helpful conclusions were drawn.
Related Concept Videos
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group with both...
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent of conjugation in the...
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:29
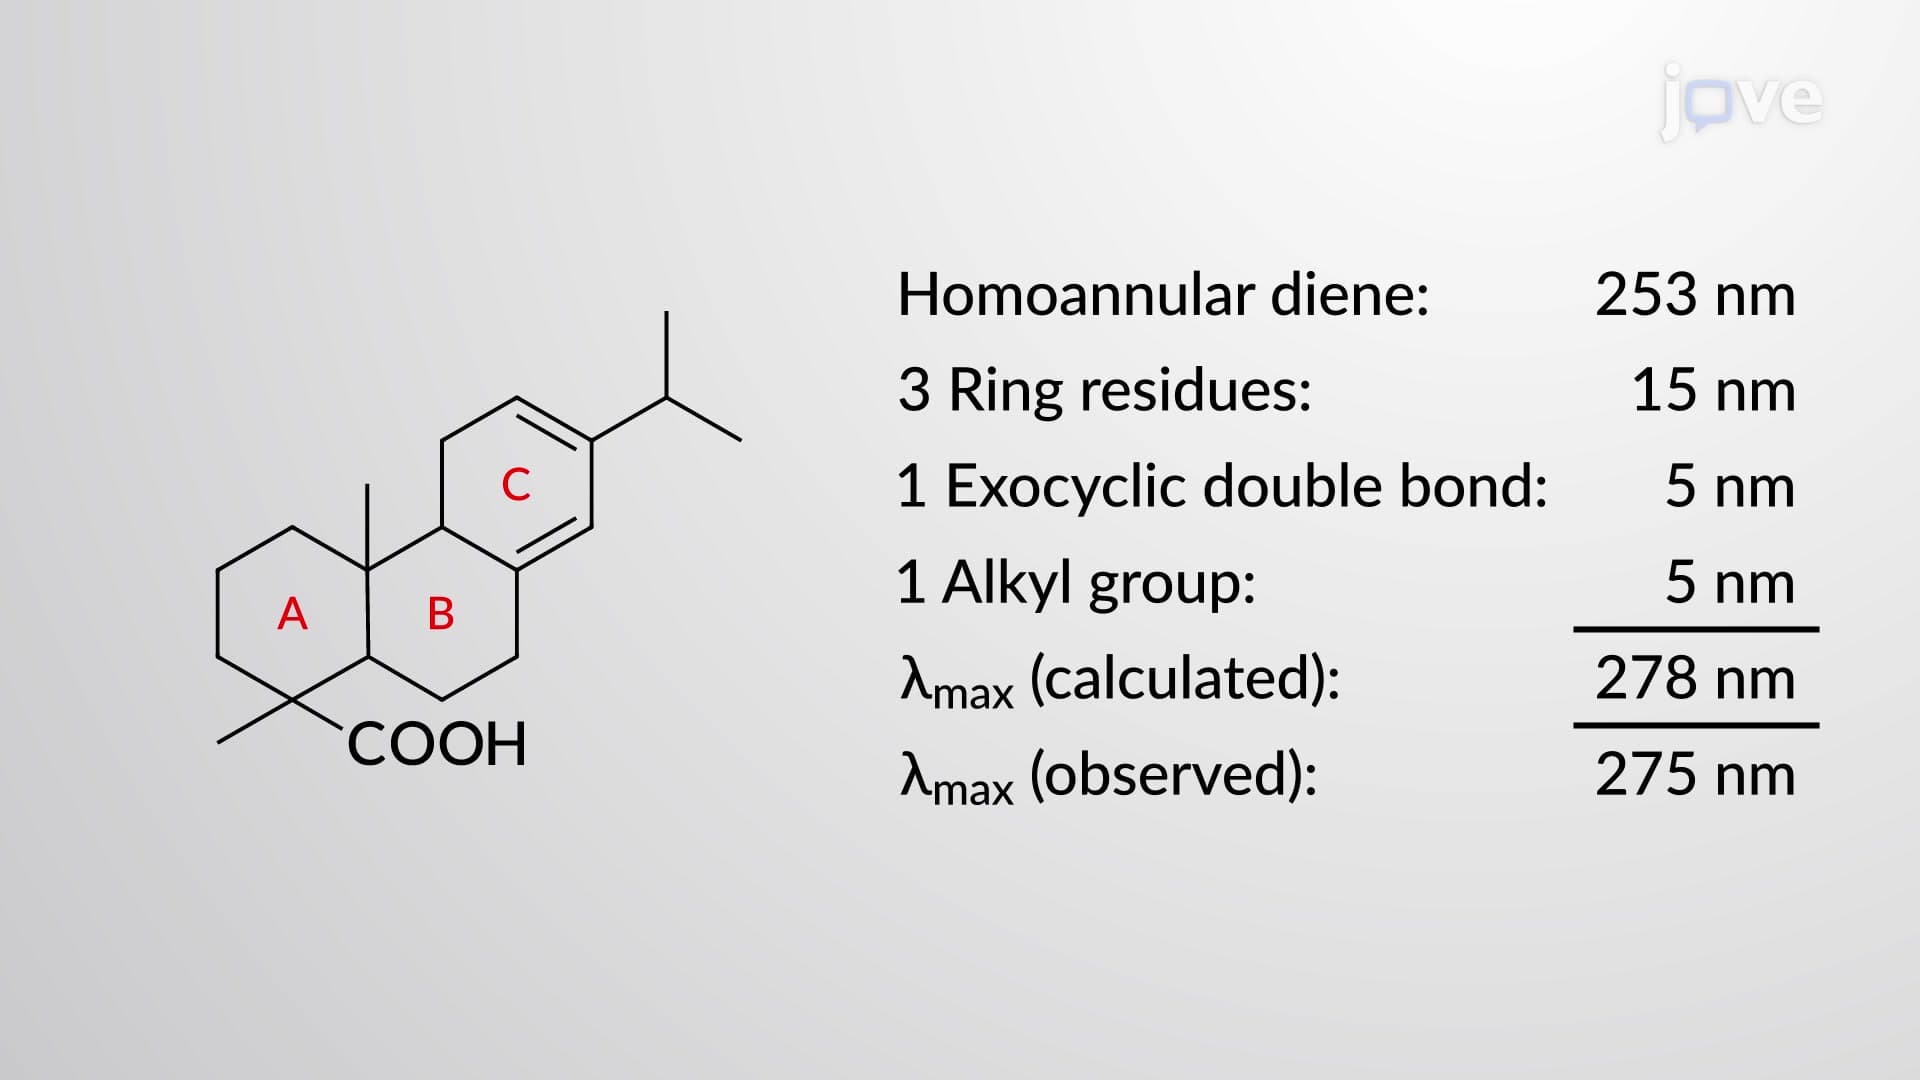
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:28
IR and UV–Vis Spectroscopy of Carboxylic Acids
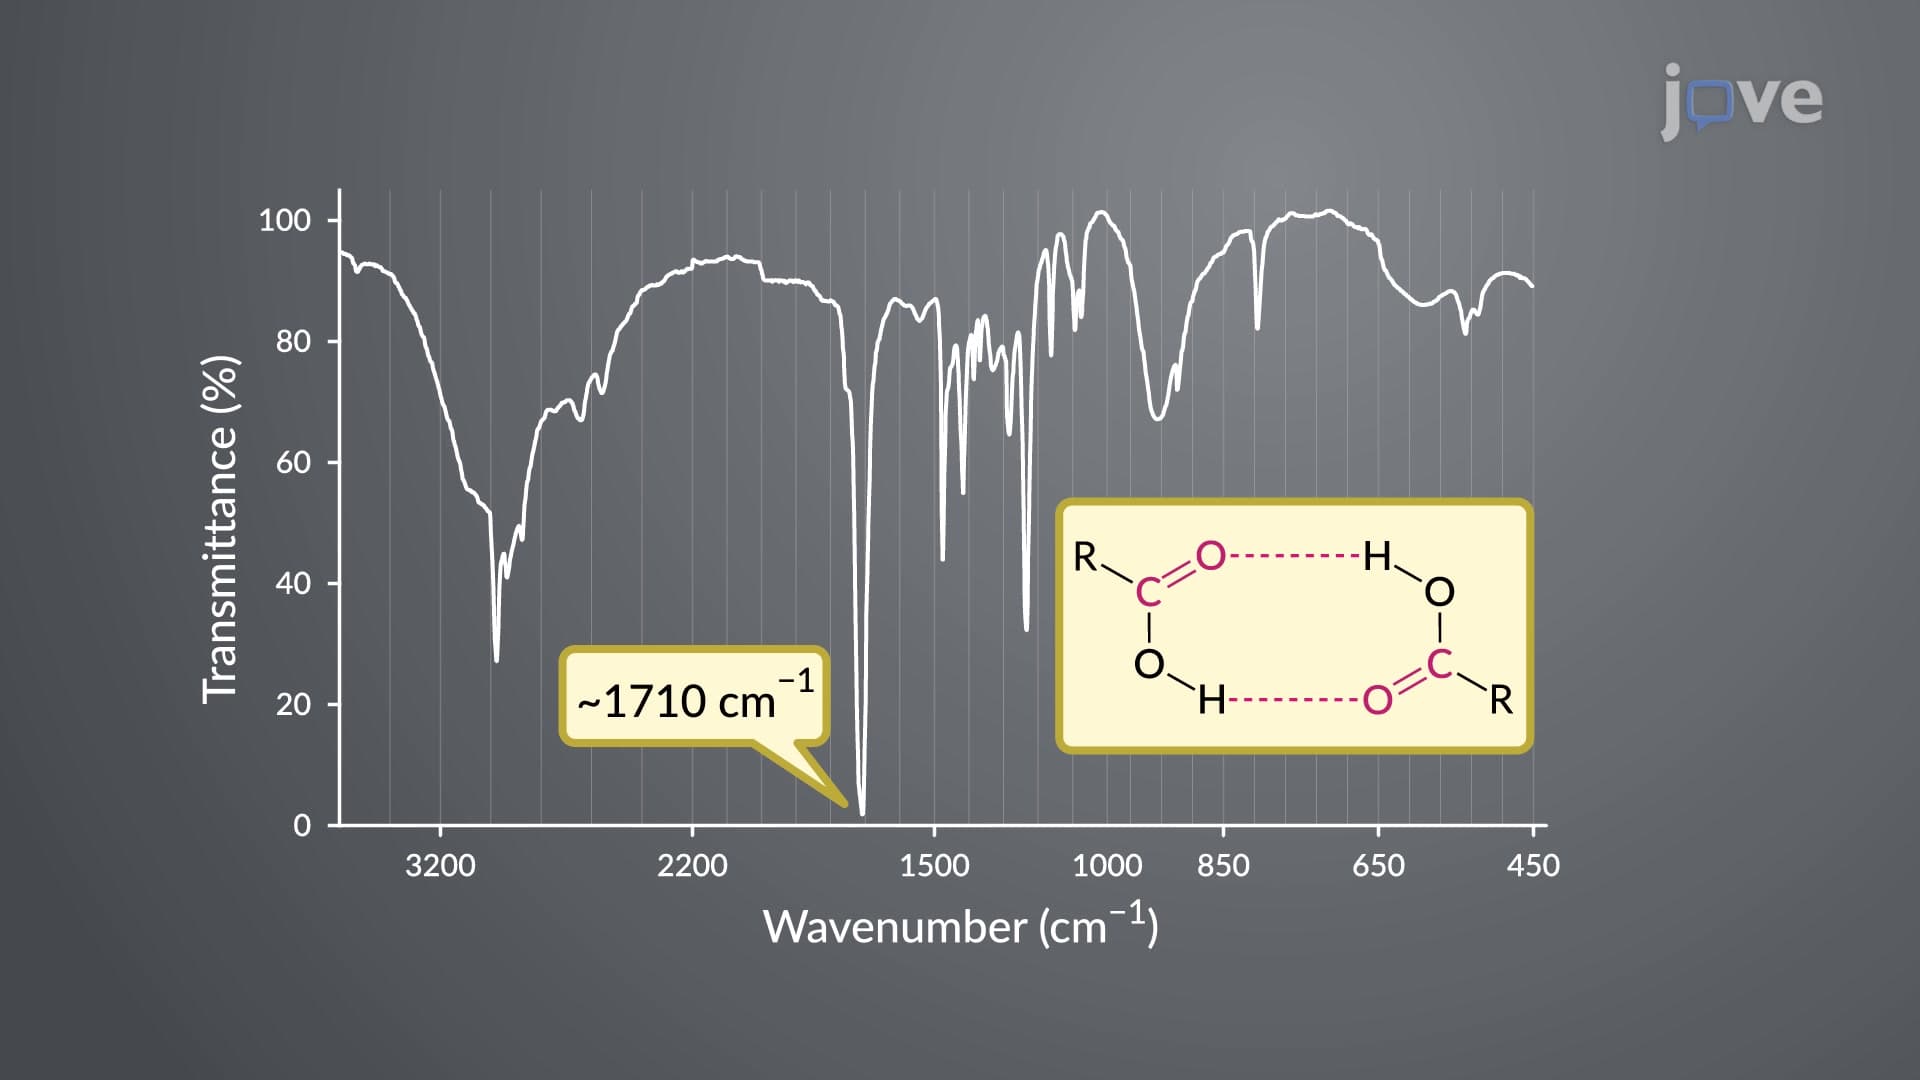
However, the stretching absorptions for the C=O bond vary depending on the structure of carboxylic acids. The C=O bond of the free carboxylic acids shows a higher stretching frequency, 1760 cm−1, while H-bonded carboxylic acids (dimers) exhibit stretching absorptions at a lower frequency, 1710 cm−1. The C=O bond of the...