Related Experiment Video
Updated: Aug 18, 2026
Spatial Separation of Molecular Conformers and Clusters
Published on: January 9, 2014
The merits of the frozen-density embedding scheme to model solvatochromic shifts
Johannes Neugebauer1, Manuel J Louwerse, Evert Jan Baerends
1Theoretical Chemistry, Vrije Universiteit Amsterdam, De Boelelaan 1083, 1081 HV Amsterdam, The Netherlands. jneugeb@chem.vu.nl

Abstract:
We investigate the usefulness of a frozen-density embedding scheme within density-functional theory [J. Phys. Chem. 97, 8050 (1993)] for the calculation of solvatochromic shifts. The frozen-density calculations, particularly of excitation energies have two clear advantages over the standard supermolecule calculations: (i) calculations for much larger systems are feasible, since the time-consuming time-dependent density functional theory (TDDFT) part is carried out in a limited molecular orbital space, while the effect of the surroundings is still included at a quantum mechanical level. This allows a large number of solvent molecules to be included and thus affords both specific and nonspecific solvent effects to be modeled. (ii) Only excitations of the system of interest, i.e., the selected embedded system, are calculated. This allows an easy analysis and interpretation of the results. In TDDFT calculations, it avoids unphysical results introduced by spurious mixings with the artificially too low charge-transfer excitations which are an artifact of the adiabatic local-density approximation or generalized gradient approximation exchange-correlation kernels currently used. The performance of the frozen-density embedding method is tested for the well-studied solvatochromic properties of the n-->pi(*) excitation of acetone. Further enhancement of the efficiency is studied by constructing approximate solvent densities, e.g., from a superposition of densities of individual solvent molecules. This is demonstrated for systems with up to 802 atoms. To obtain a realistic modeling of the absorption spectra of solvated molecules, including the effect of the solvent motions, we combine the embedding scheme with classical molecular dynamics (MD) and Car-Parrinello MD simulations to obtain snapshots of the solute and its solvent environment, for which then excitation energies are calculated. The frozen-density embedding yields estimated solvent shifts in the range of 0.20-0.26 eV, in good agreement with experimental values of between 0.19 and 0.21 eV.
More Related Videos
 07:03
Measuring Magnetically-Tuned Ferroelectric Polarization in Liquid Crystals
Published on: August 15, 2018
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:24
Freezing Point Depression and Boiling Point Elevation
 03:12
Freezing Point Depression and Boiling Point Elevation
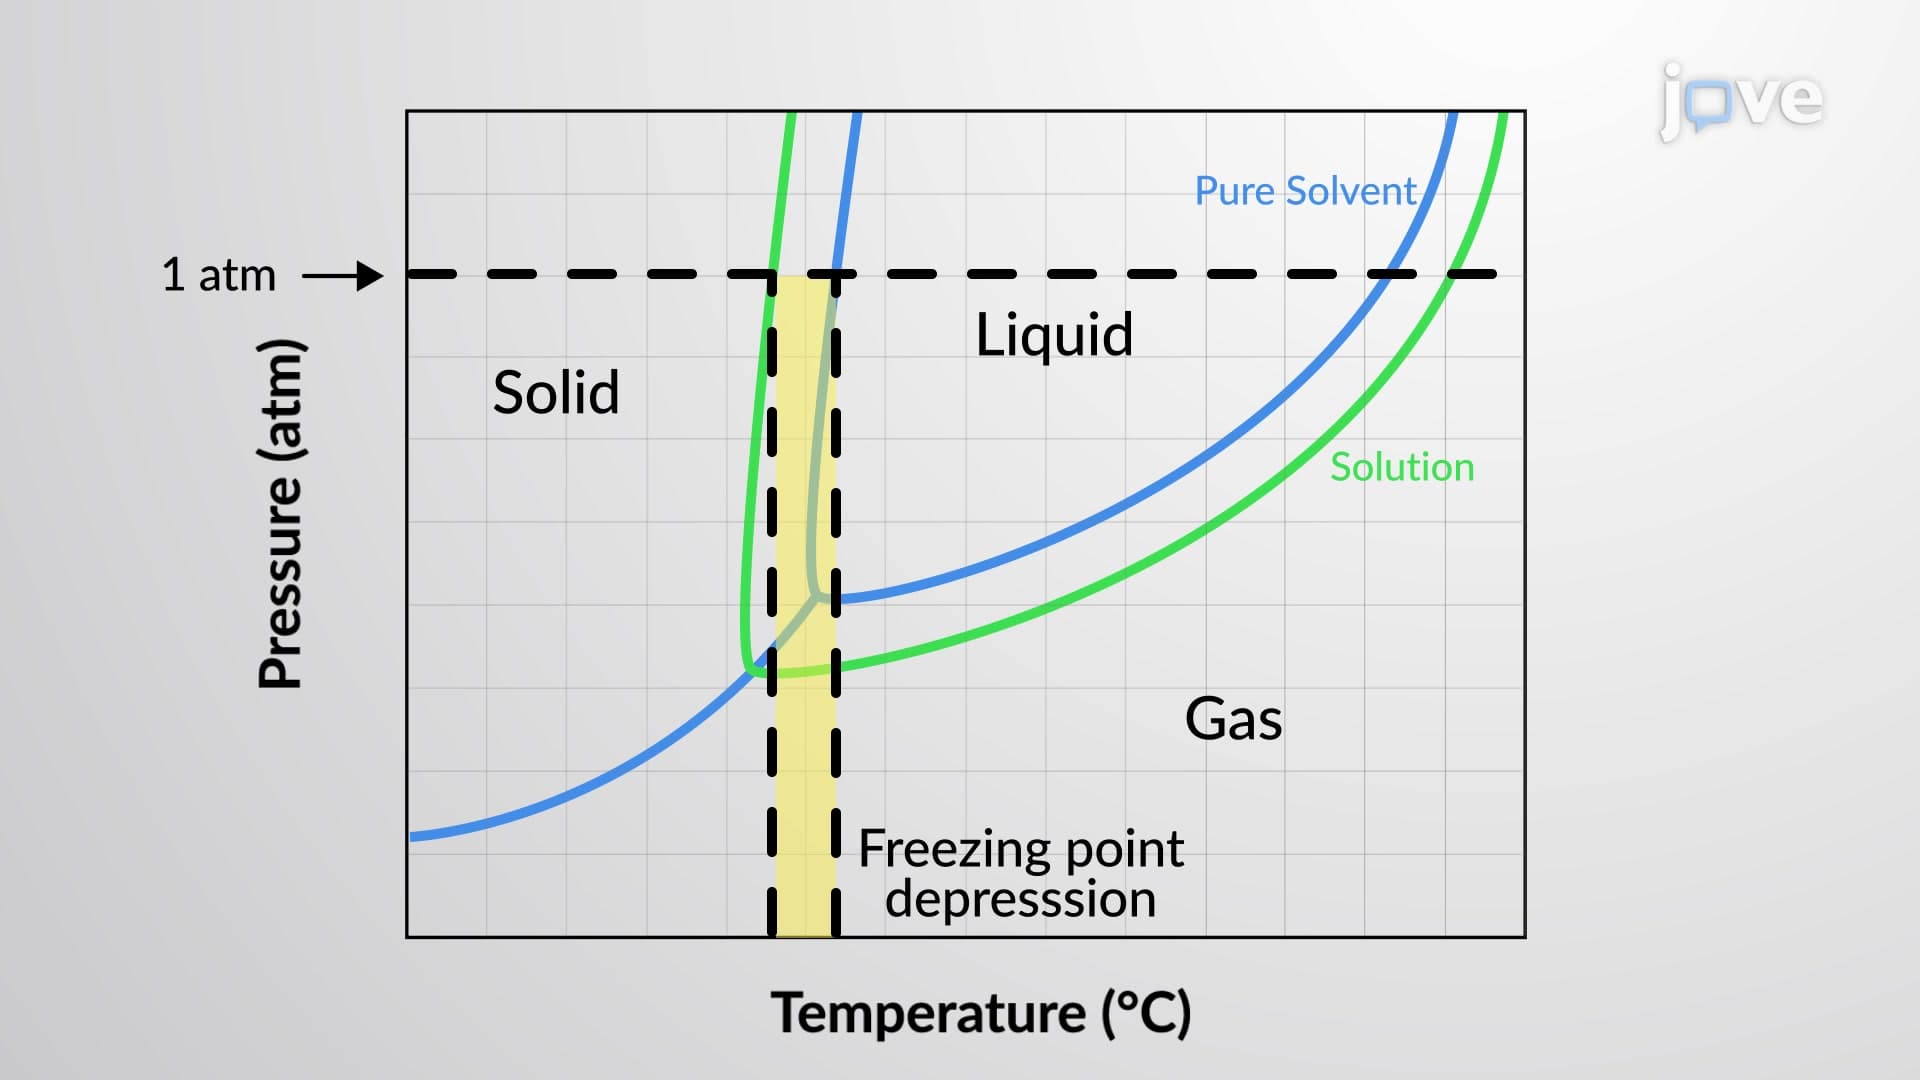
The boiling point of a liquid is the temperature at which its vapor pressure is equal to ambient atmospheric pressure. Since the vapor pressure of a solution is lowered due to the presence of nonvolatile solutes, it stands to reason that the solution’s boiling point will subsequently be increased. Vapor pressure increases with temperature, and so a solution will require a higher temperature than will pure solvent to achieve any given vapor pressure, including one...
 02:39
Phase Transitions: Melting and Freezing
 01:27
Inductive Effects on Chemical Shift: Overview
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
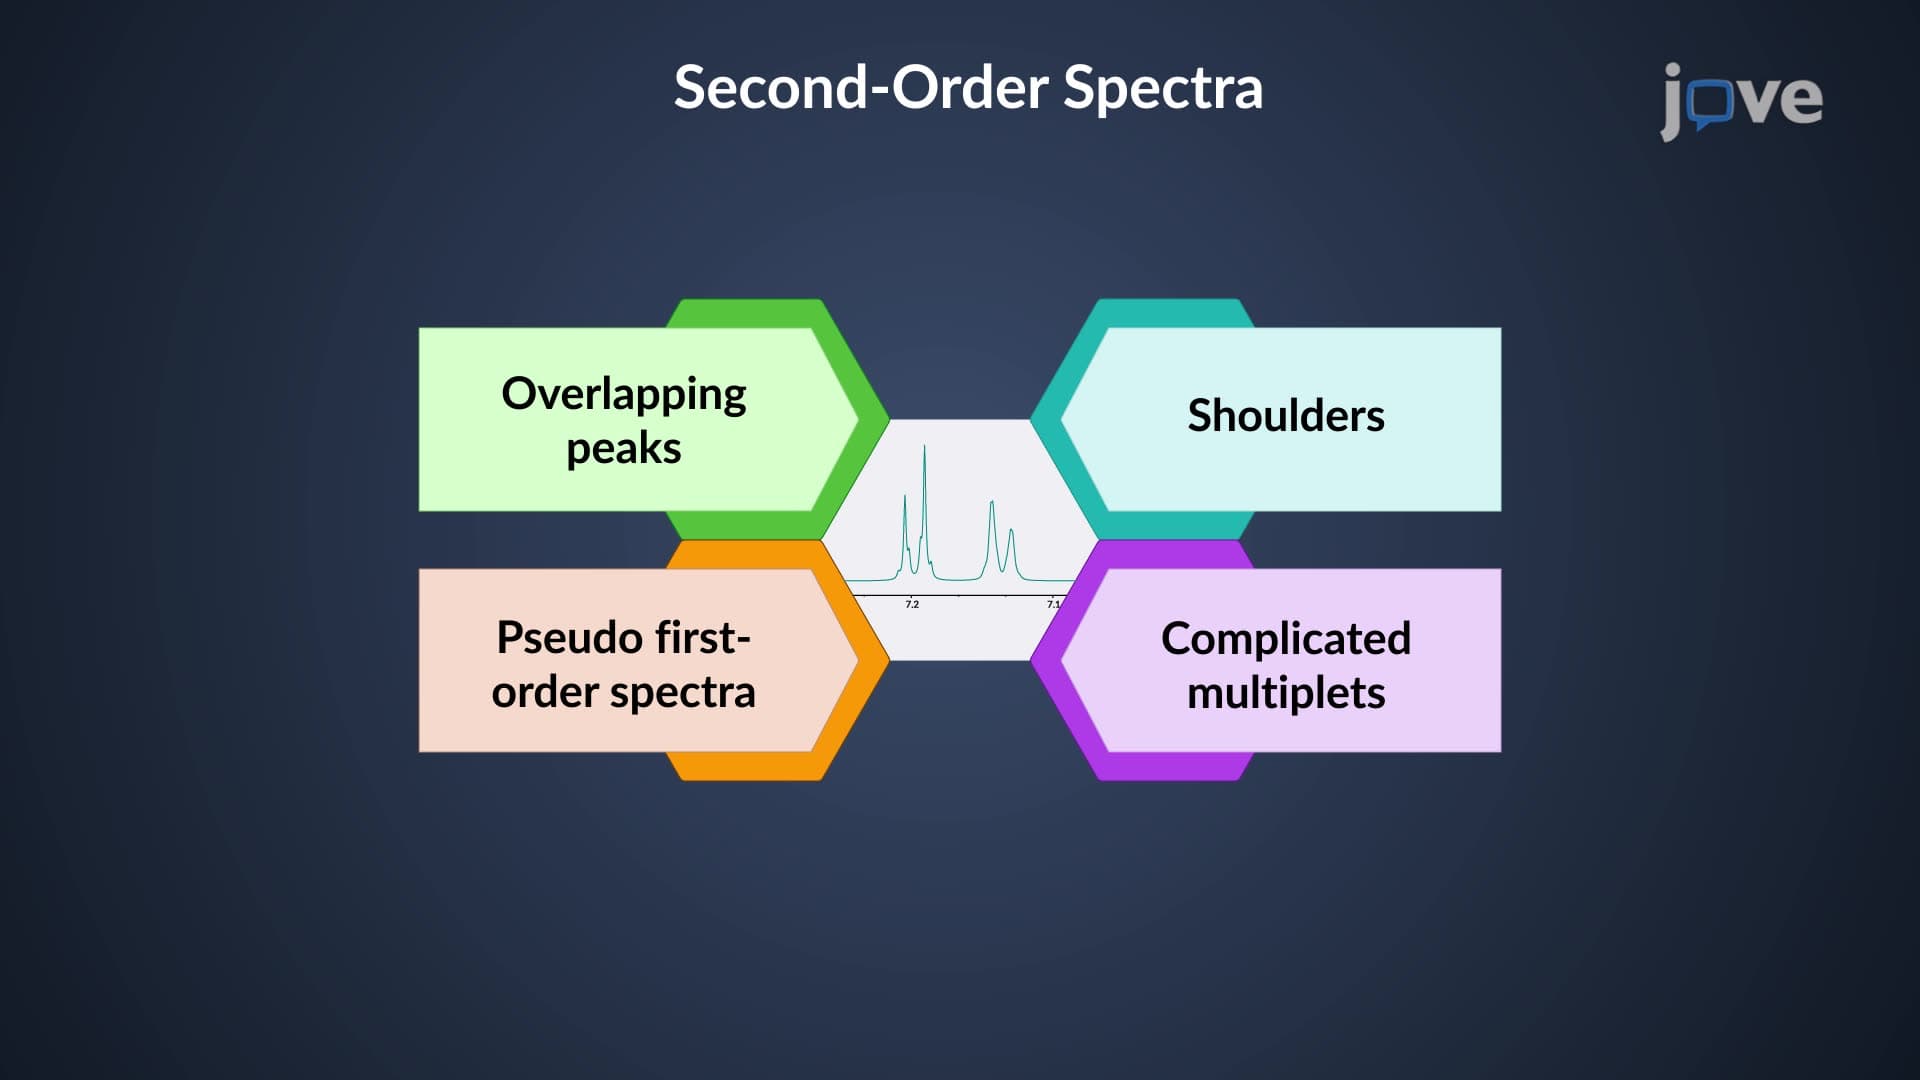
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...