Related Experiment Video
Updated: Aug 18, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A reliable ab initio potential energy surface and vibrational states for the ground electronic state of
1Institute of Theoretical and Computational Chemistry, Laboratory of Mesoscopic Chemistry, Nanjing University, Nanjing 210093, People' Republic of China.

Abstract:
A three-dimensional global potential energy surface for the ground (X (1)Sigma(+)(g))electronic state of HgH(2) is constructed from more than 13,00 ab initio points. These points are generated using an internally contracted multireference configuration interaction method with the Davidson correction and a large basis set. Low-lying vibrational energy levels of HgH(2), HHgD, and HgD(2) calculated using the Lanczos algorithm are found to be in good agreement with the available experimental band origins. The majority of the vibrational energy levels up to 9000 cm(-1) are assigned with normal mode quantum numbers. Our results indicate a gradual transition for the stretching vibrations from the normal mode regime at low energies to the local mode regime near 9000 and 8000 cm(-1) for HgH(2) and HgD(2), respectively, as evidenced by a decreasing energy gap between the (0,0,n(3)) and (1,0,n(3)-1) vibrational states and bifurcation of the corresponding wave functions.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 03:51
Molecular Orbital Theory II
 03:24
Hybridization of Atomic Orbitals I
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:13
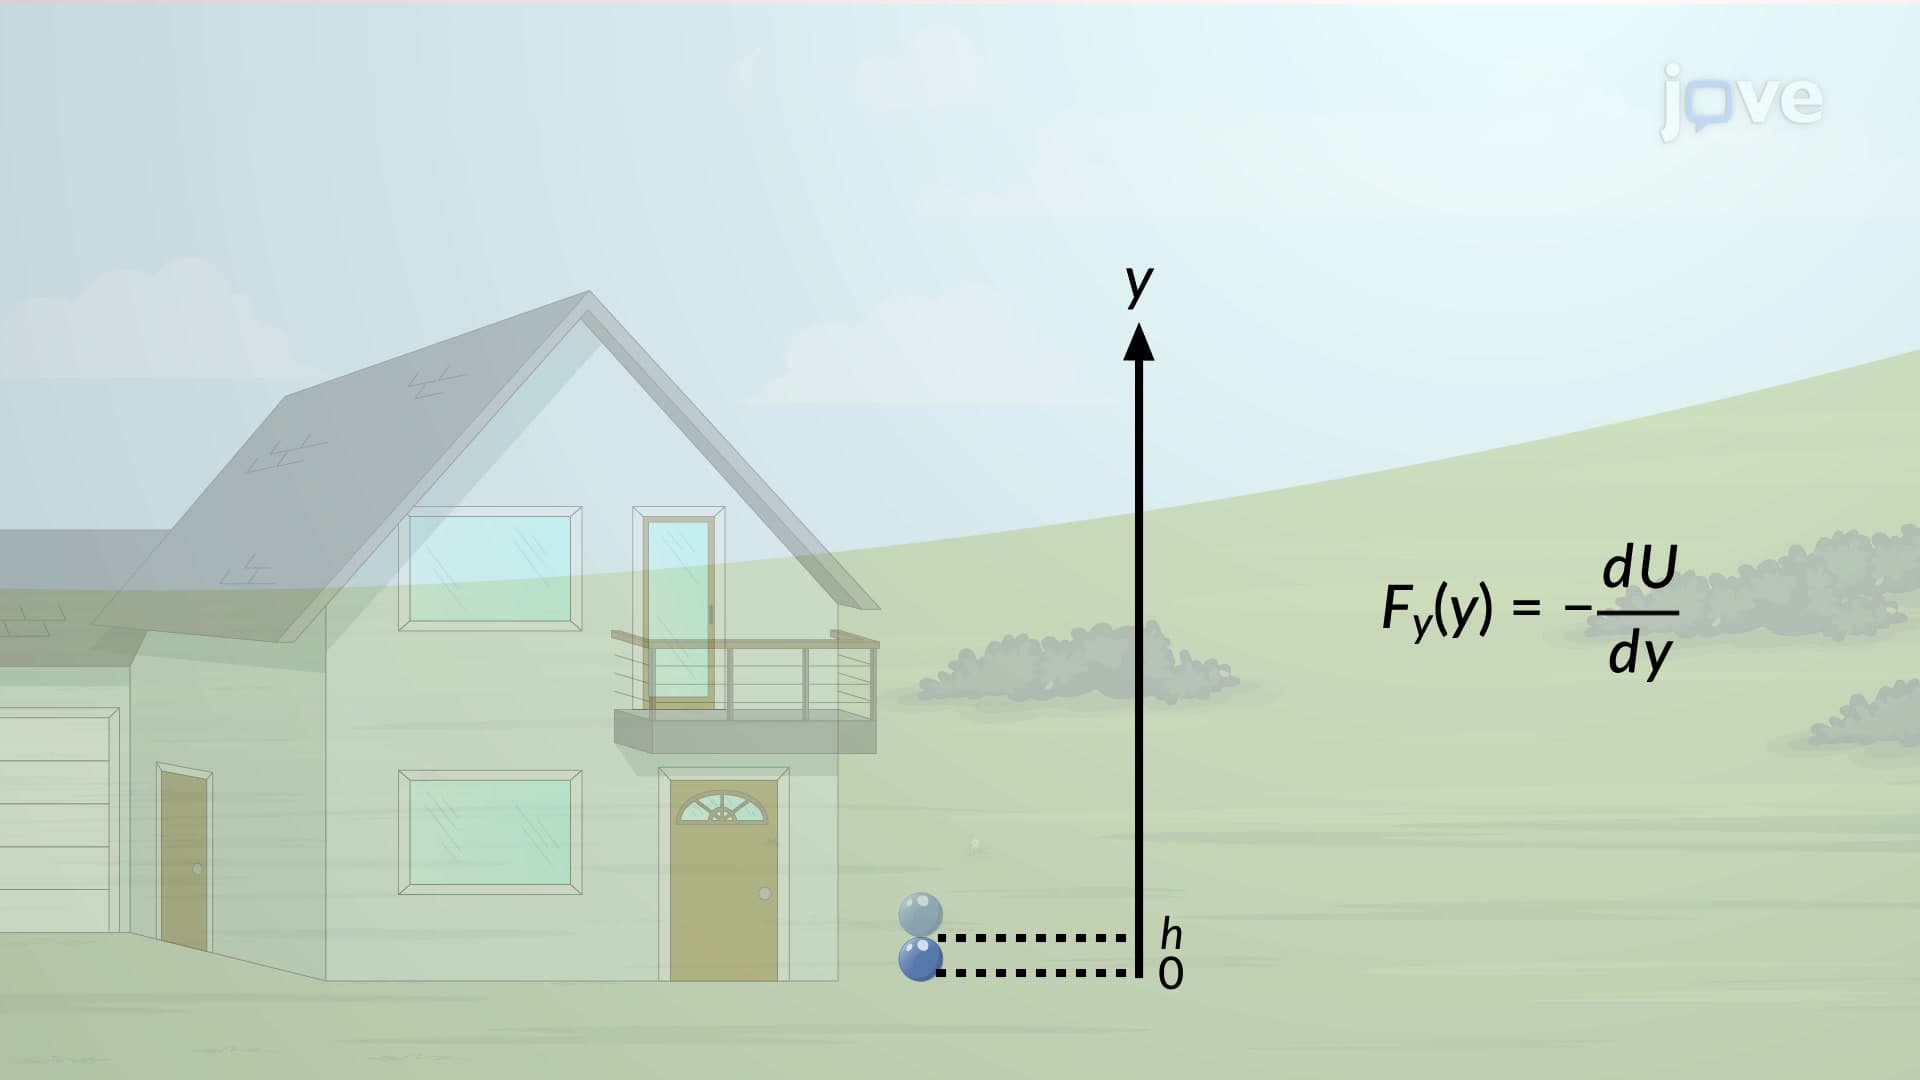
Force and Potential Energy in One Dimension
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 02:38
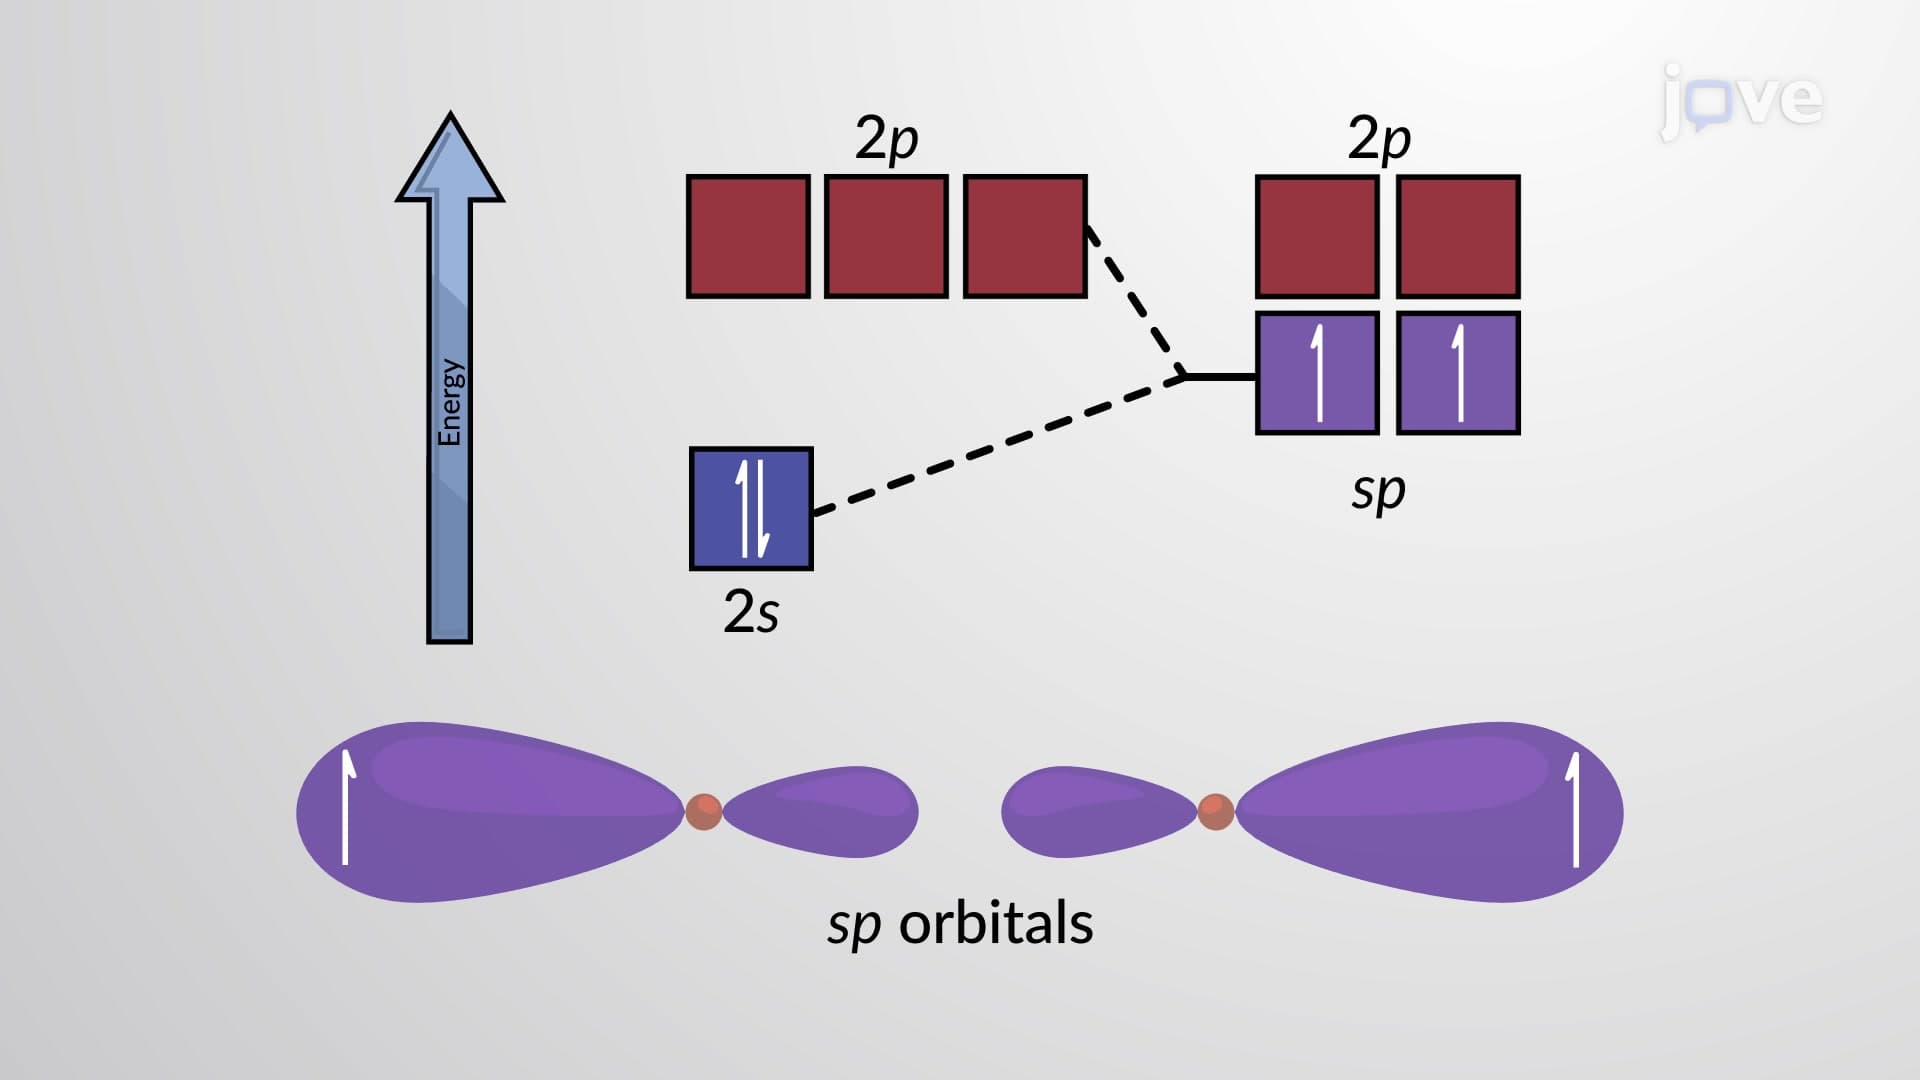
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...