Related Experiment Video
Updated: Aug 18, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Ab initio calculations of structures and interaction energies of toluene dimers including CCSD(T) level electron
Seiji Tsuzuki1, Kazumasa Honda, Tadafumi Uchimaru
1National Institute of Advanced Industrial Science and Technology, Tsukuba, Ibaraki 305-8568, Japan. s.tsuzuki@aist.go.jp

Abstract:
The intermolecular interaction energy of the toluene dimer has been calculated with the ARS-F model (a model chemistry for the evaluation of intermolecular interaction energy between ARomatic Systems using Feller's method), which was formerly called as the AIMI model III. The CCSD(T) (coupled cluster calculations with single and double substitutions with noniterative triple excitations) interaction energy at the basis set limit has been estimated from the second-order Moller-Plesset perturbation interaction energy at the basis set limit obtained by Feller's method and the CCSD(T) correction term obtained using a medium-size basis set. The cross (C(2)) dimer has the largest (most negative) interaction energy (-4.08 kcal/mol). The antiparallel (C(2h)) and parallel (C(S)) dimers (-3.77 and -3.41 kcal/mol, respectively) are slightly less stable. The dispersion interaction is found to be the major source of attraction in the toluene dimer. The dispersion interaction mainly determines the relative stability of the stacked three dimers. The electrostatic interaction of the stacked three dimers is repulsive. Although the T-shaped and slipped-parallel benzene dimers are nearly isoenergetic, the stacked toluene dimers are substantially more stable than the T-shaped toluene dimer (-2.62 kcal/mol). The large dispersion interaction in the stacked toluene dimers is the cause of their enhanced stability.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:36
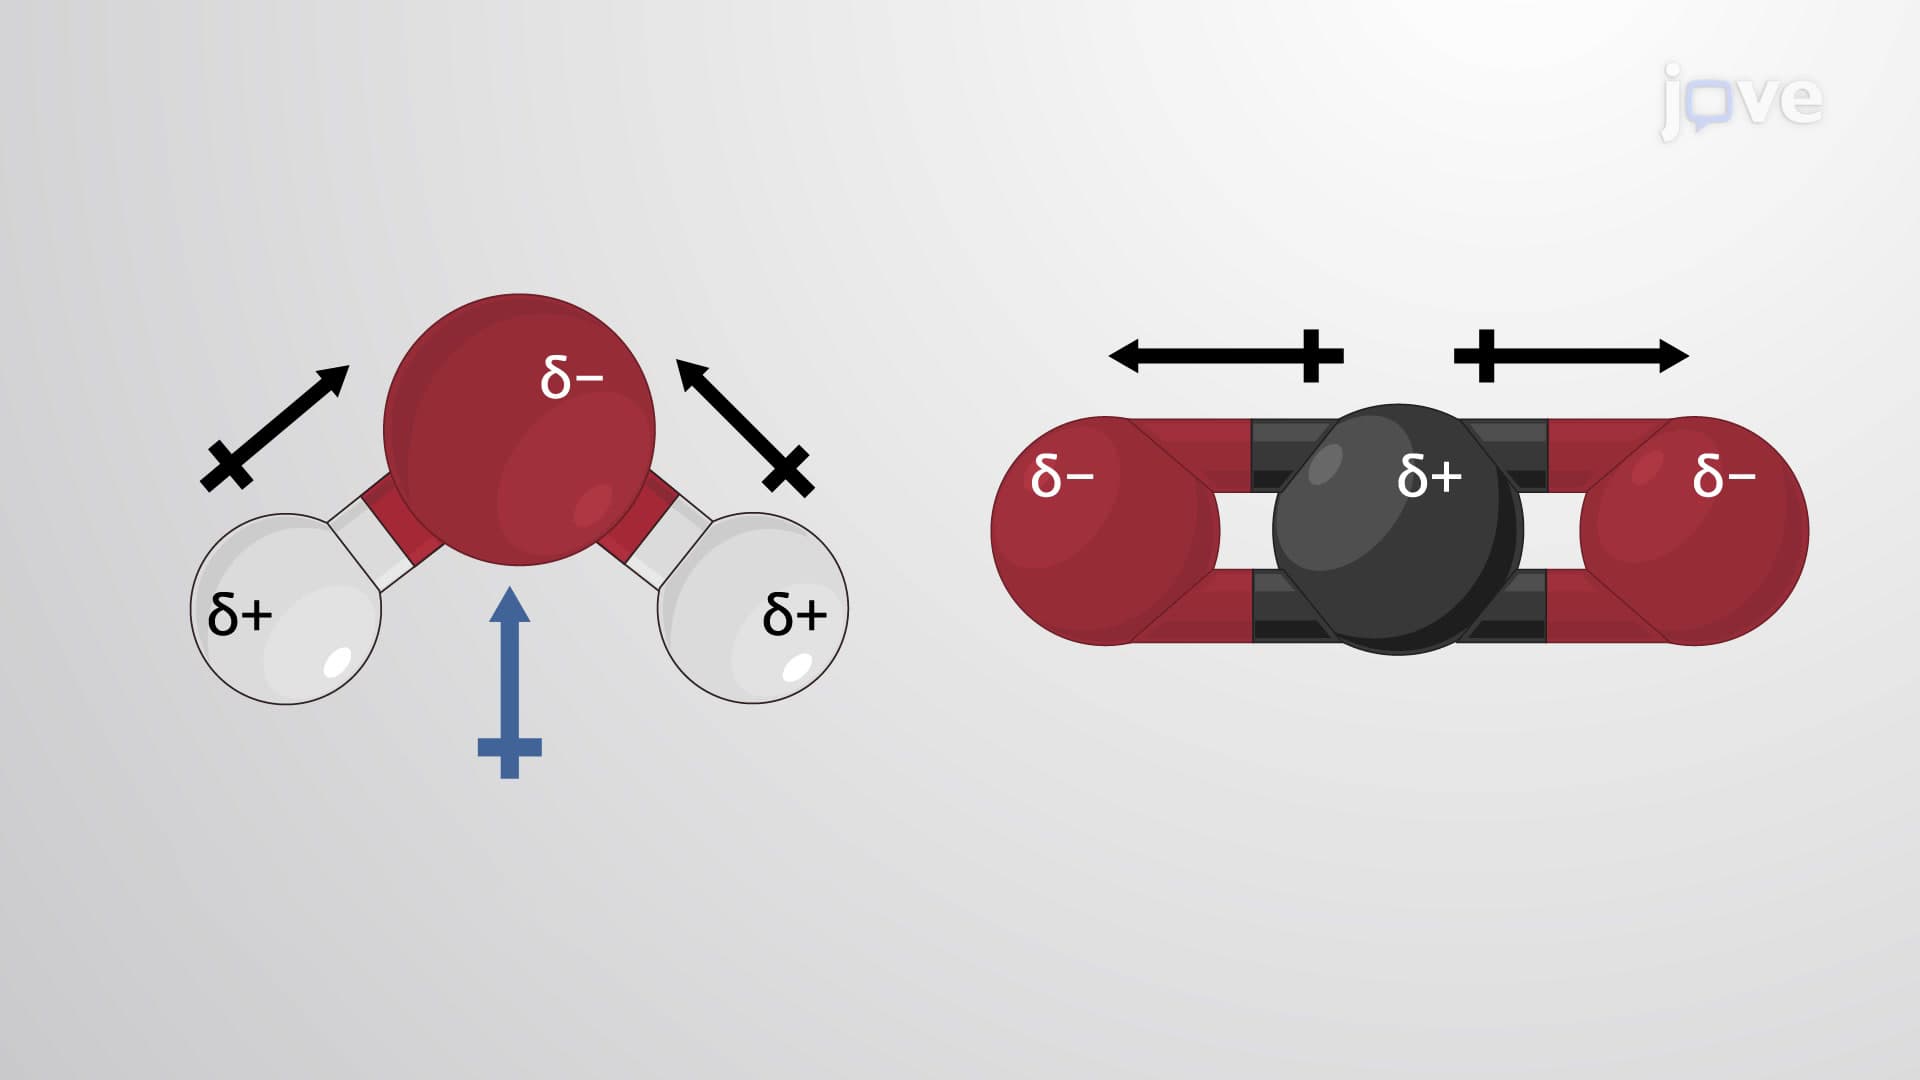
Molecular Geometry and Dipole Moments
 02:27
Predicting Molecular Geometry
 01:18
Structure of Benzene: Molecular Orbital Model
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:07
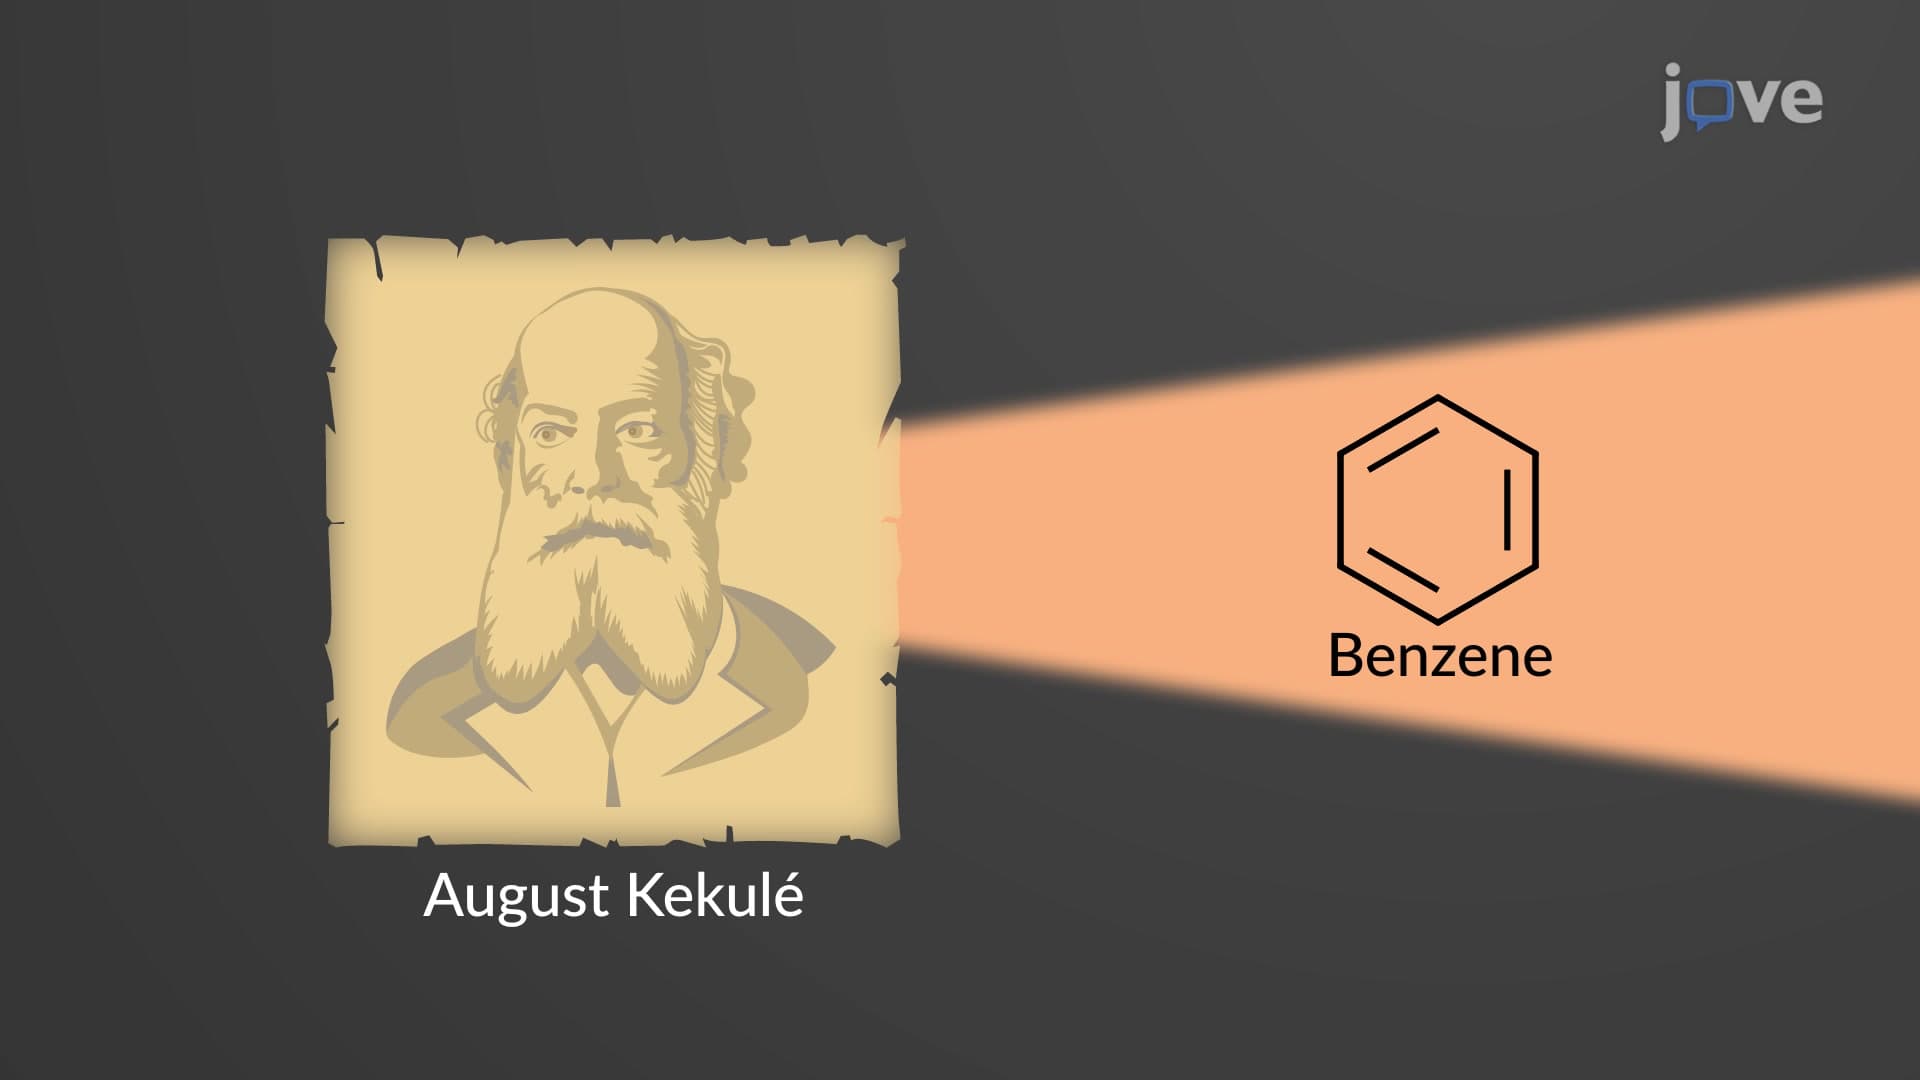
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.