A consistent set of statistical potentials for quantifying local side-chain and backbone interactions
Related Experiment Video
Updated: Aug 18, 2026
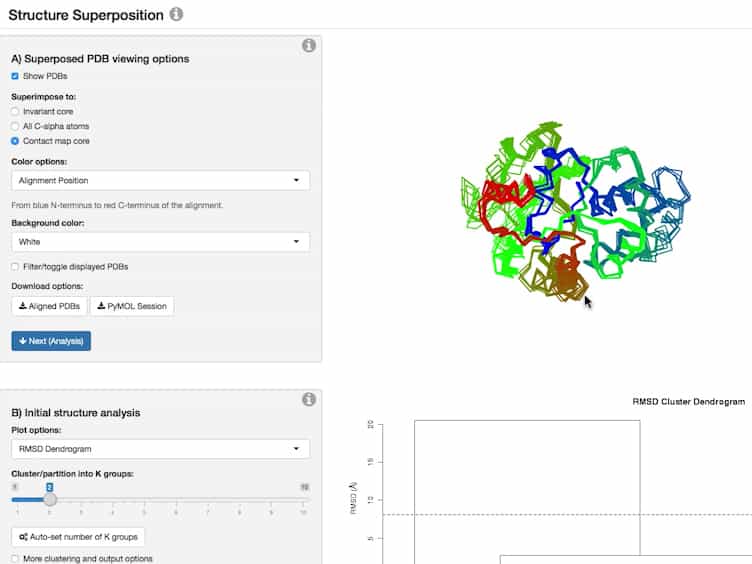
Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
1Department of Biological Chemistry, Johns Hopkins University School of Medicine, Baltimore, Maryland 21205, USA.

Abstract:
The frequencies of occurrence of atom arrangements in high-resolution protein structures provide some of the most accurate quantitative measures of interaction energies in proteins. In this report we extend our development of a consistent set of statistical potentials for quantifying local interactions between side-chains and the polypeptide backbone, as well as nearby side-chains. Starting with phi/psi/chi1 propensities that select for optimal interactions of the 20 amino acid side-chains with the 2 flanking peptide bonds, the following 3 new terms are added: (1) a distance-dependent interaction between the side-chain at i and the carbonyl oxygens and amide protons of the peptide units at i +/- 2, i +/- 3, and i +/- 4; (2) a distance-dependent interaction between the side-chain at position i and side-chains at positions i + 1 through i + 4; and (3) an orientation-dependent interaction between the side-chain at position i and side-chains at i + 1 through i + 4. The relative strengths of these 4 pseudo free energy terms are estimated by the average information content of each scoring matrix and by assessing their performance in a simple fragment threading test. They vary from -0.4 - -0.5 kcal/mole per residue for phi/psi/chi1 propensities to a range of -0.15 - -0.6 kcal/mole per residue for each of the other 3 terms. The combined energy function, containing no interactions between atoms more than 4 residues apart, identifies the correct structural fragment for randomly selected 15 mers over 40% of the time, after searching through 232,000 alternative conformations. For 14 out of 20 sets of all-atom Rosetta decoys analyzed, the native structure has a combined score lower than any of the 1700-1900 decoy conformations. The ability of this energy function to detect energetically important details of local structure is demonstrated by its power to distinguish high-resolution crystal structures from NMR solution structures.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:35
Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
02:35Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
 01:57
Protein Complexes with Interchangeable Parts
The SCF ubiquitin ligase is a protein complex of five individual proteins. This complex attaches ubiquitin to other target proteins to mark them for degradation. In order to...
01:57Protein Complexes with Interchangeable Parts
The SCF ubiquitin ligase is a protein complex of five individual proteins. This complex attaches ubiquitin to other target proteins to mark them for degradation. In order to...