Evaluating molecular similarity using reduced representations of the electron density
Related Experiment Video
Updated: Aug 18, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Nathalie Meurice1, Gerald M Maggiora, Daniel P Vercauteren
1Department of Pharmacology and Toxicology, University of Arizona, College of Pharmacy, 1703 E. Mable, Tucson, AZ 85721, USA. meurice@pharmacy.arizona.edu

Abstract:
A model system of four benzodiazepine-like ligands for the central benzodiazepine receptors (CBRs) and peripheral benzodiazepine receptors (PBRs)is examined using a genetic algorithm procedure (GAGS) designed for evaluating molecular similarity. The method is based on the alignment of reduced representations generated from the critical points of the electron density computed at medium crystallographic resolution. The results are further characterized by a comparison with alignments produced by MIMIC, a field-based superimposition method that matches both steric and electrostatic molecular fields. The alignments produced by the two methods are generally seen to be consistent. The relationships of the compounds' binding affinities for both CBRs and PBRs to the alignments determined by GAGS yield a set of structural features required for significant binding to benzodiazepine receptors. Benefits of using reduced representations for evaluating molecular similarities and for constructing pharmacophore models are discussed.
Related Concept Videos
 01:18
Molecular Shapes
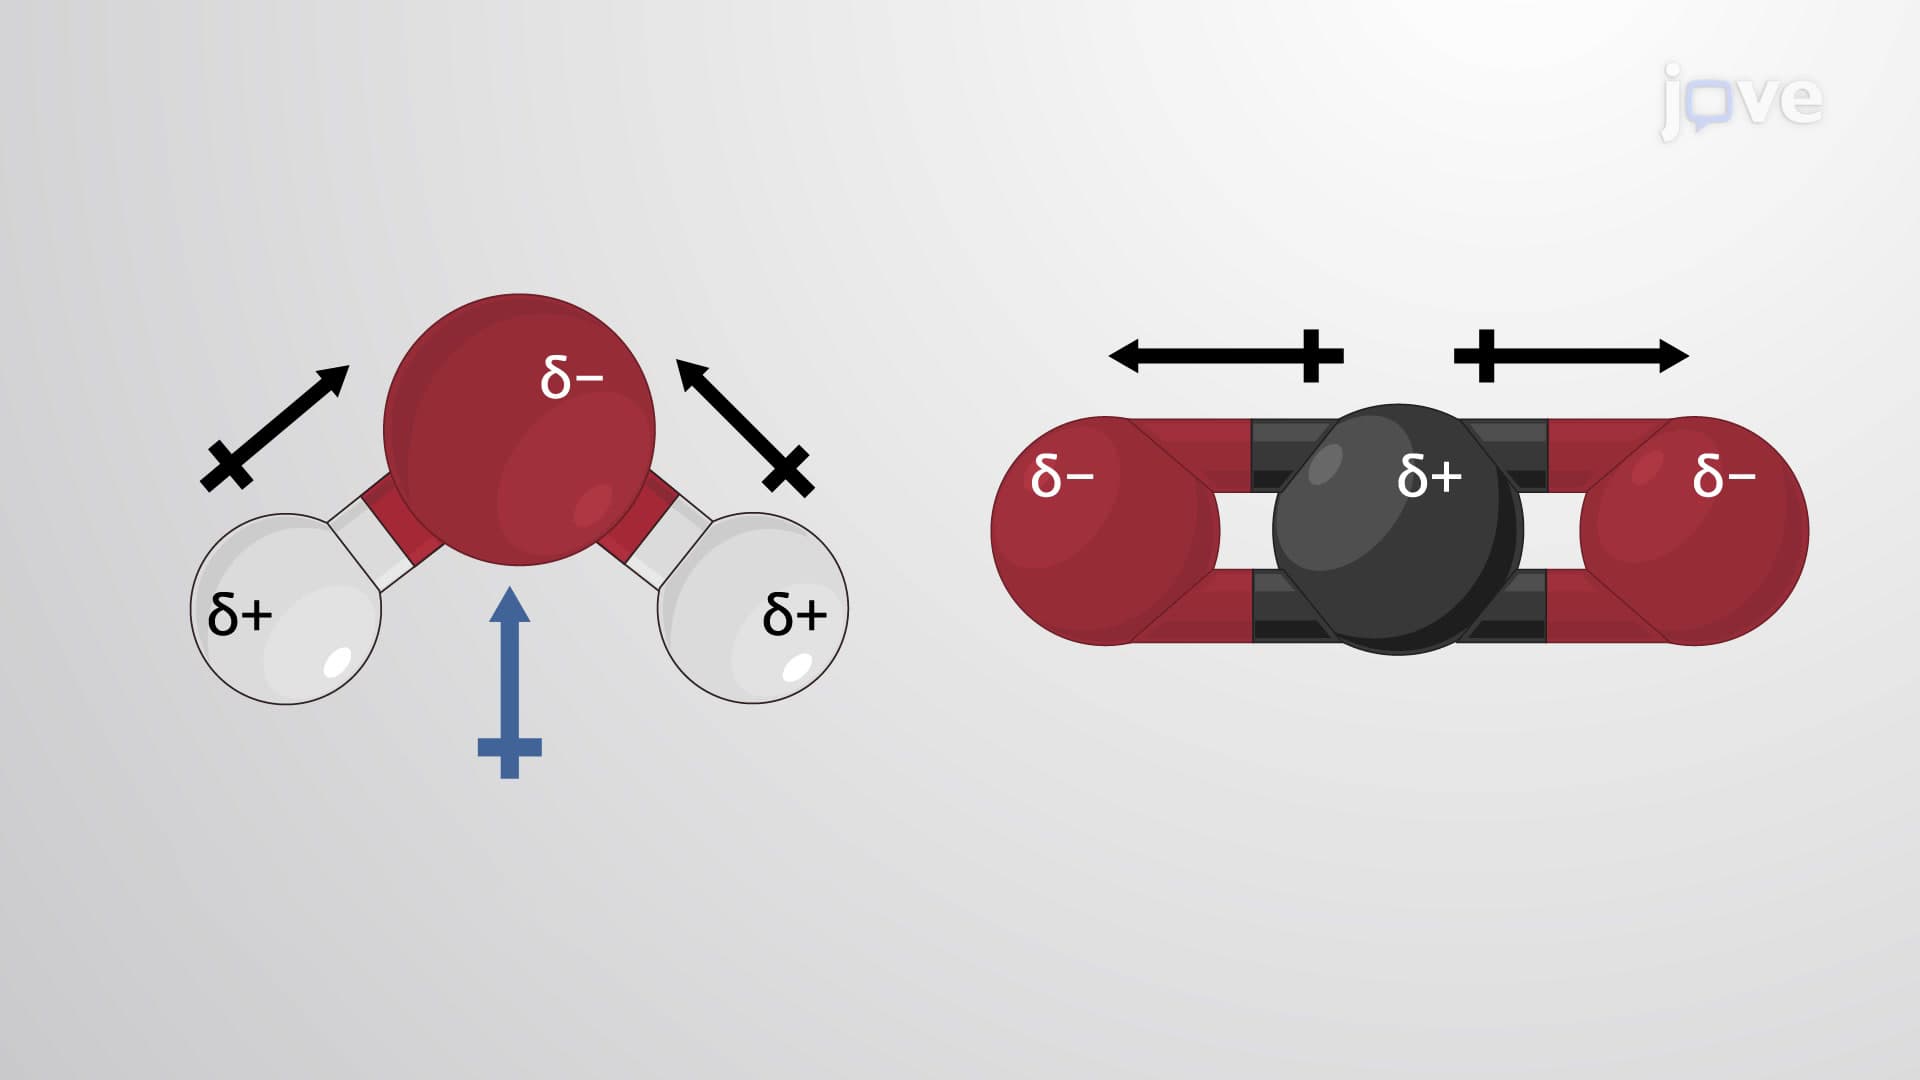 02:36
Molecular Geometry and Dipole Moments
 02:00
Molecular Models
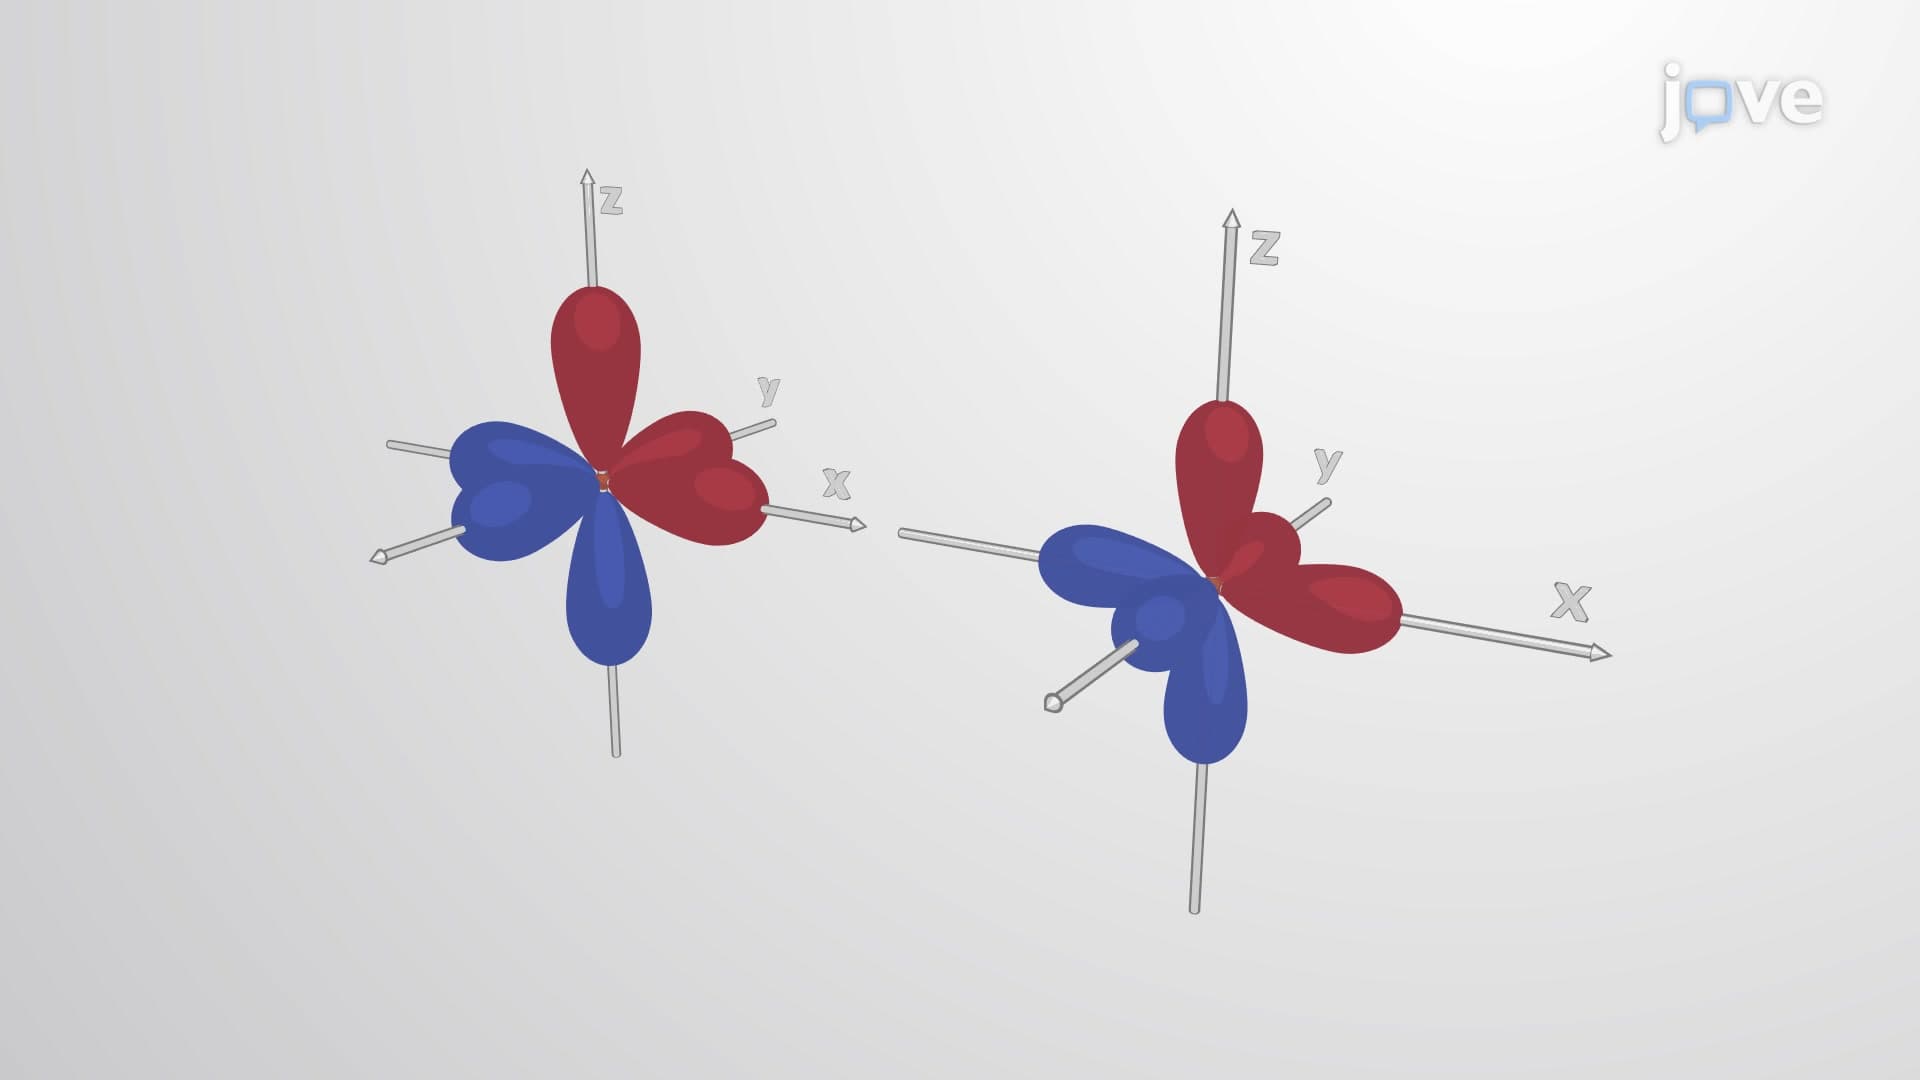 02:35
Molecular Orbital Theory I
 02:37
VSEPR Theory
 02:52
VSEPR Theory and the Basic Shapes