

Functional and basis set dependence of K-edge shake-up spectra of molecules
Barbara Brena1, Stéphane Carniato, Yi Luo
1Theoretical Chemistry, Royal Institute of Technology, SCFAB, SE-10691 Stockholm, Sweden.

Abstract:
A straightforward approach for computing the K-edge shake-up spectra of molecules based on equivalent core-hole linear response theory at both Hartree-Fock and density functional theory levels is proposed. Benchmark calculations have been performed to explore its sensitivity to different types of functionals and basis sets for the carbon 1s shake-up spectra of benzene and metal-free phthalocyanine (H2Pc). A very good agreement with previous theoretical and experimental works for the benzene molecule has been obtained for all the functionals and basis sets tested. Electron correlation is found to be essential for a good description of the H2Pc system, whose experimental C 1s shake-up spectrum is best reproduced by the hybrid density functional.
More Related Videos


Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...
 01:15
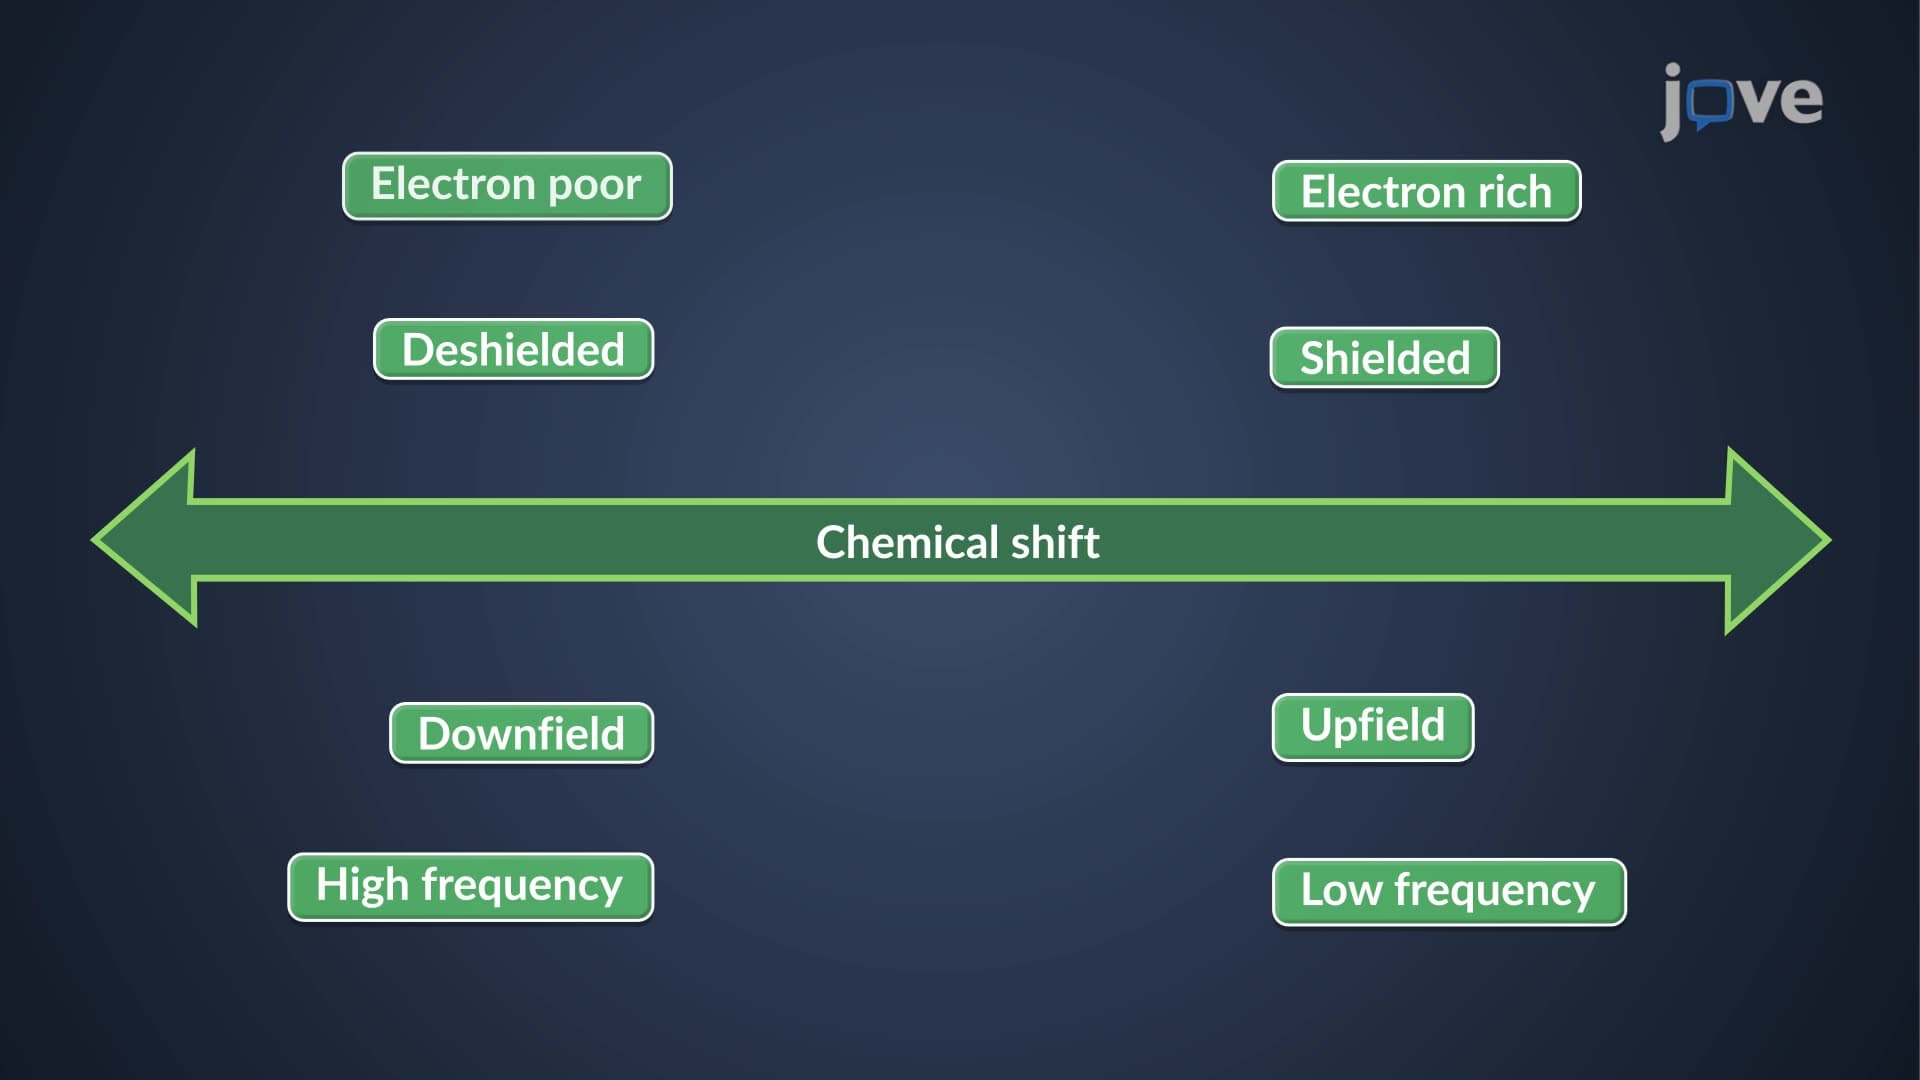
NMR Spectroscopy: Chemical Shift Overview
For instance, the proton...
 01:27
π Electron Effects on Chemical Shift: Overview
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions