

Comparative studies of the spectroscopy of CuCl2: DFT versus standard ab initio approaches
A Ramírez-Solís1, R Poteau, A Vela
1Laboratoire de Physique Quantique, IRSAMC, UMR 5626 du CNRS, Université Paul Sabatier, 118, Route de Narbonne, Toulouse, Cedex 31062, France. alex@servm.fc.uaem.mx

Abstract:
The X2Pi g-2Sigma g+, X2Pi g-2Delta g, X2Pi g-2Sigma u+, X2Pi g-2Pi u transitions on CuCl2 have been studied using several exchange-correlation functionals from the various types of density functional theory (DFT) approaches like local density approximation (LDA), generalized gradient approximation (GGA), hybrid and meta-GGA. The results are compared with the experience and with those coming from the most sophisticated nondynamic and dynamic electronic correlation treatments using the same relativistic effective core potentials and especially developed basis sets to study the electronic structure of the five lowest states and the corresponding vertical and adiabatic transition energies. The calculated transition energies for three of the hybrid functionals (B3LYP, B97-2, and PBE0) are in very good agreement with the benchmark ab initio results and experimental figures. All of the other functionals largely overestimate the X2Pi g-2Sigma g+ and X2Pi g-2Delta g transition energies, many of them even placing the 2Delta g ligand field state above the charge transfer 2Pi u and 2Sigma u+ states. The relative weight of the Hartree-Fock exchange in the definition of the functional used appears to play a key role in the accurate description of the LambdaSSigma density defined by the orientation of the 3d hole (sigma, pi, or delta) on Cu in the field of both chlorine atoms, but no simple connection of this weight with the quality of the spectra has been found. Mulliken charges and spin densities are carefully analyzed; a possible link between the extent of spin density on the metal for the X2Pi g state and the performance of the various functionals was observed, suggesting that those that lead to the largest values (close to 0.65) are the ones that best reproduce these four transitions. Most functionals lead to a remarkably low ionicity for the three ligand field states even for the best performing functionals, compared to the complete active space (SCF) (21, 14) ab initio values. These findings show that not only large variational ab initio calculations can produce reliable spectroscopic results for extremely complex systems where delicate electronic correlation effects have to be carefully dealt with. However, those functionals that were recently shown to perform best for a series of molecular properties [J. Chem. Phys. 121 3405 (2004)] are not the ones that produce the best transition energies for this complex case.

Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:29
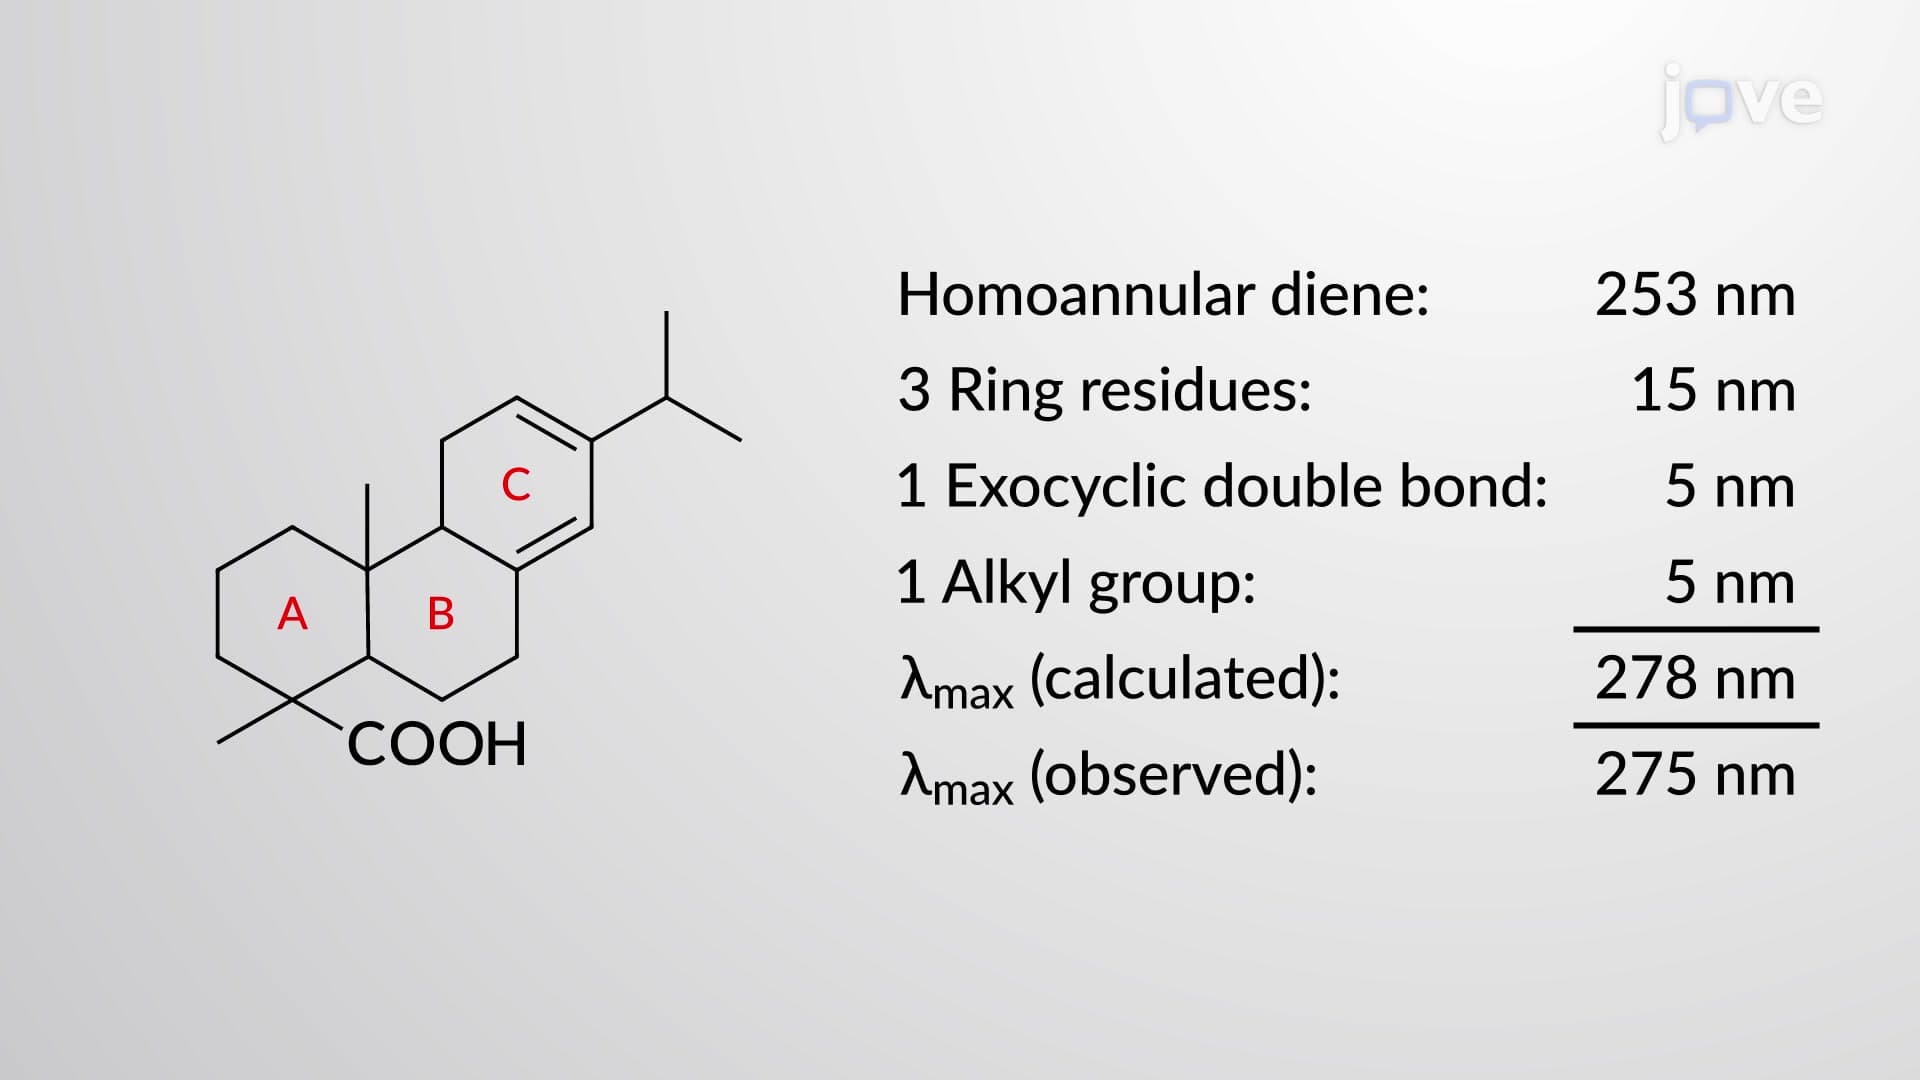
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:27
Atomic Absorption Spectroscopy: Overview
When irradiated by EMR of a particular wavelength, these...
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent of conjugation in the...
 01:21
Atomic Absorption Spectroscopy: Lab
Solutions containing organic solvents, such as low-molecular-mass alcohols, esters, or ketones, enhance absorbances by increasing nebulizer...