Related Experiment Video
Updated: Aug 17, 2026
Determining the Serum Stability of Human Adenosine Deaminase 1 Enzyme
Published on: September 27, 2024
QSAR of adenosine A3 receptor antagonist 1,2,4-triazolo[4,3-a]quinoxalin-1-one derivatives using chemometric tools
Prosenjit Bhattacharya1, Kunal Roy
1Drug Theoretics and Cheminformatics Lab, Division of Medicinal and Pharmaceutical Chemistry, Department of Pharmaceutical Technology, Jadavpur University, Kolkata 700 032, India.

Abstract:
Considering the potential of selective adenosine A3 receptor subtype ligands in the development of prospective therapeutic agents, an attempt has been made to explore physicochemical requirements of 1,2,4-triazolo[4,3-a]quinoxalin-1-one derivatives for A3 receptor binding. In this study, lipophilicity (logP), physicochemical substituent constants (pi, MR, sigma p) of phenyl ring substituents, and Wang-Ford charges of common atoms of the quinoxaline nucleus (calculated from molecular electrostatic potential surface of energy-minimized geometry using AM1 technique) were used as independent variables along with suitable dummy parameters. The best multiple linear regression (MLR) equation obtained from factor analysis (FA-MLR) as the preprocessing step could explain and predict 72.6% and 65.3%, respectively, of the variance of the binding affinity. The same equation also emerged as the best equation in the population of 100 equations obtained from genetic function approximation (GFA-MLR). The results suggested that presence of an electron-withdrawing group at the para position of the phenyl ring would be favorable for the binding affinity. Again, the presence of a nitro group at position R1 increases the binding affinity. When factor scores were used as predictor variables in the principal component regression analysis, the resultant model showed 78.6% explained variance and 63.1% predicted variance. The best equation derived from G/PLS could explain and predict 74.4% and 64.8%, respectively. The results have suggested the importance of Wang-Ford charges of atoms C15 and C19, apart from positive contributions of electron-withdrawing para substituents of the variance of the phenyl ring and nitro group at the R1 position.

Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:17
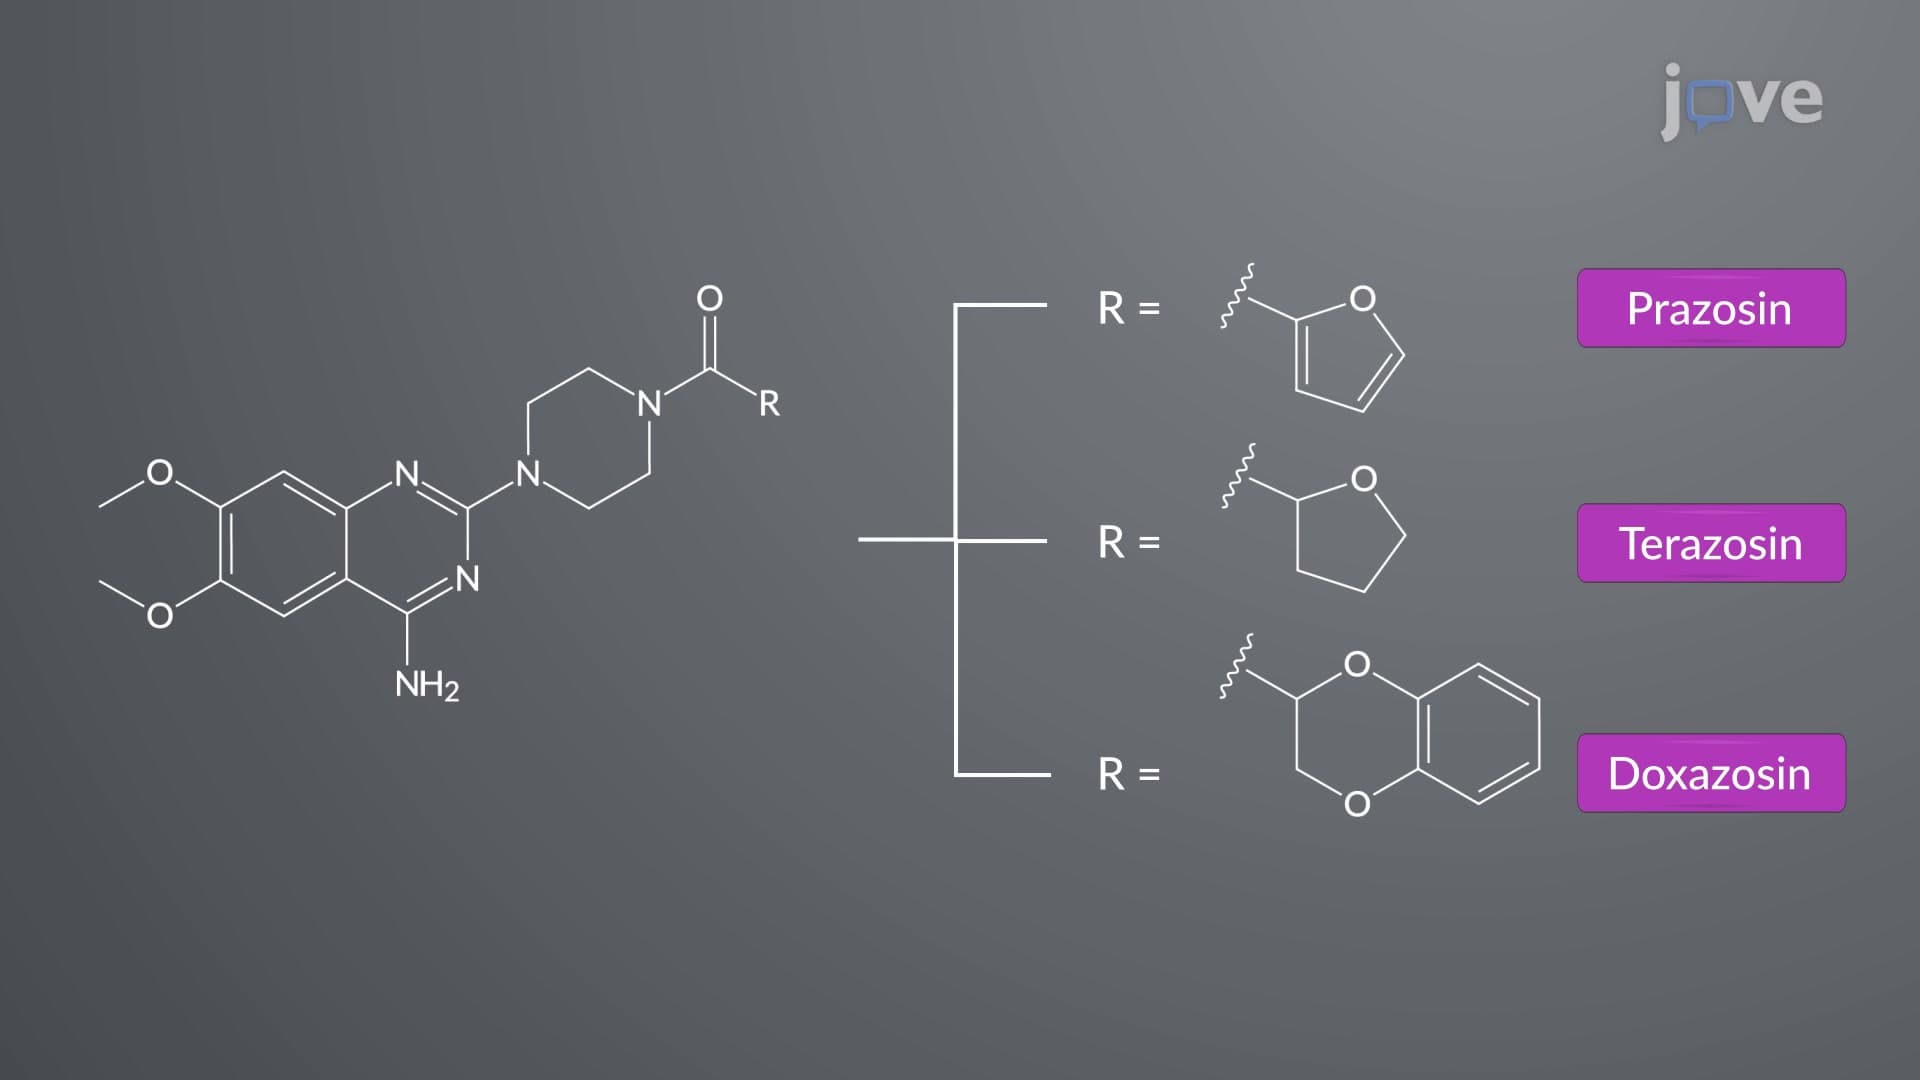
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:31
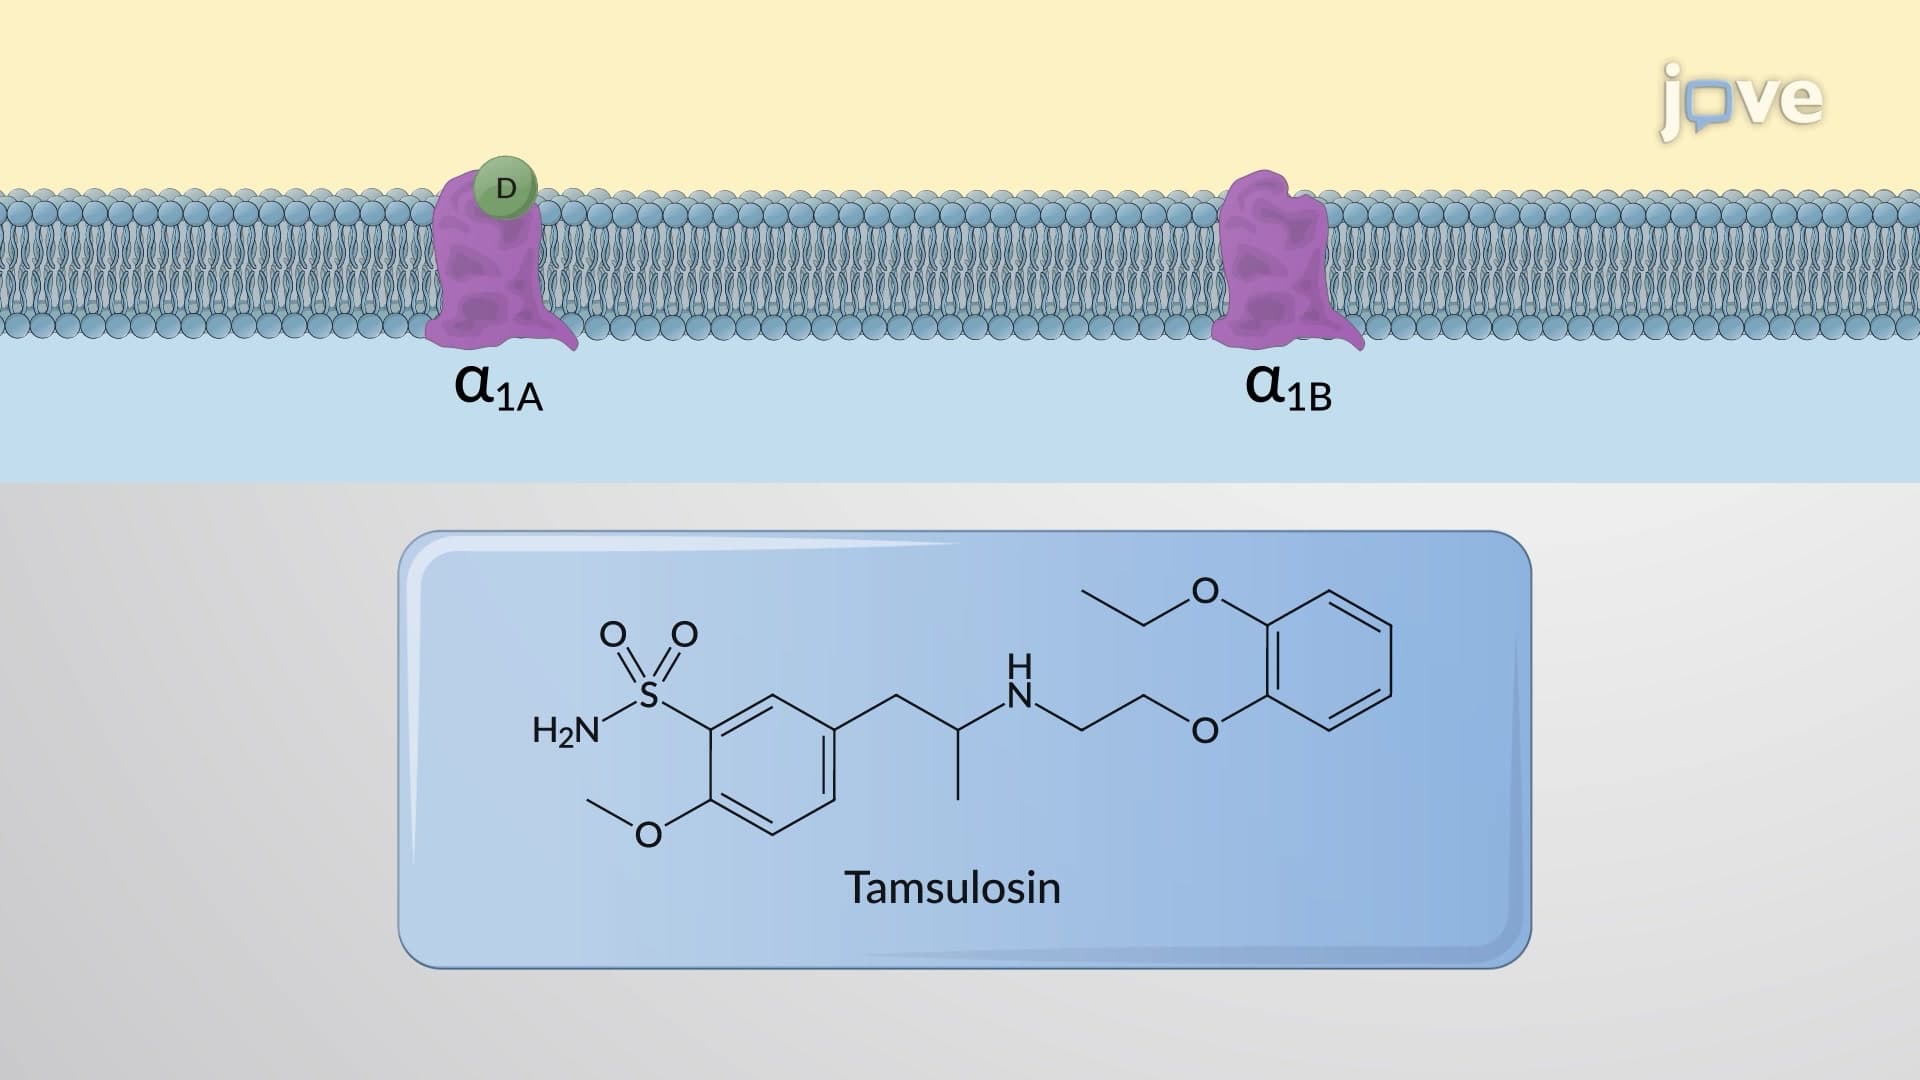
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:22
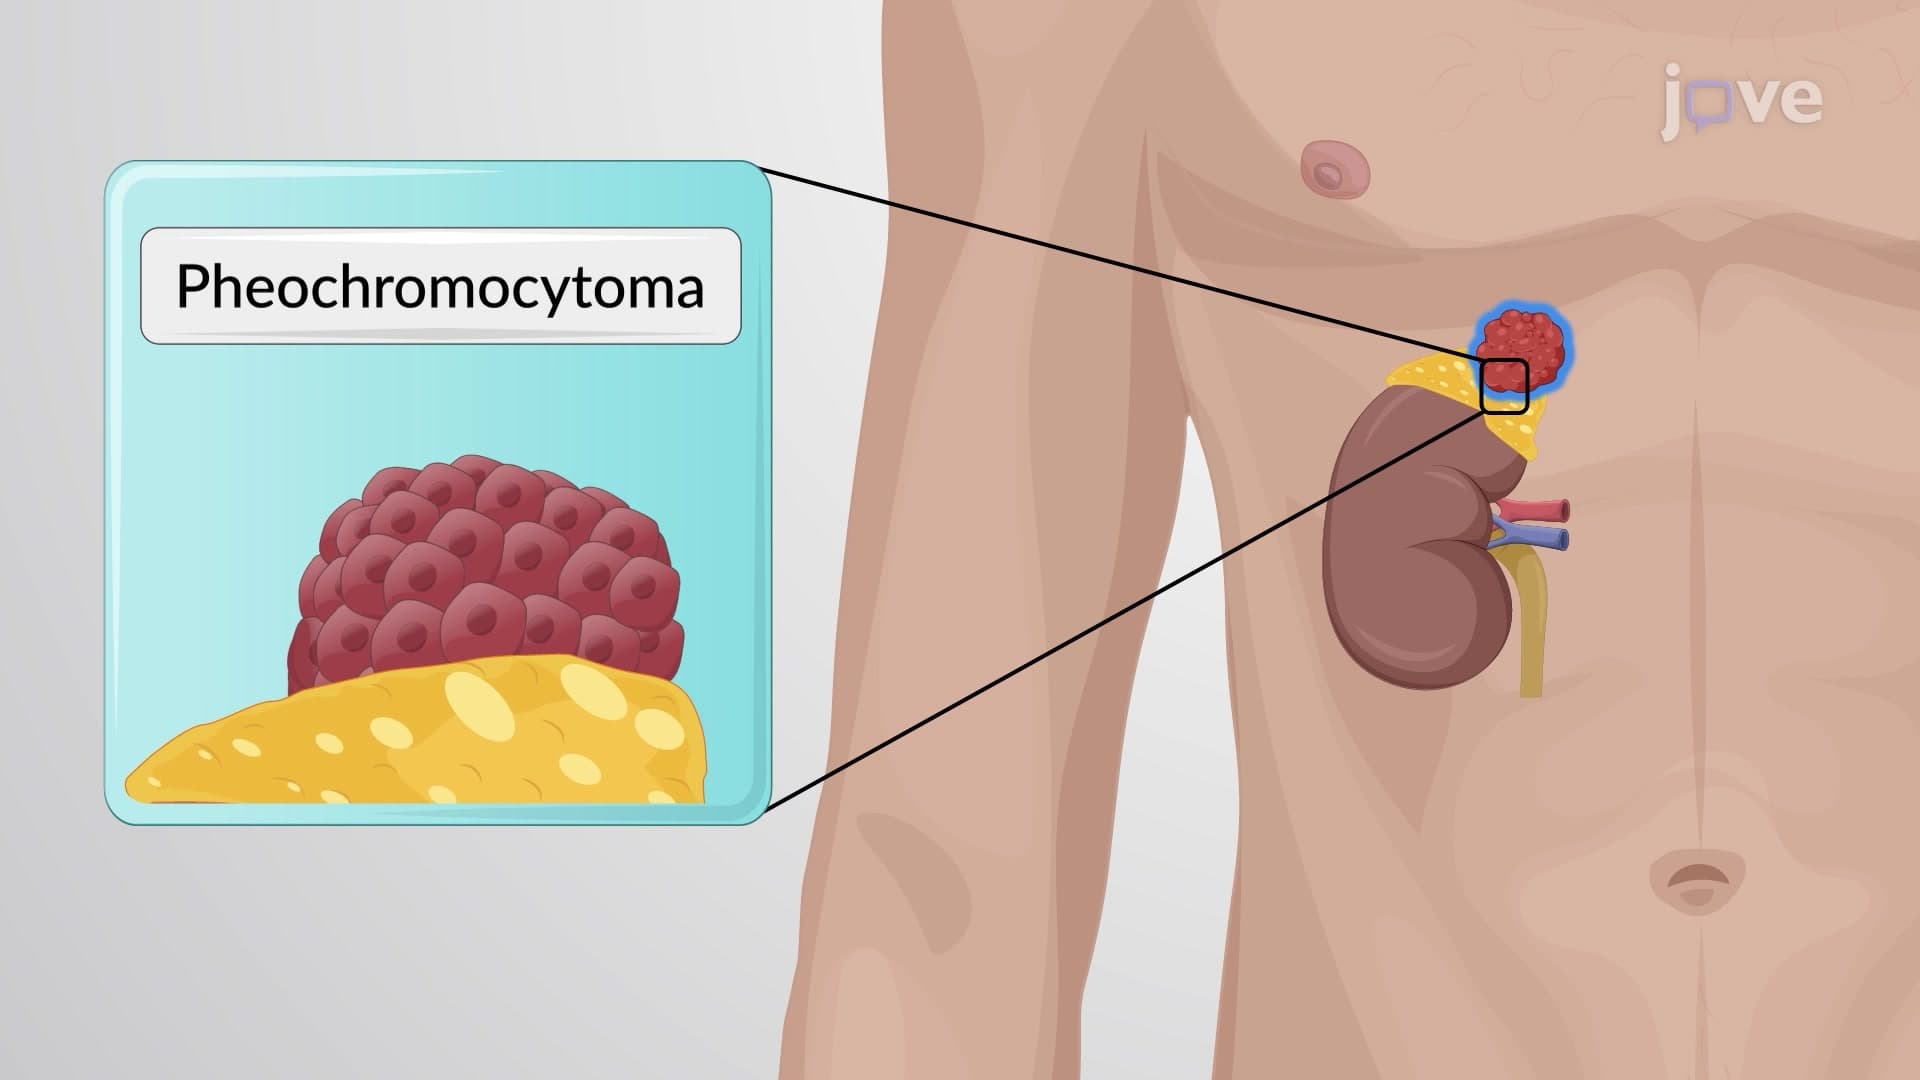
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:25
Adrenergic Antagonists: Chemistry and Classification of β-Receptor Blockers