Related Experiment Video
Updated: Aug 17, 2026

Fractionation for Resolution of Soluble and Insoluble Huntingtin Species
Published on: February 27, 2018
Mutant huntingtin represses CBP, but not p300, by binding and protein degradation
Shu-Yan Cong1, Barry A Pepers, Bernd O Evert
1CBG-Center of Human and Clinical Genetics, Leiden University Medical Center, Leiden, The Netherlands.

Abstract:
Huntington's disease can be used as a model to study neurodegenerative disorders caused by aggregation-prone proteins. It has been proposed that the entrapment of transcription factors in aggregates plays an important role in pathogenesis. We now report that the transcriptional activity of CBP is already repressed in the early time points by soluble mutant huntingtin, whereas the histone acetylase activity of CBP/p300 is gradually diminished over time. Mutant huntingtin bound much stronger to CBP than normal huntingtin, possibly contributing to repression. Especially at the later time points, CBP protein level was gradually reduced via the proteasome pathway. In sharp contrast, p300 was unaffected by mutant huntingtin. This selective degradation of CBP was absent in spinocerebellar ataxia 3. Thus, mutant huntingtin specifically affects CBP and not p300 both at the early and later time points, via multiple mechanisms. In addition to the reduction of CBP, also the altered ratio of these closely related histone acetyltransferases may affect chromatin structure and transcription and thus contribute to neurodegeneration.
Insights
Mutant huntingtin selectively degrades CBP, a key protein, impacting transcription and contributing to neurodegeneration in Huntington's disease models. This selective protein loss differs from other neurodegenerative conditions.
Area of Science:
- Neuroscience
- Molecular Biology
- Genetics
Background:
- Huntington's disease (HD) serves as a model for protein aggregation-related neurodegenerative disorders.
- Transcription factor sequestration in aggregates is a proposed pathogenic mechanism in HD.
Purpose of the Study:
- To investigate the impact of mutant huntingtin on CBP and p300, crucial transcriptional coactivators.
- To elucidate the mechanisms underlying CBP/p300 dysfunction in early and late stages of HD pathogenesis.
Main Methods:
- Assessing transcriptional and histone acetyltransferase activity of CBP/p300.
- Analyzing binding affinity of mutant huntingtin to CBP.
- Investigating CBP protein degradation pathways.
- Comparing CBP/p300 regulation in HD models versus spinocerebellar ataxia 3.
Main Results:
- Soluble mutant huntingtin represses CBP transcriptional activity early on.
- Histone acetyltransferase activity of CBP/p300 diminishes over time.
- Mutant huntingtin exhibits stronger binding to CBP, leading to its repression.
- CBP protein levels are reduced via proteasome degradation in later stages.
- p300 remains unaffected, and selective CBP degradation is absent in spinocerebellar ataxia 3.
Conclusions:
- Mutant huntingtin selectively targets CBP, not p300, through multiple mechanisms.
- Altered CBP/p300 ratios and reduced CBP levels contribute to chromatin and transcriptional dysregulation in HD.
- These molecular disruptions play a role in the neurodegenerative process of Huntington's disease.
More Related Videos
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 02:23
Abnormal Proliferation
 02:27
Nonsense-mediated mRNA Decay
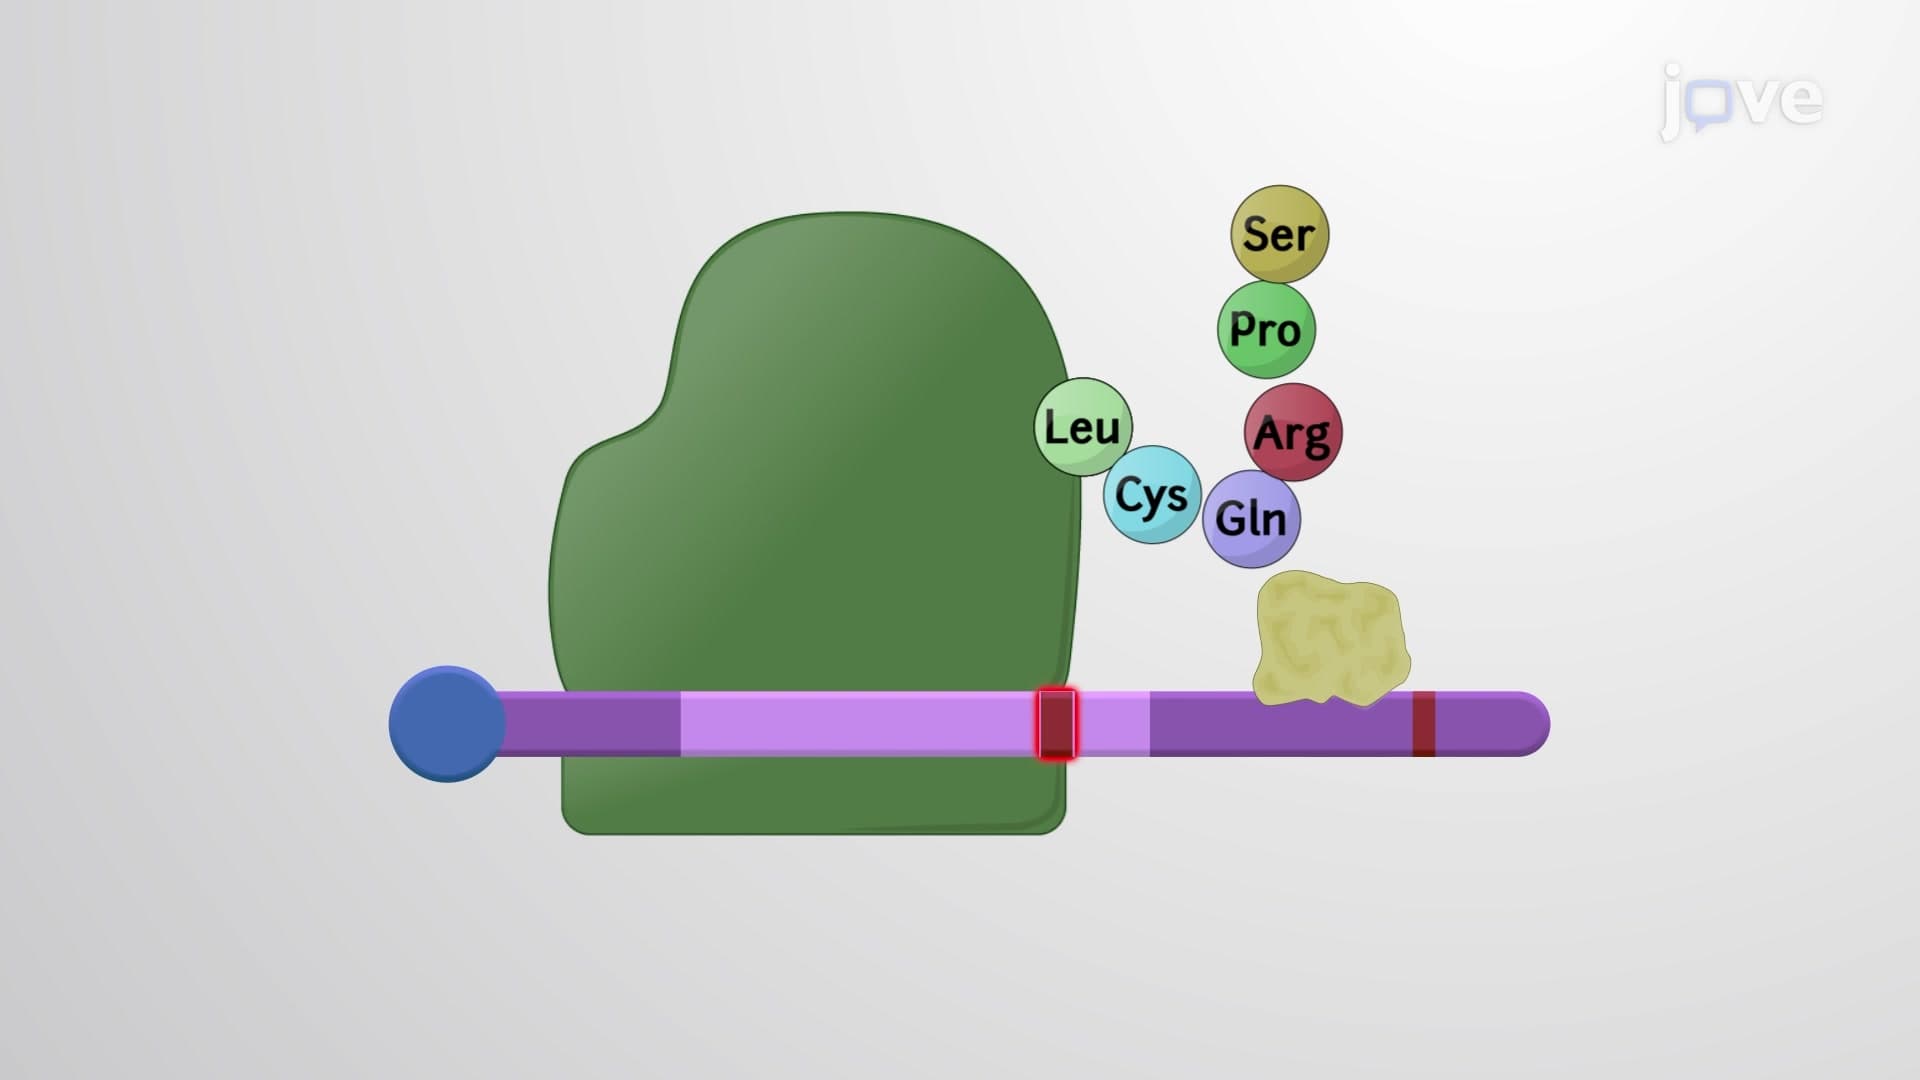
Usually, Upf3 binds to an Exon Junction Complex (EJC) at mRNA splice sites. If a ribosome fully translates the mRNA,...
 02:04
Covalently Linked Protein Regulators
These groups modify specific amino acids in a protein.
 02:38
Heterochromatin
Constitutive heterochromatin: It is a highly compact region of chromatin that is mostly concentrated in the centromere and telomere. Unlike euchromatin, the amino acid at 9th...
 02:04
Co-activators and Co-repressors