Related Experiment Video
Updated: Aug 16, 2026

Patch Clamp and Perfusion Techniques for Studying Ion Channels Expressed in Xenopus oocytes
Published on: January 10, 2011
Structural basis for competition between drug binding and Kvbeta 1.3 accessory subunit-induced N-type inactivation of
Niels Decher1, Pradeep Kumar, Teresa Gonzalez
1Nora Eccles Harrison Cardiovascular Research and Training Institute, University of Utah, Salt Lake City, UT 84112, USA.

Abstract:
Kvbeta subunits are accessory proteins that modify gating of Kv1 channels. Kvbeta1.3 subunits bind to the N termini of Kv1.5 alpha-subunits and induce fast N-type inactivation, slow the rate of deactivation, and alter the voltage dependence and kinetics of channel activation. The N terminus of a Kvbeta subunit and quaternary ammonium compounds bind to the inner pore of Kv1 channels; however, it is unknown to what extent the pore binding sites for drugs and Kvbeta subunits overlap. Here, we used site-directed Ala mutagenesis to scan residues of the Kv1.5 pore to define the binding site for Kvbeta1.3 subunits. Individual mutations of five residues in the S6 domain (Val505, Ile508, Leu510, Val512, and Val516) greatly retarded or prevented Kvbeta1.3 induced inactivation, and reduced effects on Kv1.5 deactivation. Mutation of Thr479 and Thr480 enhanced Kvbeta1.3-induced N-type inactivation. None of the Ala mutations prevented the Kvbeta1.3-induced negative shifts in the voltage dependence of activation or slow C-type inactivation, suggesting that these gating effects are mediated by an interaction other than the one for N-type inactivation. Thr479, Thr480, Val505, Ile508, and Val512, of Kv1.5 channels are also important interaction sites for the anthranilic acid S0100176 (N-benzyl-N-pyridin-3-ylmethyl-2-(toluene-4-sulfonylamino)-benzamide hydrochloride). Leu510 and V516A prevented Kvbeta1.3-induced inactivation but did not alter drug block. Block of Kv1.5 by S0100176 was reduced and voltage-dependent in the presence of Kvbeta1.3 but not in the presence of an N-truncated form of the Kvbeta subunit. Thus, residues in the pore of Kv1.5 required for N-type inactivation overlap with but are not identical to the drug binding site.
Related Concept Videos
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:29
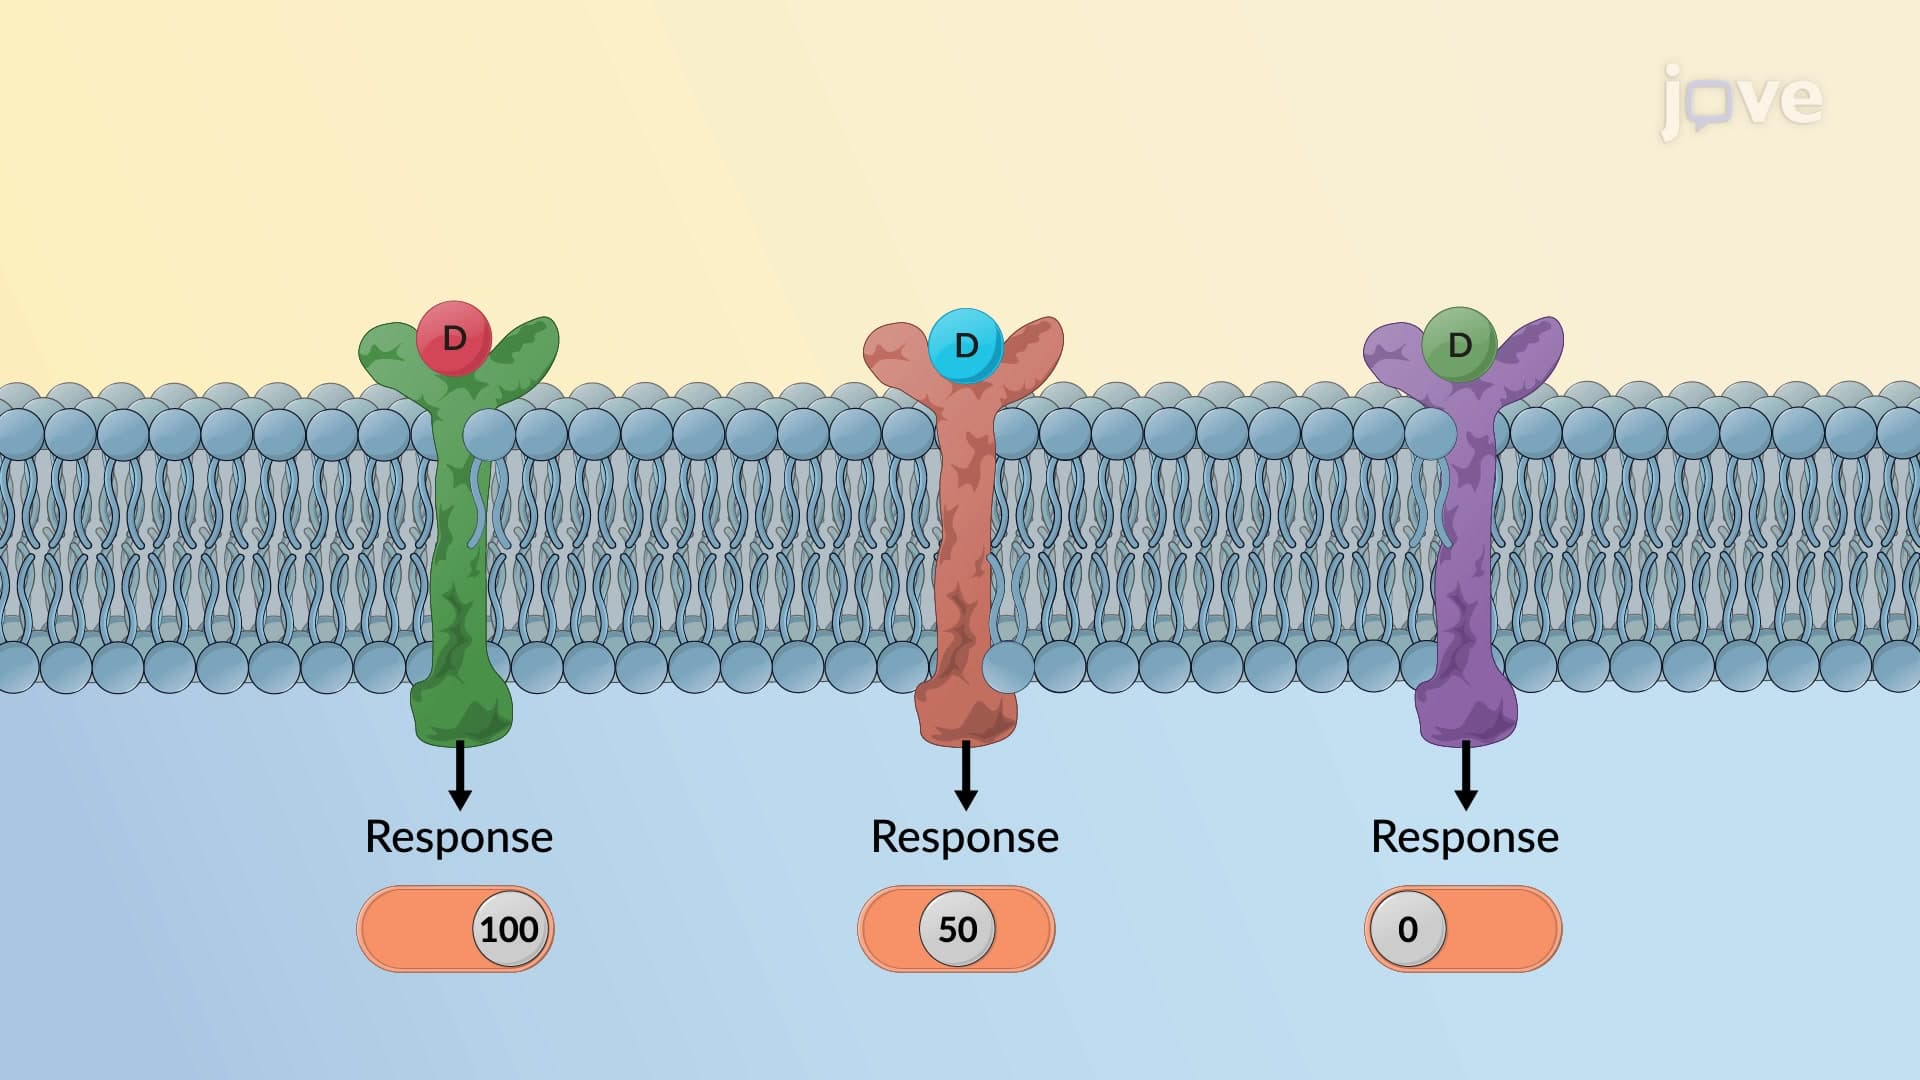
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue.
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:26
Voltage-gated Ion Channels
Generally, all voltage-gated ion channels have a 'voltage-sensing domain' that spans the lipid bilayer. The charged residues in the sensor move in response to the membrane potential changes that open the channel allowing ions movement. There are several types of...
 01:19
Ligand-gated Ion Channels
Three Subfamilies of Ligand-gated Ion Channels
Ligand-gated ion channels fall into three subfamilies. The 'Cys-loop' includes the nicotinic acetylcholine receptors, γ-aminobutyric acid (GABA), glycine, and 5-hydroxytryptamine receptors. The second one is the 'Pore-loop' channels that include the...