Related Experiment Video
Updated: Aug 16, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
"Reversine" and its 2-substituted adenine derivatives as potent and selective A3 adenosine receptor antagonists
Melissa Perreira1, Jian-Kang Jiang, Athena M Klutz
1Chemical Biology Core Facility, and Molecular Recognition Section, Laboratory of Biological Chemistry, National Institute of Diabetes and Digestive and Kidney Diseases, National Institutes of Health, Bethesda, Maryland 20892, USA.

Abstract:
The dedifferentiation agent "reversine" [2-(4-morpholinoanilino)-N(6)-cyclohexyladenine 2] was found to be a moderately potent antagonist for the human A(3) adenosine receptor (AR) with a K(i) value of 0.66 microM. This result prompted an exploration of the structure-activity relationship of related derivatives, synthesized via sequential substitution of 6-chloro-2-fluoropurine with selected nucleophiles. Optimization of substituents at these two positions identified 2-(phenylamino)-N(6)-cyclohexyladenine (12), 2-(phenylamino)-N(6)-cycloheptyladenine (19), and 2-phenylamino-N(6)-endo-norbornyladenine (21) as potent A(3) AR ligands with K(i) values of 51, 42, and 37 nM, respectively, with 30-200-fold selectivity in comparison to A(1) and A(2A) ARs. The most selective A(3) AR antagonist (>200-fold) was 2-(phenyloxy)-N(6)-cyclohexyladenine (22). 9-Methylation of 12, but not 19, was well-tolerated in A(3) AR binding. Extension of the 2-phenylamino group to 2-benzyl- and 2-(2-phenylethylamino) reduced affinity. In the series of 2-(phenylamino), 2-(phenyloxy), and 2-(phenylthio) substitutions, the order of affinity at the A(3) AR was oxy > or = amino > thio. Selected derivatives, including reversine (K(B) value of 466 nM via Schild analysis), competitively antagonized the functional effects of a selective A(3) AR agonist, i.e., inhibition of forskolin-stimulated cAMP production in stably transfected Chinese hamster ovary (CHO) cells. These results are in agreement with other studies suggesting the presence of a lipophilic pocket in the AR binding site that is filled by moderately sized cycloalkyl rings at the N(6) position of both adenine and adenosine derivatives. Thus, the compound series reported herein comprise an important new series of selective A(3) AR antagonists. We were unable to reproduce the dedifferentiation effect of reversine, previously reported, or to demonstrate any connection between A(3) AR antagonist effects and dedifferentiation.
Related Concept Videos
 01:17
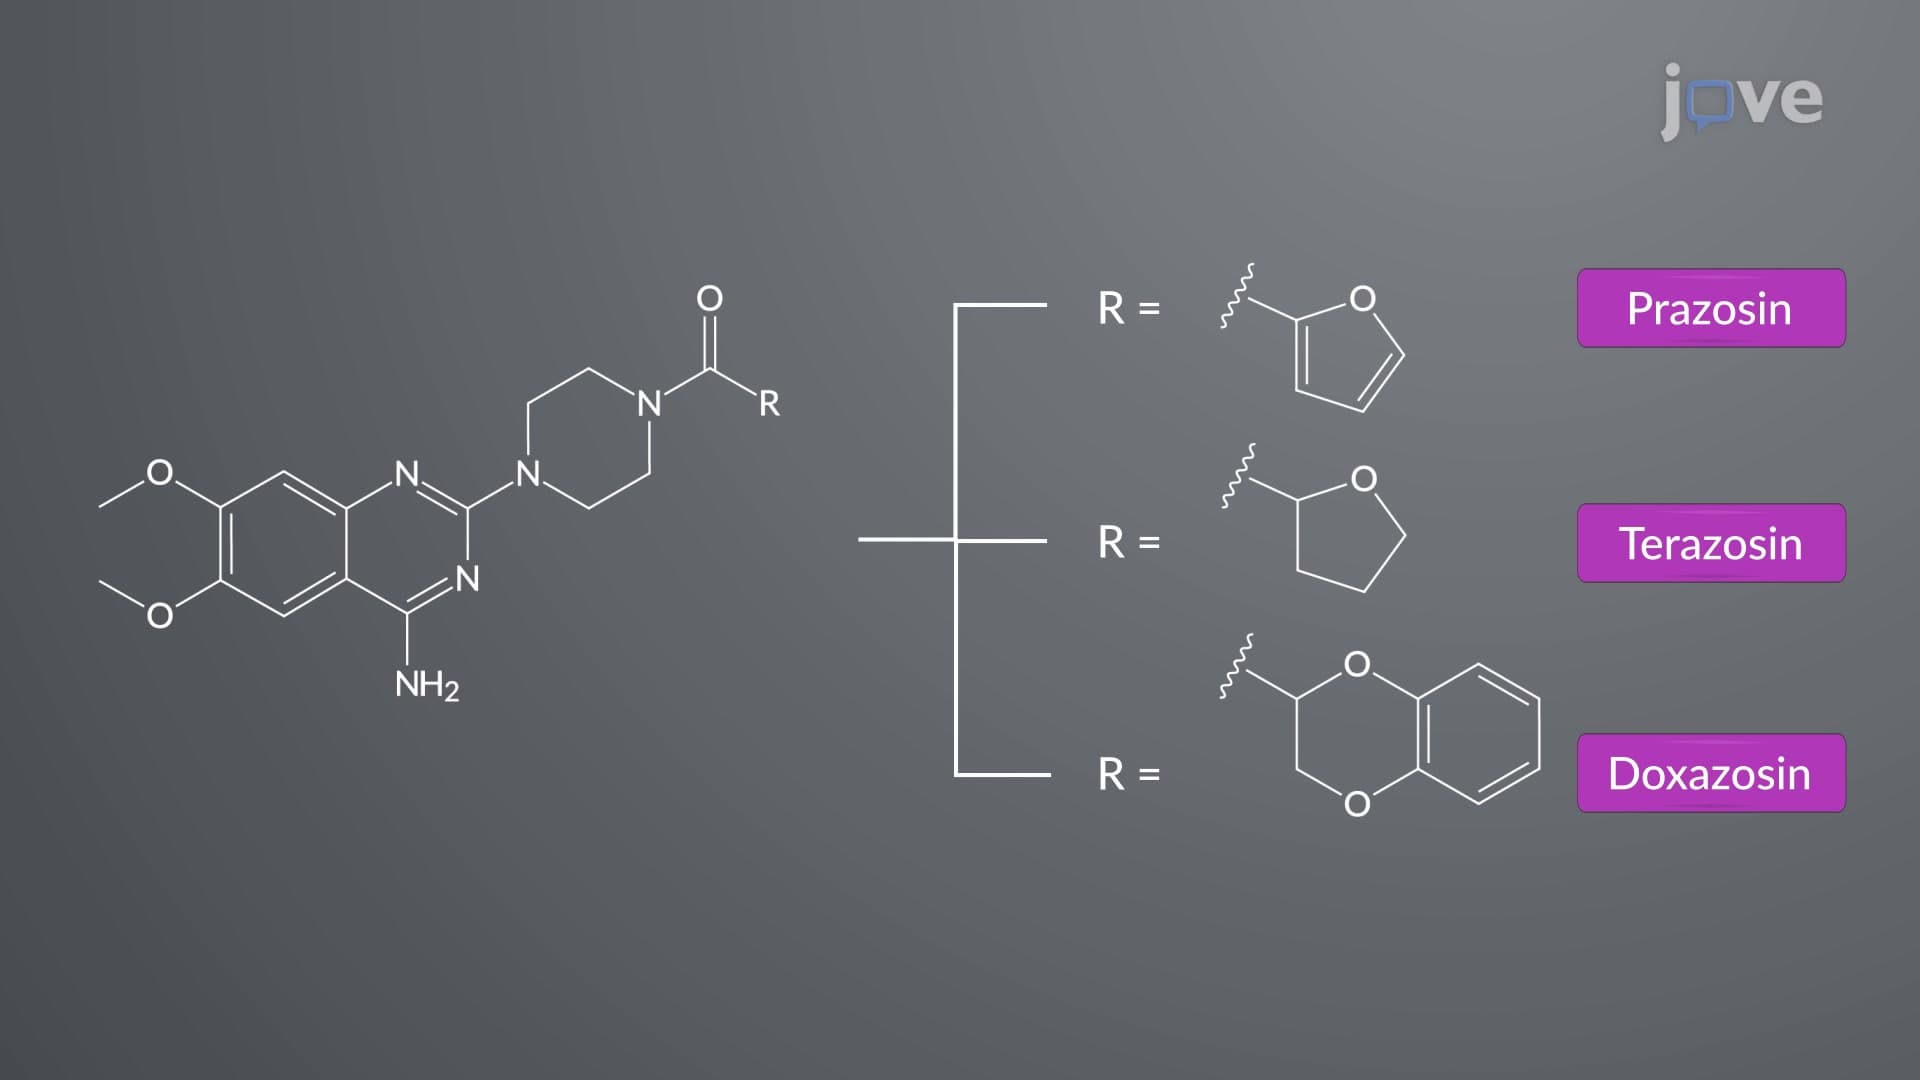
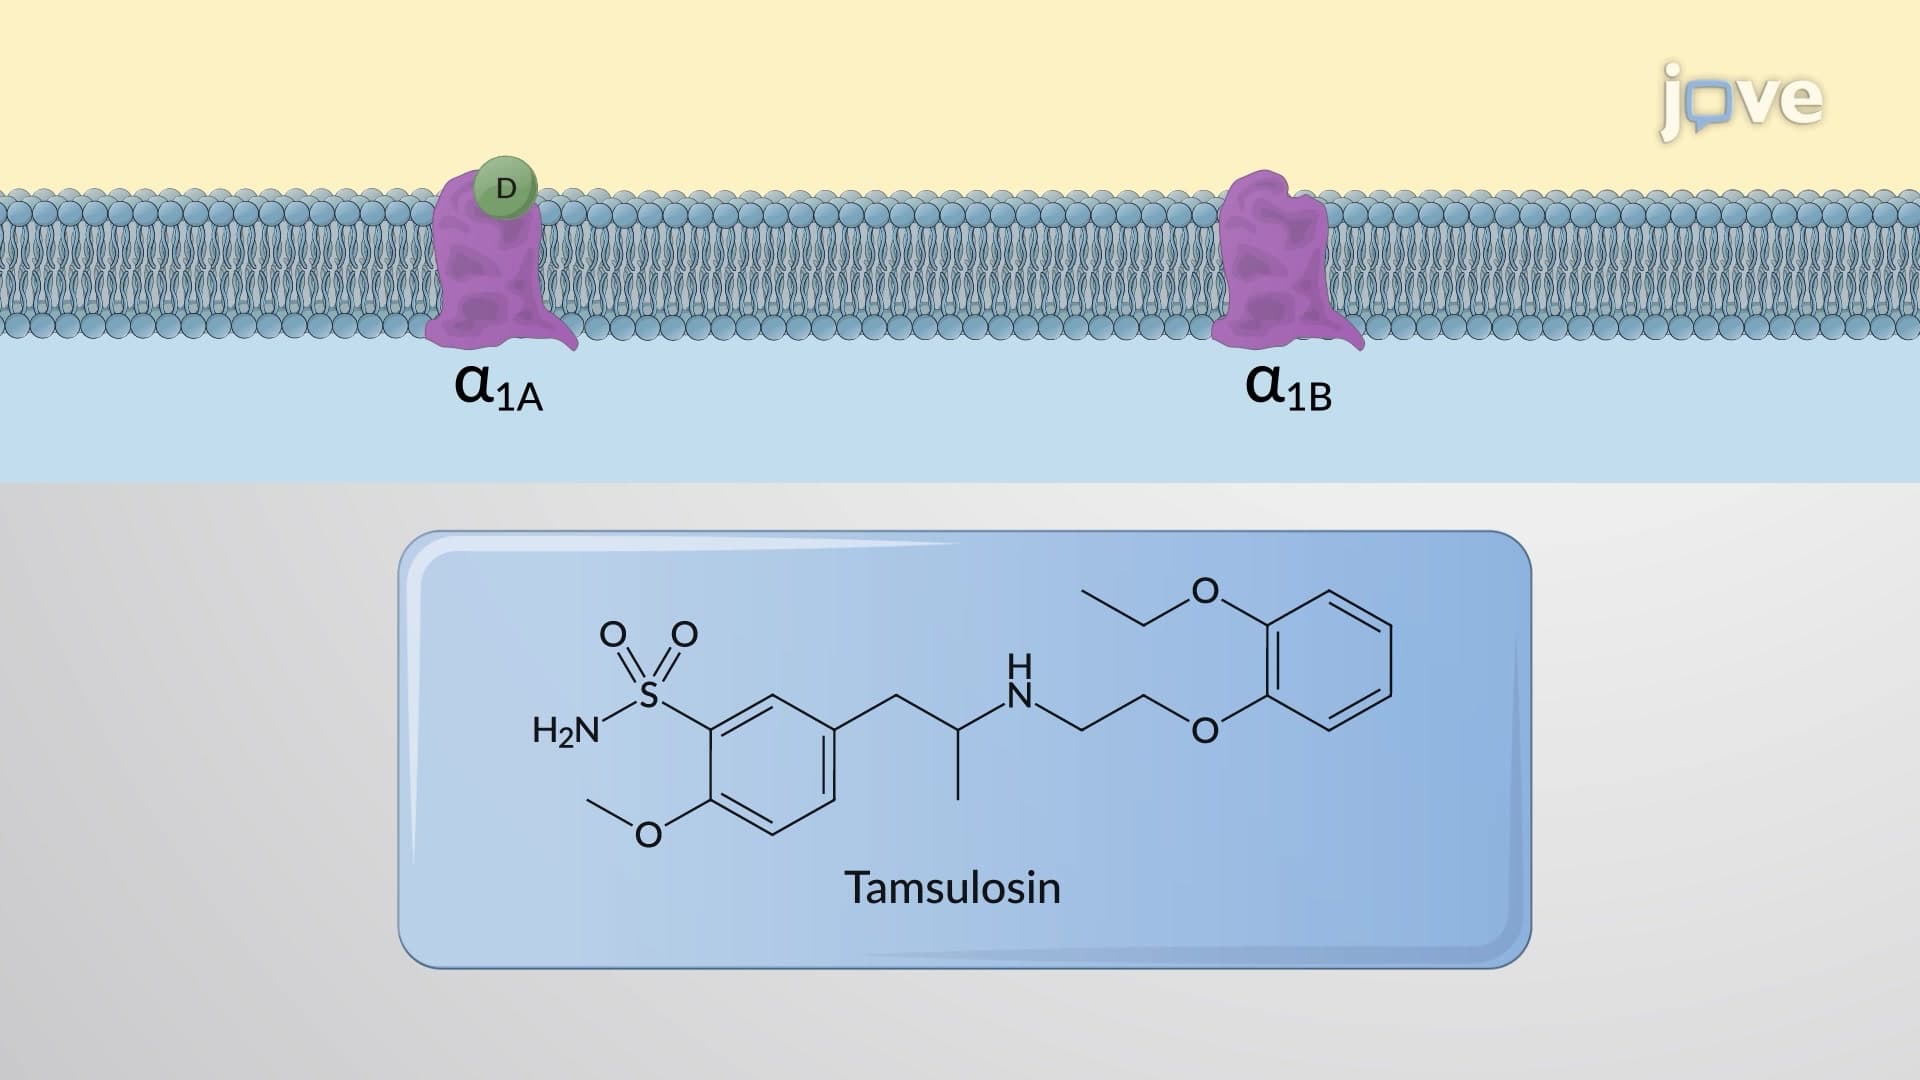
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:22
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:29
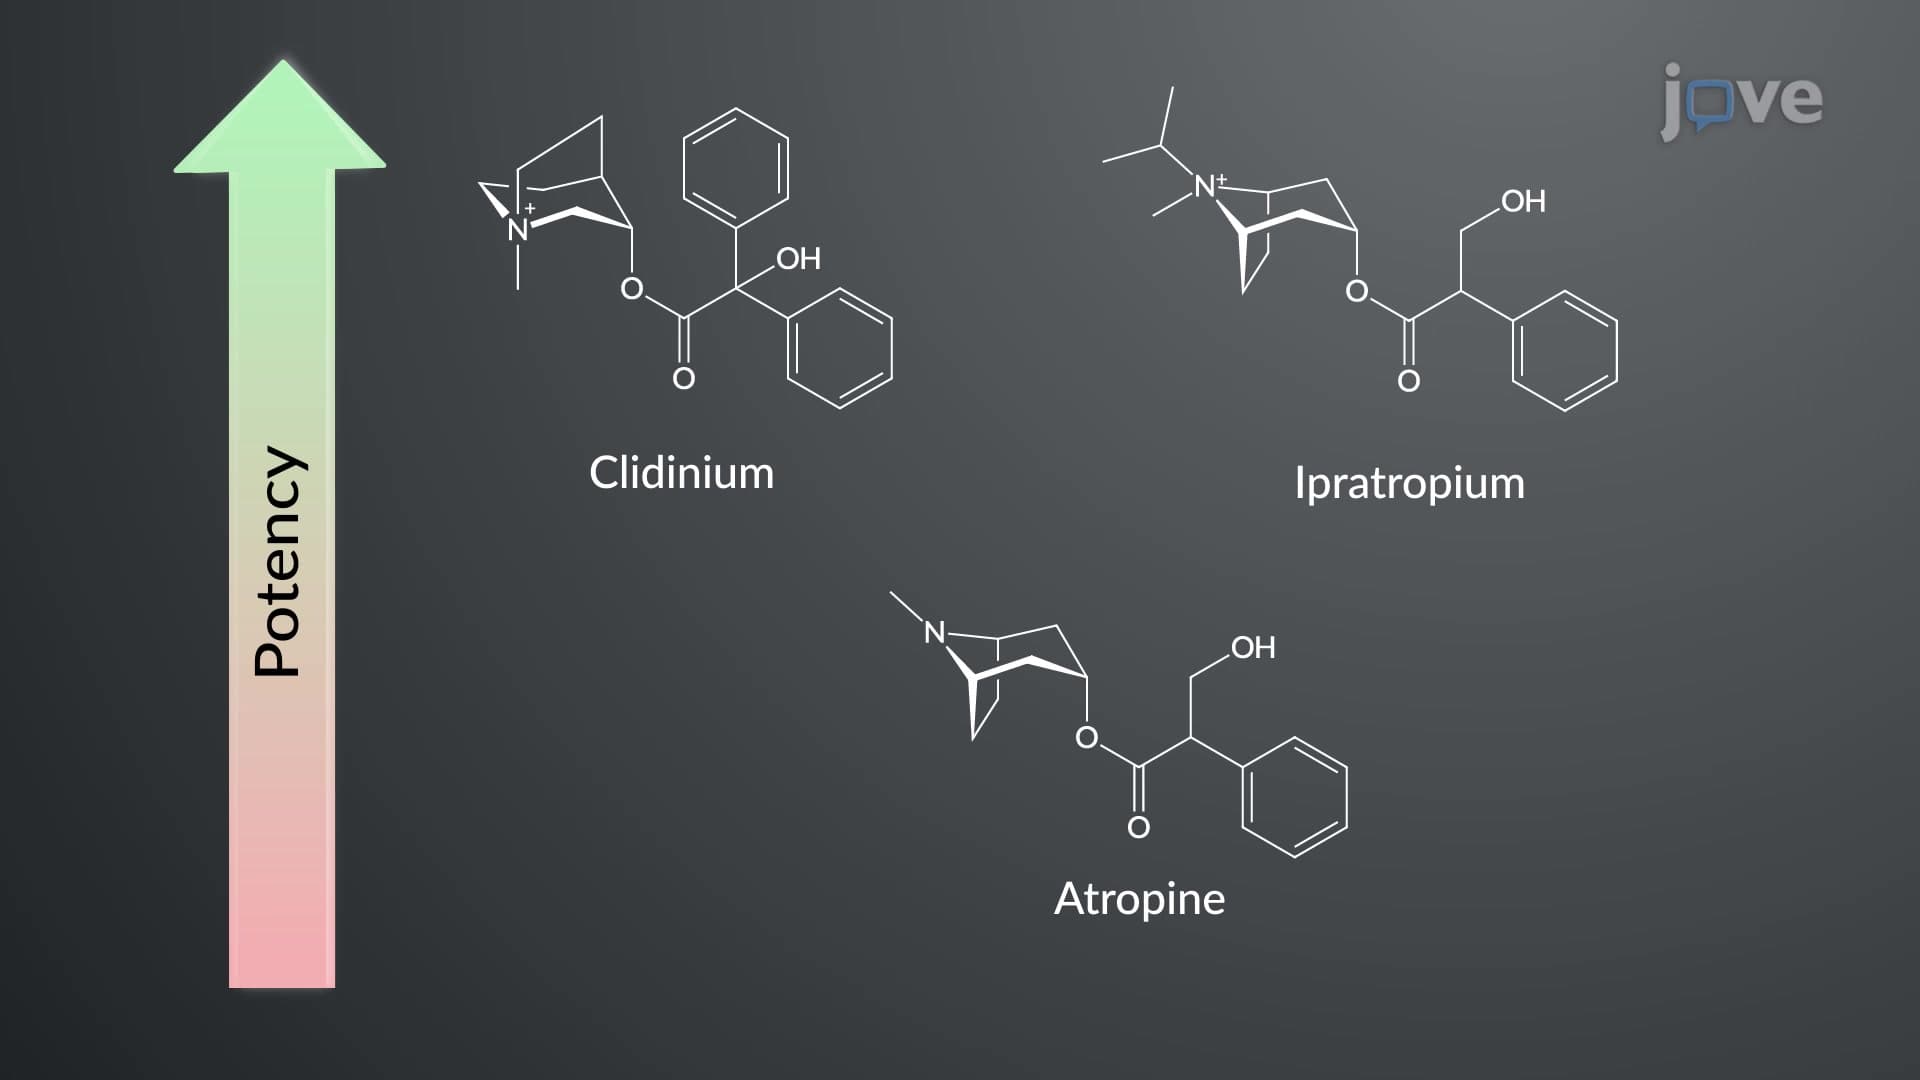
Cholinergic Antagonists: Chemistry and Structure-Activity Relationship
 01:28
Drug-Receptor Interaction: Antagonist
Antagonists can be classified as competitive or noncompetitive based on their...