Coupling between conformation and proton binding in proteins
Related Experiment Video
Updated: Aug 16, 2026
Covalent Labeling with Diethylpyrocarbonate for Studying Protein Higher-Order Structure by Mass Spectrometry
Published on: June 15, 2021
Jorge A Vila1, Daniel R Ripoll, Yelena A Arnautova
1Baker Laboratory of Chemistry and Chemical Biology, Cornell University, Ithaca, New York 14853-1301, USA.

Abstract:
Interest centers here on whether the use of a fixed charge distribution of a protein solute, or a treatment that considers proton-binding equilibria by solving the Poisson equation, is a better approach to discriminate native from non-native conformations of proteins. In this analysis of the charge distribution of 7 proteins, we estimate the solvation free energy contribution to the total free energy by exploring the 2(zeta) possible ionization states of the whole molecule, with zeta being the number of ionizable groups in the amino acid sequence, for every conformation in the ensembles of 7 proteins. As an additional consideration of the role of electrostatic interactions in determining the charge distribution of native folds, we carried out a comparison of alternative charge assignment models for the ionizable residues in a set of 21 native-like proteins. The results of this work indicate that (1) for 6 out of 7 proteins, estimation of solvent polarization based on the Generalized Born model with a fixed charge distribution provides the optimal trade-off between accuracy, with respect to the Poisson equation, and speed when compared to the accessible surface area model; for the seventh protein, consideration of all possible ionization states of the whole molecule appears to be crucial to discriminate the native from non-native conformations; (2) significant differences in the degree of ionization and hence the charge distribution for native folds are found between the different charge models examined; (3) the stability of the native state is determined by a delicate balance of all the energy components, and (4) conformational entropy, and hence the dynamics of folding, may play a crucial role for a successful ab initio protein folding prediction.
More Related Videos
 10:03
Proton Transfer and Protein Conformation Dynamics in Photosensitive Proteins by Time-resolved Step-scan Fourier-transform Infrared Spectroscopy
Published on: June 27, 2014
 09:25
NMR 15N Relaxation Experiments for the Investigation of Picosecond to Nanoseconds Structural Dynamics of Proteins
Published on: November 1, 2024
Related Concept Videos
 00:49
Ligand Binding and Linkage
 02:43
Peptide Bonds
 01:22
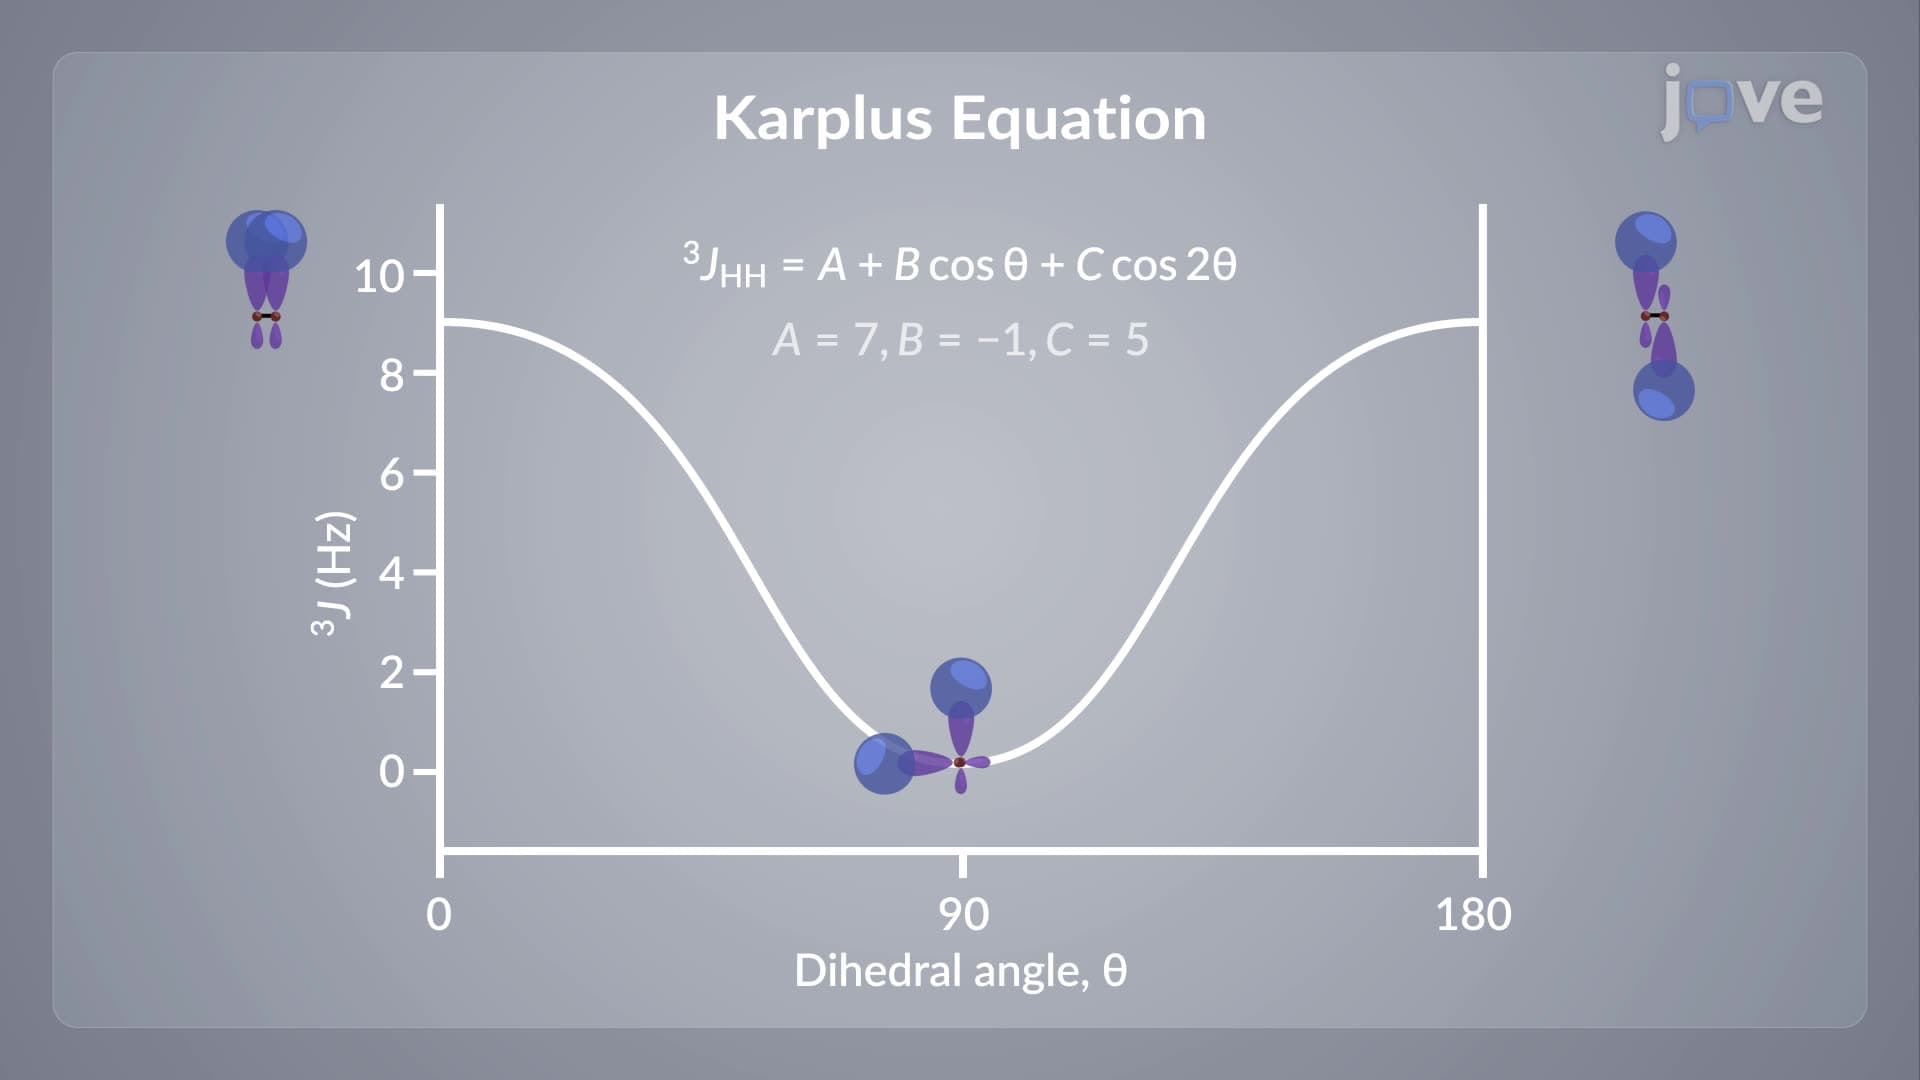
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:17
Spin–Spin Coupling: One-Bond Coupling