Related Experiment Video
Updated: Aug 16, 2026

Probing the Structure and Dynamics of Interfacial Water with Scanning Tunneling Microscopy and Spectroscopy
Published on: May 27, 2018
Photodissociation of the water dimer: three-dimensional quantum dynamics studies on diabatic potential-energy
Loredana Valenzano1, Marc C van Hemert, Geert-Jan Kroes
1Gorlaeus Laboratories, Leiden Institute of Chemistry, Leiden University, The Netherlands.

Abstract:
The results are presented of three-dimensional model studies of the photodissociation of the water dimer following excitation in the first absorption band. Diabatic potential-energy surfaces are used to investigate the photodissociation following excitation of the hydrogen bond donor molecule and of the hydrogen bond acceptor molecule. In both cases, the degrees of freedom considered are the two OH-stretch modes of the molecule being excited, and the dimer stretch vibration. The diabatic potentials are based on adiabatic potential surfaces computed with the multireference configuration-interaction method, and the dynamics of dissociation was studied using the time-dependent wave-packet method. The dynamics calculations yield a donor spectrum extending over roughly the same range of frequencies as the spectrum of the water monomer computed at the same level of theory. The acceptor spectrum has the same width as the monomer spectrum, but is shifted to the blue by 0.4-0.5 eV. The dimer spectrum obtained by averaging the donor and the acceptor spectrum is broader than the monomer spectrum, with the center of the dimer first absorption band shifted to the blue by about 0.2 eV relative to the monomer band. Our reduced dimensionality calculations do not find the red tail predicted for the dimer first absorption band by Harvey et al. [J. Chem. Phys. 109, 8747 (1998)]. This conclusion also holds if preexcitation of the dimer stretch vibration with one or two quanta is considered.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:25
Deactivation Processes: Jablonski Diagram
 02:13
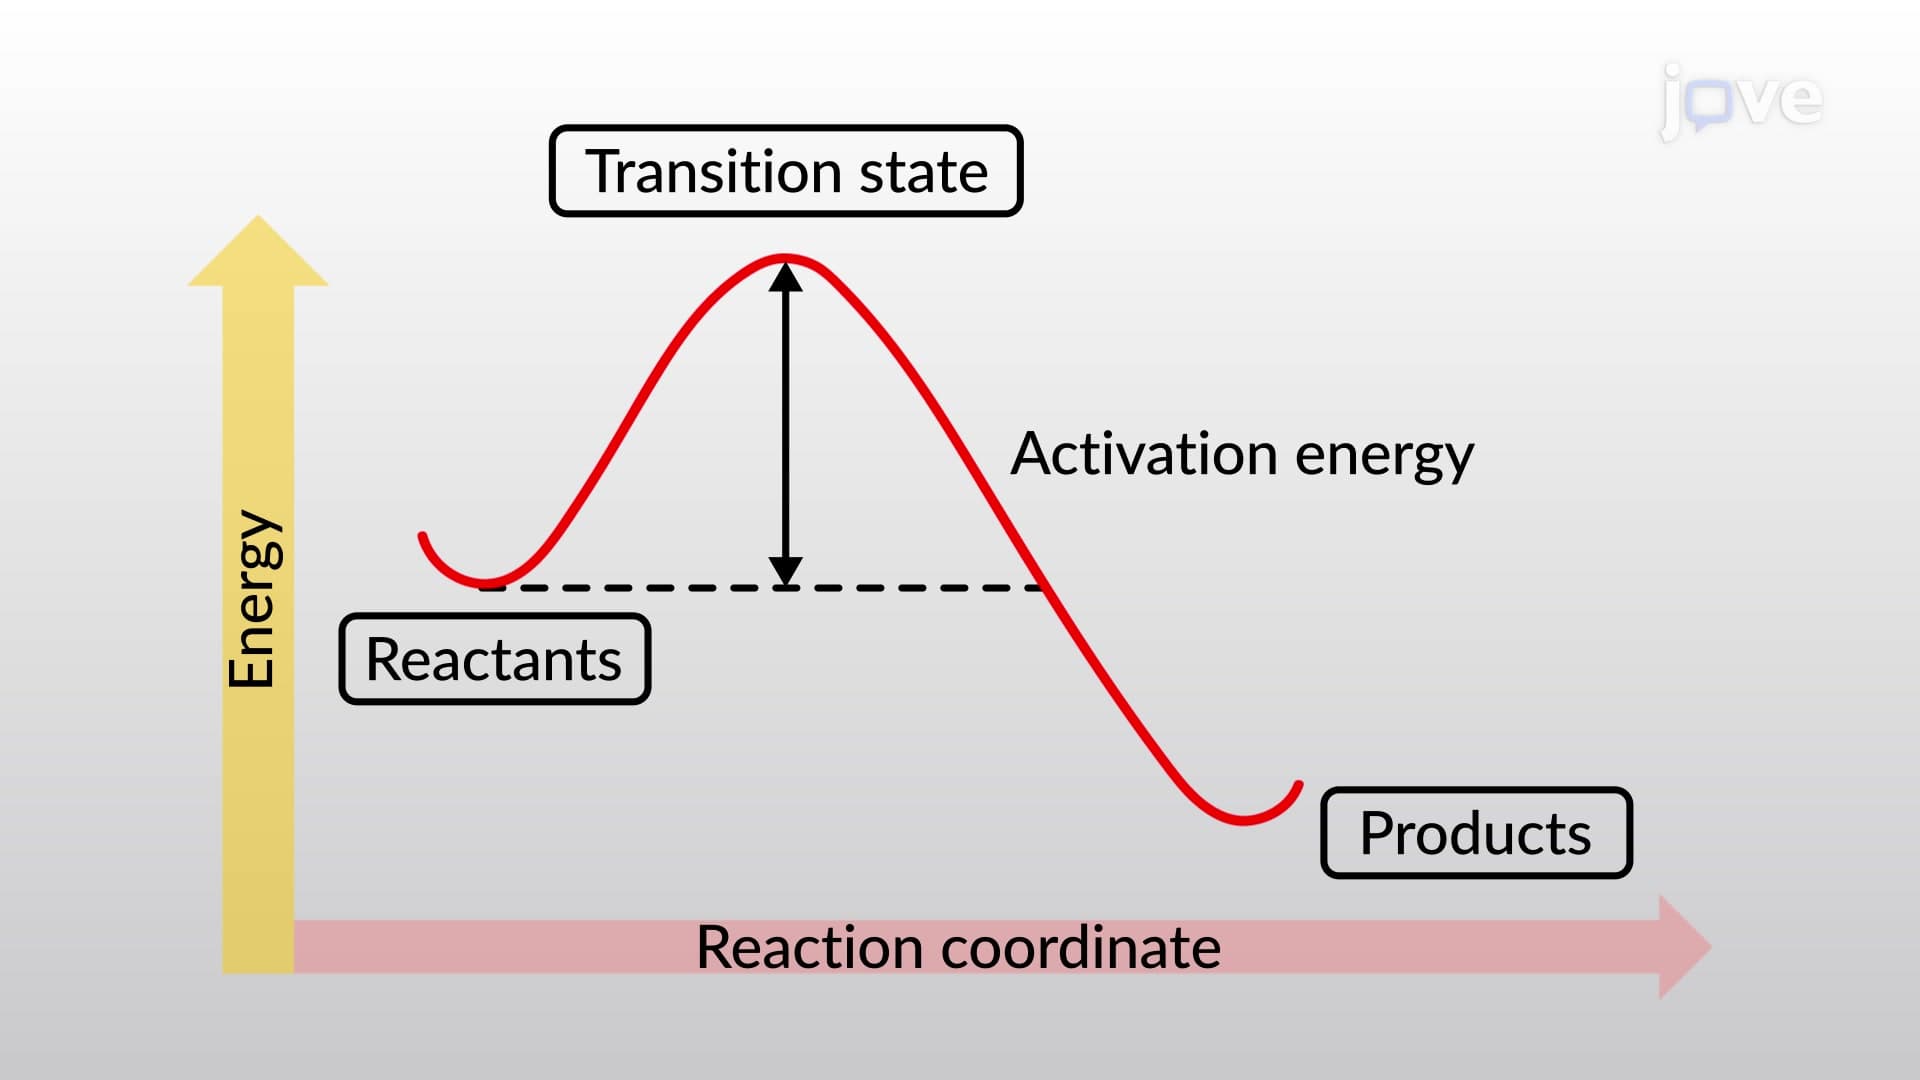
Bond Dissociation Energy and Activation Energy
 02:36
Molecular Geometry and Dipole Moments
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 00:48
¹H NMR of Labile Protons: Deuterium (²H) Substitution