Related Experiment Video
Updated: Aug 16, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Dual-scale modeling of benzene adsorption onto Ni(111) and Au(111) surfaces in explicit water
Pim Schravendijk1, Nico van der Vegt, Luigi Delle Site
1Max Planck Institute for Polymer Research, Ackermannweg 10, 55128 Mainz, Germany.

Abstract:
We present a multiscale modeling approach for studying interactions of organic molecules with metal surfaces in explicit water. The approach is based on combining adsorption energies of isolated molecules on transition metal surfaces calculated by ab initio density functional methods and classical molecular dynamics simulations using atomistically detailed force fields. The interaction of benzene with Ni(111) and Au(111) surfaces was studied. It is shown that a strong affinity of water for the hydrophilic surfaces makes benzene adsorption on Au thermodynamically unfavorable, while on Ni there is no preference. The work presented here serves as a first step in modeling the interactions of larger organic molecules with metal surfaces.
More Related Videos
Related Concept Videos
 01:25
Adsorption Isotherms II
 01:29
Adsorption Isotherms I
 01:28
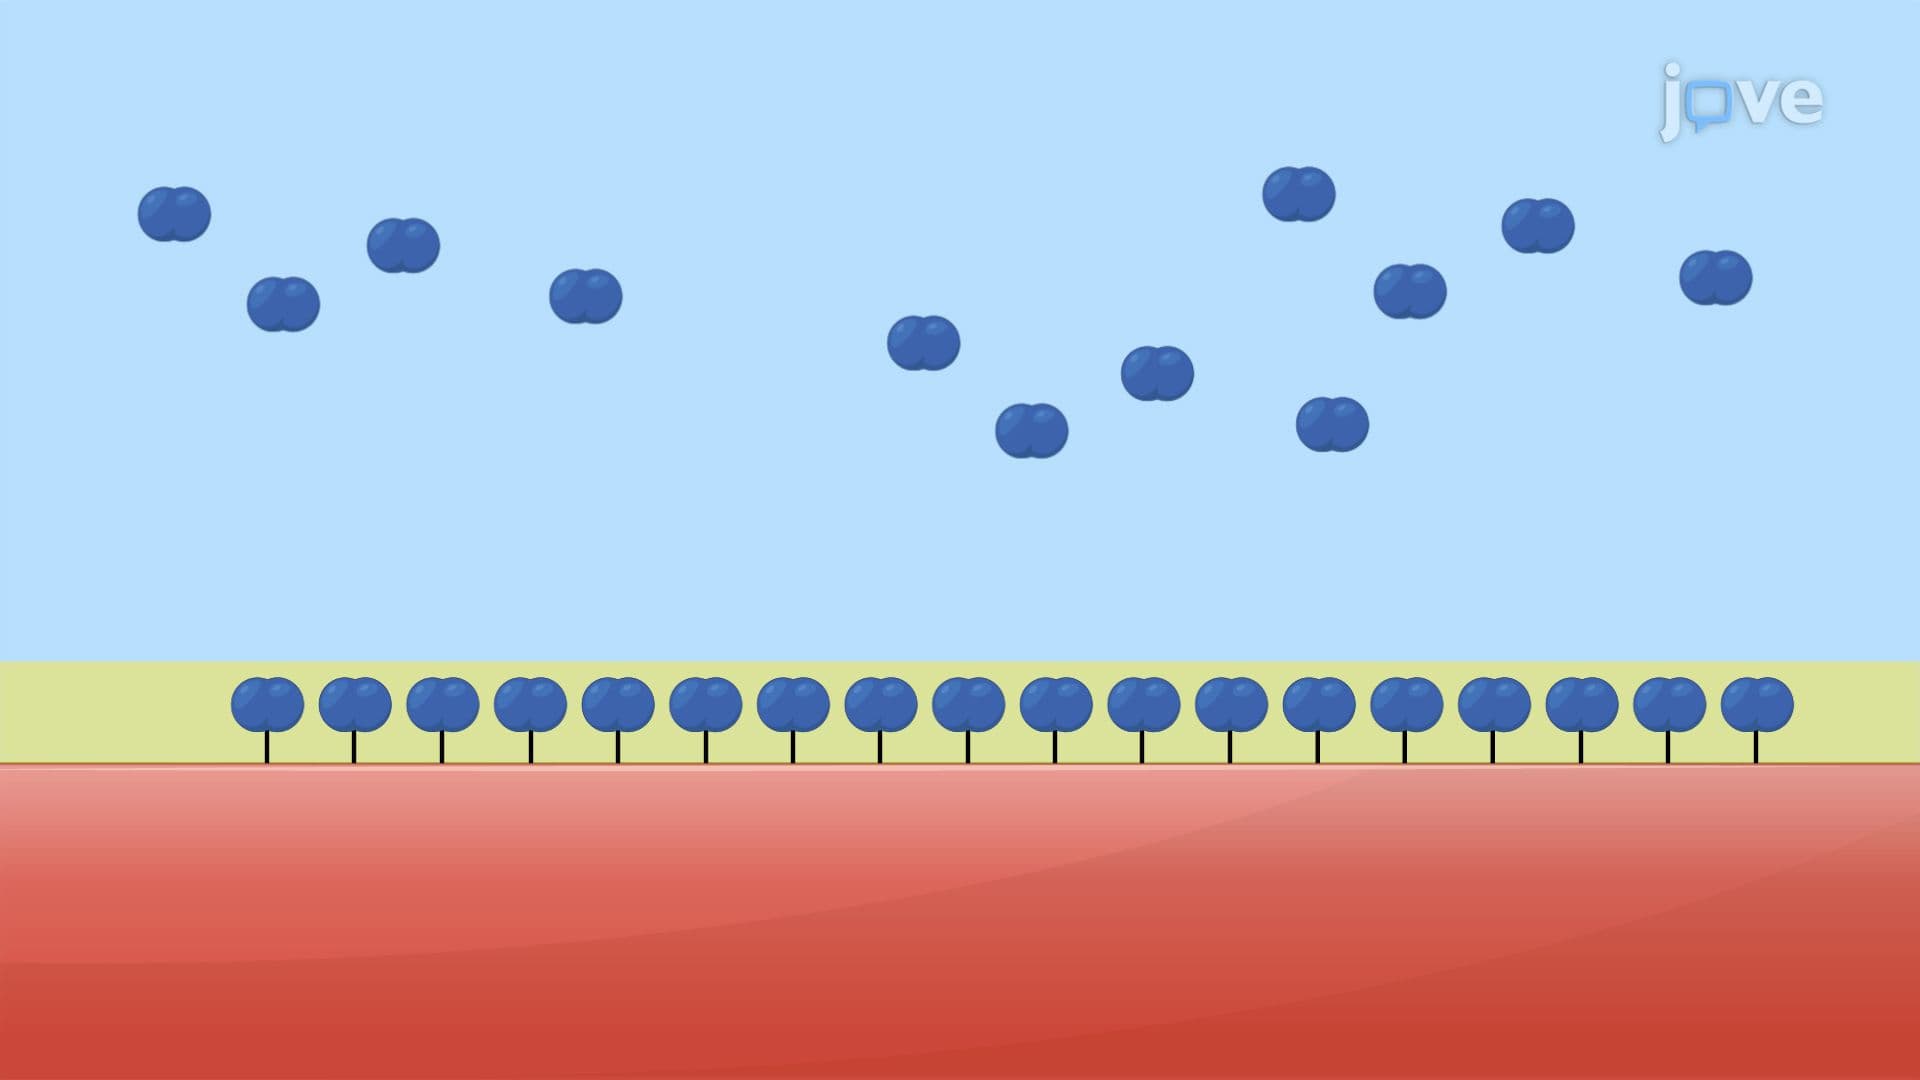
Adsorption of Gases on Solids