Related Experiment Video
Updated: Aug 16, 2026

Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Defunctionalized lobeline analogues: structure-activity of novel ligands for the vesicular monoamine transporter
Guangrong Zheng1, Linda P Dwoskin, Agripina G Deaciuc
1College of Pharmacy, University of Kentucky, Lexington, Kentucky 40536-0082, USA.

Abstract:
(-)-Lobeline (2R,6S,10S), an antagonist at nicotinic acetylcholine receptors (nAChRs), inhibits the neurochemical and behavioral effects of methamphetamine and inhibits dopamine transporter (DAT) and vesicular monoamine transporter (VMAT2) function. VMAT2 is a target for the development of treatments for methamphetamine abuse. Structural modification of lobeline affords the defunctionalized analogues meso-transdiene (MTD) and lobelane, which have high potency and selectivity for VMAT2. To establish the structure-activity relationships within this novel class of VMAT2 ligands, specific stereochemical forms of MTD, lobelane, and other structurally related analogues have been synthesized. These compounds have been evaluated for inhibition of [(3)H]nicotine ([(3)H]NIC) binding (alpha4beta2 nAChR), [(3)H]methyllycaconitine ([(3)H]MLA) binding (alpha7 nAChR), and [(3)H]dihydrotetrabenazine ([(3)H]DTBZ) binding (VMAT2). Generally, all of these analogues had lower affinities at alpha4beta2 and alpha7 nAChRs compared to lobeline, thereby increasing selectivity for VMAT2. The following structural modifications resulted in only modest changes in affinity for VMAT2, affording analogues that were less potent than the lead compound, lobelane: (1) altering the stereochemistry at the C-2 and C-6 positions of the piperidino ring, (2) varying unsaturation in the piperidino C-2 and C-6 substituents, (3) introducing unsaturation into the piperidine ring, (4) ring-opening or eliminating the piperidine ring, and (5) removing the piperidino N-methyl group. Furthermore, incorporating a quaternary ammonium group into defunctionalized lobeline molecules in the cis-series resulted in significant loss of affinity for VMAT2, whereas only a modest change in affinity was obtained in the trans-series. The most potent (K(i) = 630 nM) and VMAT2-selective compound evaluated was the N-methyl-2,6-cis-bis(naphthaleneethyl)piperidine analogue (1-NAP-lobelane), in which the phenyl groups of lobelane were replaced with 1-naphthyl moieties. Thus, initial structure-activity relationship studies reveal that the most promising structural changes to the lobeline molecule that lead to enhancement of VMAT2 affinity and selectivity are defunctionalization, affording lobelane and MTD, and replacement of the phenyl rings of lobelane with other aromatic moieties that have a pi-extended structure.

Related Concept Videos
 01:25
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral bioavailability, and...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:21
Drugs Affecting Neurotransmitter Release or Uptake
 01:29
Indirect-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...
 01:22
Direct-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
The direct-acting...
 01:25
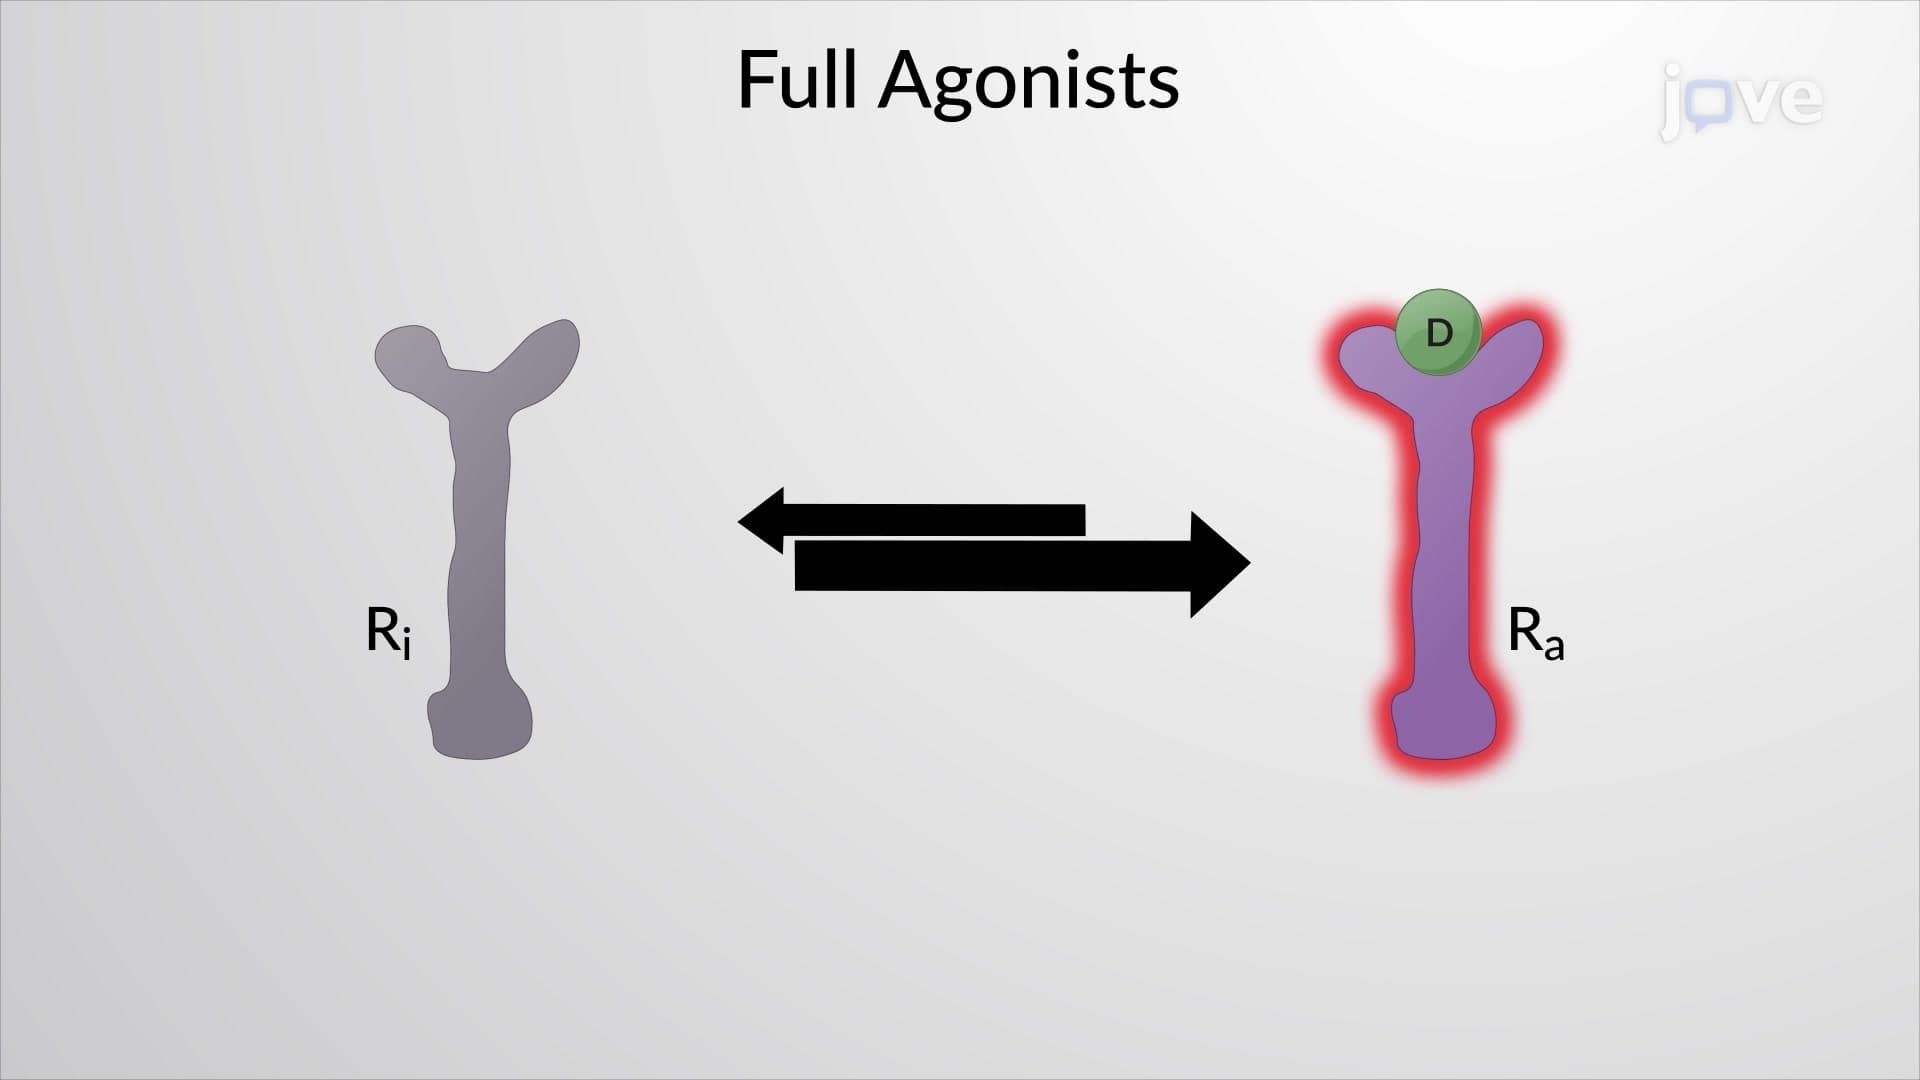
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous ligand's action.