Related Experiment Video
Updated: Aug 16, 2026


Realistic Membrane Modeling Using Complex Lipid Mixtures in Simulation Studies
Published on: September 1, 2023
Detailed considerations for a balanced and broadly applicable force field: a study of substituted benzenes modeled
Daniel J Price1, Charles L Brooks
1Department of Molecular Biology, The Scripps Research Institute, La Jolla, California 92037, USA.

Abstract:
Modern classical force fields have been traditionally parameterized by attempting to maximize agreement to any number of experimental and/or quantum mechanical target properties. As these force fields are pushed towards obtaining quantitative estimates of often subtle energetic differences, stringent and consistent parameterization criteria, particularly in regard to charge distributions, are required to ensure that systematic errors cancel, that parameters are transferable between molecules, and that performance does not significantly deteriorate when using more approximate methods, such as with continuum solvent models. Relative free energies of hydration are presented here for 40 mono- and disubstituted benzenes modeled with the OPLS-AA force field; heats of vaporization and pure liquid densities at standard conditions are presented when experimental data is available. Overall agreement between OPLS-AA and experiment is remarkable (average error = 0.5 kcal/mol for DeltaDeltaG(hydration), 1.0 kcal/mol for DeltaH(vap) (0), 0.02 g/mL for densities), yet several functional groups are identified as having consistent and correctable errors (alkyl-, nitro-, and thiobenzenes). Relative free energies of hydration obtained with rigorous free energy perturbations using explicit solvent are also compared with energies from minimizations using a generalized Born model (GB). There is high correlation between these estimates (R = 0.99), and as demonstrated here, reparameterization of the aforementioned groups can be guided with rapid GB calculations.
More Related Videos
 05:57
Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 03:51
Molecular Orbital Theory II
 01:18
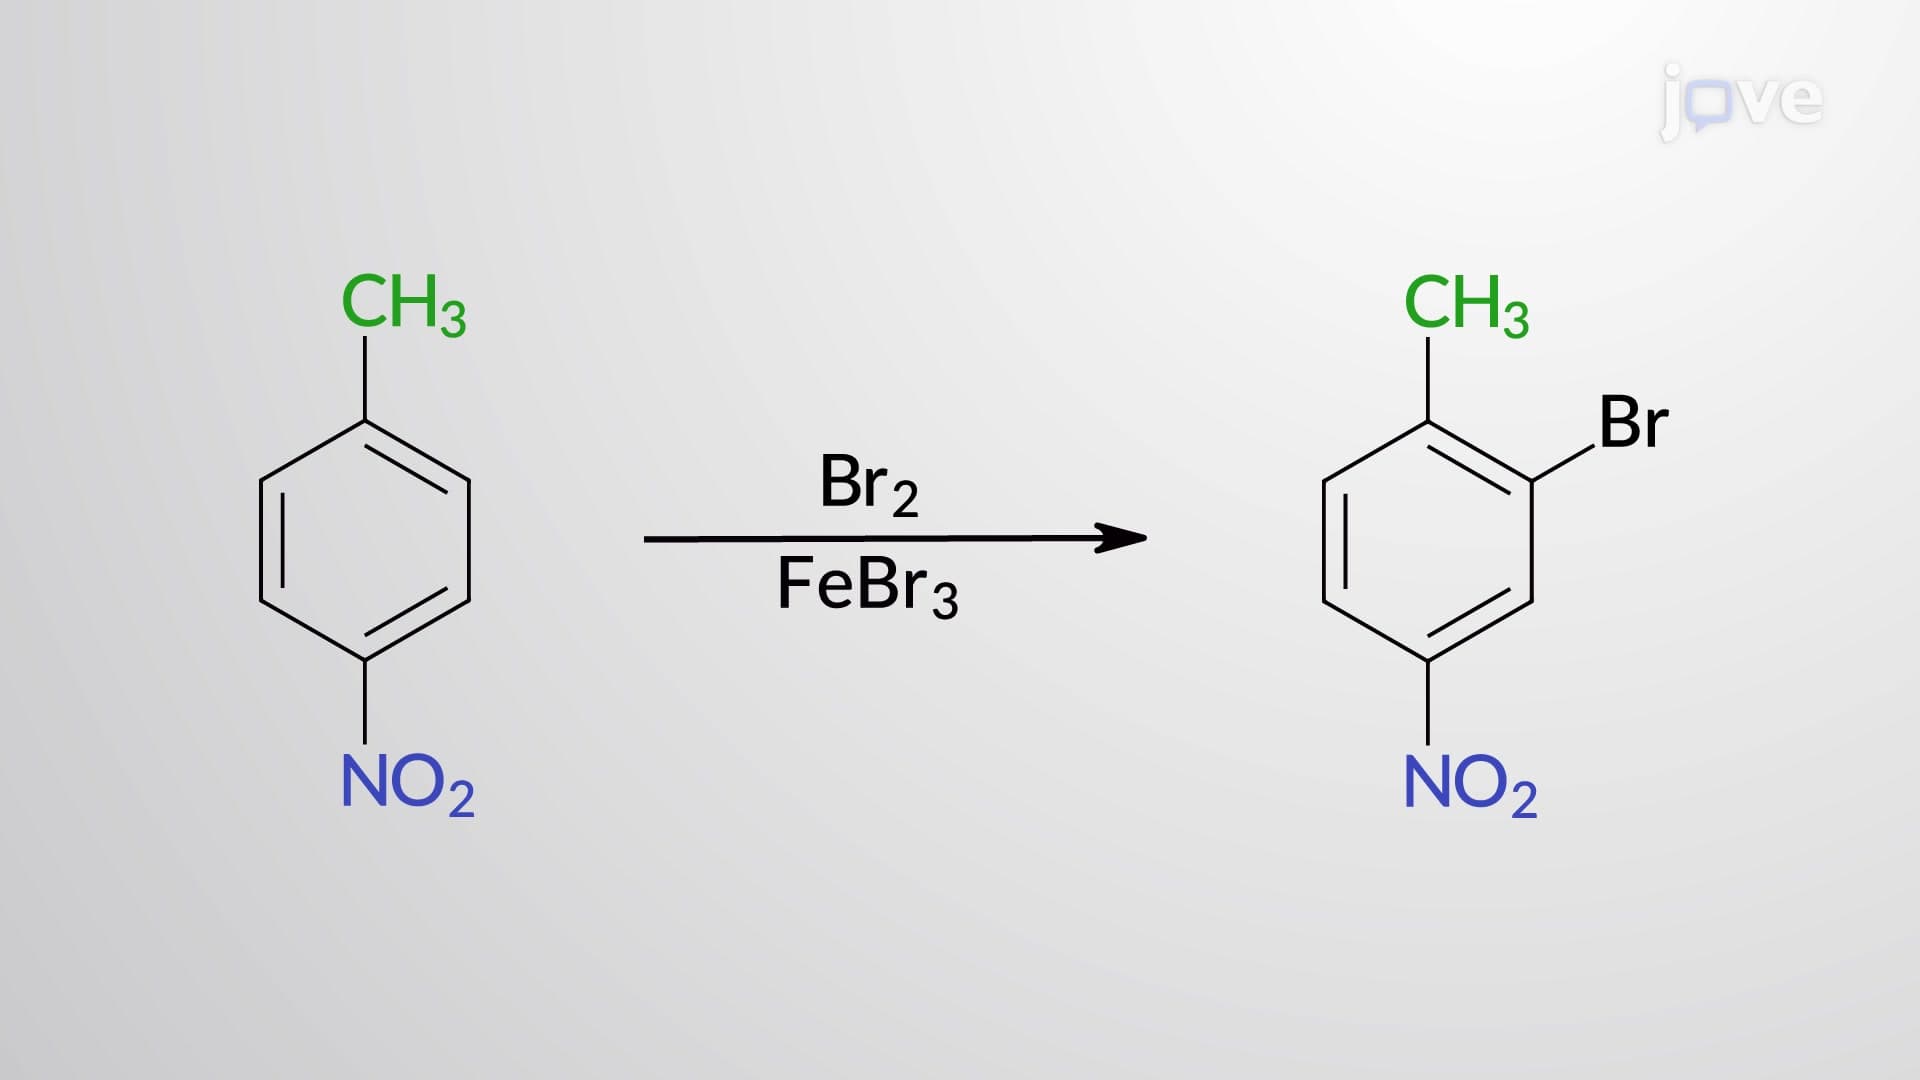
Directing and Steric Effects in Disubstituted Benzene Derivatives
 02:40
MO Theory and Covalent Bonding
 02:00
Molecular Models
 01:27
π Electron Effects on Chemical Shift: Overview