Related Experiment Video
Updated: Aug 16, 2026
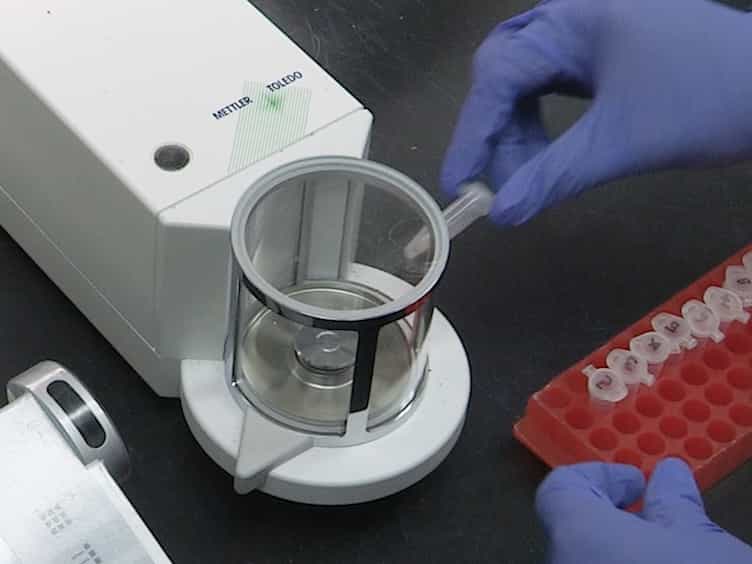

Deep Proteome Profiling by Isobaric Labeling, Extensive Liquid Chromatography, Mass Spectrometry, and Software-assisted Quantification
Published on: November 15, 2017
Data pre-processing in liquid chromatography-mass spectrometry-based proteomics
Xiang Zhang1, John M Asara, Jiri Adamec
1Bindley Bioscience Center, Purdue University West Lafayette, IN, USA. zhang100@purdue.edu

Motivation:
In a liquid chromatography-mass spectrometry (LC-MS)-based expressional proteomics, multiple samples from different groups are analyzed in parallel. It is necessary to develop a data mining system to perform peak quantification, peak alignment and data quality assurance.
Results:
We have developed an algorithm for spectrum deconvolution. A two-step alignment algorithm is proposed for recognizing peaks generated by the same peptide but detected in different samples. The quality of LC-MS data is evaluated using statistical tests and alignment quality tests.
Availability:
Xalign software is available upon request from the author.
Related Concept Videos
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:19
MALDI-TOF Mass Spectrometry