
Estrogenic compounds inhibit gap junctional intercellular communication in mouse Leydig TM3 cells
Yumiko Iwase1, Hideki Fukata, Chisato Mori
1Department of Bioenvironmental Medicine, Graduate School of Medicine, Chiba University, Chiba 260-8670, Japan. Iwase.Yumiko@mg.m-pharma.co.jp

Abstract:
Some estrogenic compounds are reported to cause testicular disorders in humans and/or experimental animals by direct action on Leydig cells. In carcinogenesis and normal development, gap junctional intercellular communication (GJIC) plays an essential role in maintaining homeostasis. In this study, we examine the effects of diethylstilbestrol (DES, a synthetic estrogen), 17beta-estradiol (E(2), a natural estrogen), and genistein (GEN, a phytoestrogen) on GJIC between mouse Leydig TM3 cells using Lucifer yellow microinjection. The three compounds tested produced GJIC inhibition in the TM3 cells after 24 h. Gradually, 10 microM DES began to inhibit GJIC for 24 h and this effect was observed until 72 h. On the other hand, both 20 microM E(2) and 25 microM GEN rapidly inhibited GJIC in 6 h and 2 h, respectively. The effects continued until 24 h, but weakened by 72 h. Furthermore, a combined effect at microM level between DES and E(2) on GJIC inhibition was observed, but not between GEN and E(2). DES and E(2) showed GJIC inhibition at low dose levels (nearly physiological estrogen levels) after 72 h, but GEN did not. DES-induced GJIC inhibition at 10 pM and 10 microM was completely counteracted by ICI 182,780 (ICl), an estrogen receptor antagonist. On the other hand, the inhibitory effects on GJIC with E(2) (10 pM and 20 microM) and GEN (25 microM) were partially blocked by ICI or calphostin C, a protein kinase C (PKC) inhibitor, and were completely blocked by the combination of ICI and calphostin C. These results demonstrate that DES inhibits GJIC between Leydig cells via the estrogen receptor (ER), and that E(2) and GEN inhibit GJIC via ER and PKC. These estrogenic compounds may have different individual non-genotoxic mechanism including PKC pathway on testicular carcinogenesis or development.
Insights
Estrogenic compounds like DES, E2, and GEN inhibit gap junctional intercellular communication (GJIC) in mouse Leydig cells. These compounds may impact testicular function through estrogen receptor and protein kinase C pathways.
Area of Science:
- Endocrinology
- Cell Biology
- Toxicology
Background:
- Estrogenic compounds can disrupt testicular function by affecting Leydig cells.
- Gap junctional intercellular communication (GJIC) is crucial for maintaining homeostasis in Leydig cells during development and carcinogenesis.
Purpose of the Study:
- To investigate the effects of diethylstilbestrol (DES), 17beta-estradiol (E2), and genistein (GEN) on GJIC in mouse Leydig TM3 cells.
- To elucidate the mechanisms underlying the observed effects on GJIC.
Main Methods:
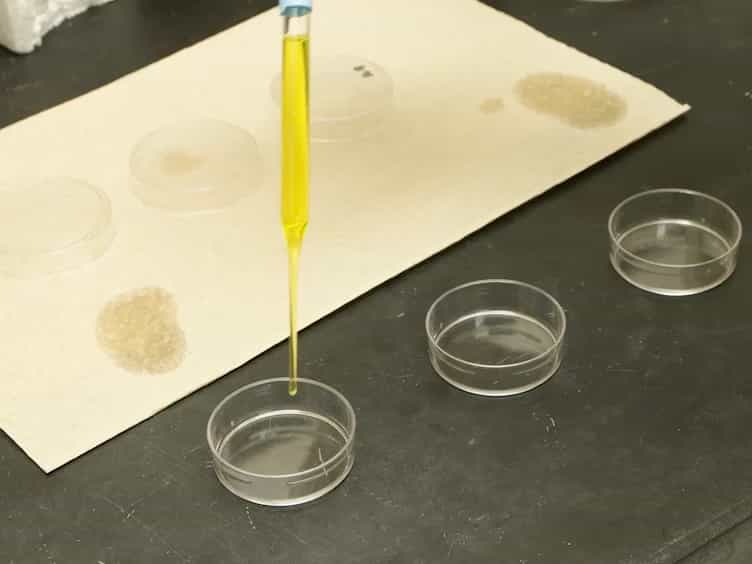
- Lucifer yellow microinjection was used to assess GJIC in mouse Leydig TM3 cells.
- Cells were treated with varying concentrations and time points of DES, E2, and GEN.
- The involvement of estrogen receptor (ER) and protein kinase C (PKC) was examined using specific inhibitors (ICI 182,780 and calphostin C).
Main Results:
- DES, E2, and GEN all inhibited GJIC in TM3 cells.
- DES exhibited sustained inhibition, while E2 and GEN showed rapid but less persistent inhibition.
- DES-induced inhibition was ER-dependent, whereas E2 and GEN involved both ER and PKC pathways.
- Combined effects and dose-dependent inhibition at near-physiological estrogen levels were observed for DES and E2.
Conclusions:
- Estrogenic compounds differentially affect GJIC in Leydig cells.
- DES acts primarily via the estrogen receptor, while E2 and GEN utilize both estrogen receptor and protein kinase C pathways.
- These distinct mechanisms may contribute to non-genotoxic effects of these compounds on testicular carcinogenesis and development.
More Related Videos
Related Concept Videos
 01:37
Gap Junctions
 01:27
Gap Junctions
 01:19
Contact-dependent Signaling
Gap Junctions
In animal cells, gap junctions are formed...
 01:45
Endocrine Signaling