Related Experiment Video
Updated: Jul 21, 2026

Modification and Functionalization of the Guanidine Group by Tailor-made Precursors
Published on: April 27, 2017
Angiotensin-converting enzyme inhibitors: a new mechanism of action
Hongmei Peng1, Oscar A Carretero, Nikola Vuljaj
1Hypertension and Vascular Research Division, Henry Ford Hospital, Detroit, MI 48202-2689, USA.

Background:
Angiotensin-converting enzyme (ACE) inhibitors are valuable agents for the treatment of hypertension, heart failure, and other cardiovascular and renal diseases. The cardioprotective effects of ACE inhibitors are mediated by blockade of both conversion of angiotensin (Ang) I to Ang II and kinin hydrolysis. Here, we report a novel mechanism that may explain the cardiac antifibrotic effect of ACE inhibition, involving blockade of the hydrolysis of N-acetyl-seryl-aspartyl-lysyl-proline (Ac-SDKP).
Methods And Results:
To study the role of Ac-SDKP in the therapeutic effects of the ACE inhibitor captopril, we used a model of Ang II-induced hypertension in rats treated with the ACE inhibitor either alone or combined with a blocking monoclonal antibody (mAb) to Ac-SDKP. These hypertensive rats had left ventricular hypertrophy (LVH) as well as increases in cardiac fibrosis, cell proliferation, transforming growth factor-beta (TGF-beta) expression, and phosphorylation of Smad2 (P-Smad2), a signaling mediator of the effects of TGF-beta. The ACE inhibitor did not decrease either blood pressure or LVH; however, it significantly decreased LV collagen from 13.3+/-0.9 to 9.6+/-0.6 microg/mg dry wt (P<0.006), and this effect was blocked by the mAb (12.1+/-0.6; P<0.034, ACE inhibitor versus ACE inhibitor+mAb). In addition, analysis of interstitial collagen volume fraction and perivascular collagen (picrosirius red staining) showed a very similar tendency. Likewise, the ACE inhibitor significantly decreased LV monocyte/macrophage infiltration, cell proliferation, and TGF-beta expression, and these effects were blocked by the mAb. Ang II increased Smad2 phosphorylation 3.2+/-0.9-fold; the ACE inhibitor lowered this to 0.6+/-0.1-fold (P<0.001), and the mAb blocked this decrease to 2.1+/-0.3 (P<0.001, ACE inhibitor versus ACE inhibitor+mAb). Similar findings were seen when the ACE inhibitor was replaced by Ac-SDKP.
Conclusions:
We concluded that in Ang II-induced hypertension, the cardiac antifibrotic effect of ACE inhibitors is a result of the inhibition of Ac-SDKP hydrolysis, resulting in a decrease in cardiac cell proliferation (probably fibroblasts), inflammatory cell infiltration, TGF-beta expression, Smad2 activation, and collagen deposition.
Related Concept Videos
 01:25
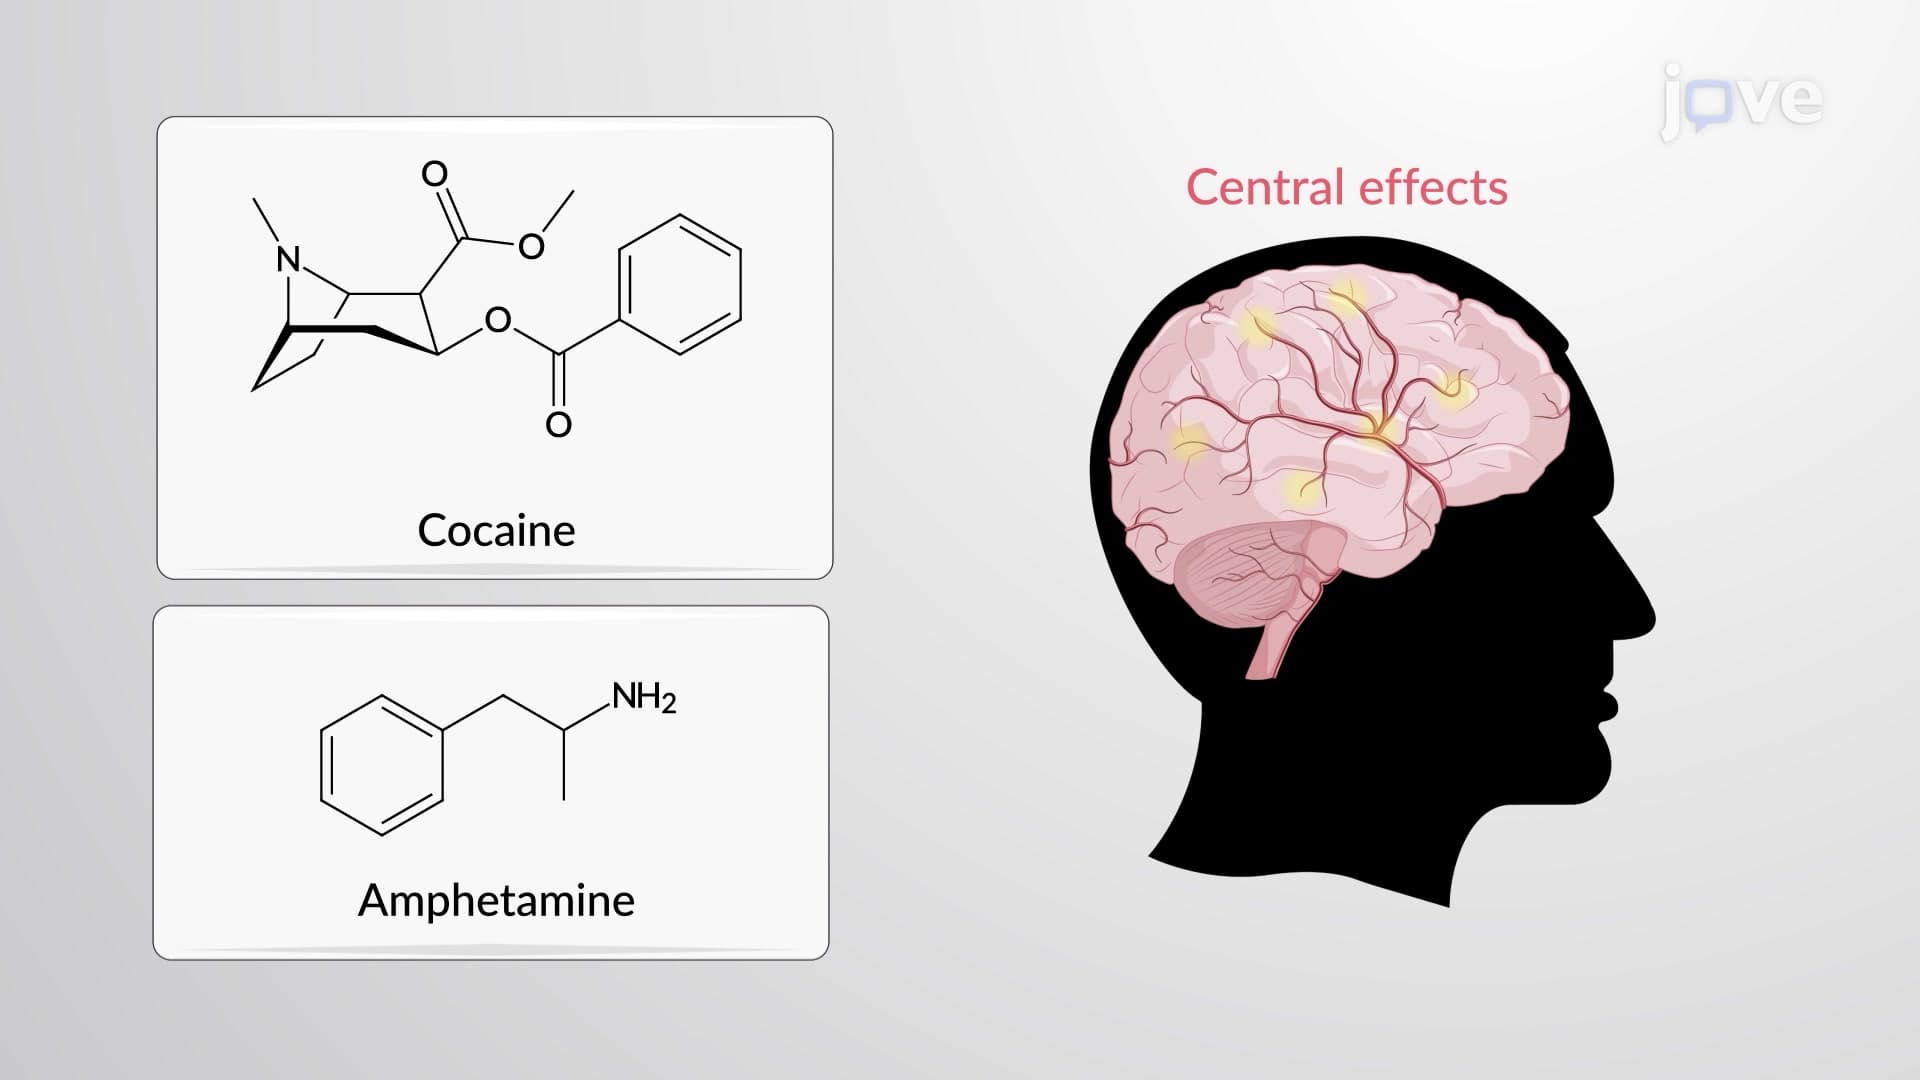
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral bioavailability, and...
 01:29
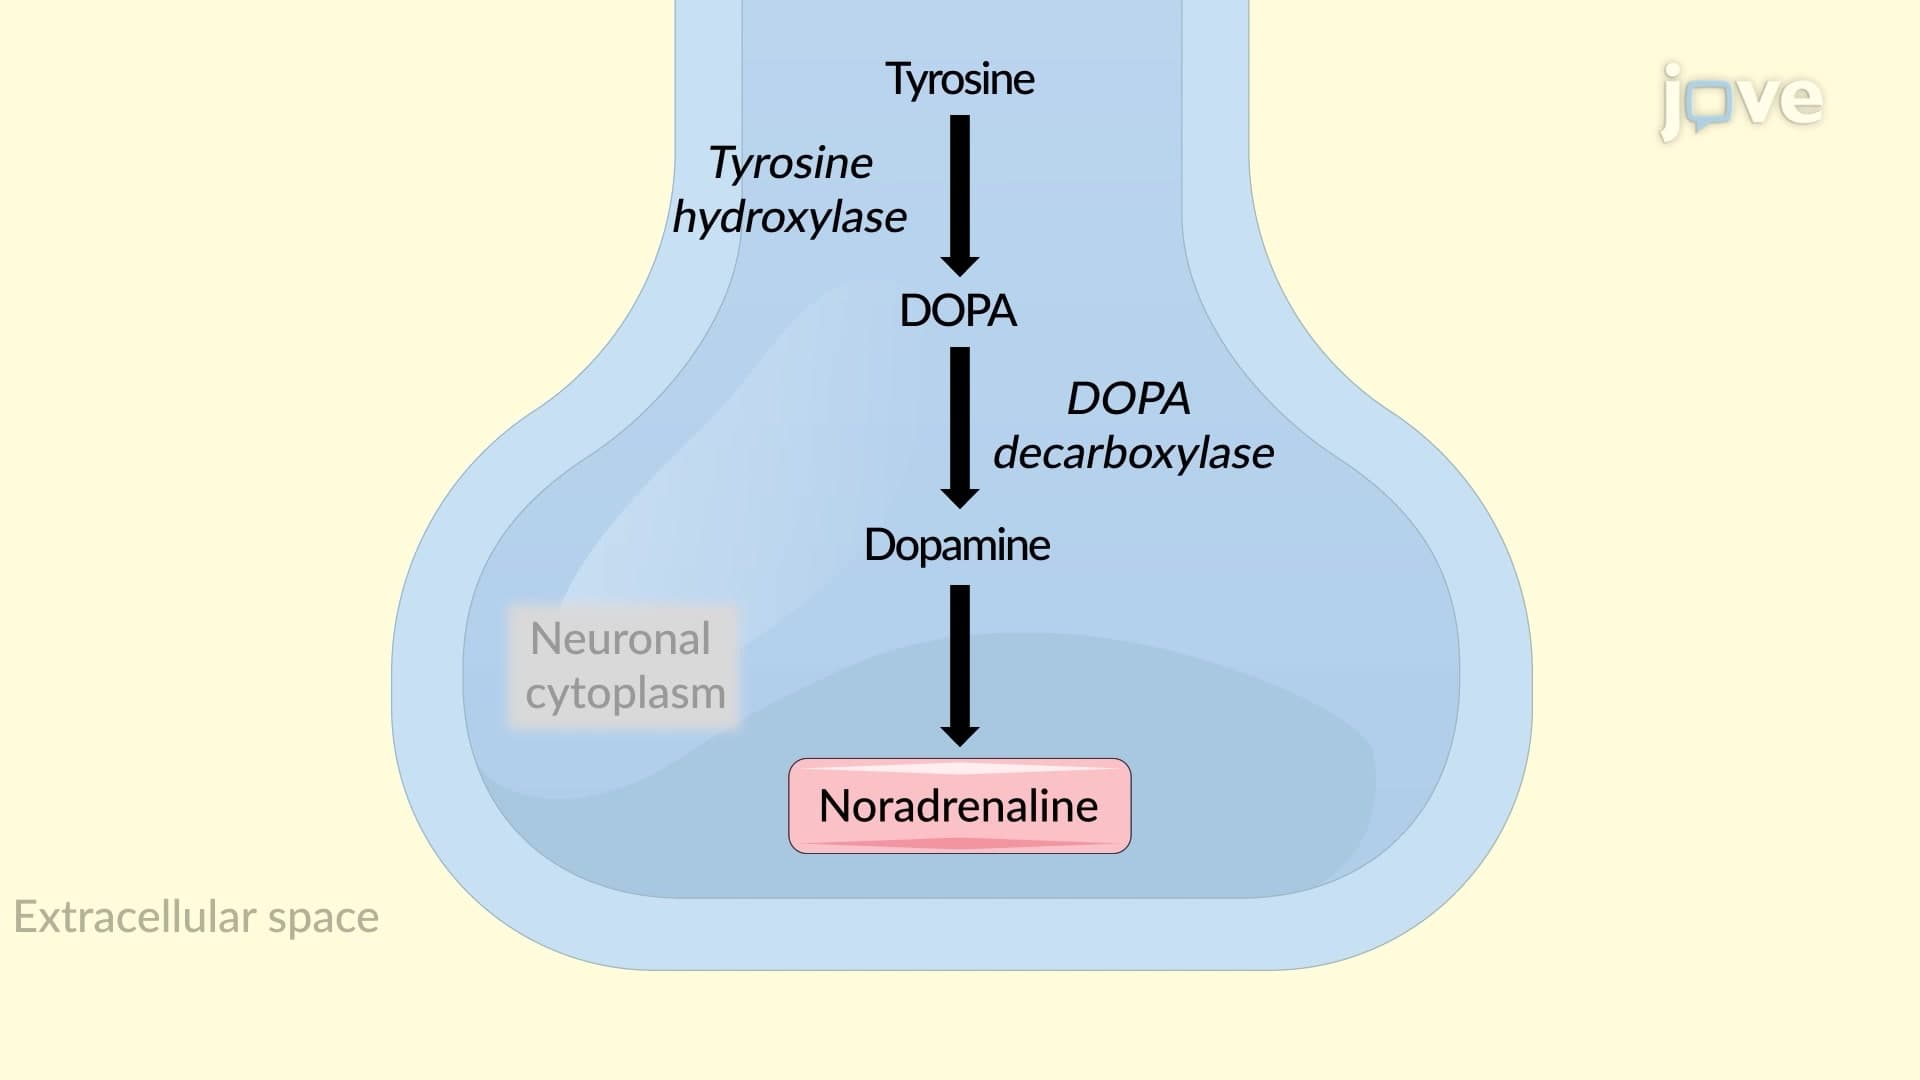
Drugs Affecting Neurotransmitter Synthesis
 01:30
Antihypertensive Drugs: Angiotensin-Converting Enzyme Inhibitors
 01:30
Antihypertensive Drugs: Angiotensin II Receptor Blockers
 01:25
Antihypertensive Drugs: Direct Renin Inhibitors
 01:18
Treatment for Pulmonary Arterial Hypertension: Endothelin Receptor Antagonists
ETs are synthesized through a complex sequence of enzymatic steps, primarily involving an enzyme referred to as endothelin-converting enzyme (ECE). Of...