Related Experiment Video
Updated: Aug 15, 2026


Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
Better prediction of protein contact number using a support vector regression analysis of amino acid sequence
1Institute for Molecular Bioscience, ARC Centre in Bioinformatics, The University of Queensland, St. Lucia, 4072, Australia. z.yuan@imb.uq.edu.au

Background:
Protein tertiary structure can be partly characterized via each amino acid's contact number measuring how residues are spatially arranged. The contact number of a residue in a folded protein is a measure of its exposure to the local environment, and is defined as the number of Cbeta atoms in other residues within a sphere around the Cbeta atom of the residue of interest. Contact number is partly conserved between protein folds and thus is useful for protein fold and structure prediction. In turn, each residue's contact number can be partially predicted from primary amino acid sequence, assisting tertiary fold analysis from sequence data. In this study, we provide a more accurate contact number prediction method from protein primary sequence.
Results:
We predict contact number from protein sequence using a novel support vector regression algorithm. Using protein local sequences with multiple sequence alignments (PSI-BLAST profiles), we demonstrate a correlation coefficient between predicted and observed contact numbers of 0.70, which outperforms previously achieved accuracies. Including additional information about sequence weight and amino acid composition further improves prediction accuracies significantly with the correlation coefficient reaching 0.73. If residues are classified as being either "contacted" or "non-contacted", the prediction accuracies are all greater than 77%, regardless of the choice of classification thresholds.
Conclusion:
The successful application of support vector regression to the prediction of protein contact number reported here, together with previous applications of this approach to the prediction of protein accessible surface area and B-factor profile, suggests that a support vector regression approach may be very useful for determining the structure-function relation between primary protein sequence and higher order consecutive protein structural and functional properties.


Related Concept Videos
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:04
Protein-protein Interfaces
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 02:15
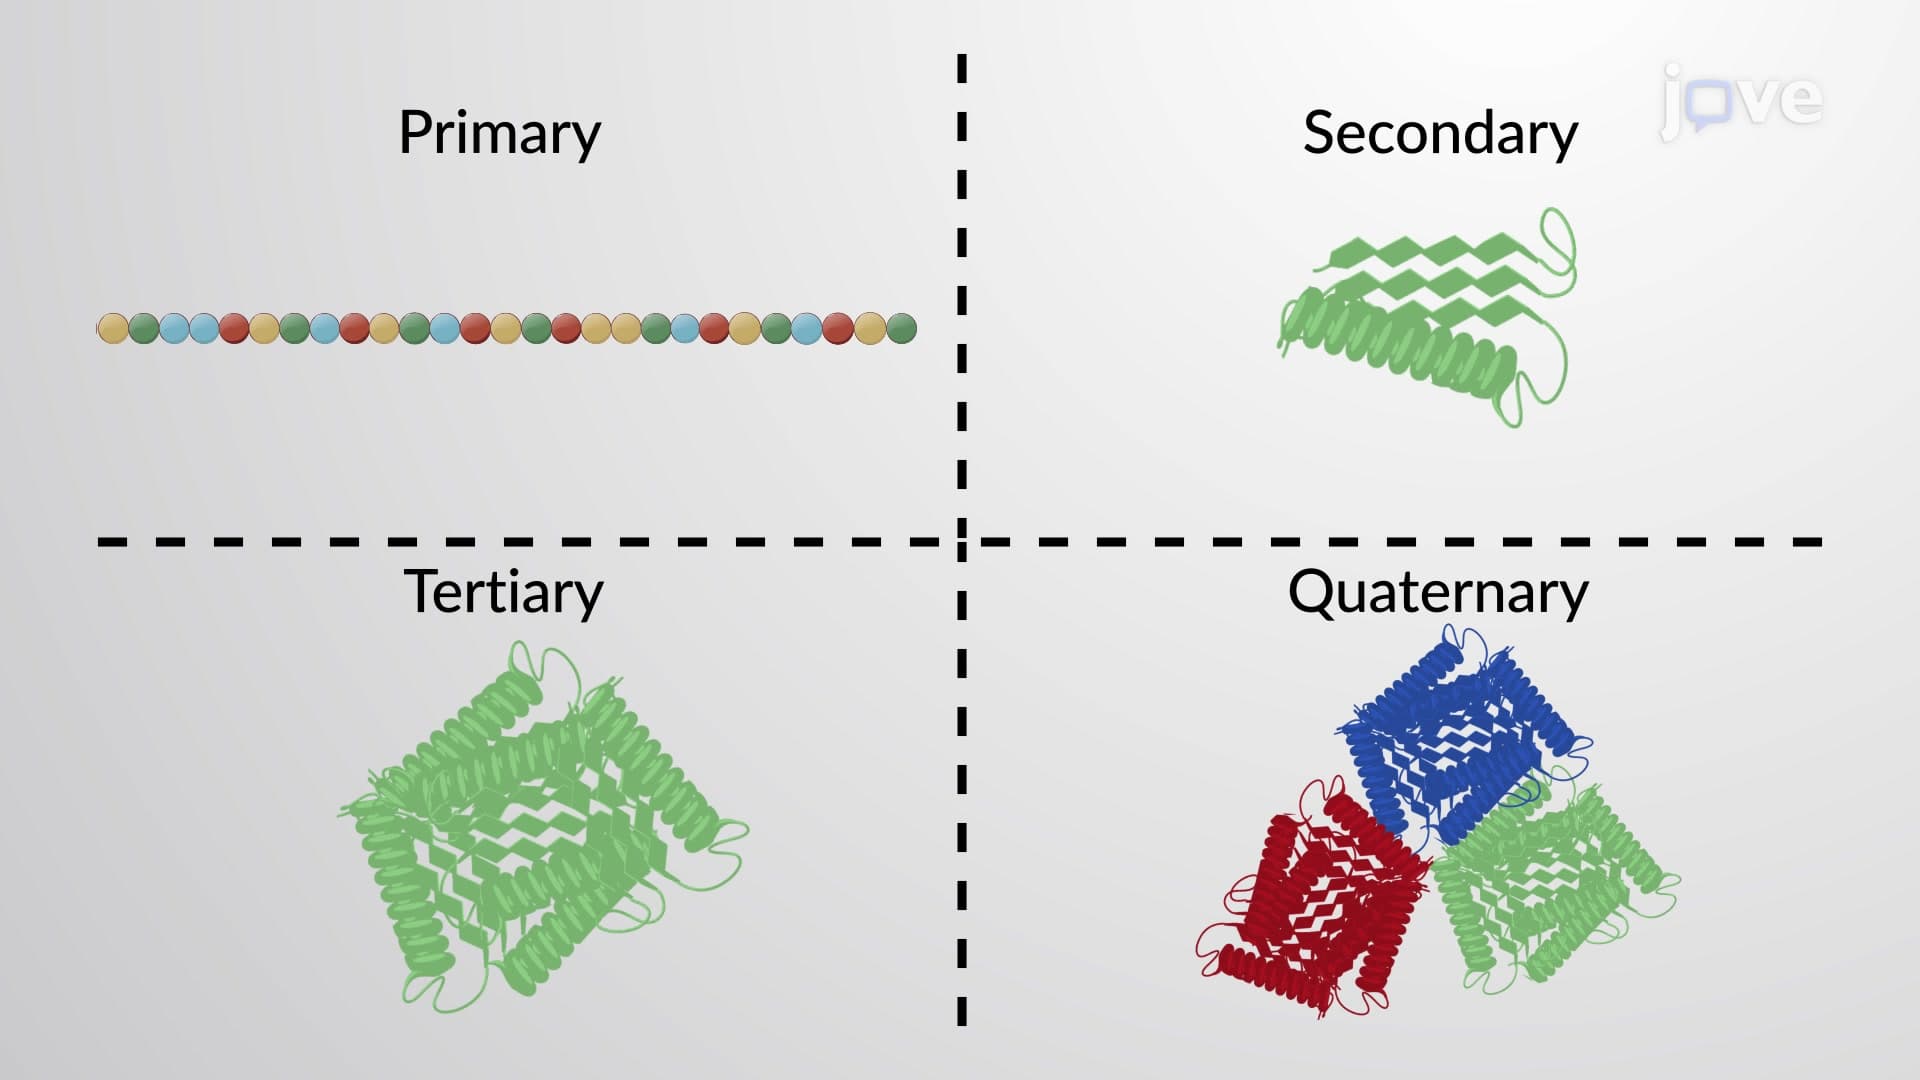
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme can...
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence.