
Density functional theory calculations for endohedral complexes of non-pi C60H60 cage with small guest molecules
1Department of Chemical and Biological Engineering, State University of New York at Buffalo, Amherst, New York 14260, USA. yhu@buffalo.edu

Abstract:
Hybrid density-functional theory (B3LYP) calculations were carried out to determine the structures and energies of endohedral complexes of non-pi C(60)H(60) with H(2), CO, and LiH. It was demonstrated that the endohedral complexes of C(60)H(60) with the above three guest molecules are more stable than the corresponding complexes with C(60). Furthermore, the interaction between C(60)H(60) and the inside H(2) or CO is negligible, but the formation of the LiH-C(60)H(60) complex is exothermic with a stabilization energy of -6.0 kcal/mol. While the bond lengths of H(2) and CO changed a little when placed inside the cages, that of the LiH molecule increased and decreased inside C(60)H(60) and C(60), respectively.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:42
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:40
MO Theory and Covalent Bonding
 01:18
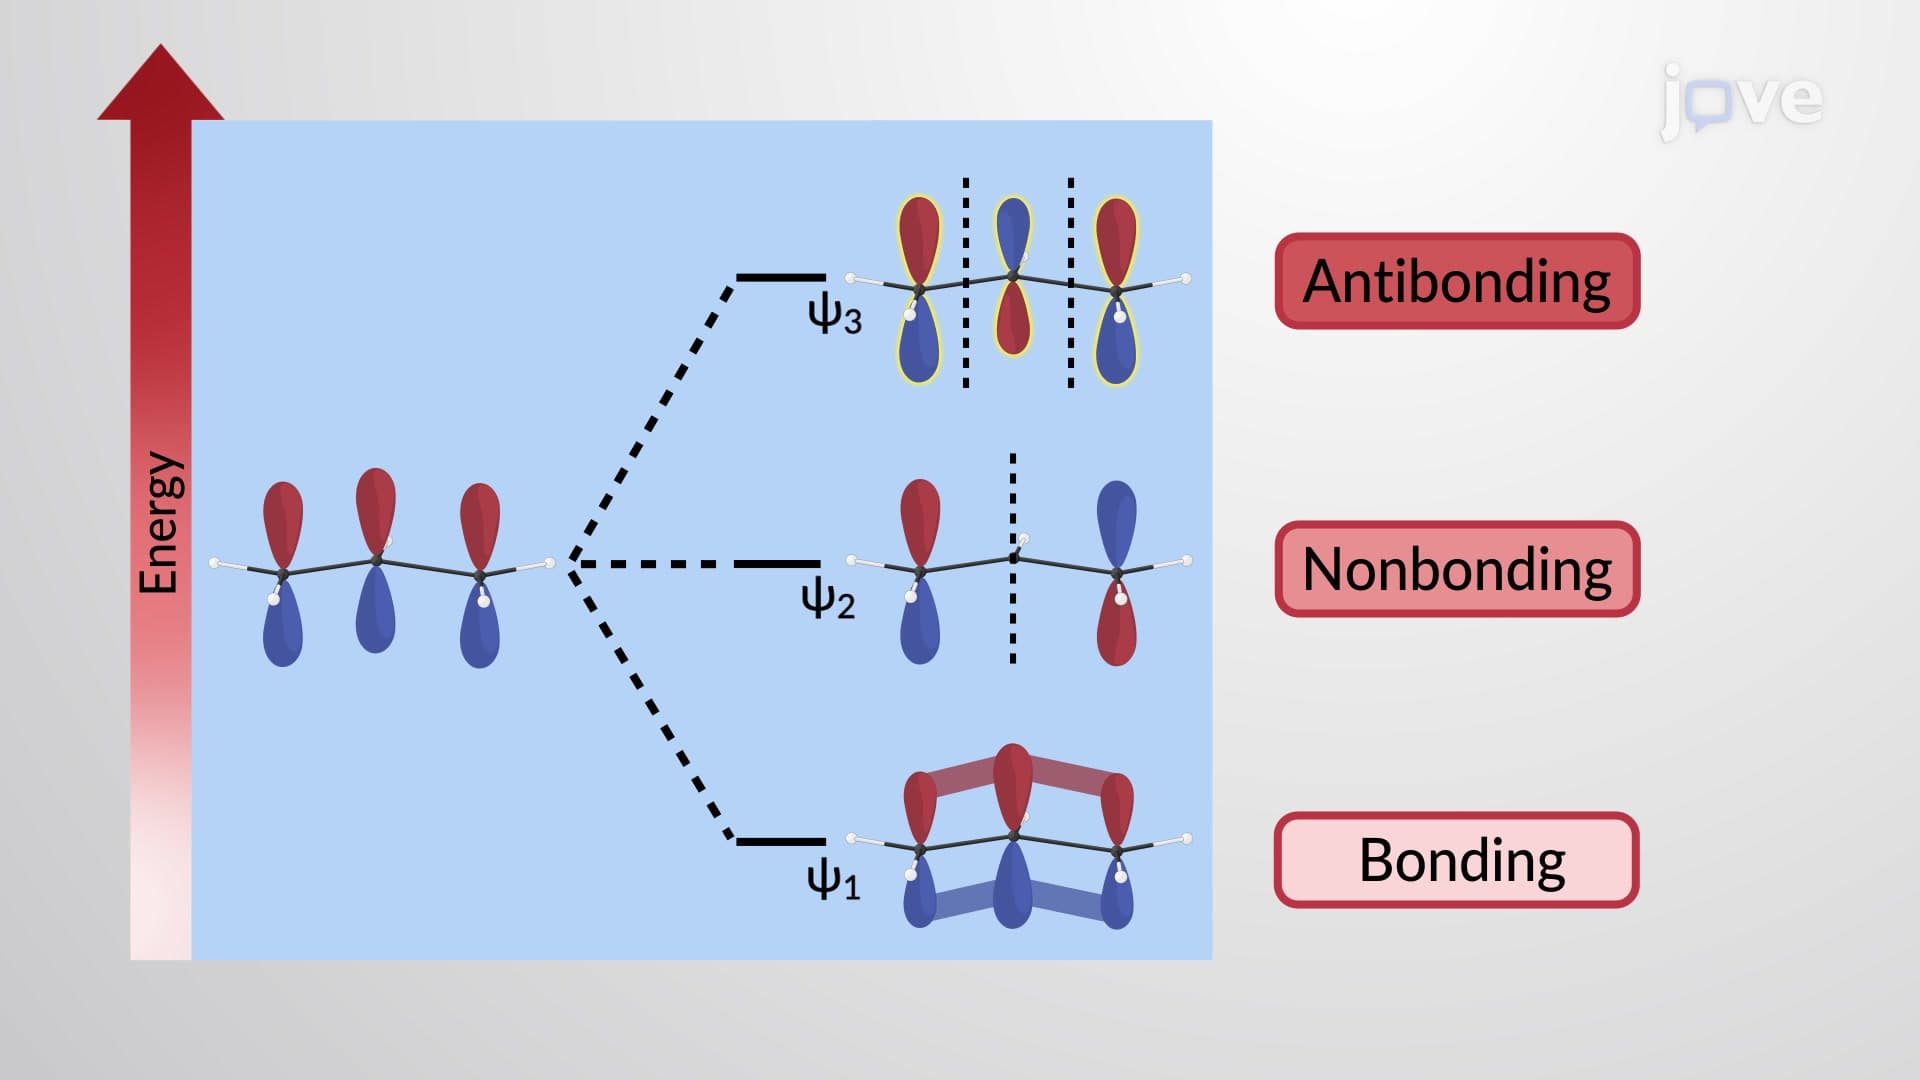
π Molecular Orbitals of the Allyl Cation and Anion