A simplified relativistic time-dependent density-functional theory formalism for the calculations of excitation energies including spin-orbit coupling effect
Related Experiment Video
Updated: Jul 9, 2026
Recombination Dynamics in Thin-film Photovoltaic Materials via Time-resolved Microwave Conductivity
Published on: March 6, 2017
1Department of Chemistry, University of Calgary, Calgary, Alberta T2N 1N4, Canada.

Abstract:
In the present work we have proposed an approximate time-dependent density-functional theory (TDDFT) formalism to deal with the influence of spin-orbit coupling effect on the excitation energies for closed-shell systems. In this formalism scalar relativistic TDDFT calculations are first performed to determine the lowest single-group excited states and the spin-orbit coupling operator is applied to these single-group excited states to obtain the excitation energies with spin-orbit coupling effects included. The computational effort of the present method is much smaller than that of the two-component TDDFT formalism and this method can be applied to medium-size systems containing heavy elements. The compositions of the double-group excited states in terms of single-group singlet and triplet excited states are obtained automatically from the calculations. The calculated excitation energies based on the present formalism show that this formalism affords reasonable excitation energies for transitions not involving 5p and 6p orbitals. For transitions involving 5p orbitals, one can still obtain acceptable results for excitations with a small truncation error, while the formalism will fail for transitions involving 6p orbitals, especially 6p1/2 spinors.
More Related Videos


Related Concept Videos
 02:18
The Bohr Model
 02:45
The Quantum-Mechanical Model of an Atom
 03:21
The Energies of Atomic Orbitals
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 02:38
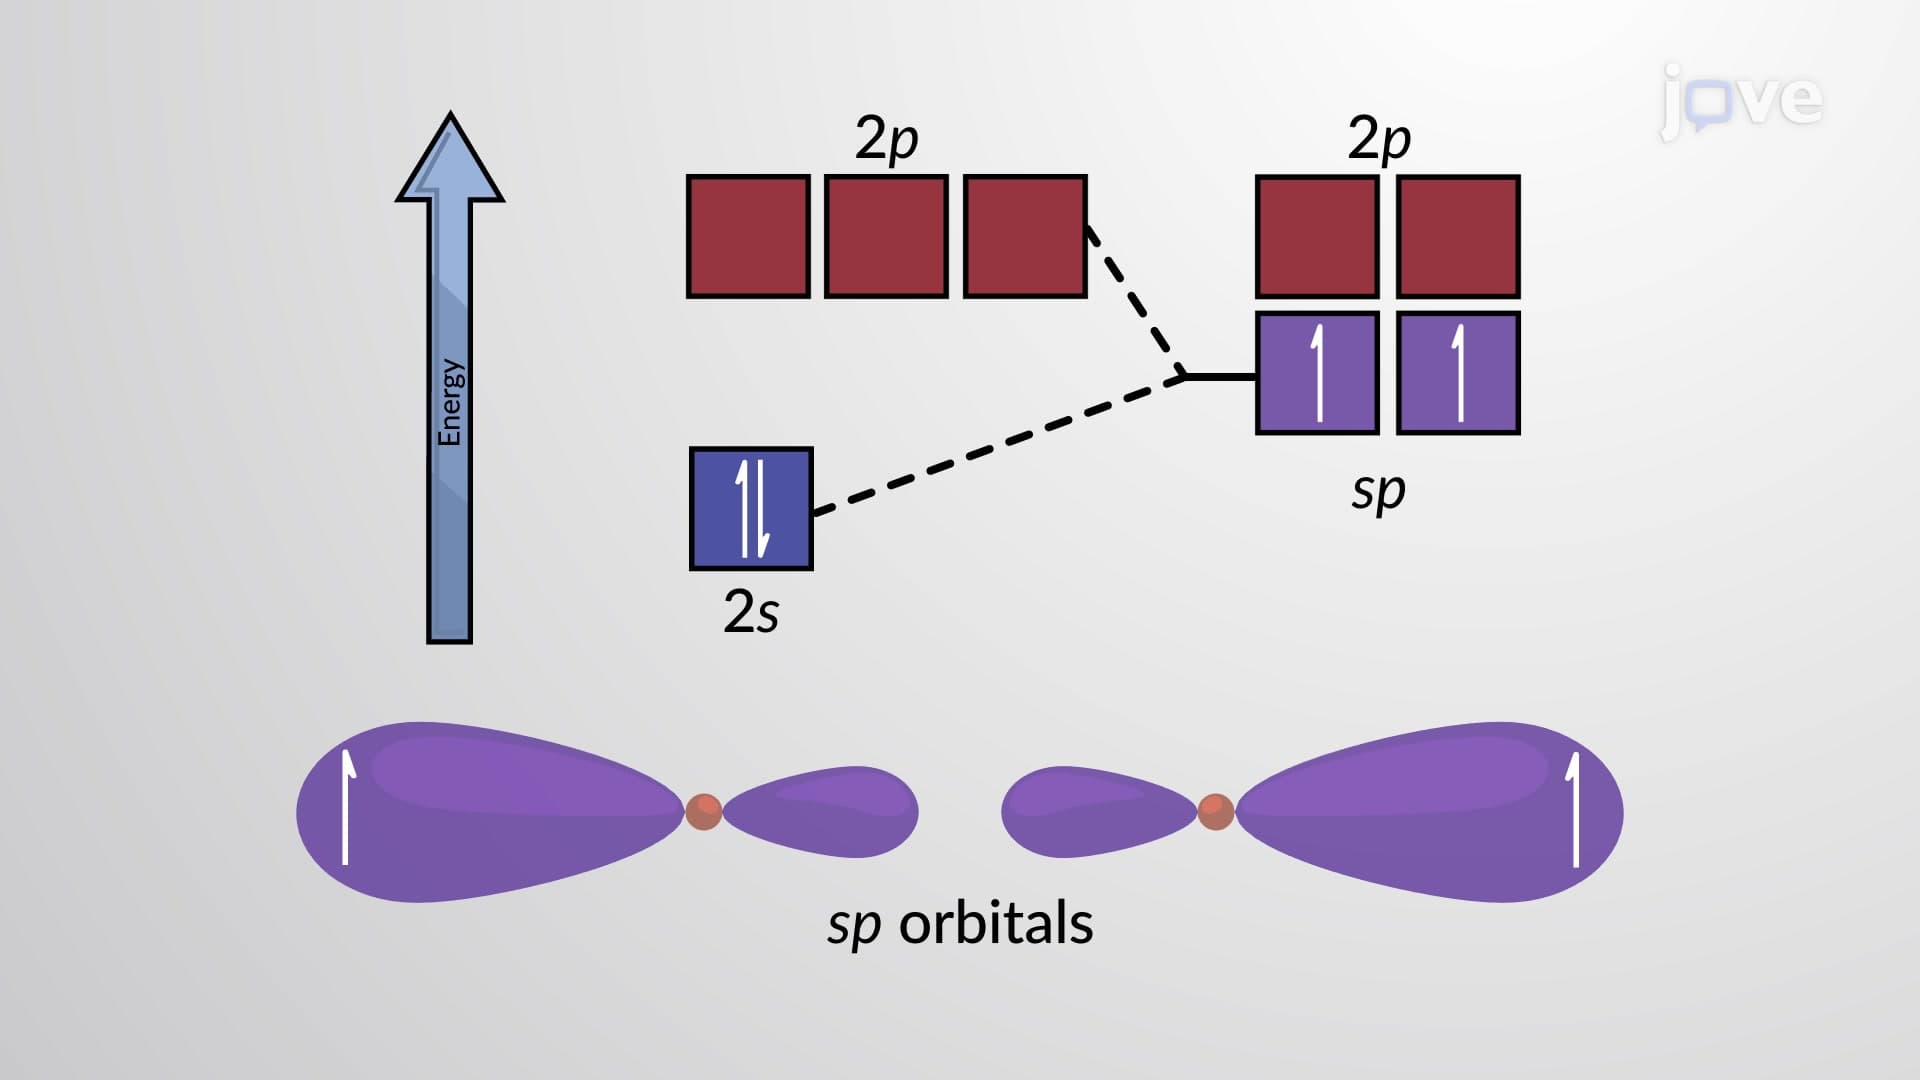
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...