Related Experiment Video
Updated: Aug 15, 2026

Visualization of Pseudomonas aeruginosa within the Sputum of Cystic Fibrosis Patients
Published on: July 16, 2020
Positive expiratory pressure changes aerosol distribution in patients with cystic fibrosis
Beth L Laube1, David E Geller, Ta-Chun Lin
1Department of Pediatrics, Johns Hopkins University, Baltimore, Maryland 21287-2533, USA. blaube@jhmi.edu

Hypothesis:
We hypothesized that aerosol distribution in the lungs of patients with cystic fibrosis changes with positive expiratory pressure (PEP).
Methods:
Eight patients were randomized to one of 2 conditions. On one study day, patients inhaled saline aerosol containing 99mtechnetium generated by a Pari LC Plus nebulizer and exhaled through a Pari PEP device. On another day, the same patients exhaled through a low-resistance Pari filter (no PEP). Afterwards, they underwent gamma-camera lung imaging. Images were analyzed for lung deposition fraction, expressed as a percent of the initial nebulizer activity, and deposition pattern, expressed in terms of inner-outer and apical-basal ratios.
Results:
Lung deposition fraction was significantly lower with the Pari PEP device; the mean + SD deposition fraction was 6.10 + 3.05% (median 6.20%) with PEP, compared to 10.76 + 4.52% (median 10.32%) (p = 0.0078) without PEP. The inner-outer ratio was 2.01 + 0.69 (median 2.23) with PEP, which was significantly lower than without PEP (2.76 + 1.33, median 2.55) (p = 0.004). The apical-basal ratio was 0.82 + 0.31 (median 0.80) with PEP, which was not significantly different from no PEP (1.00 + 0.49, median 0.90).
Conclusion:
These results indicate that less aerosol is deposited in the lungs of patients with cystic fibrosis when the Pari LC Plus nebulizer is used with the Pari PEP device, as described in these experiments. Nevertheless, aerosol administration with this nebulizer and PEP device also results in a proportional redistribution of aerosol to the peripheral airways, compared to nebulization without the PEP device. The clinical relevance of this subtle redistribution of aerosol in cystic fibrosis patients will probably depend on the drug administered and disease severity.
Related Concept Videos
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:24
Chest Physiotherapy
Purpose
CPT is primarily used for patients with excessive bronchial secretions who have difficulty clearing...
 01:24
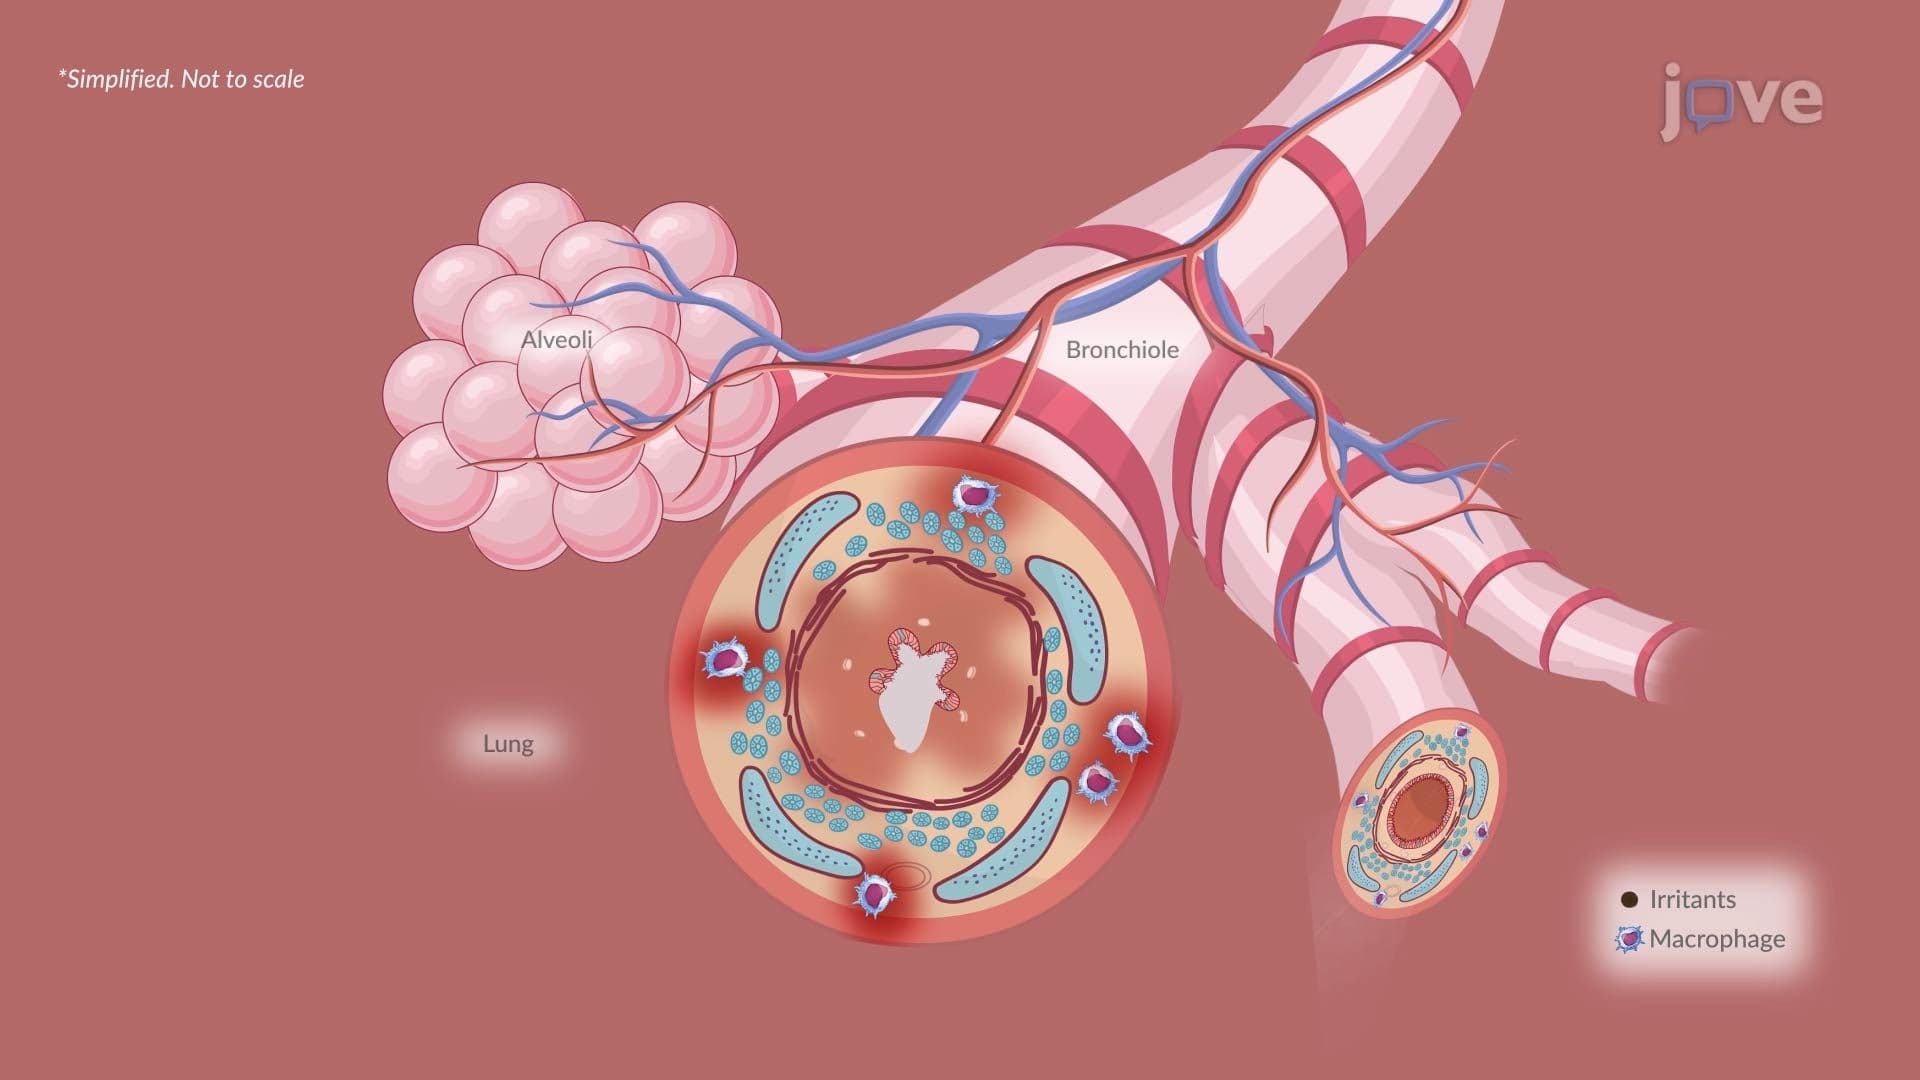
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features
 01:23
Mechanical Ventilation III: Noninvasive Ventilation
Noninvasive Positive-Pressure Ventilation (NIPPV)
 01:25
Pulmonary Function Tests
Pulmonary Function Tests are crucial diagnostic tools for assessing respiratory function, particularly in patients with chronic respiratory disorders. They comprehensively evaluate lung volumes, ventilatory function, breathing mechanics, diffusion, and gas exchange. These tests help diagnose pulmonary diseases and play a significant role in monitoring disease progression, evaluating disability, and assessing response to therapy.
PFTs involve using a spirometer, a...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...