Related Experiment Video
Updated: Aug 15, 2026
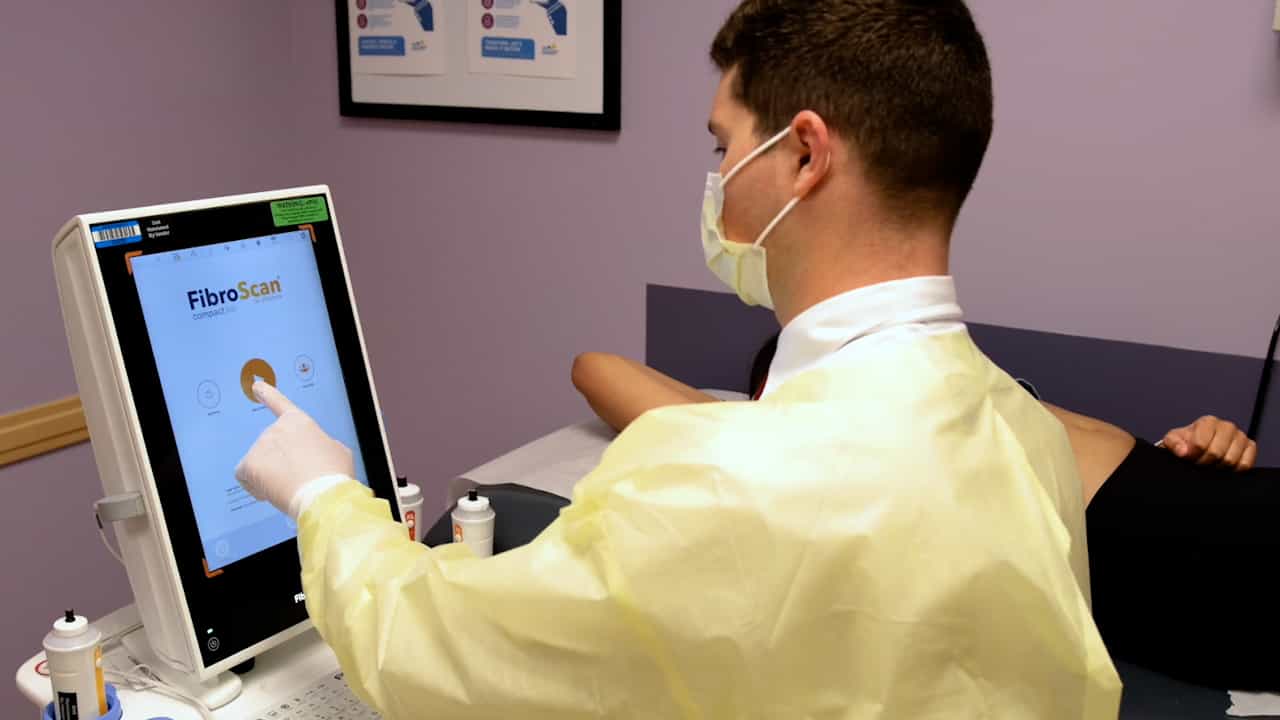

Implementation of Non-invasive Point of Care Transient Elastography for Evaluation of Liver Disease in Pediatric Populations with Cystic Fibrosis
Published on: August 29, 2025
Surgical considerations in cystic fibrosis: a 32-year evaluation of outcomes
Mauricio A Escobar1, Jay L Grosfeld, Justin J Burdick
1Section of Pediatric Surgery, Department of Surgery, Indiana University School of Medicine, Indianapolis, Indiana, USA.

Insights
Cystic fibrosis (CF) patients undergoing surgery show improved long-term survival rates, reaching up to 89% into their fourth decade. Early meconium ileus (MI) can lead to later complications, and pneumothorax indicates a poor prognosis in CF patients.
Area of Science:
- Pediatric Surgery
- Cystic Fibrosis Research
- Gastrointestinal Surgery
Background:
- Limited data exists on long-term surgical outcomes for cystic fibrosis (CF) patients.
- Operative interventions are increasingly necessary for CF patients.
Purpose of the Study:
- To analyze long-term operative outcomes, including morbidity and survival, in pediatric patients with cystic fibrosis (CF).
- To identify risk factors and late complications associated with surgery in CF patients.
Main Methods:
- Retrospective review of CF patients who underwent operations between 1972 and 2004 at a tertiary children's hospital.
- Analysis of patient demographics, operative procedures, long-term morbidity, and survival rates.
- Extended follow-up evaluation for up to 35 years.
Main Results:
- 226 CF patients underwent 422 operations; 109 were neonates.
- Meconium ileus (MI) and its complications (jejunoileal atresia, meconium ileus equivalent, fibrosing colonopathy) required surgical intervention in a significant proportion.
- Long-term operative morbidity was low (<1% after 10 years).
- Overall survival was 89%, with 22 deaths primarily due to CF-related pulmonary complications, notably pneumothorax (50% mortality in affected children).
Conclusions:
- Long-term survival for cystic fibrosis (CF) patients undergoing surgery has significantly improved, with many reaching their fourth decade.
- Meconium ileus (MI) may predispose to late gastrointestinal complications.
- Pneumothorax is a critical indicator of mortality in CF patients.
- Children with CF are viable surgical candidates but require lifelong monitoring for exocrine dysfunction.
Background:
Information concerning long-term operative outcomes in patients with cystic fibrosis (CF) is relatively sparse in the operative literature.
Methods:
A retrospective review of CF patients with operative conditions was performed (1972-2004) at a tertiary children's hospital to analyze outcomes including long-term morbidity and survival.
Results:
A total of 226 patients with CF presented with an operative diagnosis (113 men, 113 women). A total of 422 operations were performed in 213 patients (94%). The mean age at operation was 4.1 +/- 6.2 years (range, 1 d to 26 y) and 109 were neonates. Fifteen of 42 (36%) babies with simple meconium ileus (MI) were treated nonoperatively with hypertonic enemas, 27 of 42 and all 45 patients with complicated MI required operation, including 15 with jejunoileal atresia (17%). Seventeen of 27 (63%) patients with meconium ileus equivalent had MI as neonates; 7 of 27 (26%) required operation. Eight of 9 (89%) with fibrosing colonopathy required operation. Organ transplantation was required in 21 patients. Follow-up evaluation was possible in 204 of 213 (96%) patients. The duration of follow-up evaluation was 14.9 +/- 8.5 years (range, 2 mo to 35 y). Operative morbidity was 11% at 1 year, 2% at 2 to 4 years, 1% at 5 to 10 years, and less than 1% at more than 10 years. There were 24 deaths (11%); 22 followed CF-related pulmonary complications and included 8 of 16 (50%) children with pneumothorax.
Conclusions:
Long-term survival in CF patients has improved significantly (89%), with many surviving into the fourth decade. MI may predispose to late complications including meconium ileus equivalent and fibrosing colonopathy. Pneumothorax in CF patients is an ominous predictor of mortality. Children with CF are living longer and are good candidates for operation, but require long-term follow-up evaluation because of ongoing exocrine dysfunction.
Related Concept Videos
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...