Related Experiment Video
Updated: Aug 11, 2026

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Aminoacylase I deficiency: a novel inborn error of metabolism
R N Van Coster1, E A Gerlo, T G Giardina
1Department of Pediatrics, Division of Neurology and Metabolism, University Hospital, Ghent, Belgium. rudy.vancoster@ugent.be

Abstract:
This is the first report of a patient with aminoacylase I deficiency. High amounts of N-acetylated amino acids were detected by gas chromatography-mass spectrometry in the urine, including the derivatives of serine, glutamic acid, alanine, methionine, glycine, and smaller amounts of threonine, leucine, valine, and isoleucine. NMR spectroscopy confirmed these findings and, in addition, showed the presence of N-acetylglutamine and N-acetylasparagine. In EBV transformed lymphoblasts, aminoacylase I activity was deficient. Loss of activity was due to decreased amounts of aminoacylase I protein. The amount of mRNA for the aminoacylase I was decreased. DNA sequencing of the encoding ACY1 gene showed a homozygous c.1057 C>T transition, predicting a p.Arg353Cys substitution. Both parents were heterozygous for the mutation. The mutation was also detected in 5/161 controls. To exclude the possibility of a genetic polymorphism, protein expression studies were performed showing that the mutant protein had lost catalytic activity.
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:22
Lysosomal Hydrolases
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:20
Inborn Errors of Metabolism
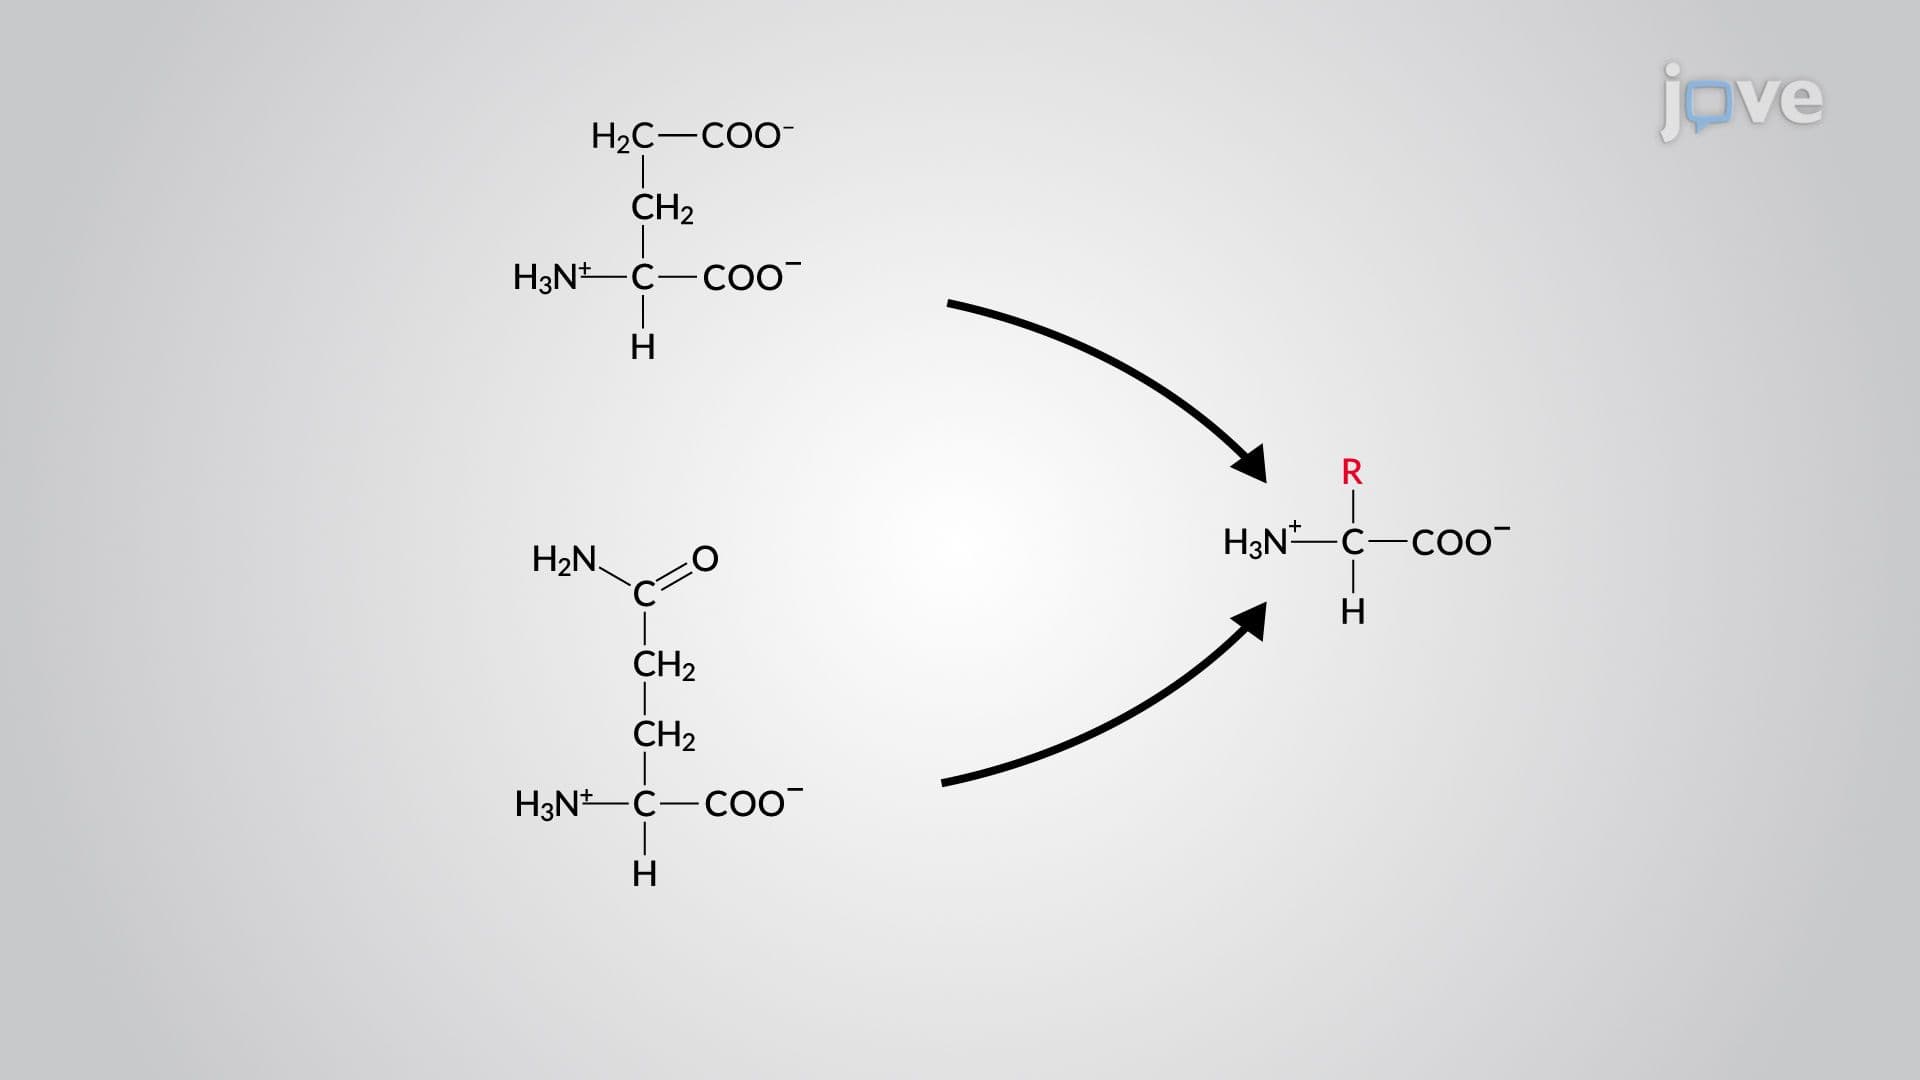 01:29
Amino Acid Biosynthetic Pathways