Related Experiment Video
Updated: Aug 14, 2026


A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Interstrand pairing patterns in beta-barrel membrane proteins: the positive-outside rule, aromatic rescue, and strand
1Department of Bioengineering, SEO, MC-063, University of Illinois at Chicago, 851 S. Morgan Street, Room 218, Chicago, IL 60607-7052, USA.

Abstract:
beta-Barrel membrane proteins are found in the outer membrane of Gram-negative bacteria, mitochondria, and chloroplasts. Little is known about how residues in membrane beta-barrels interact preferentially with other residues on adjacent strands. We have developed probabilistic models to quantify propensities of residues for different spatial locations and for interstrand pairwise contact interactions involving strong H-bonds, side-chain interactions, and weak H-bonds. Using the reference state of exhaustive permutation of residues within the same beta-strand, the propensity values and p-values measuring statistical significance are calculated exactly by analytical formulae we have developed. Our findings show that there are characteristic preferences of residues for different membrane locations. Contrary to the "positive-inside" rule for helical membrane proteins, beta-barrel membrane proteins follow a significant albeit weaker "positive-outside" rule, in that the basic residues Arg and Lys are disproportionately favored in the extracellular cap region and disfavored in the periplasmic cap region. We find that different residue pairs prefer strong backbone H-bonded interstrand pairings (e.g. Gly-aromatic) or non-H-bonded pairings (e.g. aromatic-aromatic). In addition, we find that Tyr and Phe participate in aromatic rescue by shielding Gly from polar environments. We also show that these propensities can be used to predict the registration of strand pairs, an important task for the structure prediction of beta-barrel membrane proteins. Our accuracy of 44% is considerably better than random (7%). It also significantly outperforms a comparable registration prediction for soluble beta-sheets under similar conditions. Our results imply several experiments that can help to elucidate the mechanisms of in vitro and in vivo folding of beta-barrel membrane proteins. The propensity scales developed in this study will also be useful for computational structure prediction and for folding simulations.
More Related Videos
 07:08
Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
 06:50
Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
Related Concept Videos
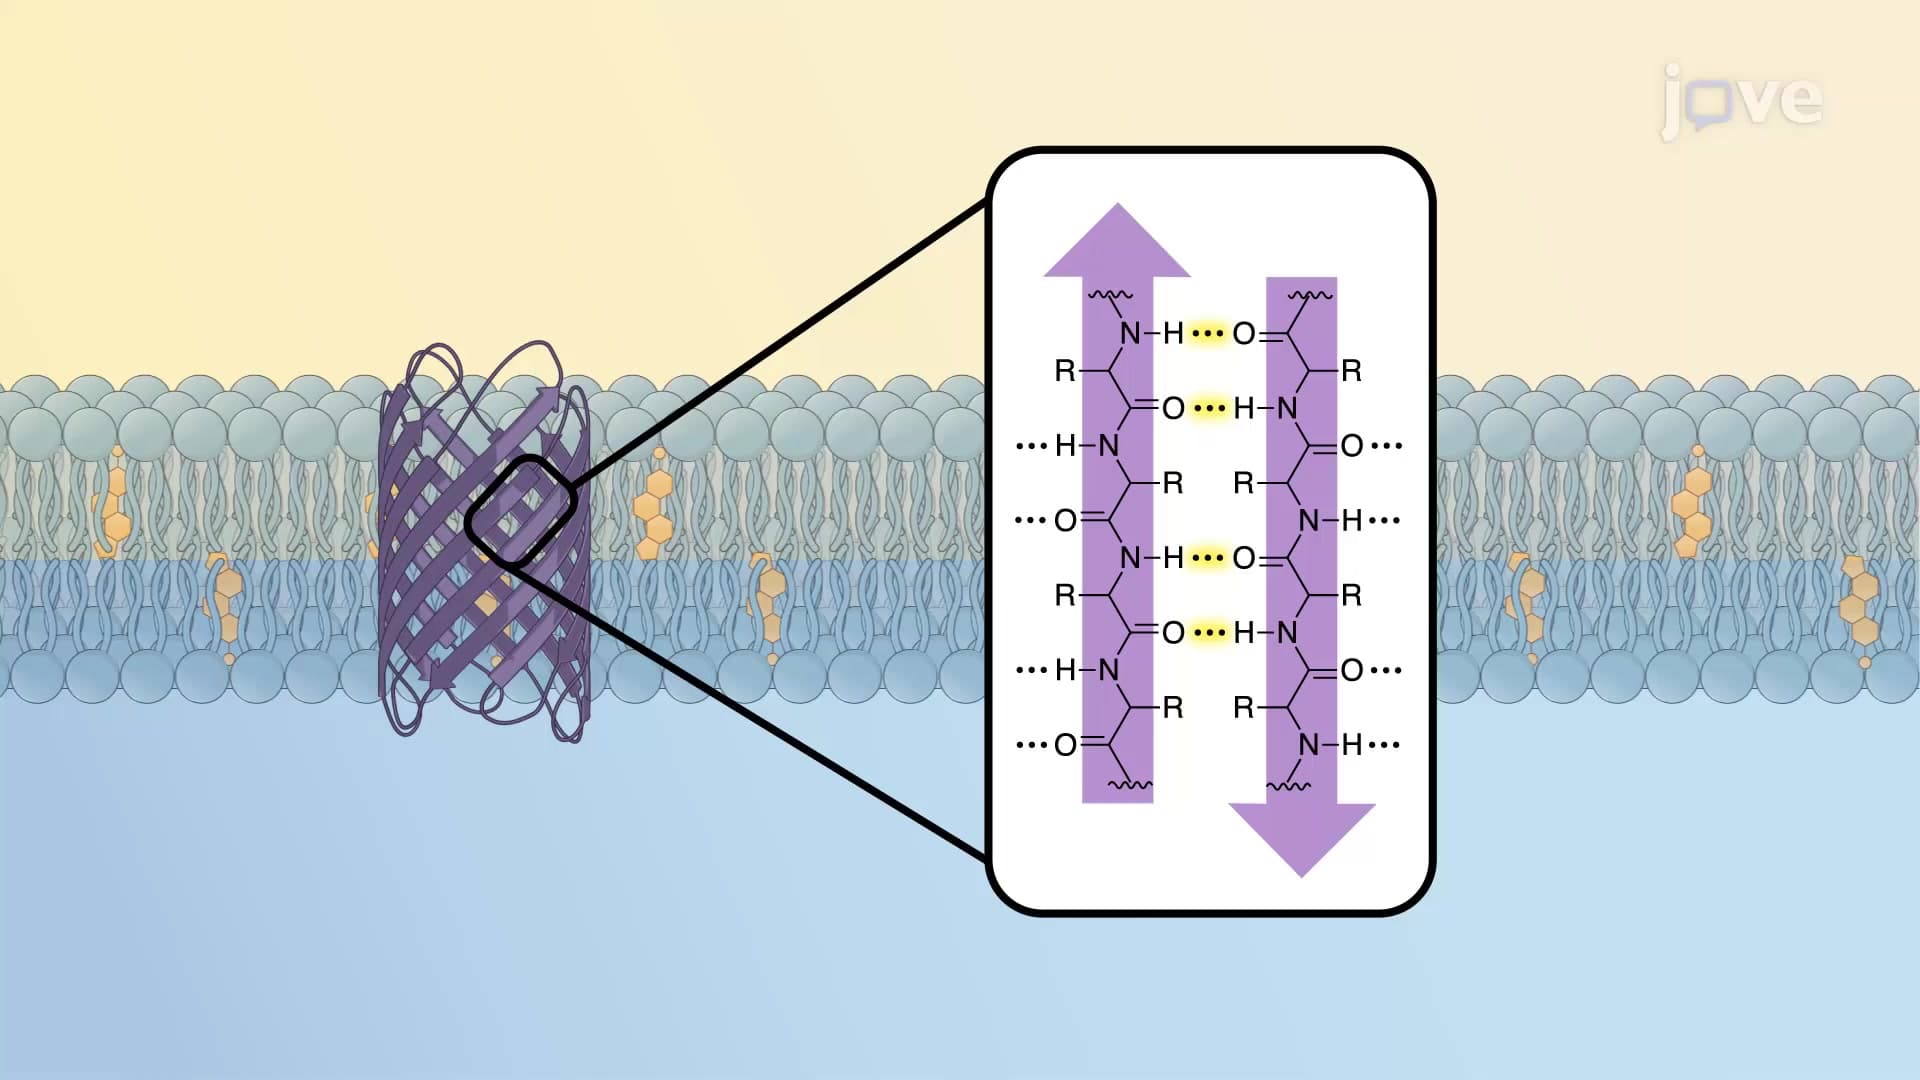 01:09
Multi-pass Transmembrane Proteins and β-barrels
α-Helix containing multi-pass transmembrane proteins
Multi-pass transmembrane proteins such as G-protein-linked receptors (GPCRs) and...
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
01:22Protein Folding
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...