Related Experiment Video
Updated: Aug 14, 2026

An Aptamer-based Sensor for Unchelated Gadolinium(III)
Published on: January 9, 2017
Quantum chemical investigation of hyperfine coupling constants on first coordination sphere water molecule of
Oleg V Yazyev1, Lothar Helm, Vladimir G Malkin
1Laboratoire de Chimie Inorganique et Bioinorganique, Ecole Polytechnique Fédérale de Lausanne, EPFL-BCH, CH-1015 Lausanne, Switzerland.

Abstract:
Hyperfine interactions (HFI) on the nuclei of the first coordination sphere water molecules in a model [Gd(H(2)O)(8)](3+) aqua complex and in the magnetic resonance imaging contrast agent [Gd(DOTA)(H(2)O)](-) were studied theoretically. Density functional theory (DFT) calculations combined with classical molecular dynamics (MD) simulations have been used in order to take into account dynamic effects in aqueous solution. DFT relativistic calculations show a strong spin-polarization of the first coordination sphere water molecules. This spin-polarization leads to a positive (17)O isotropic hyperfine coupling constant (A(iso)((17)O) = 0.58 +/- 0.11 MHz) and to a significant increase of the effective distance (r(eff)(Gd-O) = 2.72 +/- 0.06 A) of dipolar interaction compared to the mean internuclear distance (r(Gd-O) = 2.56 +/- 0.06 A) obtained from the MD trajectory of [Gd(DOTA)(H(2)O)](-) in aqueous solution. The point-dipole model for anisotropic hyperfine interaction overestimates therefore the longitudinal relaxation rate of the (17)O nucleus by approximately 45%. The (1)H isotropic hyperfine coupling constant of the bound water molecule is predicted to be very small (A(iso)((1)H) = 0.03 +/- 0.02 MHz), and the point-dipole approximation for first coordination sphere water protons holds. The calculated hyperfine parameters are in good agreement with available experimental data.
Related Concept Videos
 02:42
Valence Bond Theory
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human eye.
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:17
Spin–Spin Coupling: One-Bond Coupling
 00:43
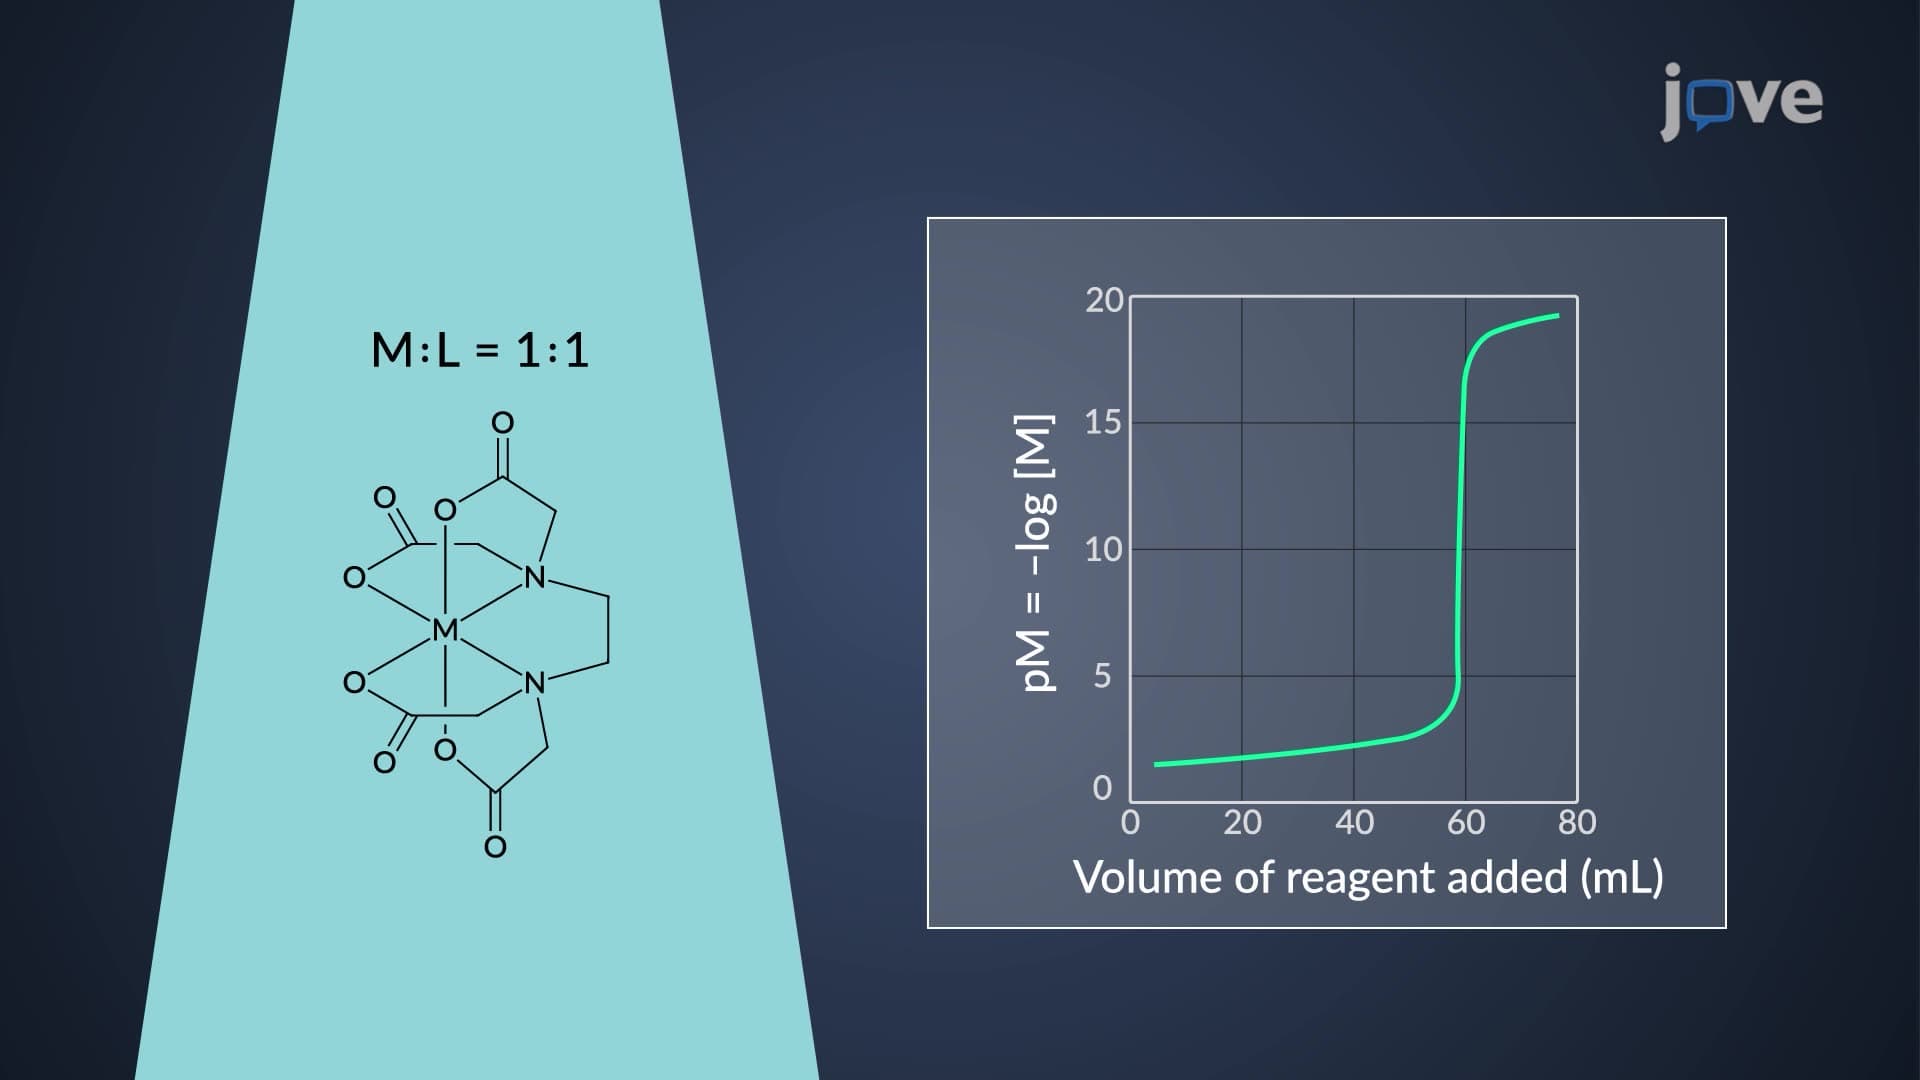
Complexometric Titration: Ligands