Related Experiment Video
Updated: Aug 14, 2026
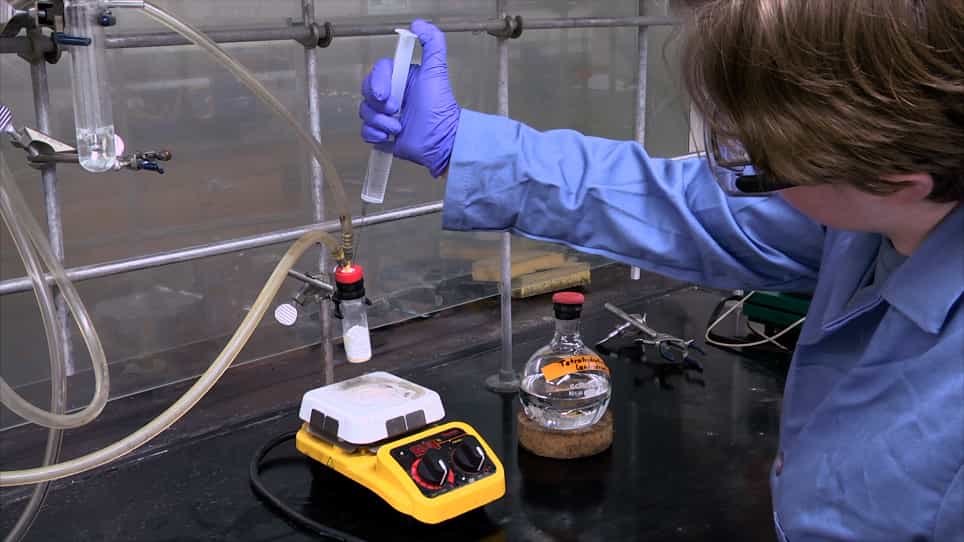

Synthesis of a Borylated Ibuprofen Derivative Through Suzuki Cross-Coupling and Alkene Boracarboxylation Reactions
Published on: November 30, 2022
Synthetic and reaction chemistry of heteroatom stabilized boryl and cationic borylene complexes
Deborah L Kays Née Coombs1, Andrea Rossin, Joanne K Day
1Centre for Fundamental and Applied Main Group Chemistry, School of Chemistry, Cardiff University, UK.

Abstract:
The synthesis, spectroscopic and structural characterization of the aryloxy and amino functionalized chloroboryl complexes (eta(5)-C(5)R(5))Fe(CO)(2)B(OMes)Cl (R = H, 2a ; R = Me, 3a) and (eta(5)-C(5)H(5))Fe(CO)(2)B(N(i)Pr(2))Cl (7a) are reported. Compound 2ais shown to be a versatile substrate for further boron-centred substitution chemistry leading to the asymmetric boryl complexes (eta(5)-C(5)H(5))Fe(CO)(2)B(OMes)ER(n) [ER(n) = OC(6)H(4)(t)Bu-4, 2c; ER(n) = SPh, 2d] with retention of the metal-boron bond. The reactivities of 2a, 3a and 7a towards the halide abstraction agent Na[BAr(f)(4)] have also been examined, in order to investigate the potential for the generation of cationic heteroatom-stabilized terminal borylene complexes. The application of this methodology to the mesityloxy derivatives and gives rise to B-F containing products, presumably via fluoride abstraction from the [BAr(f)(4)](-) counter-ion. By contrast, amino-functionalized complex 7a is more amenable to this approach, and the thermally robust terminal aminoborylene complex [(eta(5)-C(5)H(5))Fe(CO)(2)B(N(i)Pr(2))][BAr(f)(4)] (9) can be isolated in ca. 50% yield. The reactivity of 9towards a range of nucleophilic and/or unsaturated reagents has been examined, with examples of addition, protonolysis and metathesis chemistries having been established.
More Related Videos
Related Concept Videos
 03:08
Hydroboration-Oxidation of Alkenes
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 02:47
Alkynes to Aldehydes and Ketones: Hydroboration-Oxidation
One of the convenient methods for the preparation of aldehydes and ketones is via hydration of alkynes. Hydroboration-oxidation of alkynes is an indirect hydration reaction in which an alkyne is treated with borane followed by oxidation with alkaline peroxide to form an enol that rapidly converts into an aldehyde or a ketone. Terminal alkynes form aldehydes, whereas internal alkynes give ketones as the final product.
 02:46
Halogenation of Alkenes
Consider the bromination of cyclopentene. Molecular bromine is polarized in the proximity of the π electrons of cyclopentene. An electrophilic bromine atom adds across the double bond, forming a cyclic bromonium ion intermediate.
 01:16
Cycloaddition Reactions: MO Requirements for Thermal Activation
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.