Related Experiment Video
Updated: Jan 3, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
DFT-based chemical reactivity indices in the Hartree-Fock method. II. Fukui function, chemical potential, and
Robert Balawender1, Paul Geerlings
1Institute of Physical Chemistry of the Polish Academy of Sciences (PAS), Kasprzaka 44/52, PL-01-224 Warsaw, Poland. balawend@ichf.edu.pl

Abstract:
A derivation of the density-functional-theory- (DFT) based reactivity indices in the ensemble unrestricted Hartree-Fock (eUHF) method is presented. The comparison between the properties of the reactivity indices evaluated in one and two sets of spin-orbital approach of the eUHF and hyper-unrestricted Hartree-Fock (UHF) methods are shown. All approaches give similar Fukui function irrespective of methodology used, but significantly differ for the global indices, containing important chemical information, and so their interpretation in terms of DFT- based indices can be questionable. The calculation scheme for the indices using the first- and second-order coupled perturbed eHF equations is proposed. A method for the identification of the spinorbitals involved in the change of the total number of electrons is included. The illustrative examples (water and hydrogen cyanide) show that the ground-state (GS) properties of the (Z +/- 1)-electron systems can be predicted from the GS properties of the Z-electron systems with an accuracy comparable with the UHF calculations. The relaxation effect, important for the HCN system in which a change in the symmetry of the highest-occupied spin-orbital occurs, is effectively predicted.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 03:02
Acid/Base Strengths and Dissociation Constants
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:10
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 01:13
Relative Reactivity of Carboxylic Acid Derivatives
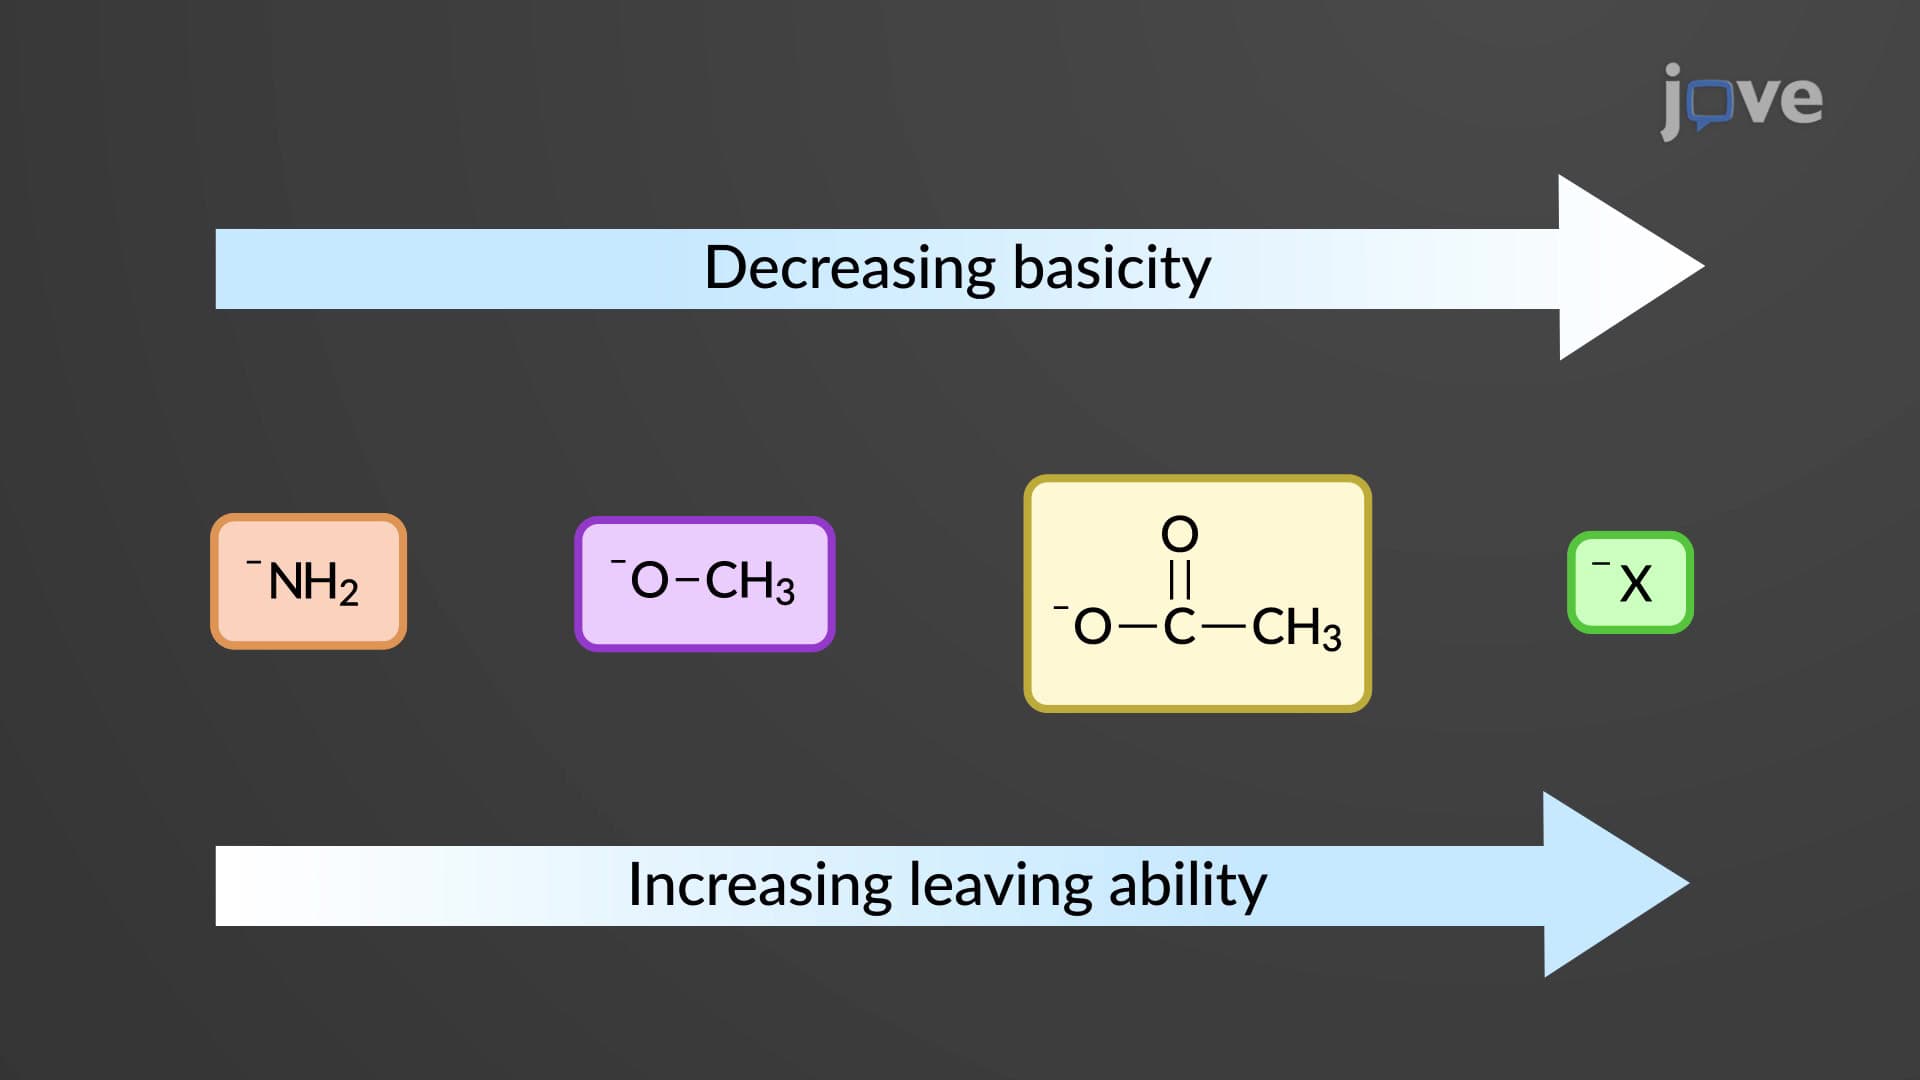
A key factor in assessing the reactivity of the acid derivatives is the basicity of the substituent or the leaving group. The lower the basicity of the leaving group, the higher the reactivity of the derivative. The basicity of the leaving group follows this order:
Halide ions < Acyloxy ions < Alkoxy ions < Amine ions
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:27
π Electron Effects on Chemical Shift: Overview