Related Experiment Video
Updated: Aug 13, 2026

Spatial Separation of Molecular Conformers and Clusters
Published on: January 9, 2014
Coexistence of 1,3-butadiene conformers in ionization energies and Dyson orbitals
Saumitra Saha1, Feng Wang, Chantal T Falzon
1Centre for Molecular Simulation, Swinburne University of Technology, P.O. Box 218, Hawthorn, Melbourne, Victoria 3122, Australia.

Abstract:
The minimum-energy structures on the torsional potential-energy surface of 1,3-butadiene have been studied quantum mechanically using a range of models including ab initio Hartree-Fock and second-order Møller-Plesset theories, outer valence Green's function, and density-functional theory with a hybrid functional and statistical average orbital potential model in order to understand the binding-energy (ionization energy) spectra and orbital cross sections observed by experiments. The unique full geometry optimization process locates the s-trans-1,3-butadiene as the global minimum structure and the s-gauche-1,3-butadiene as the local minimum structure. The latter possesses the dihedral angle of the central carbon bond of 32.81 degrees in agreement with the range of 30 degrees-41 degrees obtained by other theoretical models. Ionization energies in the outer valence space of the conformer pair have been obtained using Hartree-Fock, outer valence Green's function, and density-functional (statistical average orbital potentials) models, respectively. The Hartree-Fock results indicate that electron correlation (and orbital relaxation) effects become more significant towards the inner shell. The spectroscopic pole strengths calculated in the Green's function model are in the range of 0.85-0.91, suggesting that the independent particle picture is a good approximation in the present study. The binding energies from the density-functional (statisticaly averaged orbital potential) model are in good agreement with photoelectron spectroscopy, and the simulated Dyson orbitals in momentum space approximated by the density-functional orbitals using plane-wave impulse approximation agree well with those from experimental electron momentum spectroscopy. The coexistence of the conformer pair under the experimental conditions is supported by the approximated experimental binding-energy spectra due to the split conformer orbital energies, as well as the orbital momentum distributions of the mixed conformer pair observed in the orbital cross sections of electron momentum spectroscopy.
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 01:18
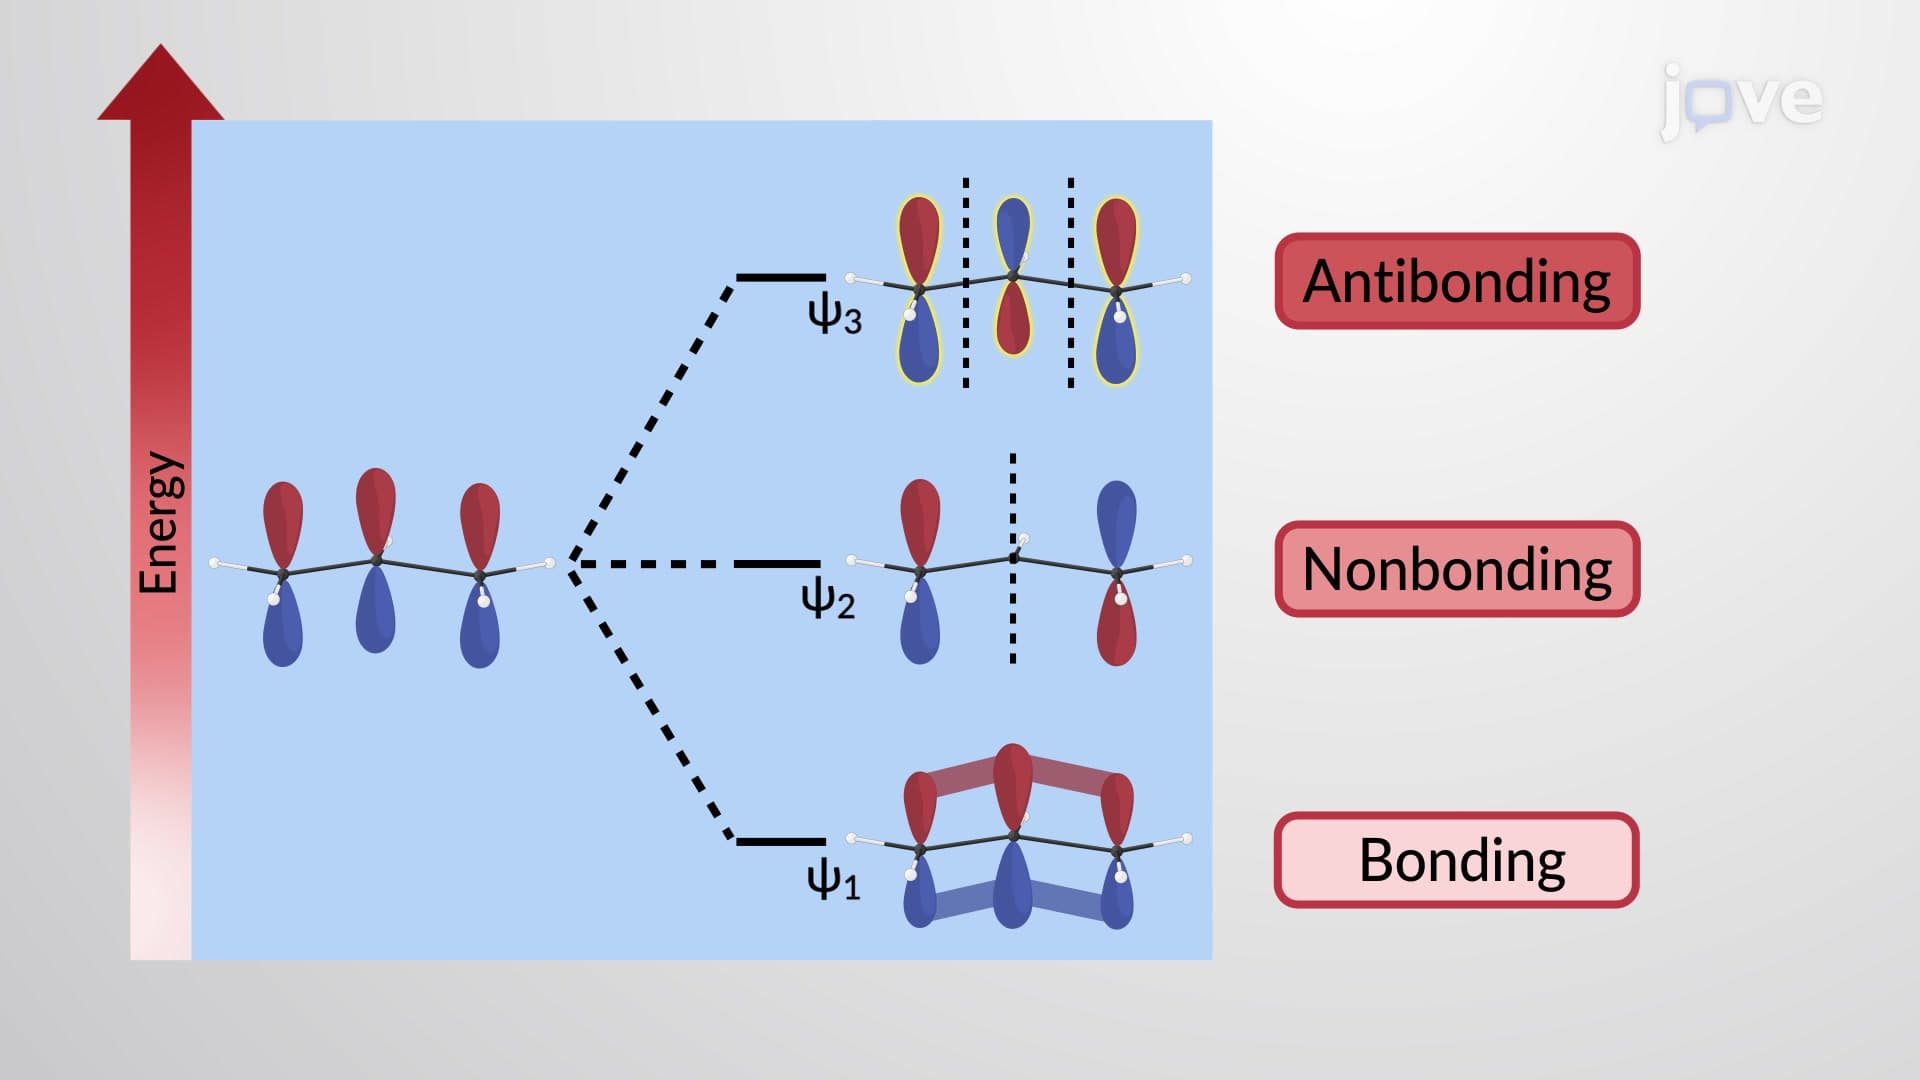
π Molecular Orbitals of the Allyl Cation and Anion
 01:16
Structure of Conjugated Dienes
Conjugated dienes are compounds characterized by the presence of alternating double and single bonds. In a conjugated system like 1,3-butadiene, the unhybridized 2p orbital on each carbon overlaps continuously, allowing the π electrons to be delocalized across the entire molecule. In contrast, this type of overlap does not occur in cumulated and isolated dienes, such as 2,3-pentadiene and 1,4-pentadiene, respectively. Instead, the π electrons remain localized between the double...
 02:20
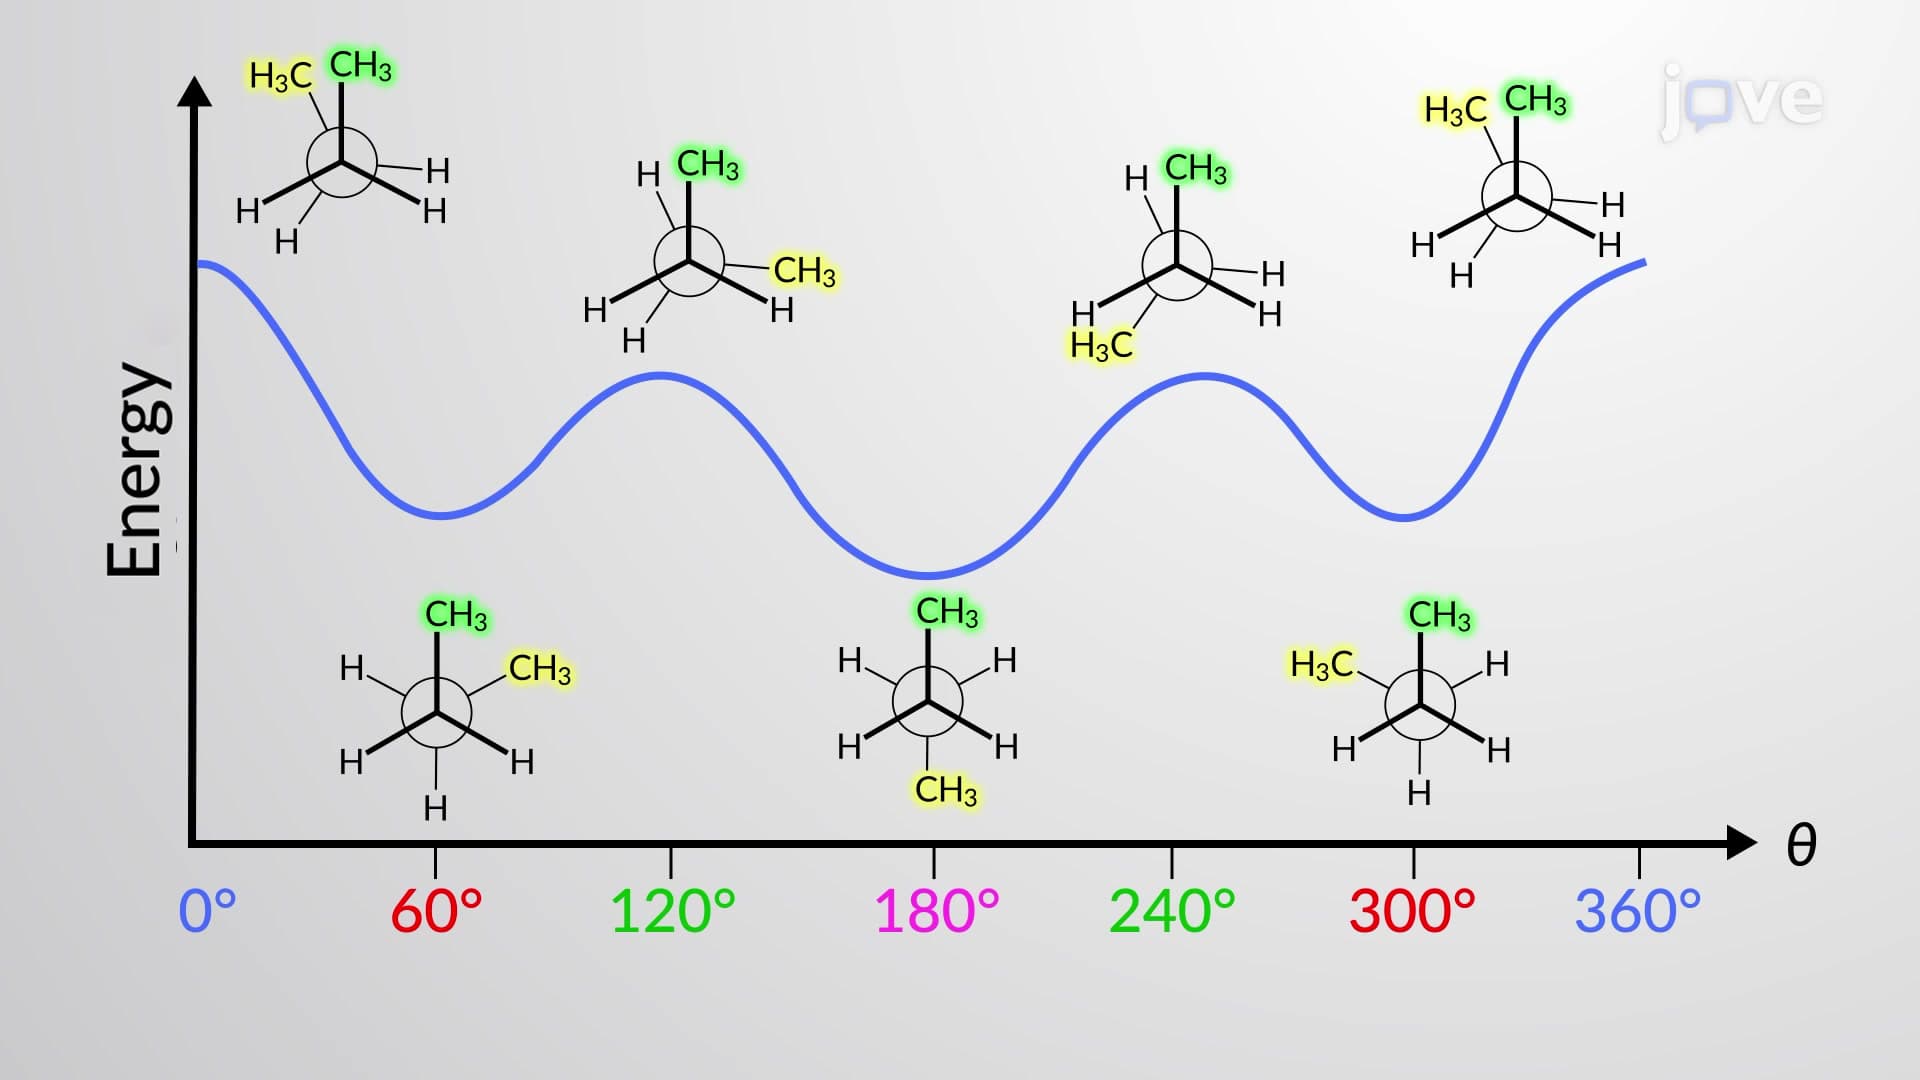
Conformations of Butane
 02:18
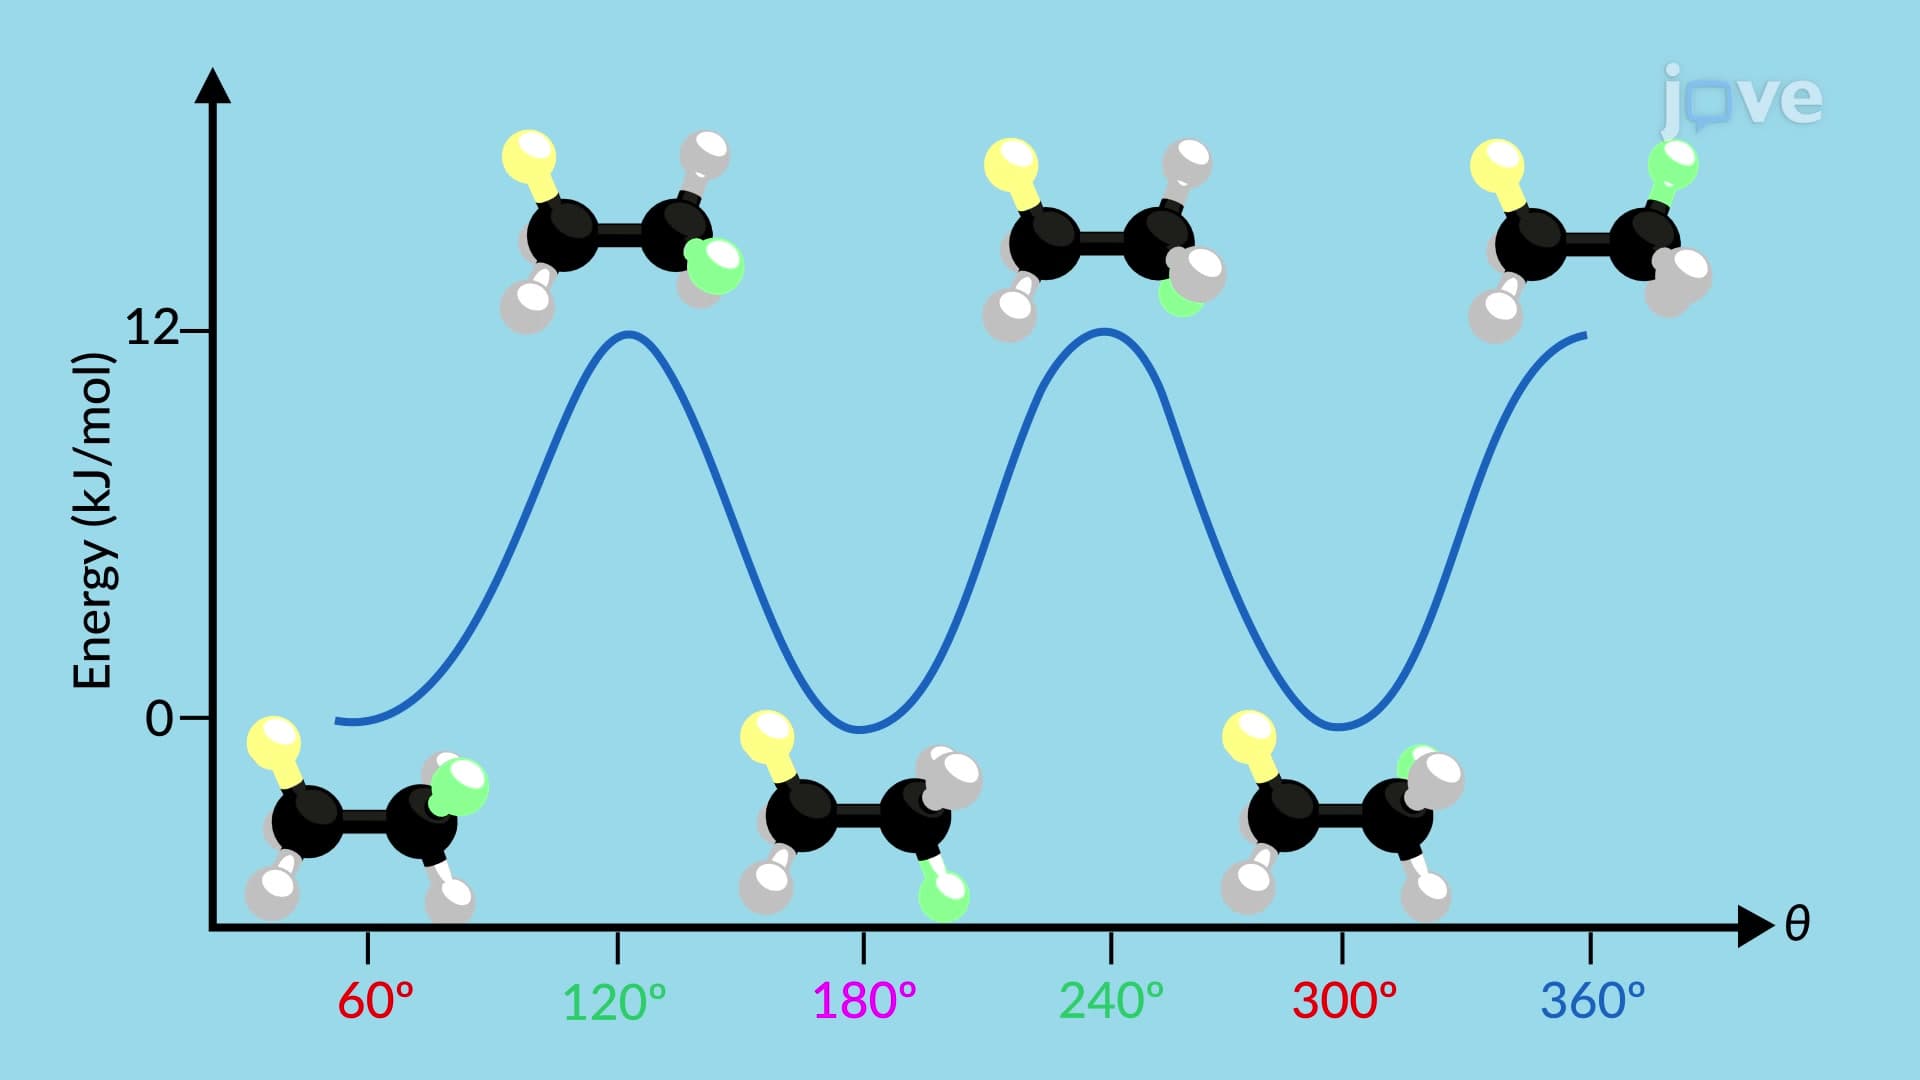
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...