PSI-DOCK: towards highly efficient and accurate flexible ligand docking
Related Experiment Video
Updated: Aug 13, 2026
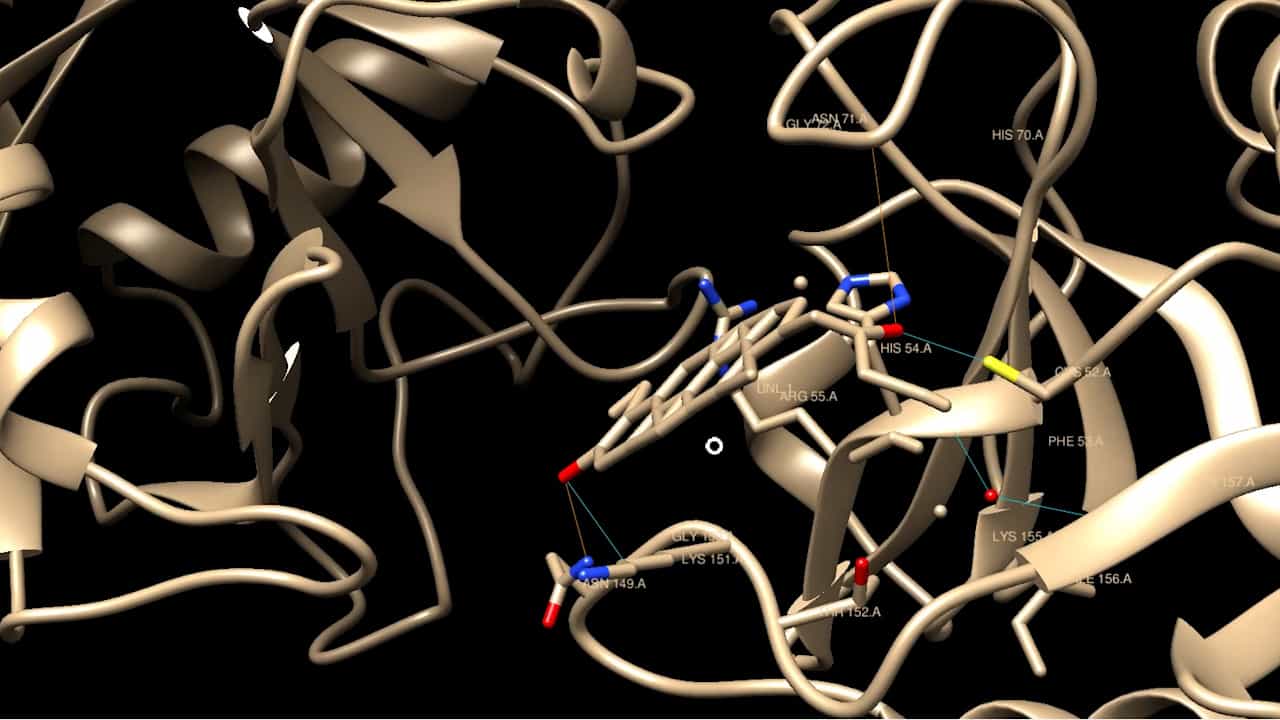
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Jianfeng Pei1, Qi Wang, Zhenming Liu
1State Key Laboratory for Structural Chemistry of Stable and Unstable Species, College of Chemistry and Molecular Engineering, Peking University, Beijing, People's Republic of China.

Abstract:
We have developed a new docking method, Pose-Sensitive Inclined (PSI)-DOCK, for flexible ligand docking. An improved SCORE function has been developed and used in PSI-DOCK for binding free energy evaluation. The improved SCORE function was able to reproduce the absolute binding free energies of a training set of 200 protein-ligand complexes with a correlation coefficient of 0.788 and a standard error of 8.13 kJ/mol. For ligand binding pose exploration, a unique searching strategy was designed in PSI-DOCK. In the first step, a tabu-enhanced genetic algorithm with a rapid shape-complementary scoring function is used to roughly explore and store potential binding poses of the ligand. Then, these predicted binding poses are optimized and compete against each other by using a genetic algorithm with the accurate SCORE function to determine the binding pose with the lowest docking energy. The PSI-DOCK 1.0 program is highly efficient in identifying the experimental binding pose. For a test dataset of 194 complexes, PSI-DOCK 1.0 achieved a 67% success rate (RMSD < 2.0 A) for only one run and a 74% success rate for 10 runs. PSI-DOCK can also predict the docking binding free energy with high accuracy. For a test set of 64 complexes, the correlation between the experimentally observed binding free energies and the docking binding free energies for 64 complexes is r = 0.777 with a standard deviation of 7.96 kJ/mol. Moreover, compared with other docking methods, PSI-DOCK 1.0 is extremely easy to use and requires minimum docking preparations. There is no requirement for the users to add hydrogen atoms to proteins because all protein hydrogen atoms and the flexibility of the terminal protein atoms are intrinsically taken into account in PSI-DOCK. There is also no requirement for the users to calculate partial atomic charges because PSI-DOCK does not calculate an electrostatic energy term. These features are not only convenient for the users but also help to avoid the influence of different preparation methods.
More Related Videos
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:18
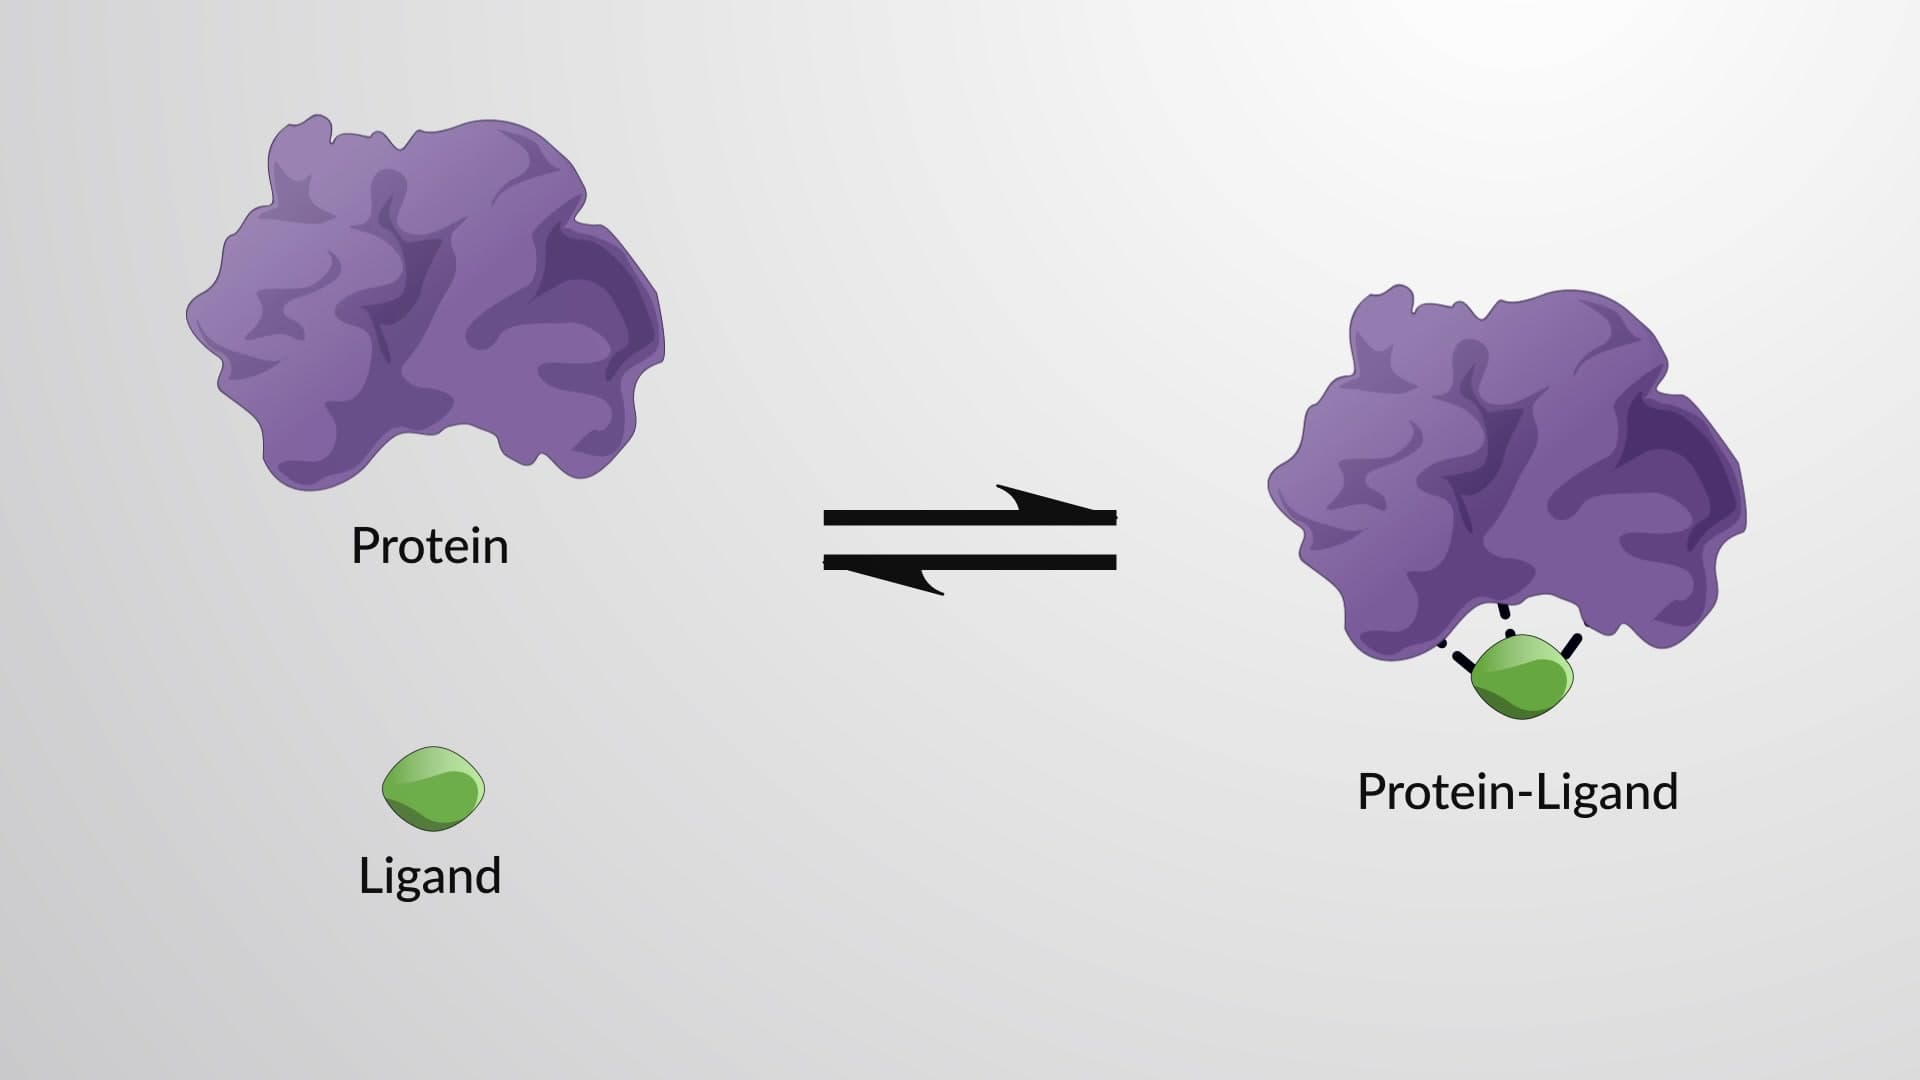
The Equilibrium Binding Constant and Binding Strength
 00:49
Ligand Binding and Linkage