Related Experiment Video
Updated: Aug 13, 2026


Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Extending, and repositioning, a thermochemical ladder: high-level quantum chemical calculations on the sodium cation
Jolyon Bloomfield1, Erin Davies, Phillip Gatt
1Department of Chemistry, the Faculties, the Australian National University, Canberra ACT 0200, Australia.

Abstract:
High-level ab initio quantum chemical calculations, at the CP-dG2thaw level of theory, are reported for coordination of Na+ to a wide assortment of small organic and inorganic ligands. The ligands range in size from H to C6H6, and include 22 of the ligands for which precise relative sodium ion binding free energies have been determined by recent Fourier transform ion cyclotron resonance and guided ion beam studies. Agreement with the relative experimental values is excellent (+/-1.1 kJ mol(-1)), and agreement with the absolute scale (obtained when these relative values are pegged to the CH3NH2 "anchor" value measured in a high-pressure mass spectrometric study) is only marginally poorer, with CP-dG2thaw values exceeding the absolute experimental DeltaG(298) values by an average of 2.1 kJ mol(-1). The excellent agreement between experiment and the CP-dG2thaw technique also suggests that the additional 97 ligands surveyed here (which, in many cases, are not readily susceptible to laboratory investigation) can also be reliably fitted to the existing experimental scale. However, while CP-dG2thaw and the experimental ladder are in close accord, a small set of higher level ab initio calculations on sodium ion/ligand complexes (including several values obtained here using the W1 protocol) suggests that the CP-dG2thaw values are themselves too low by approximately 2.5 kJ mol(-1), thereby implying that the accepted laboratory values are typically 4.6 kJ mol(-1) too low. The present work also highlights the importance of Na+/ligand binding energy determinations (whether by experimental or theoretical approaches) on a case-by-case basis: trends in increasing binding energy along homologous series of compounds are not reliably predictable, nor are binding site preferences or chelating tendencies in polyfunctional compounds.

Related Concept Videos
 01:07
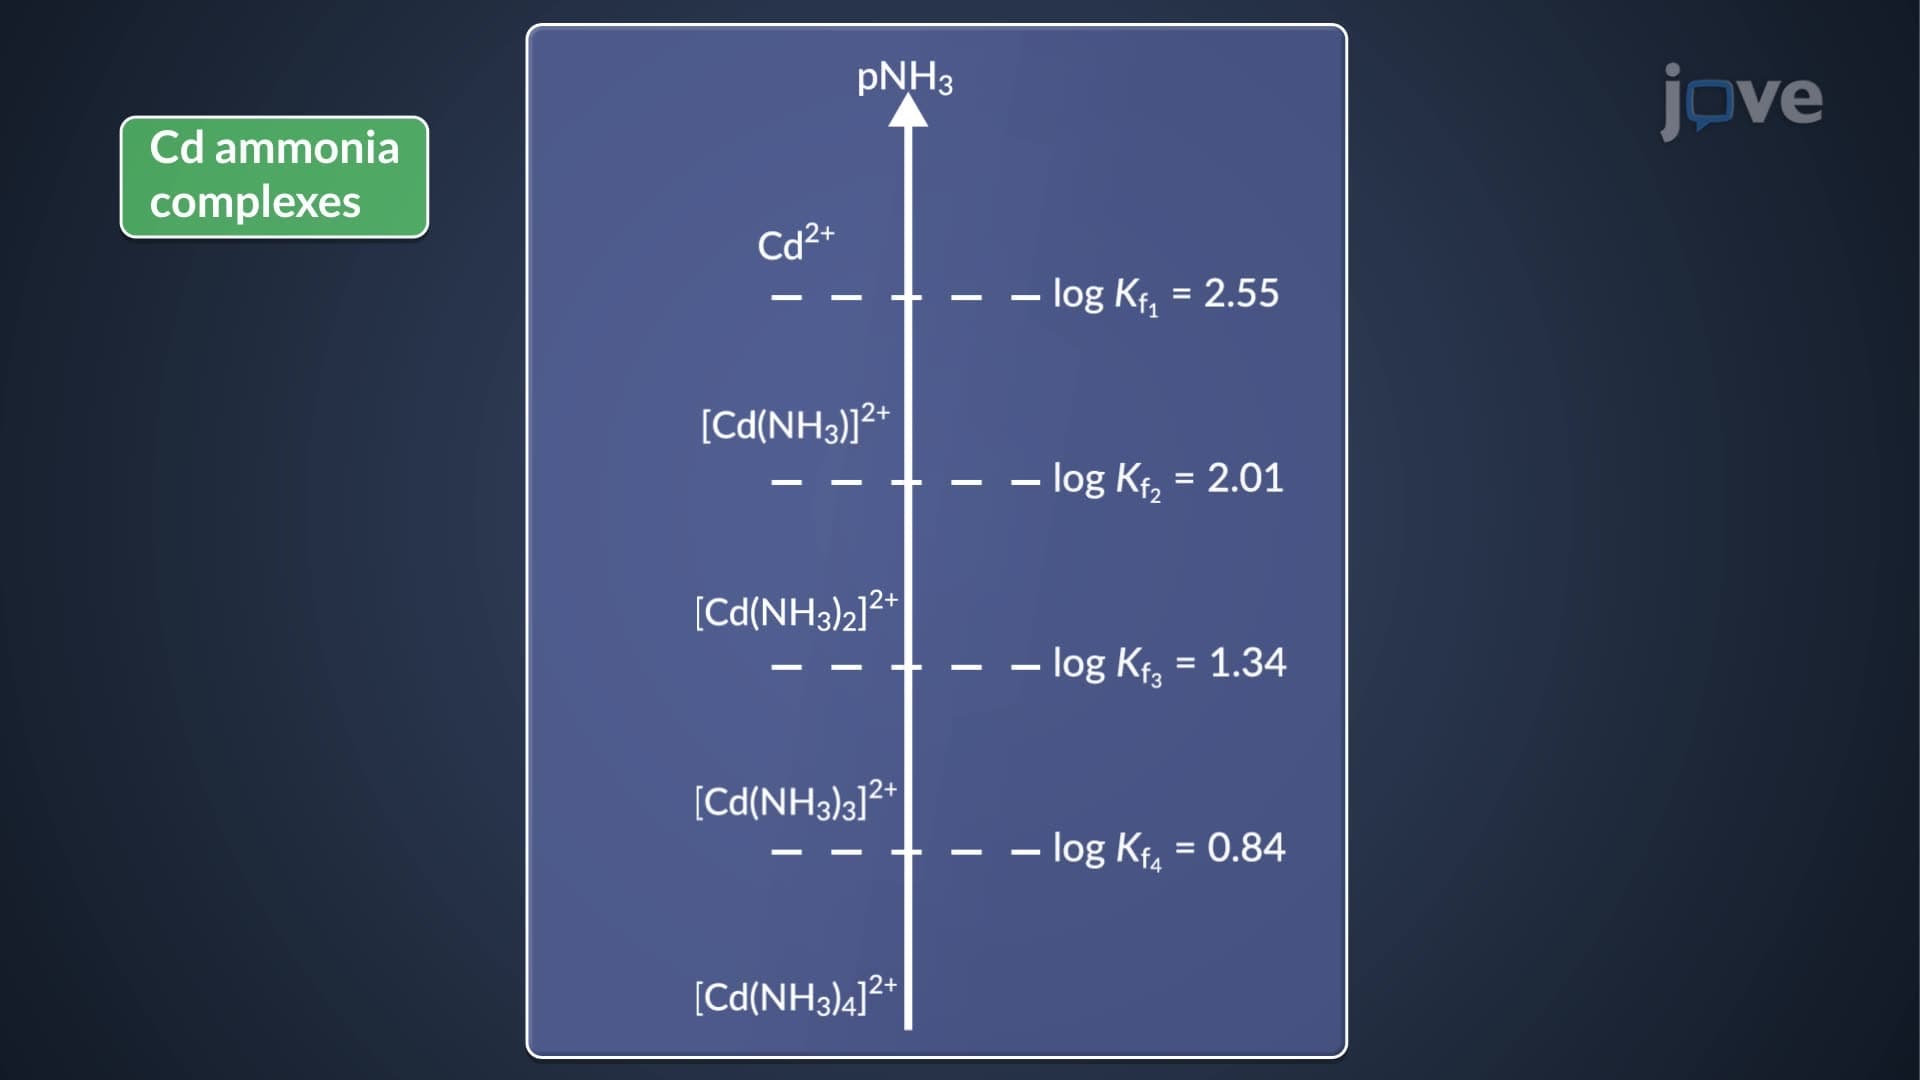
Ladder Diagrams: Complexation Equilibria
The formation constant, K1, for the formation of Cd(NH3)2+ complex from cadmium and ammonia is 3.55 × 10². Log K1 (i.e. pNH3) is 2.55, and...
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:44
The Born-Haber Cycle
 02:48
Ionic Bonding and Electron Transfer
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...