Related Experiment Video
Updated: Aug 13, 2026
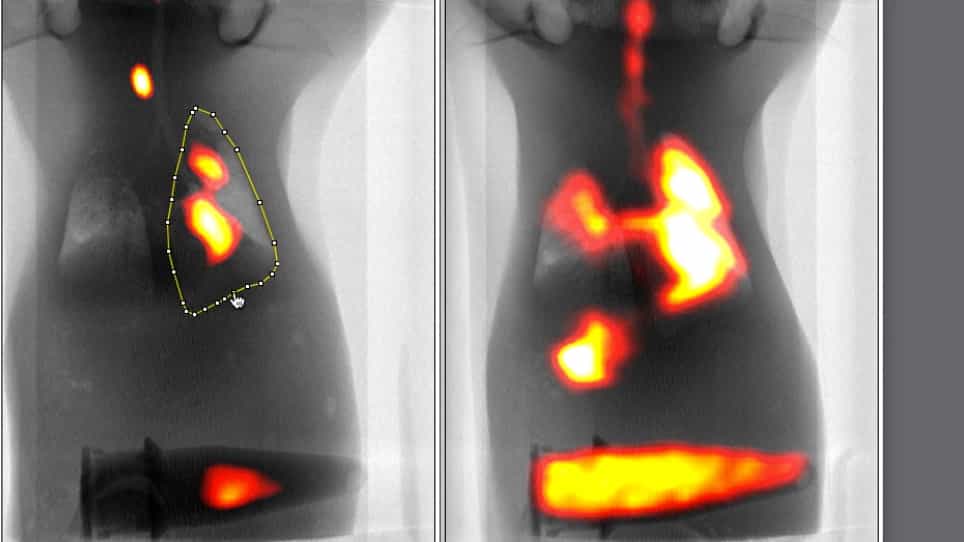
In vivo Evaluation of Mucociliary Clearance in Mice
Published on: December 18, 2020
Mucus properties in children with primary ciliary dyskinesia: comparison with cystic fibrosis
Andrew Bush1, Donald Payne, Sarah Pike
1Pediatric Pulmonology, Imperial School of Medicine at National Heart and Lung Institute, London, UK.

Objective:
It has been assumed that cystic fibrosis (CF) lung disease is due in part to abnormal airway mucus. Primary ciliary dyskinesia (PCD) is a form of bronchiectasis that is similar to CF in many ways but is caused by congenital defects in mucociliary clearance. Our objective was to compare the biophysical and transport properties of CF and PCD sputa in subjects matched for age and degree of lung function impairment.
Design, Setting, Participants:
PCD patients (n = 19; mean age, 9.5 +/- 3.0 years [+/- SD]; FEV1, 65.0 +/- 7.8 L) were recruited from the clinic at the Royal Brompton Hospital. Patients with CF (n = 30, mean age, 10.8 +/- 2.6 years; FEV1, 61.8 +/- 22.8 L) were identified from the Wake Forest University School of Medicine CF Center. Pulmonary function testing and sputum collection were performed as part of routine, scheduled clinic visits.
Measurements:
Pulmonary function was measured by spirometry, and sputum was collected during the pulmonary function test maneuver. Some patients were longitudinally assessed at visits during the course of 3 years. Sputum properties measured were dynamic viscoelasticity, wettability, cohesivity, interfacial (surface) tension, solids composition, DNA and interleukin (IL)-8 concentration, in vitro mucociliary transportability, and cough transportability.
Results:
Inflammation as measured by IL-8 concentration was three times greater in the PCD sputa (p < 0.0001). There were no significant differences in the sputum biophysical or transport properties comparing CF with PCD sputum.
Conclusions:
It is unlikely that established CF lung disease is principally due to abnormal sputum properties, and it is more likely that the biophysical and transport properties reflect disease severity regardless of whether bronchiectasis is due to CF or PCD.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:16
Drugs Used in Upper Respiratory Disorders: Overview
Antihistamines (e.g., Benadryl) block histamines from binding. Histamines are chemicals released during an allergic reaction in the body. As a...
 01:24
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features
 01:23
Upper Respiratory Drugs: Antitussives, Expectorants, and Mucolytics
Antitussives include codeine, dextromethorphan (Robitussin), and benzonatate (Tessalon). Codeine and dextromethorphan exert their effects centrally by suppressing the cough reflex center in the medulla. Benzonatate operates peripherally within the respiratory tract by anesthetizing...
 01:23
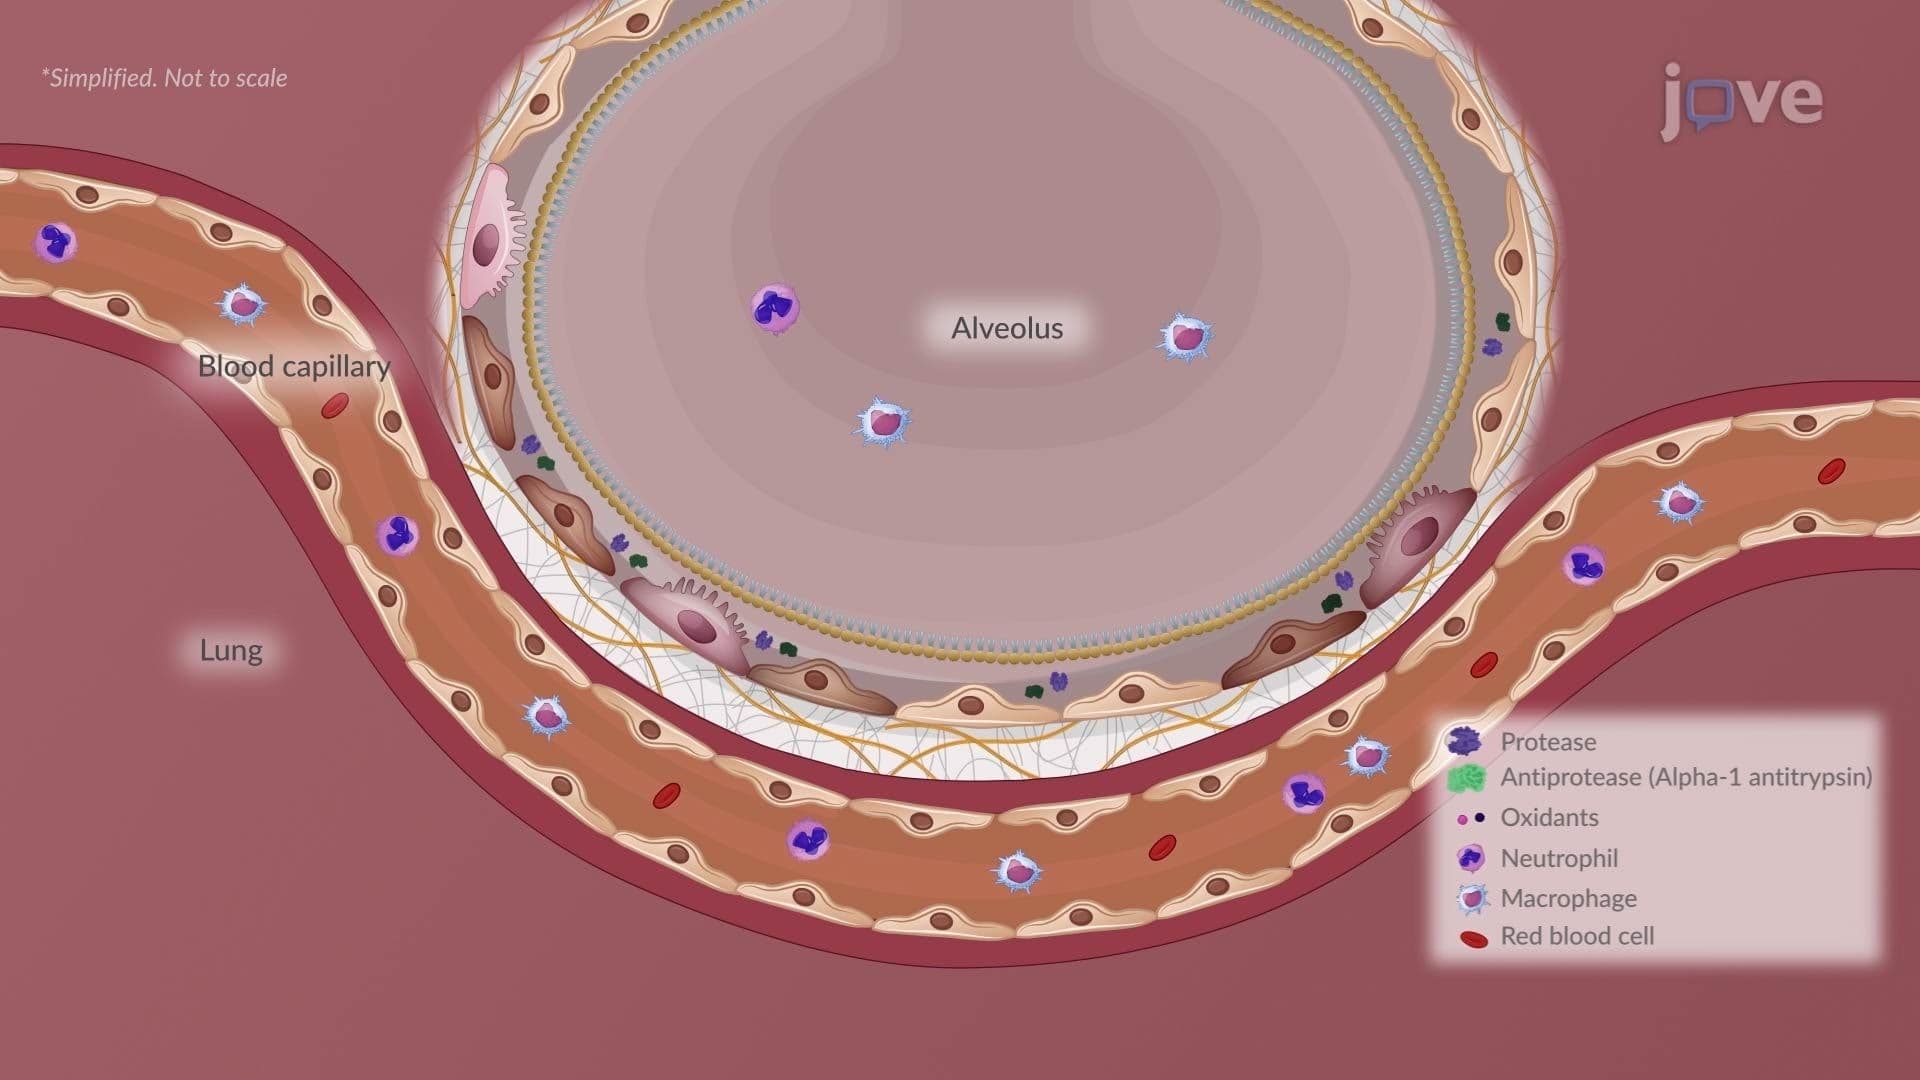
Chronic Obstructive Pulmonary Disease II: Emphysema