Related Experiment Video
Updated: Aug 13, 2026

The Preparation and Properties of Thermo-reversibly Cross-linked Rubber Via Diels-Alder Chemistry
Published on: August 25, 2016
Molecular structure and benzene ring deformation of three ethynylbenzenes from gas-phase electron diffraction and
Anna Rita Campanelli1, Antonio Arcadi, Aldo Domenicano
1Department of Chemistry, University of Rome La Sapienza, I-00185 Rome, Italy.

Abstract:
The molecular structures of ethynylbenzene and s-triethynylbenzene have been accurately determined by gas-phase electron diffraction and ab initio/DFT MO calculations and are compared to that of p-diethynylbenzene from a previous study [Domenicano, A.; Arcadi, A.; Ramondo, F.; Campanelli, A. R.; Portalone, G.; Schultz, G.; Hargittai, I. J. Phys. Chem. 1996, 100, 14625]. Although the equilibrium structures of the three molecules have C2v, D3h, and D2h symmetry, respectively, the corresponding average structures in the gaseous phase are best described by nonplanar models of Cs, C3v, and C2v symmetry, respectively. The lowering of symmetry is due to the large-amplitude motions of the substituents out of the plane of the benzene ring. The use of nonplanar models in the electron diffraction analysis yields ring angles consistent with those from MO calculations. The molecular structure of ethynylbenzene reported from microwave spectroscopy studies is shown to be inaccurate in the ipso region of the benzene ring. The variations of the ring C-C bonds and C-C-C angles in p-diethynylbenzene and s-triethynylbenzene are well interpreted as arising from the superposition of independent effects from each substituent. In particular, experiments and calculations consistently show that the mean length of the ring C-C bonds increases by about 0.002 A per ethynyl group. MO calculations at different levels of theory indicate that though the length of the C[triple bond]C bond of the ethynyl group is unaffected by the pattern of substitution, the C(ipso)-C(ethynyl) bonds in p-diethynylbenzene are 0.001-0.002 A shorter than the corresponding bonds in ethynylbenzene and s-triethynylbenzene. This small effect is attributed to conjugation of the two substituents through the benzene ring. Comparison of experimental and MO results shows that the differences between the lengths of the C(ipso)-C(ethynyl) and C(ipso)-C(ortho) bonds in the three molecules, 0.023-0.027 A, are correctly computed at the MP2 and B3LYP levels of theory but are overestimated by a factor of 2 when calculated at the HF level.
More Related Videos
Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 02:18
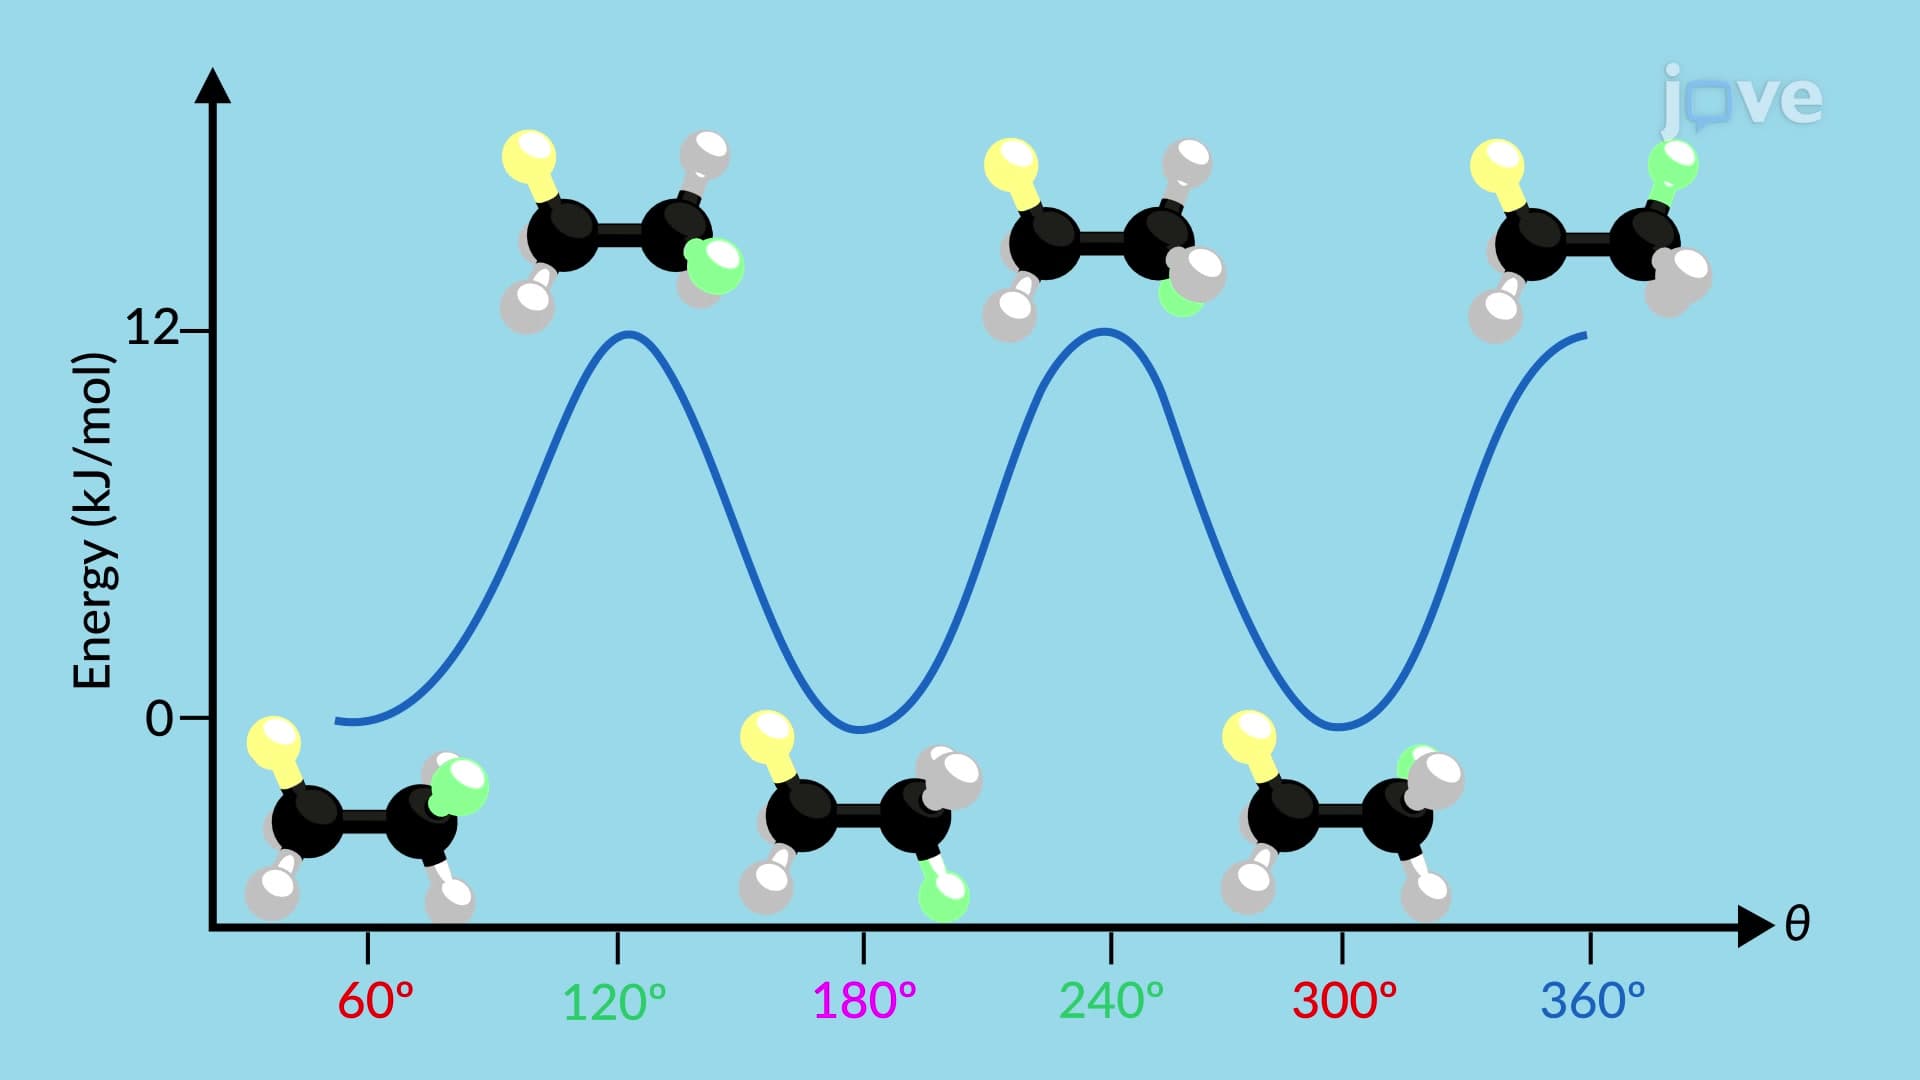
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...
 01:07
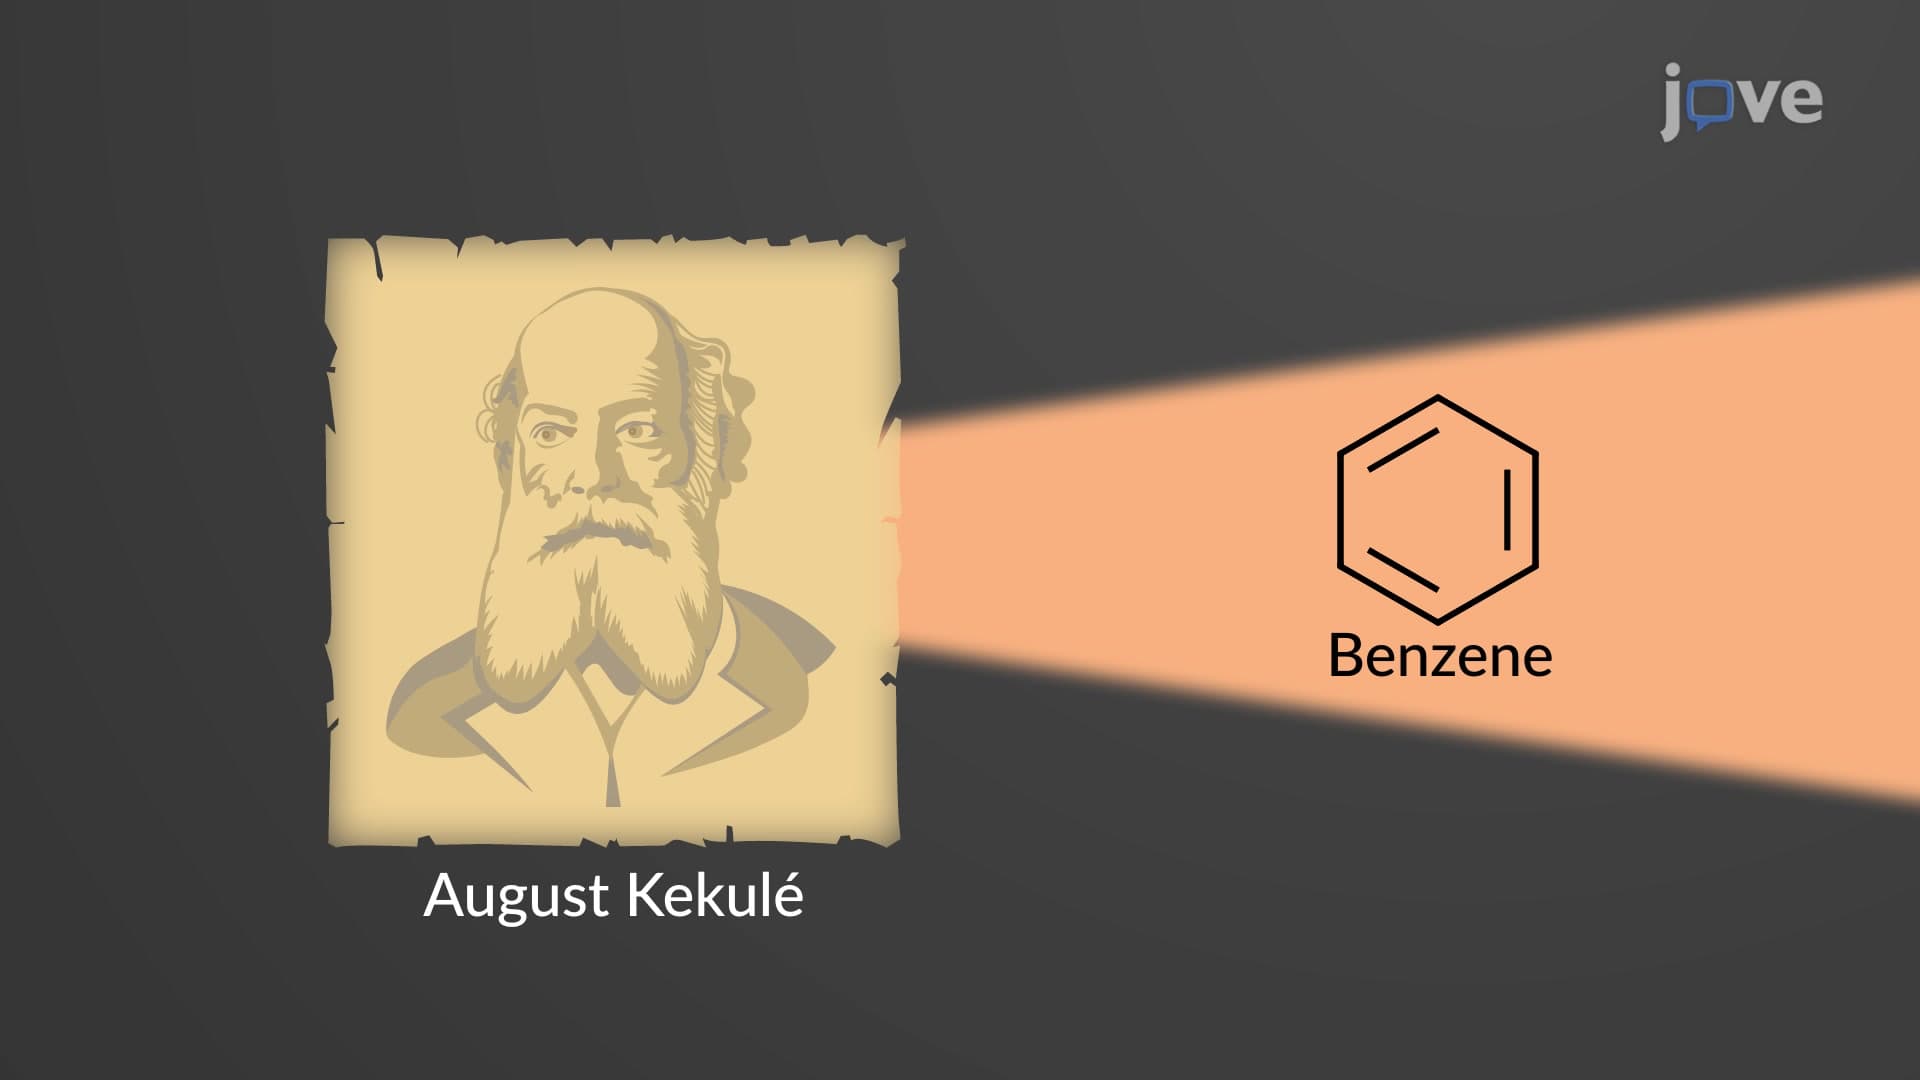
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 02:52
VSEPR Theory and the Basic Shapes
 02:29
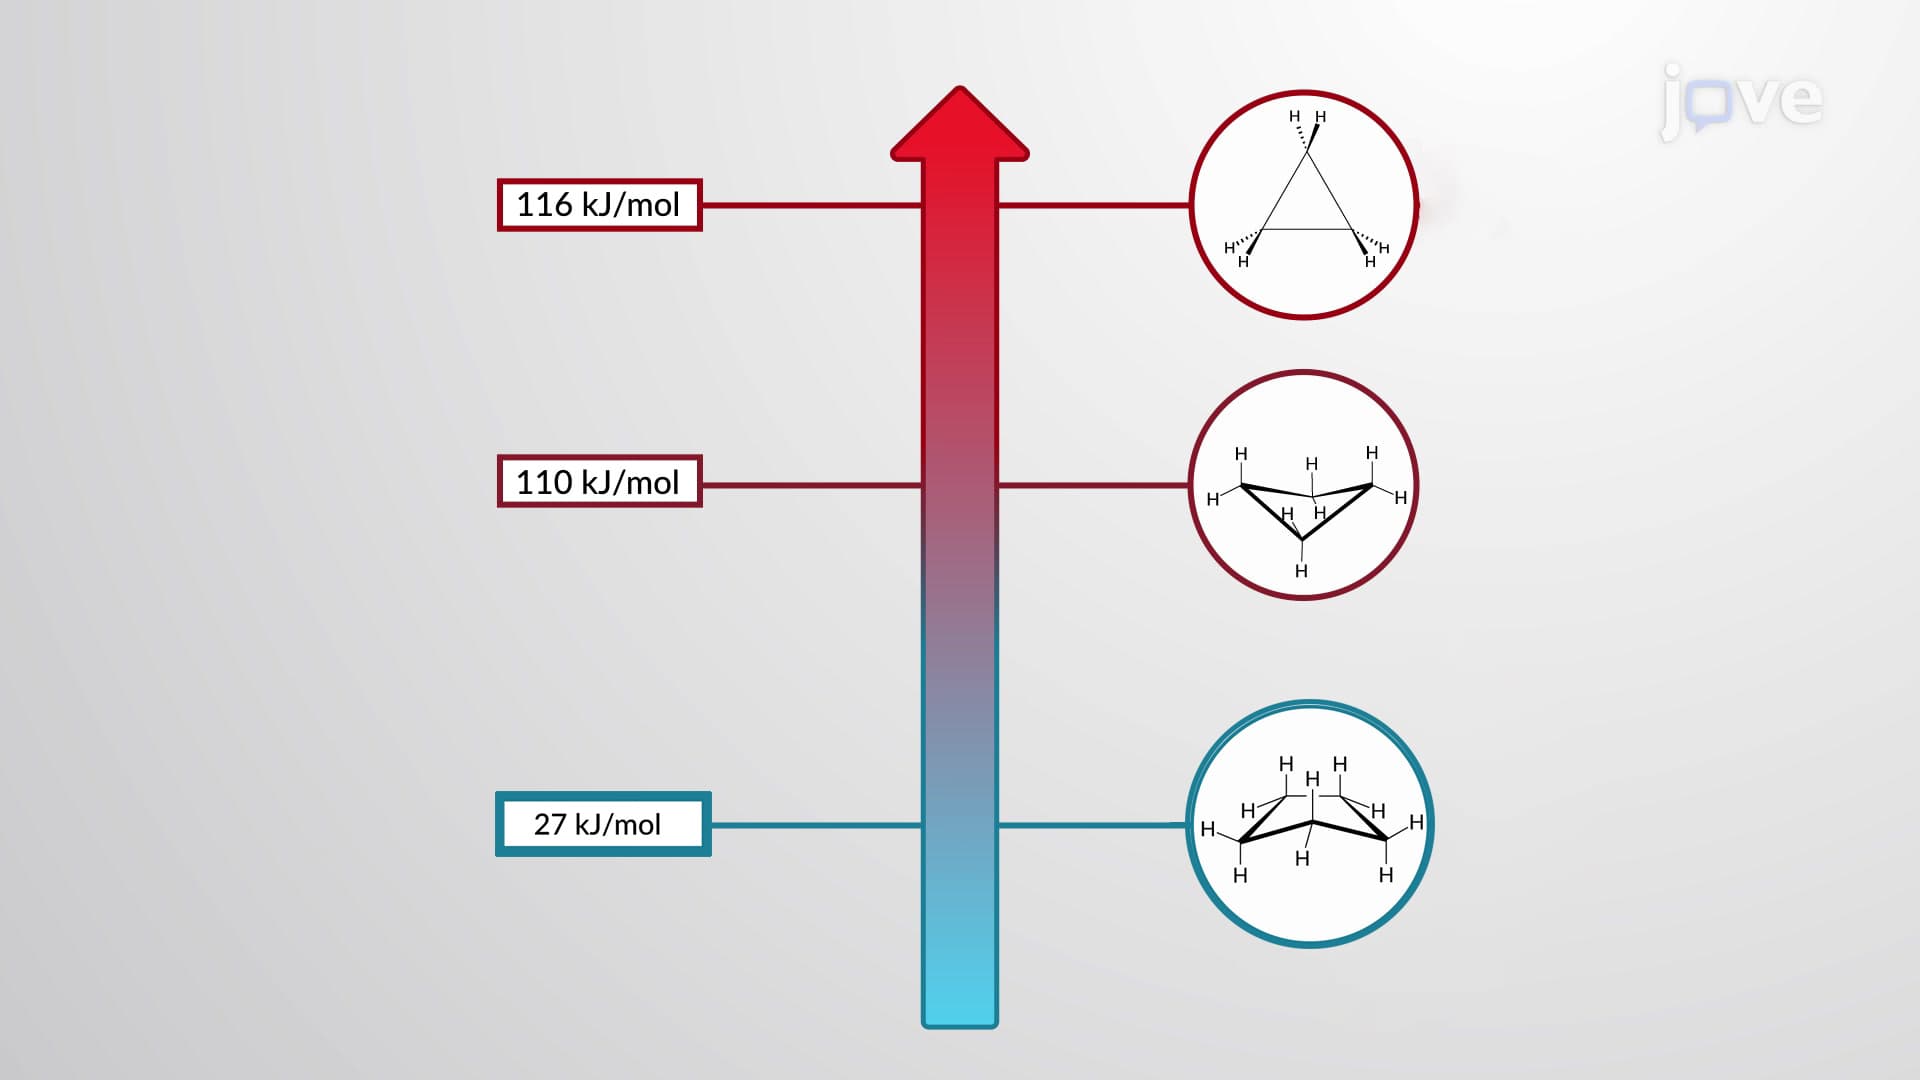
Conformations of Cycloalkanes
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...