Related Experiment Video
Updated: Aug 11, 2026
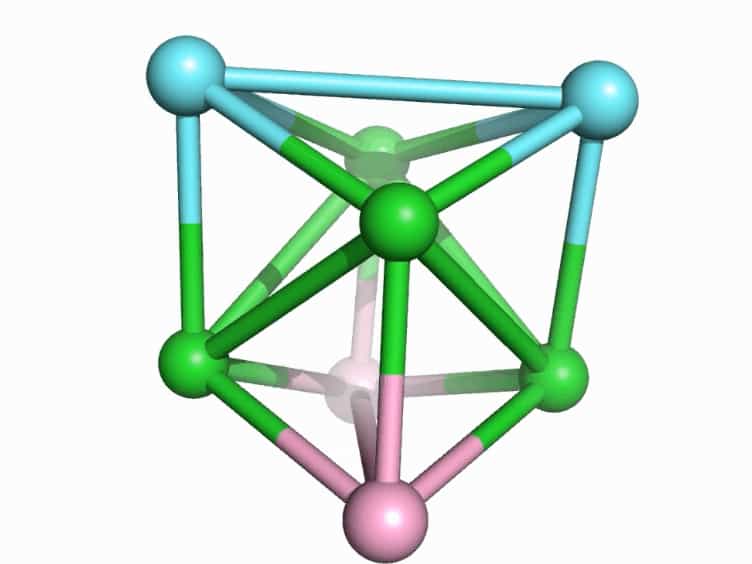
Construction and Systematical Symmetric Studies of a Series of Supramolecular Clusters with Binary or Ternary Ammonium Triphenylacetates
Published on: February 15, 2016
Pseudo-Jahn-Teller origin of geometry and pseudorotations in second row tetra-atomic clusters X4 (X=Na, Mg, Al, Si,
Pablo Garcia-Fernandez1, Isaac B Bersuker, James E Boggs
1Institute for Theoretical Chemistry, The University of Texas at Austin, Chemistry and Biochemistry Department, Austin, Texas 78712-0165, USA. garciapa@mail.utexas.edu

Abstract:
Experimentally determined or ab initio calculated molecular geometries carry no information about their origin. Employing the Jahn-Teller (JT) vibronic coupling effects as the only source of instability and consequent distortions of high-symmetry molecular configurations, we have worked out a procedure that allows us to trace the origin of particular geometries and determine the detailed electronic mechanism of their formation. This procedure is illustrated by considering a series of X(4) clusters with X=Na, Mg, Al, Si, P, and S. It shows explicitly why Na(4), Si(4), and Al(4) have a rhombic geometry in the ground state, while Mg(4) and P(4) are tetrahedral, whereas S(4) is a trapezium. Even when the minimum-energy geometries are the same (as in the case of rhombic Na(4), Si(4), and Al(4)), the electronic mechanism of their formation is quite different. In particular, in Na(4) and Si(4) the rhombic minima are produced by a strong pseudo JT coupling between two excited states in the square-planar configuration (different in the two cases) that stabilizes one of them and makes it the ground state by rhombic distortions. The rhombic configuration of Al(4) is due to the pseudo JT effect in its ground-state square-planar configuration, and the trapezium in S(4) is formed by two pseudo JT couplings essentially involving excited states. In several cases this analysis shows also the tunneling paths between equivalent configurations.
Related Concept Videos
 02:27
Predicting Molecular Geometry
 04:01
VSEPR Theory and the Effect of Lone Pairs
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:57
Coordination Number and Geometry
 02:52
VSEPR Theory and the Basic Shapes
 03:24
Hybridization of Atomic Orbitals I