Related Experiment Video
Updated: Aug 11, 2026

Formation of Ordered Biomolecular Structures by the Self-assembly of Short Peptides
Published on: November 21, 2013
Solvation model based on order parameters and a fast sampling method for the calculation of the solvation free
Chong Gu1, Steve Lustig, Bernhardt L Trout
1Department of Chemical Engineering, Massachusetts Institute of Technology, 77 Massachusetts Avenue E19-502B, Cambridge Massachusetts 02139, USA.

Abstract:
An analytical solvation model is proposed as a function of an order parameter, which represents the local arrangement of water molecules in the first solvation shell of peptide atoms. The model is combined with a fast sampling method, rotational isomeric state Monte Carlo, to sample efficiently the torsional degrees of freedom on a peptide backbone. This order parameter solvation model is shown to reproduce without ad hoc fitting parameters the solvation free energies of single amino acids and tripeptides with slightly better accuracy than the generalized Born model but with several orders of magnitude improvement in efficiency. This method is a potential candidate for efficiently and accurately tackling some important issues in biophysical chemistry that are related to solvation, for example, protein folding, ligand binding, etc. Our results also present fundamental new insights into solvation. Specifically, the local water geometry, represented in this work by a properly defined order parameter, carries the majority, if not all, of the energetic information of solvation, including solute-solvent interactions and solvent reorganization in the presence of the solute.
Related Concept Videos
 02:21
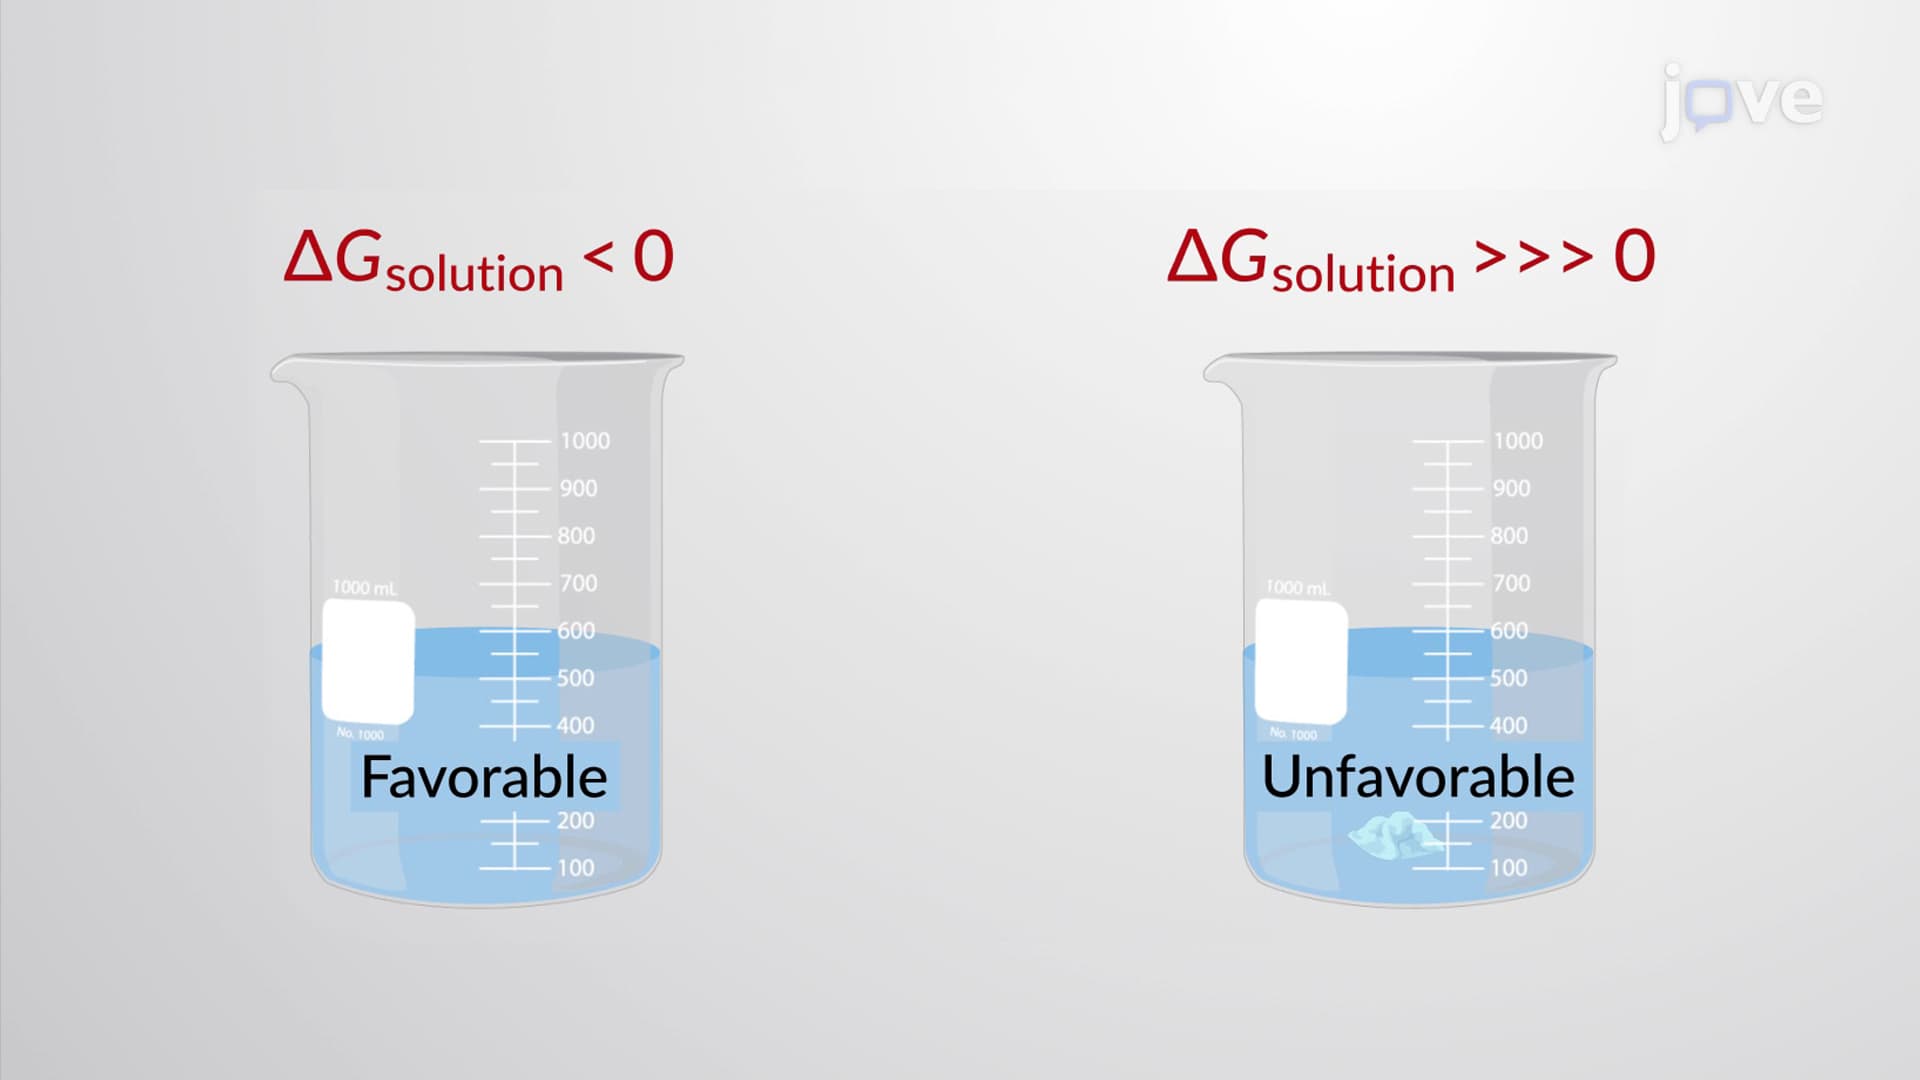
Chemical and Solubility Equilibria
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding