Related Experiment Video
Updated: Aug 11, 2026

Fabrication of Amyloid-β-Secreting Alginate Microbeads for Use in Modelling Alzheimer's Disease
Published on: July 6, 2019
A partial failure of membrane protein turnover may cause Alzheimer's disease: a new hypothesis
Kumar Sambamurti1, Anitha Suram, Chitra Venugopal
1Department of Neurosciences, Medical University of South Carolina, 173 Ashley Avenue, BSB 403, Charleston, SC 29425, USA. sambak@musc.edu

Abstract:
The amyloid hypothesis has dominated the thinking in our attempts to understand, diagnose and develop drugs for Alzheimer's disease (AD). This article presents a new hypothesis that takes into account the numerous familial AD (FAD) mutations in the amyloid precursor protein (APP) and its processing pathways, but suggests a new perspective beyond toxicity of forms of the amyloid beta-peptide (Abeta). Clearly, amyloid deposits are an invariable feature of AD. Moreover, although APP is normally processed to secreted and membrane-bound fragments, sAPPbeta and CTFbeta, by BACE, and the latter is subsequently processed by gamma-secretase to Abeta and CTFgamma, this pathway mostly yields Abeta of 40 residues, and increases in the levels of the amyloidogenic 42-residue Abeta (Abeta42) are seen in the majority of the mutations linked to the disease. The resulting theory is that the disease is caused by amyloid toxicity, which impairs memory and triggers deposition of the microtubule associated protein, Tau, as neurofibrillary tangles. Nevertheless, a few exceptional FAD mutations and the presence of large amounts of amyloid deposits in a group of cognitively normal elderly patients suggest that the disease process is more complex. Indeed, it has been hard to demonstrate the toxicity of Abeta42 and the actual target has been shifted to small oligomers of the peptide, named Abeta derived diffusible ligands (ADDLs). Our hypothesis is that the disease is more complex and caused by a failure of APP metabolism or clearance, which simultaneously affects several other membrane proteins. Thus, a traffic jam is created by failure of important pathways such as gamma-secretase processing of residual intramembrane domains released from the metabolism of multiple membrane proteins, which ultimately leads to a multiple system failure. In this theory, toxicity of Abeta42 will only contribute partially, if at all, to neurodegeneration in AD. More significantly, this theory would predict that focussing on specific reagents such as gamma-secretase inhibitors that hamper metabolism of APP, may initially show some beneficial effects on cognitive performance by elimination of acutely toxic ADDLs, but over the longer term may exacerbate the disease process by reducing membrane protein turnover.
Insights
A new Alzheimer's disease (AD) hypothesis suggests APP metabolism failure, not just amyloid beta-peptide (Abeta) toxicity, causes disease. This challenges current drug targets and predicts long-term risks of some AD therapies.
Area of Science:
- Neuroscience
- Biochemistry
- Pathology
Background:
- The amyloid hypothesis, focusing on amyloid beta-peptide (Abeta) toxicity, has long dominated Alzheimer's disease (AD) research.
- Familial AD (FAD) mutations in amyloid precursor protein (APP) processing pathways increase amyloidogenic Abeta42, supporting the amyloid hypothesis.
- However, exceptions and cognitively normal individuals with amyloid deposits suggest AD pathogenesis is more complex than solely Abeta toxicity.
Purpose of the Study:
- To propose an alternative hypothesis for Alzheimer's disease (AD) pathogenesis.
- To re-evaluate the role of amyloid beta-peptide (Abeta) toxicity in AD.
- To explore the implications of a novel hypothesis on AD therapeutic strategies.
Main Methods:
- Conceptual review and hypothesis formulation based on existing literature and FAD mutation data.
- Analysis of APP processing pathways and the impact of mutations.
- Consideration of alternative mechanisms beyond direct Abeta toxicity, including membrane protein metabolism.
Main Results:
- A new hypothesis posits that impaired APP metabolism and clearance cause a 'traffic jam' affecting multiple membrane proteins, leading to system failure.
- This model suggests Abeta42 toxicity contributes partially, if at all, to neurodegeneration.
- The hypothesis predicts that gamma-secretase inhibitors, while initially clearing toxic Abeta oligomers, may worsen AD long-term by hindering membrane protein turnover.
Conclusions:
- Alzheimer's disease may result from a broader failure in APP metabolism affecting multiple membrane proteins, rather than solely Abeta toxicity.
- This revised perspective challenges the central role of Abeta and suggests current therapeutic targets may be insufficient or even detrimental.
- Further research is needed to validate the proposed 'traffic jam' hypothesis and its implications for developing effective AD treatments.
More Related Videos
 15:04
Interactions with and Membrane Permeabilization of Brain Mitochondria by Amyloid Fibrils
Published on: September 28, 2019
 06:41
Quantitative Analysis of Mitochondria-Associated Endoplasmic Reticulum Membrane (MAM) Stabilization in a Neural Model of Alzheimer's Disease (AD)
Published on: January 10, 2025
Related Concept Videos
 01:23
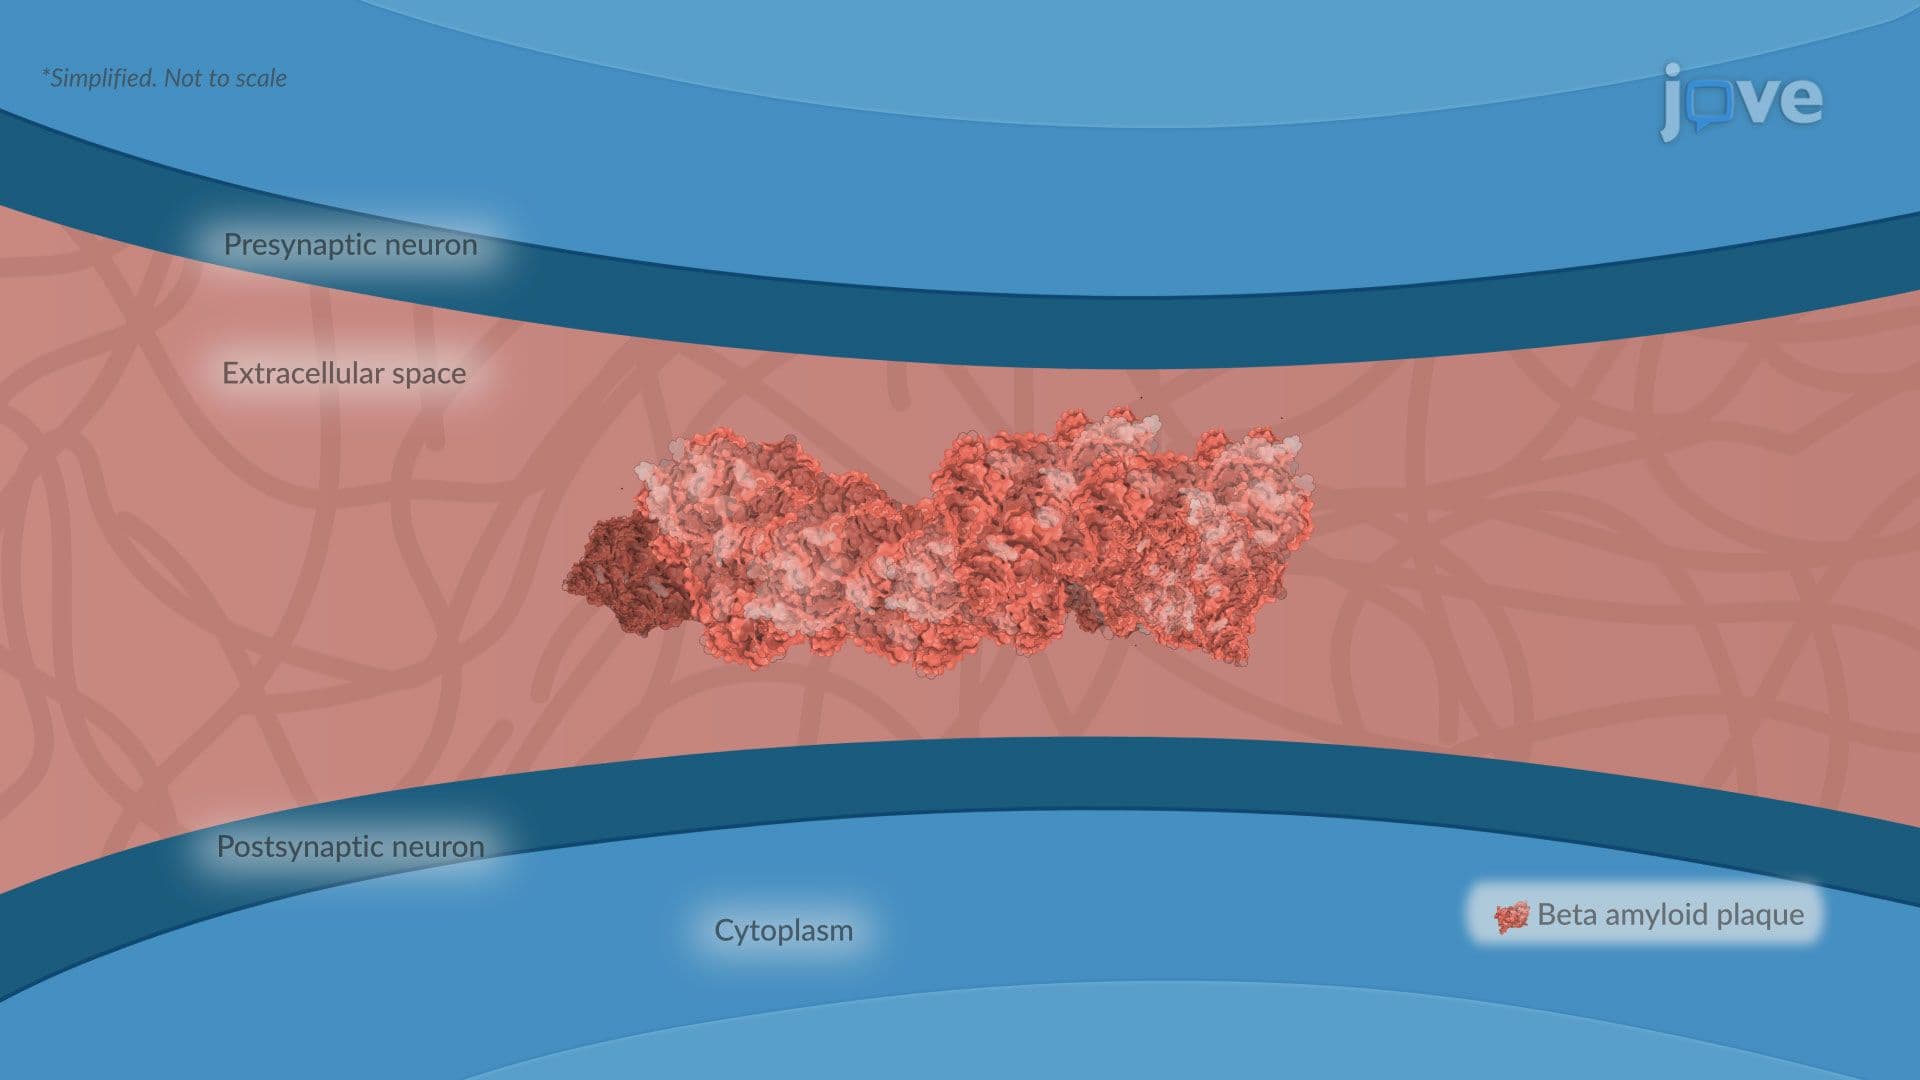
Alzheimer Disease ll: Pathophysiology
 01:29
Alzheimer Disease l: Introduction
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
03:03Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:26
Alzheimer's Disease: Overview
The clinical diagnosis of AD hinges on the presence of memory and other cognitive impairments. Biomarkers, such as changes in Aβ and tau...
 01:22
Dementia l: Introduction