Related Experiment Video
Updated: Aug 11, 2026
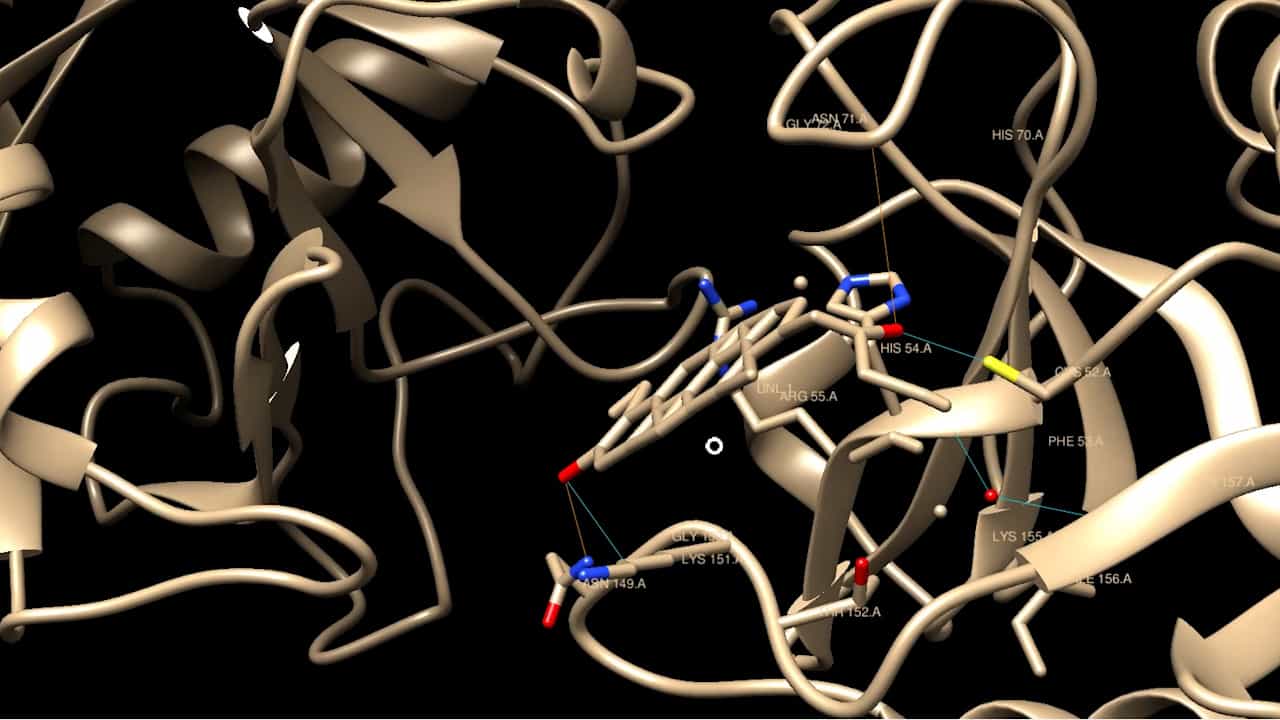

Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Probing molecular docking in a charged model binding site
Ruth Brenk1, Stefan W Vetter, Sarah E Boyce
1University of California San Francisco, QB3 Building, Department of Pharmaceutical Chemistry, 1700 4th Street, San Francisco, CA 94143-2550, USA.

Abstract:
A model binding site was used to investigate charge-charge interactions in molecular docking. This simple site, a small (180A(3)) engineered cavity in cyctochrome c peroxidase (CCP), is negatively charged and completely buried from solvent, allowing us to explore the balance between electrostatic energy and ligand desolvation energy in a system where many of the common approximations in docking do not apply. A database with about 5300 molecules was docked into this cavity. Retrospective testing with known ligands and decoys showed that overall the balance between electrostatic interaction and desolvation energy was captured. More interesting were prospective docking scre"ens that looked for novel ligands, especially those that might reveal problems with the docking and energy methods. Based on screens of the 5300 compound database, both high-scoring and low-scoring molecules were acquired and tested for binding. Out of 16 new, high-scoring compounds tested, 15 were observed to bind. All of these were small heterocyclic cations. Binding constants were measured for a few of these, they ranged between 20microM and 60microM. Crystal structures were determined for ten of these ligands in complex with the protein. The observed ligand geometry corresponded closely to that predicted by docking. Several low-scoring alkyl amino cations were also tested and found to bind. The low docking score of these molecules owed to the relatively high charge density of the charged amino group and the corresponding high desolvation penalty. When the complex structures of those ligands were determined, a bound water molecule was observed interacting with the amino group and a backbone carbonyl group of the cavity. This water molecule mitigates the desolvation penalty and improves the interaction energy relative to that of the "naked" site used in the docking screen. Finally, six low-scoring neutral molecules were also tested, with a view to looking for false negative predictions. Whereas most of these did not bind, two did (phenol and 3-fluorocatechol). Crystal structures for these two ligands in complex with the cavity site suggest reasons for their binding. That these neutral molecules do, in fact bind, contradicts previous results in this site and, along with the alkyl amines, provides instructive false negatives that help identify weaknesses in our scoring functions. Several improvements of these are considered.

Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:18
The Equilibrium Binding Constant and Binding Strength
 00:49
Ligand Binding and Linkage